Related Experiment Video
Updated: Apr 18, 2026

Characterizing Lewis Pairs Using Titration Coupled with In Situ Infrared Spectroscopy
Published on: February 20, 2020
A single theoretical descriptor for the bond-dissociation energy of substituted phenols
Carolina Aliaga1, Iriux Almodovar, Marcos Caroli Rezende
1Facultad de Química y Biología, Universidad de Santiago de Chile, Casilla 40 Correo 33, Santiago, Chile.

Abstract:
Relative to the corresponding value of phenol, the bond-dissociation energies (BDE) of substituted phenols correlate well with a single descriptor: the Mulliken charge on the oxygen atom of the phenoxyl radical. However, the correlation fails for phenols ortho-substituted with polar groups. Internal reaction coordinates (IRC) for the model reaction of hydrogen abstraction by the hydroperoxyl radical from various 2- and 4-substituted phenols were calculated in order to investigate the role of intra-molecular hydrogen bonds and steric effects on the process. Calculations yielded theoretical values in good agreement with experimental ΔBDE values. The hydrogen-abstraction process was further analyzed in terms of density functional theory (DFT)-based reactivity indices such as local electrophilicity, the Fukui function for nucleophilic attack, and dual descriptor values of the phenolic hydroxyl oxygen along the IRC.
Related Concept Videos
 02:36
Acidity and Basicity of Alcohols and Phenols
 02:32
Physical Properties of Alcohols and Phenols
Alcohols possess a higher boiling point than aliphatic hydrocarbons of...
 02:49
Bond Energies and Bond Lengths
 02:13
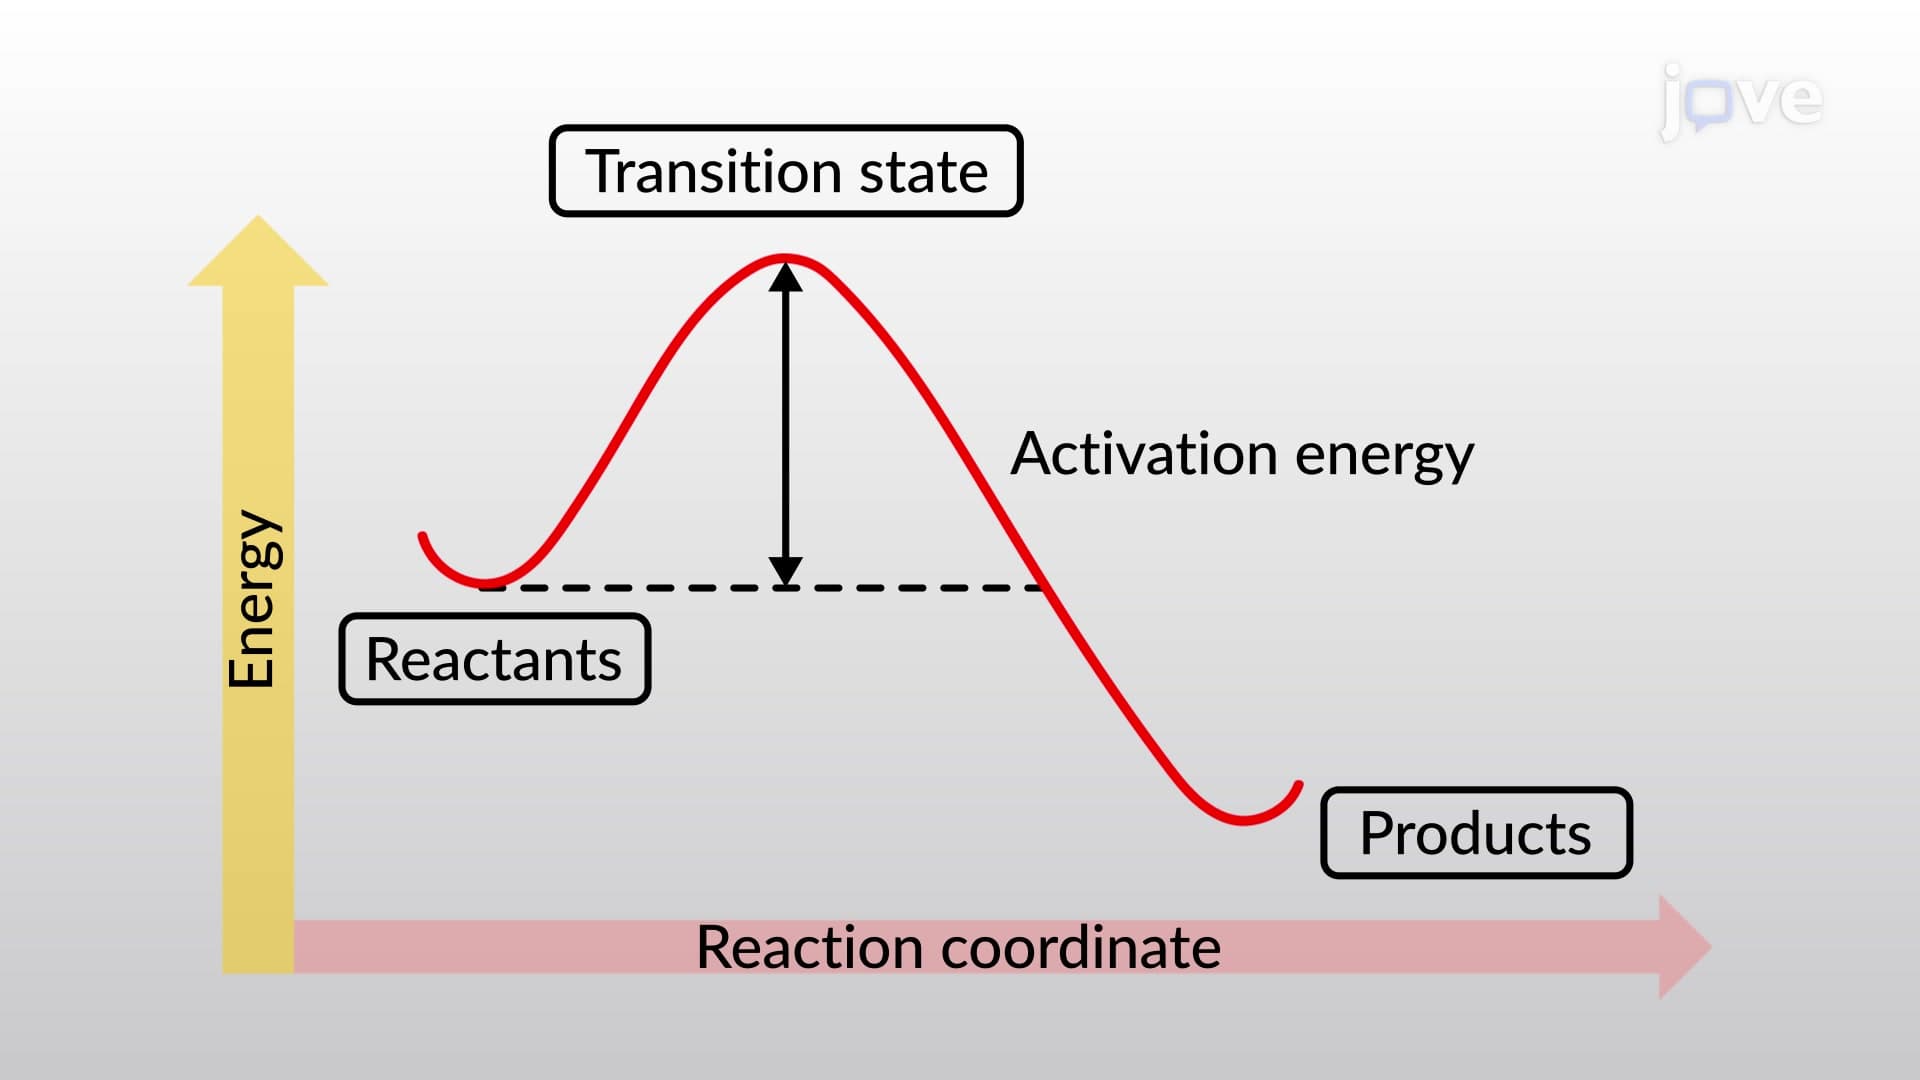
Bond Dissociation Energy and Activation Energy
 01:10
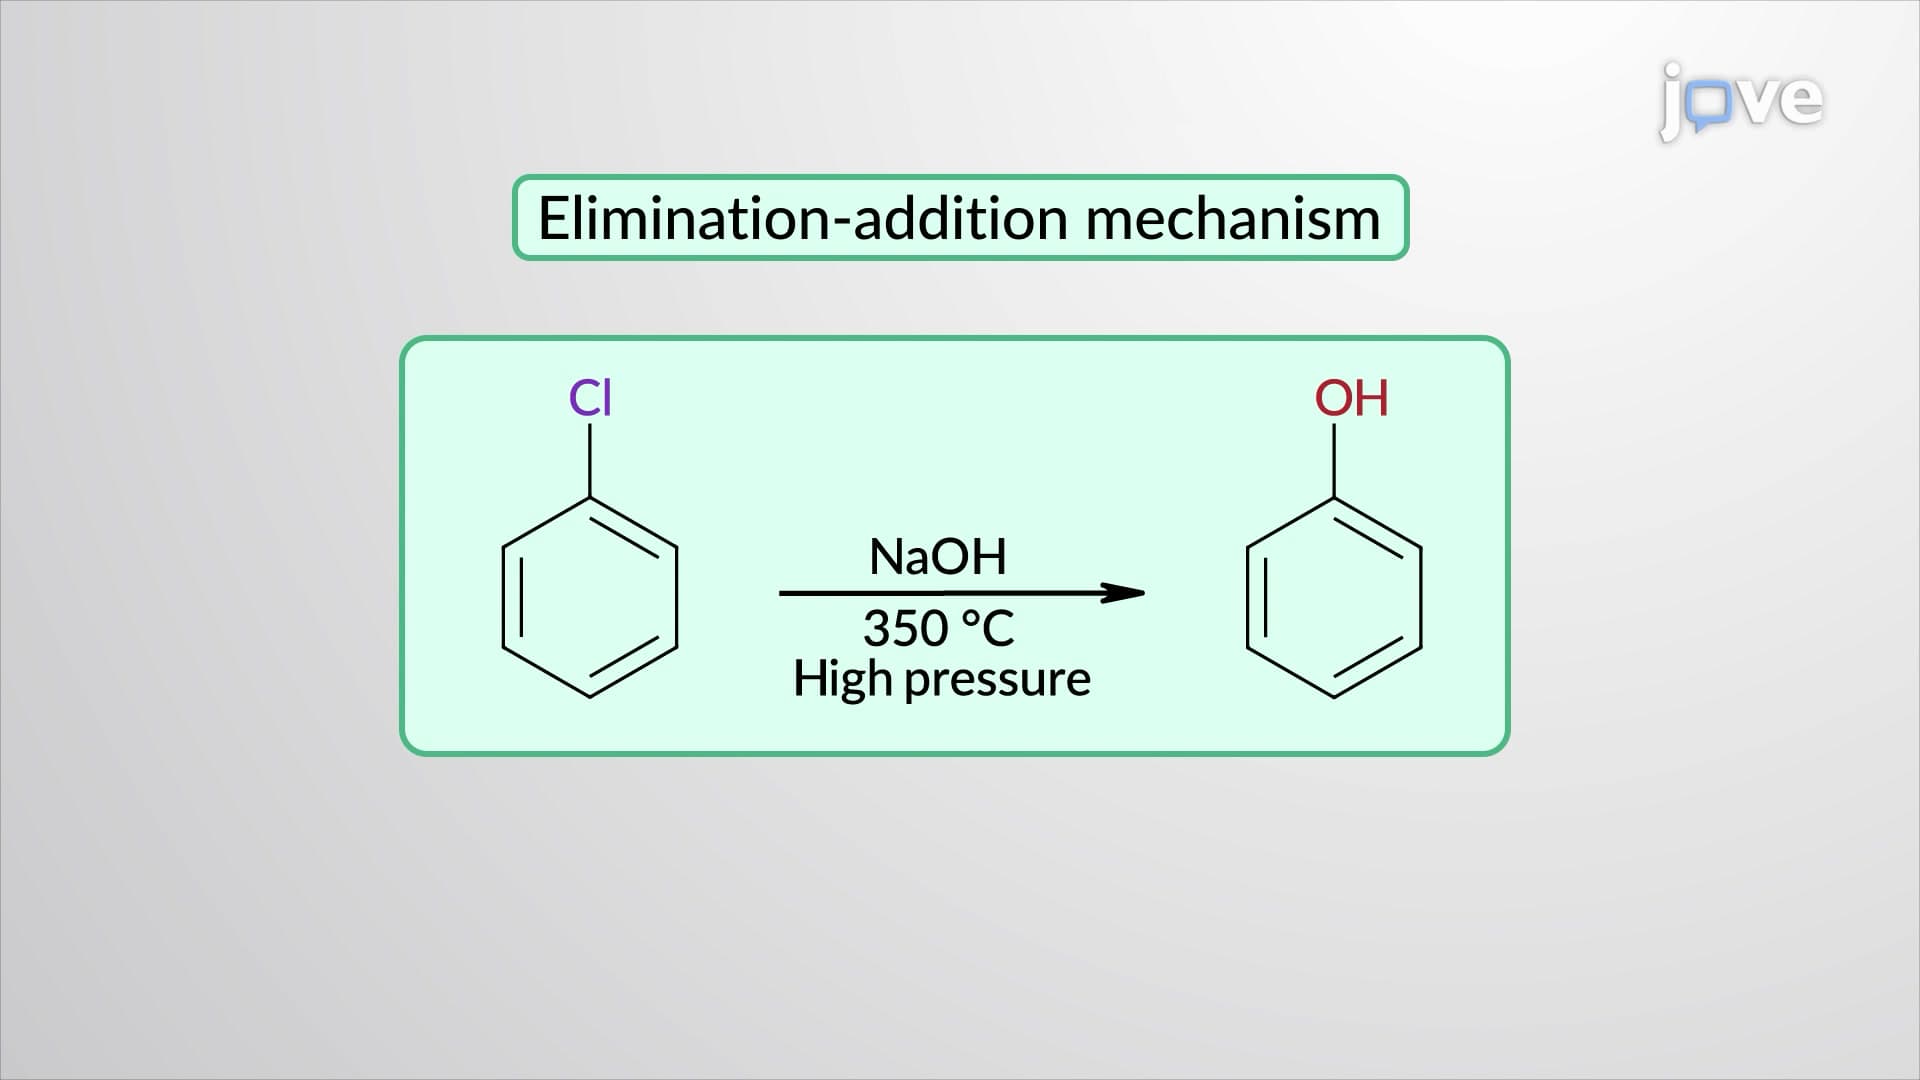
Hydrolysis of Chlorobenzene to Phenol: Dow Process
 01:23
IR Spectrum Peak Broadening: Hydrogen Bonding
However, the extent of hydrogen bonding influences the observed stretching frequency and band broadening. Intermolecular or intramolecular...