Related Experiment Video
Updated: Apr 18, 2026

Protection of H9c2 Myocardial Cells from Oxidative Stress by Crocetin via PINK1/Parkin Pathway-Mediated Mitophagy
Published on: May 26, 2023
Thioredoxin-2 inhibits mitochondrial reactive oxygen species generation and apoptosis stress kinase-1 activity to
Qunhua Huang1, Huanjiao Jenny Zhou1, Haifeng Zhang1
1From Interdepartmental Program in Vascular Biology and Therapeutics, Department of Pathology, University School of Medicine, New Haven, CT (Q.H., H.J.Z., H.Z., Y.H., F.J.G., W.M.); Center for Translational Medicine, The First Affiliated Hospital, Sun Yat-Sen University, Guangzhou, China (H.J.Z., W.M.); Gilead Sciences Inc, Foster City, CA (F.H.-K., P.F., L.Y., L.B., G.R.B.); and Department of Surgery, Yale University School of Medicine, New Haven, CT (G.T.).

Background:
Thioredoxin 2 (Trx2) is a key mitochondrial protein that regulates cellular redox and survival by suppressing mitochondrial reactive oxygen species generation and by inhibiting apoptosis stress kinase-1 (ASK1)-dependent apoptotic signaling. To date, the role of the mitochondrial Trx2 system in heart failure pathogenesis has not been investigated.
Methods And Results:
Western blot and histological analysis revealed that Trx2 protein expression levels were reduced in hearts from patients with dilated cardiomyopathy, with a concomitant increase in ASK1 phosphorylation/activity. Cardiac-specific Trx2 knockout mice develop spontaneous dilated cardiomyopathy at 1 month of age with increased heart size, reduced ventricular wall thickness, and a progressive decline in left ventricular contractile function, resulting in mortality due to heart failure by ≈4 months of age. The progressive decline in cardiac function observed in cardiac-specific Trx2 knockout mice was accompanied by the disruption of mitochondrial ultrastructure, mitochondrial membrane depolarization, increased mitochondrial reactive oxygen species generation, and reduced ATP production, correlating with increased ASK1 signaling and increased cardiomyocyte apoptosis. Chronic administration of a highly selective ASK1 inhibitor improved cardiac phenotype and reduced maladaptive left ventricular remodeling with significant reductions in oxidative stress, apoptosis, fibrosis, and cardiac failure. Cellular data from Trx2-deficient cardiomyocytes demonstrated that ASK1 inhibition reduced apoptosis and reduced mitochondrial reactive oxygen species generation.
Conclusions:
Our data support an essential role for mitochondrial Trx2 in preserving cardiac function by suppressing mitochondrial reactive oxygen species production and ASK1-dependent apoptosis. Inhibition of ASK1 represents a promising therapeutic strategy for the treatment of dilated cardiomyopathy and heart failure.
Insights
Mitochondrial Thioredoxin 2 (Trx2) deficiency causes heart failure by increasing oxidative stress and apoptosis. Inhibiting apoptosis stress kinase-1 (ASK1) shows promise for treating dilated cardiomyopathy and heart failure.
Area of Science:
- Cardiovascular Biology
- Mitochondrial Medicine
- Cellular Redox Signaling
Background:
- Thioredoxin 2 (Trx2) is a vital mitochondrial protein regulating cellular redox balance and survival.
- Trx2 suppresses mitochondrial reactive oxygen species (ROS) and apoptosis stress kinase-1 (ASK1)-dependent apoptosis.
- The role of mitochondrial Trx2 in heart failure pathogenesis was previously uninvestigated.
Purpose of the Study:
- To investigate the role of the mitochondrial Trx2 system in the pathogenesis of heart failure.
- To determine the therapeutic potential of inhibiting ASK1 in dilated cardiomyopathy.
Main Methods:
- Western blot and histological analysis of human heart samples and Trx2 knockout mice.
- Assessment of cardiac function, mitochondrial integrity, ROS production, and ATP levels.
- Evaluation of ASK1 signaling and cardiomyocyte apoptosis.
- Treatment with a selective ASK1 inhibitor in Trx2 knockout mice and cardiomyocytes.
Main Results:
- Reduced Trx2 expression and increased ASK1 activity were observed in hearts with dilated cardiomyopathy.
- Cardiac-specific Trx2 knockout mice developed spontaneous dilated cardiomyopathy, mitochondrial dysfunction, and premature mortality.
- ASK1 inhibition improved cardiac function, reduced oxidative stress, apoptosis, and fibrosis in Trx2 knockout mice.
- ASK1 inhibition reduced apoptosis and mitochondrial ROS in Trx2-deficient cardiomyocytes.
Conclusions:
- Mitochondrial Trx2 is essential for preserving cardiac function by mitigating ROS production and ASK1-dependent apoptosis.
- Inhibiting ASK1 is a potential therapeutic strategy for dilated cardiomyopathy and heart failure.
Related Concept Videos
 01:46
Electron Transport Chain: Complex I and II
ROS generation is regulated and maintained at moderate levels necessary...
 01:30
The Electron Transport Chain
Inhibitors of the electron transport chain
Rotenone, a widely used pesticide, prevents electron transfer from Fe-S cluster to ubiquinone or Q...
 01:43
Electron Transport Chain: Complex III and IV
 01:41
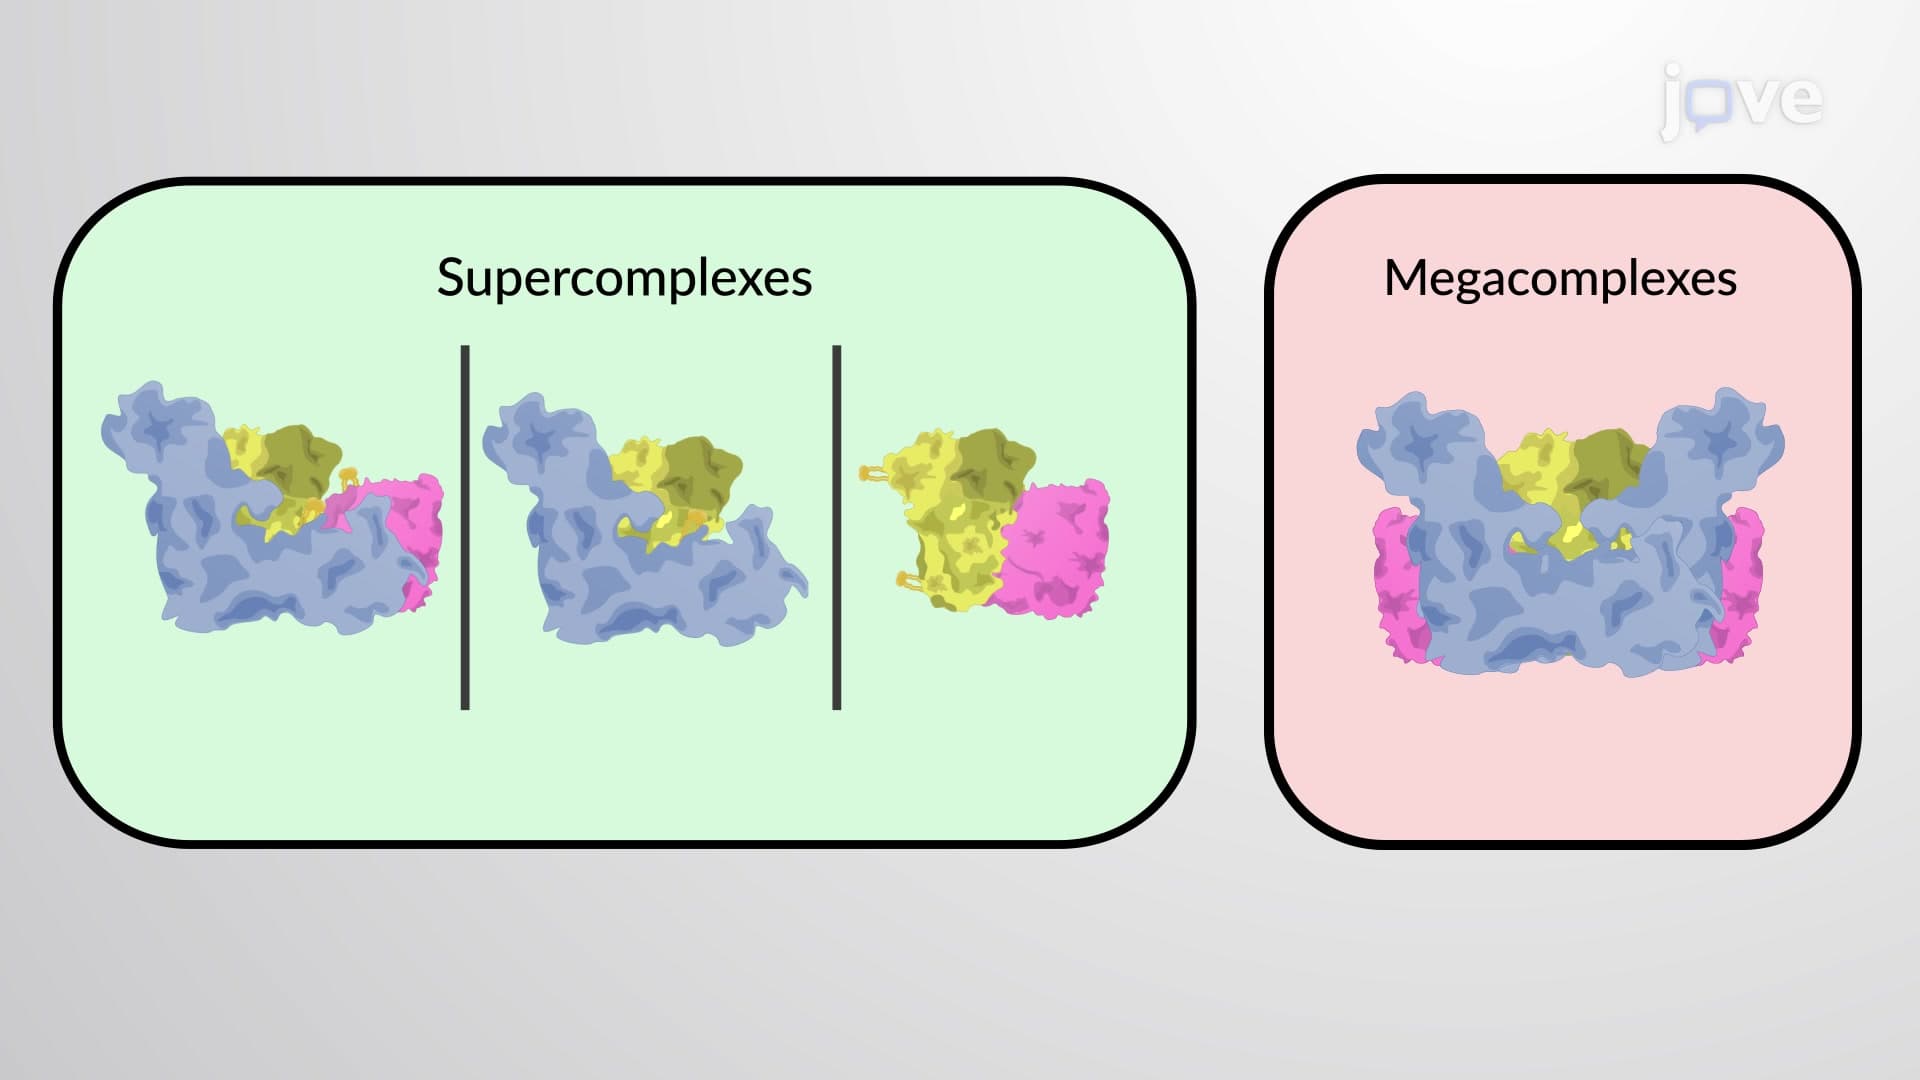
The Supercomplexes in the Crista Membrane
 01:31
Regulation of the Unfolded Protein Response
 01:45
Mitochondrial Membranes