Related Experiment Video
Updated: Apr 18, 2026

Kinase Inhibitor Screening In Self-assembled Human Protein Microarrays
Published on: October 23, 2019
Can structural features of kinase receptors provide clues on selectivity and inhibition? A molecular modeling study
Sarangan Ravichandran1, Brian T Luke1, Jack R Collins1
1Advanced Biomedical Computing Center, Frederick National Laboratory for Cancer Research (FNLCR), P.O. Box B, Frederick, MD 21702, USA.

Abstract:
Cancer is a complex disease resulting from the uncontrolled proliferation of cell signaling events. Protein kinases have been identified as central molecules that participate overwhelmingly in oncogenic events, thus becoming key targets for anticancer drugs. A majority of studies converged on the idea that ligand-binding pockets of kinases retain clues to the inhibiting abilities and cross-reacting tendencies of inhibitor drugs. Even though these ideas are critical for drug discovery, validating them using experiments is not only difficult, but also in some cases infeasible. To overcome these limitations and to test these ideas at the molecular level, we present here the results of receptor-focused in-silico docking of nine marketed drugs to 19 different wild-type and mutated kinases chosen from a wide range of families. This investigation highlights the need for using relevant models to explain the correct inhibition trends and the results are used to make predictions that might be able to influence future experiments. Our simulation studies are able to correctly predict the primary targets for each drug studied in majority of cases and our results agree with the existing findings. Our study shows that the conformations a given receptor acquires during kinase activation, and their micro-environment, defines the ligand partners. Type II drugs display high compatibility and selectivity for DFG-out kinase conformations. On the other hand Type I drugs are less selective and show binding preferences for both the open and closed forms of selected kinases. Using this receptor-focused approach, it is possible to capture the observed fold change in binding affinities between the wild-type and disease-centric mutations in ABL kinase for Imatinib and the second-generation ABL drugs. The effects of mutation are also investigated for two other systems, EGFR and B-Raf. Finally, by including pathway information in the design it is possible to model kinase inhibitors with potentially fewer side-effects.
Insights
This study used in-silico docking to predict anticancer drug targets, revealing that kinase conformations and micro-environments dictate drug binding. This approach accurately identifies drug targets and informs the design of more selective kinase inhibitors with fewer side effects.
Area of Science:
- Biochemistry
- Pharmacology
- Computational Biology
Background:
- Protein kinases are crucial in cancer due to uncontrolled cell signaling.
- Kinase inhibitors are key anticancer drug targets.
- Understanding drug binding to kinase pockets is vital for drug discovery.
Purpose of the Study:
- To perform in-silico docking of marketed drugs to wild-type and mutated kinases.
- To validate the role of kinase conformations and micro-environments in drug selectivity.
- To predict drug-target interactions and guide future experimental drug design.
Main Methods:
- Receptor-focused in-silico docking of nine marketed drugs.
- Analysis of 19 wild-type and mutated kinases across various families.
- Investigation of drug binding to different kinase conformations (DFG-out, open, closed).
Main Results:
- Accurate prediction of primary drug targets for most studied drugs.
- Demonstration that kinase conformations and micro-environments determine ligand binding.
- Type II drugs show selectivity for DFG-out conformations; Type I drugs are less selective.
- Accurate modeling of binding affinity changes for ABL kinase mutations with Imatinib.
Conclusions:
- Kinase conformation and micro-environment are critical determinants of drug selectivity.
- In-silico methods can accurately predict drug-target interactions and guide inhibitor design.
- This approach can help develop kinase inhibitors with improved efficacy and reduced side effects.
Related Concept Videos
 01:15
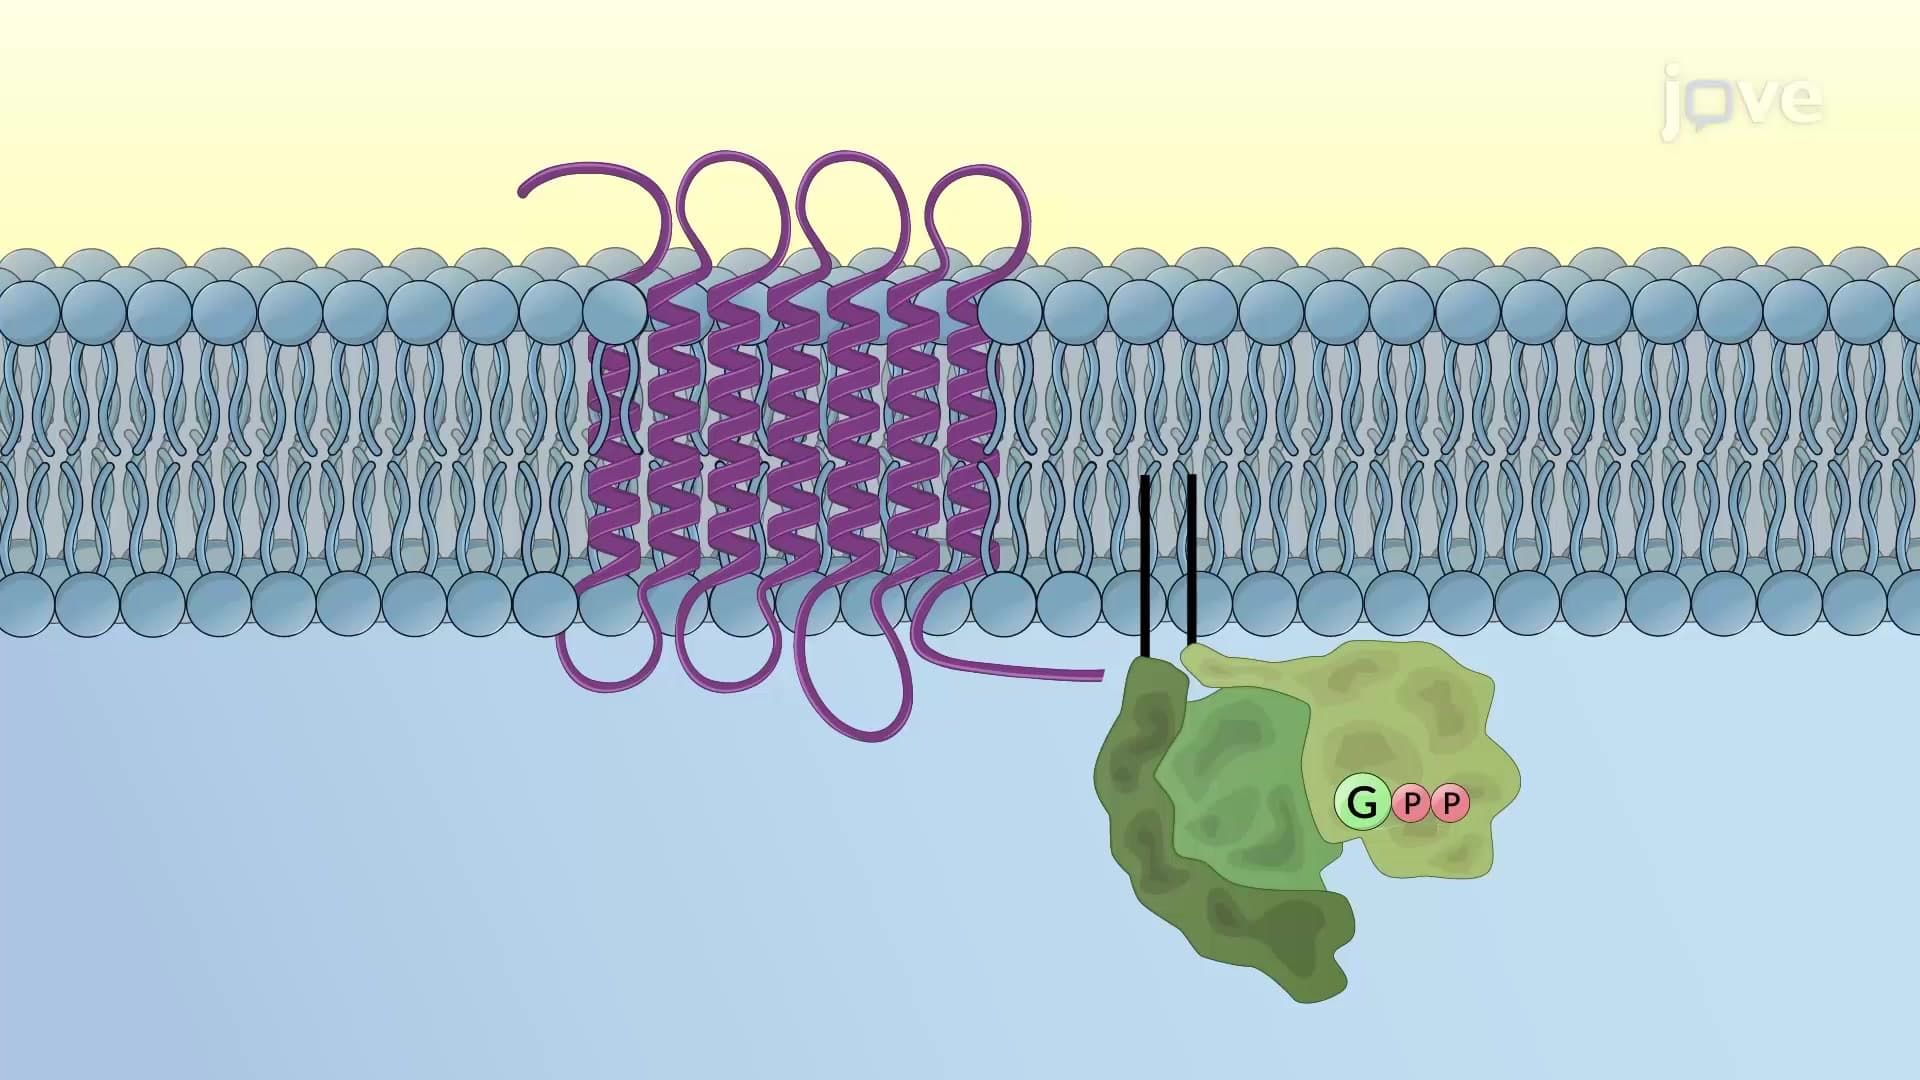
G Protein-coupled Receptors
GPCRs are also called heptahelical, 7TM, or serpentine receptors, and consist of seven (H1-H7) transmembrane alpha-helices that span the bilayer to form a cylindrical core. The transmembrane helices are connected by three extracellular loops and three...
 01:26
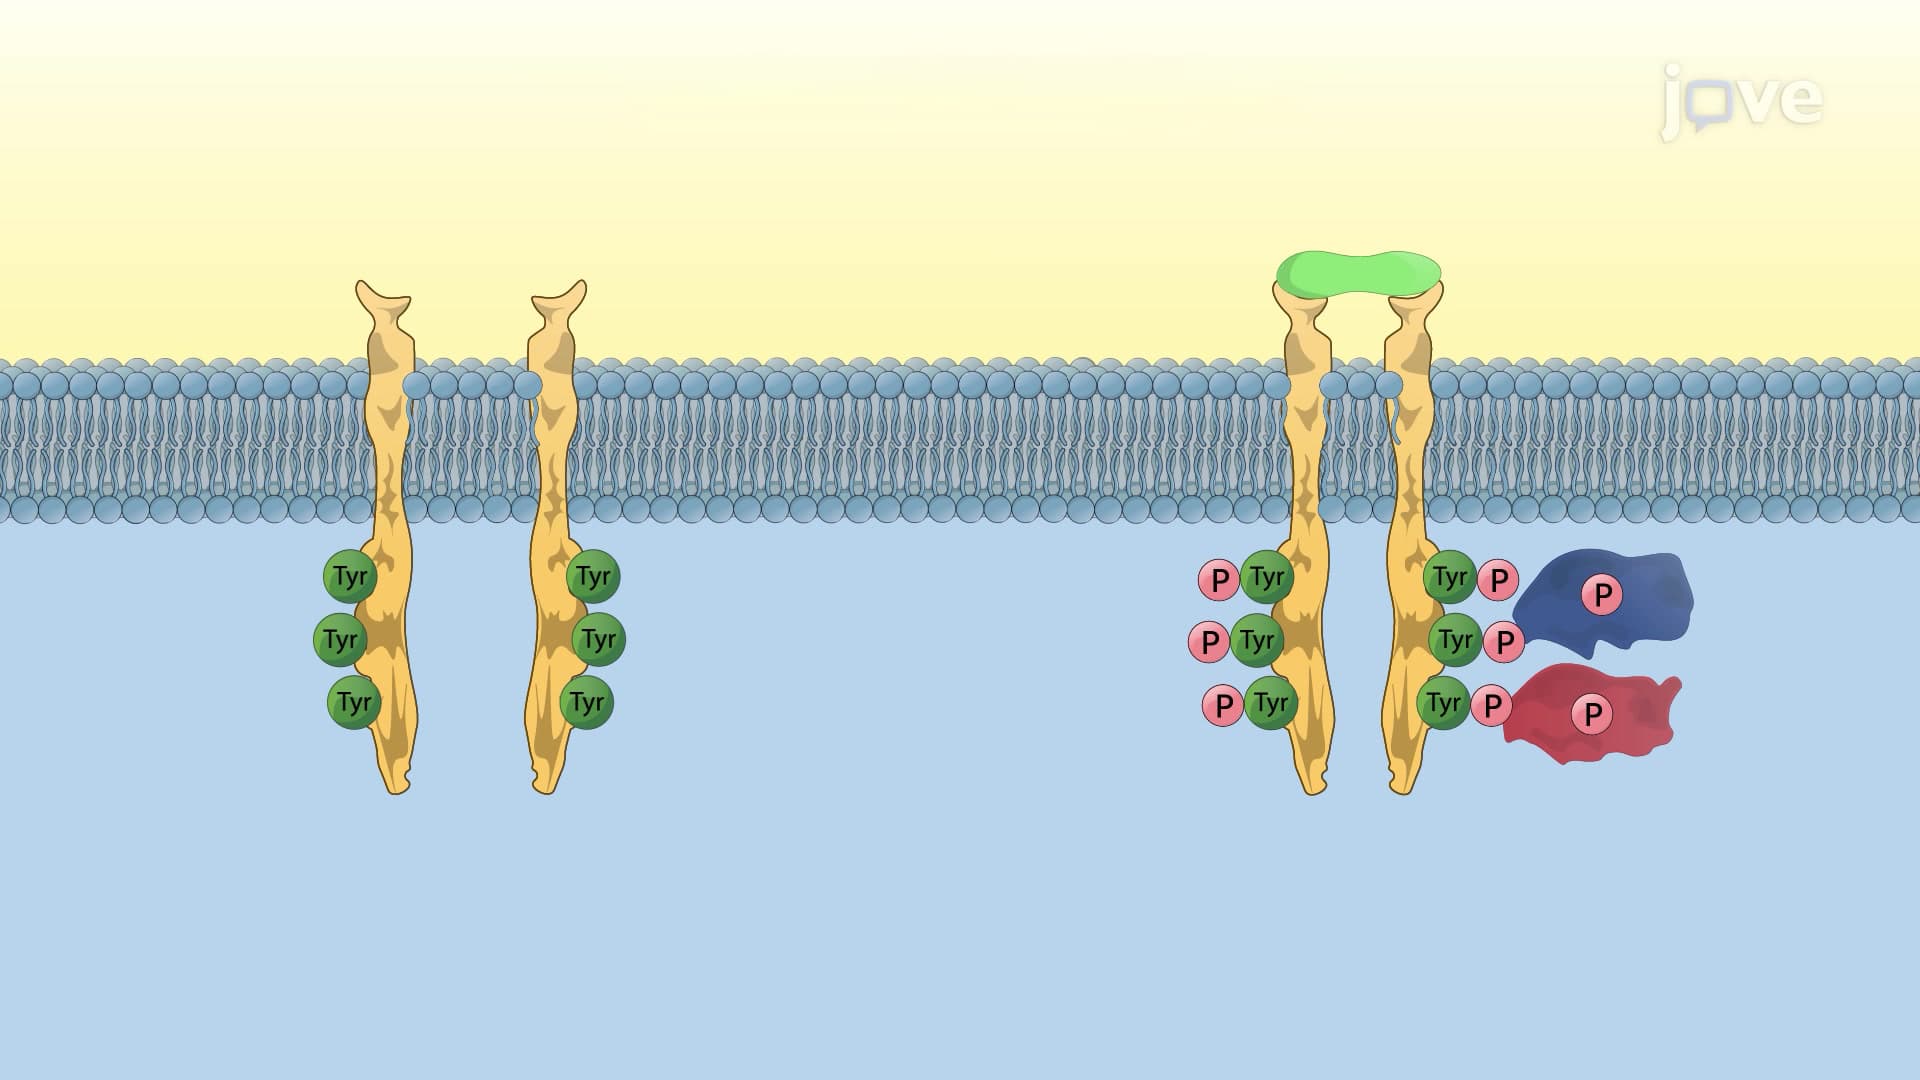
Receptor Tyrosine Kinases
 01:29
The Two-State Receptor Model
The binding affinity of a drug determines its interaction with...
 01:22
Amplifying Signals via Enzymatic Cascade
 01:27
Transducer Mechanism: Enzyme-Linked Receptors
Major types that are helpful drug targets include:
 01:30
Assembly of Signaling Complexes
Interaction domains in cell signaling
Interaction domains recognize exposed features of their binding partners containing post-translationally modified sequences,...