Related Experiment Video
Updated: Apr 18, 2026


Isotopic Effect in Double Proton Transfer Process of Porphycene Investigated by Enhanced QM/MM Method
Published on: July 19, 2019
van der Waals interactions are critical in Car-Parrinello molecular dynamics simulations of porphyrin-fullerene dyads
Topi Karilainen1, Oana Cramariuc, Mikael Kuisma
1Department of Physics, Tampere University of Technology, Tampere, Finland.

Abstract:
The interplay between electrostatic and van der Waals (vdW) interactions in porphyrin-C60 dyads is still under debate despite its importance in influencing the structural characteristics of such complexes considered for various applications in molecular photovoltaics. In this article, we sample the conformational space of a porphyrin-C60 dyad using Car-Parrinello molecular dynamics simulations with and without empirical vdW corrections. Long-range vdW interactions, which are poorly described by the commonly used density functional theory functionals, prove to be essential for a proper dynamics of the dyad moieties. Inclusion of vdW corrections brings porphyrin and C60 close together in an orientation that is in agreement with experimental observations. The structural differences arising from the vdW corrections are shown to be significant for several properties and potentially less important for others. Additionally, our Mulliken population analysis reveals that contrary to the common belief, porphyrin is not the primary electron donating moiety for C60 . In the considered dyad, fullerene's affinity for electrons is primarily satisfied by charge transfer from the amide group of the linker. However, we show that in the absence of another suitable bound donor, C60 can withdraw electrons from porphyrin if it is sufficiently close.
More Related Videos
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
 13:56
Probe Type II Band Alignment in One-Dimensional Van Der Waals Heterostructures Using First-Principles Calculations
Published on: October 12, 2019
Related Concept Videos
 01:24
Van der Waals Interactions
 02:35
Noncovalent Attractions in Biomolecules
Four types of noncovalent interactions are hydrogen bonds, van der Waals forces, ionic bonds, and hydrophobic interactions.
Hydrogen bonding results from the electrostatic attraction of a hydrogen atom covalently bonded to a strong-electronegative atom like oxygen,...
02:35Noncovalent Attractions in Biomolecules
 01:24
π Molecular Orbitals of 1,3-Butadiene
The simplest conjugated diene is 1,3-butadiene: a four-carbon system where each carbon is sp2-hybridized and has an unhybridized p orbital that contains an unpaired electron. According to molecular orbital theory, atomic orbitals combine to form molecular orbitals such that the...
 01:10
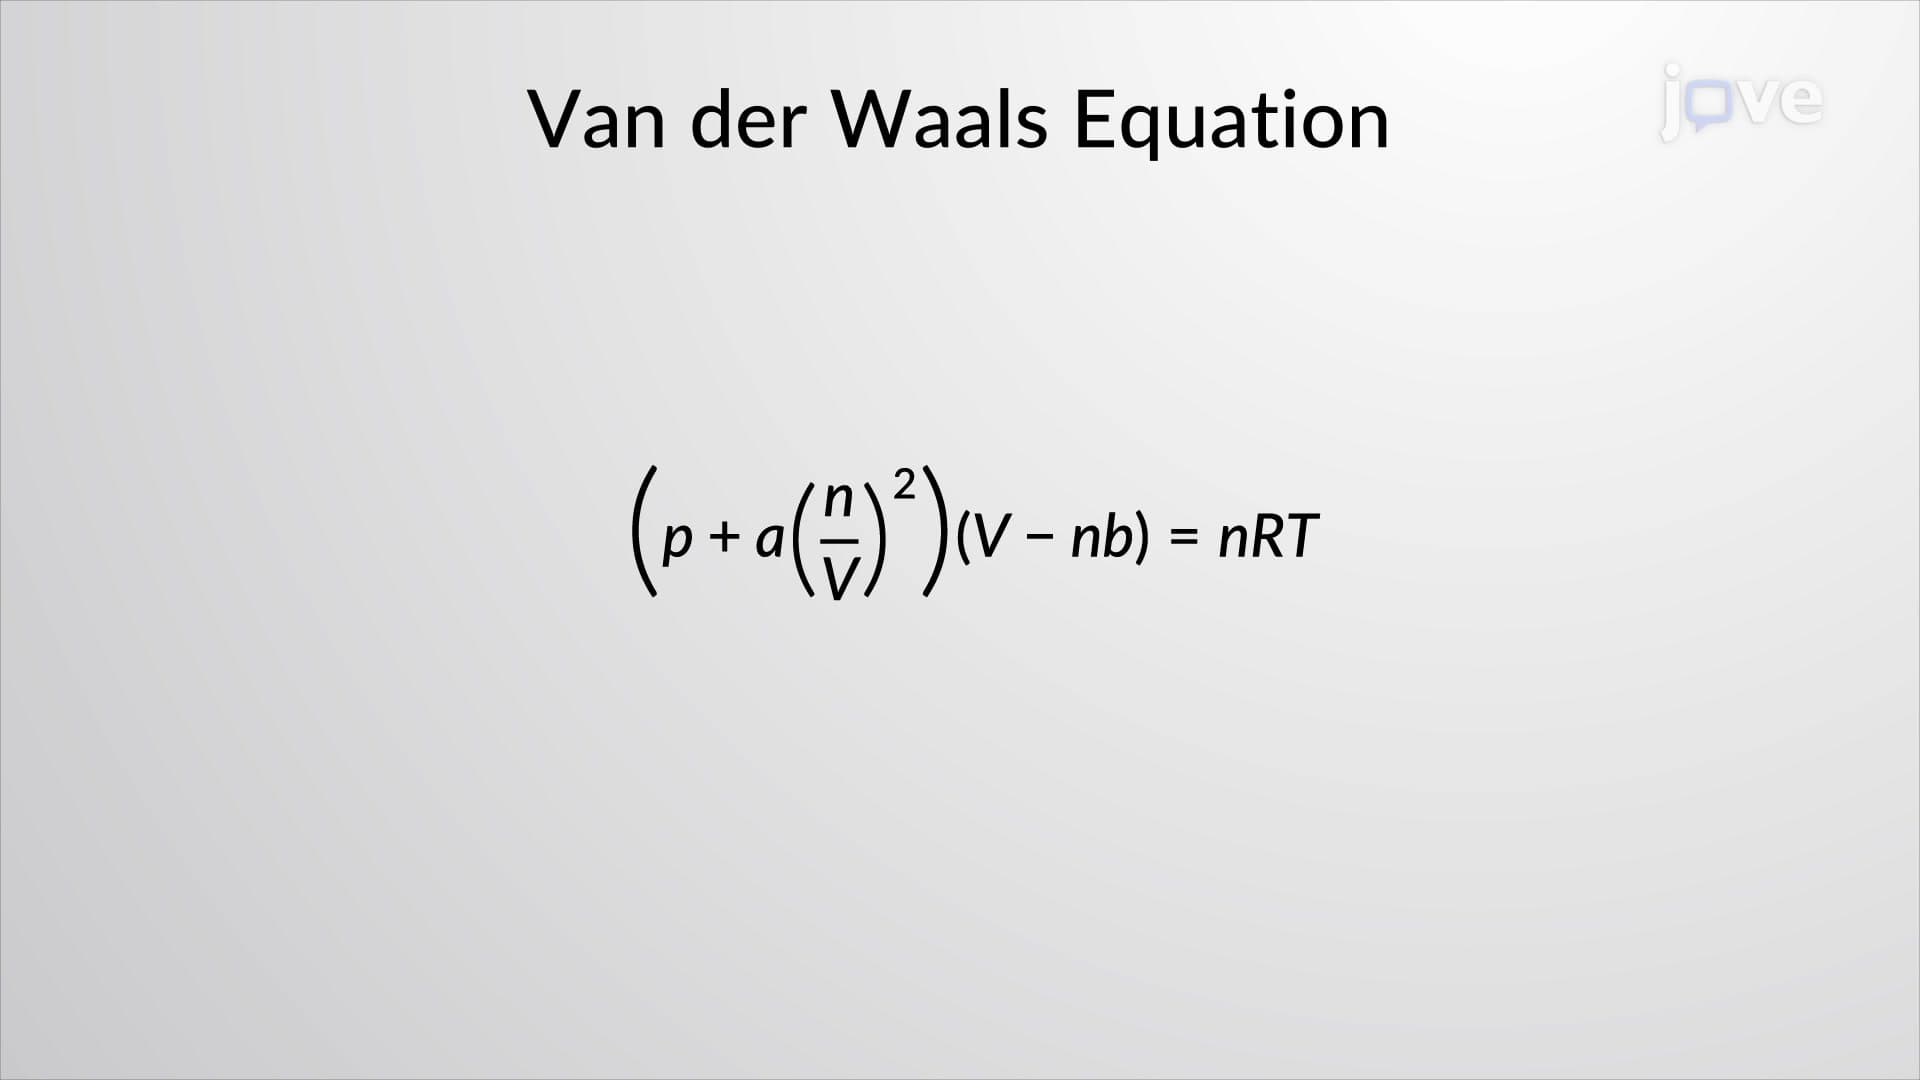
Van der Waals Equation
First, the attractive forces between molecules, which are stronger at higher densities and reduce the pressure, are considered by adding to the pressure a term equal to the square of the molar density multiplied by a positive coefficient a. Second, the...
 01:22
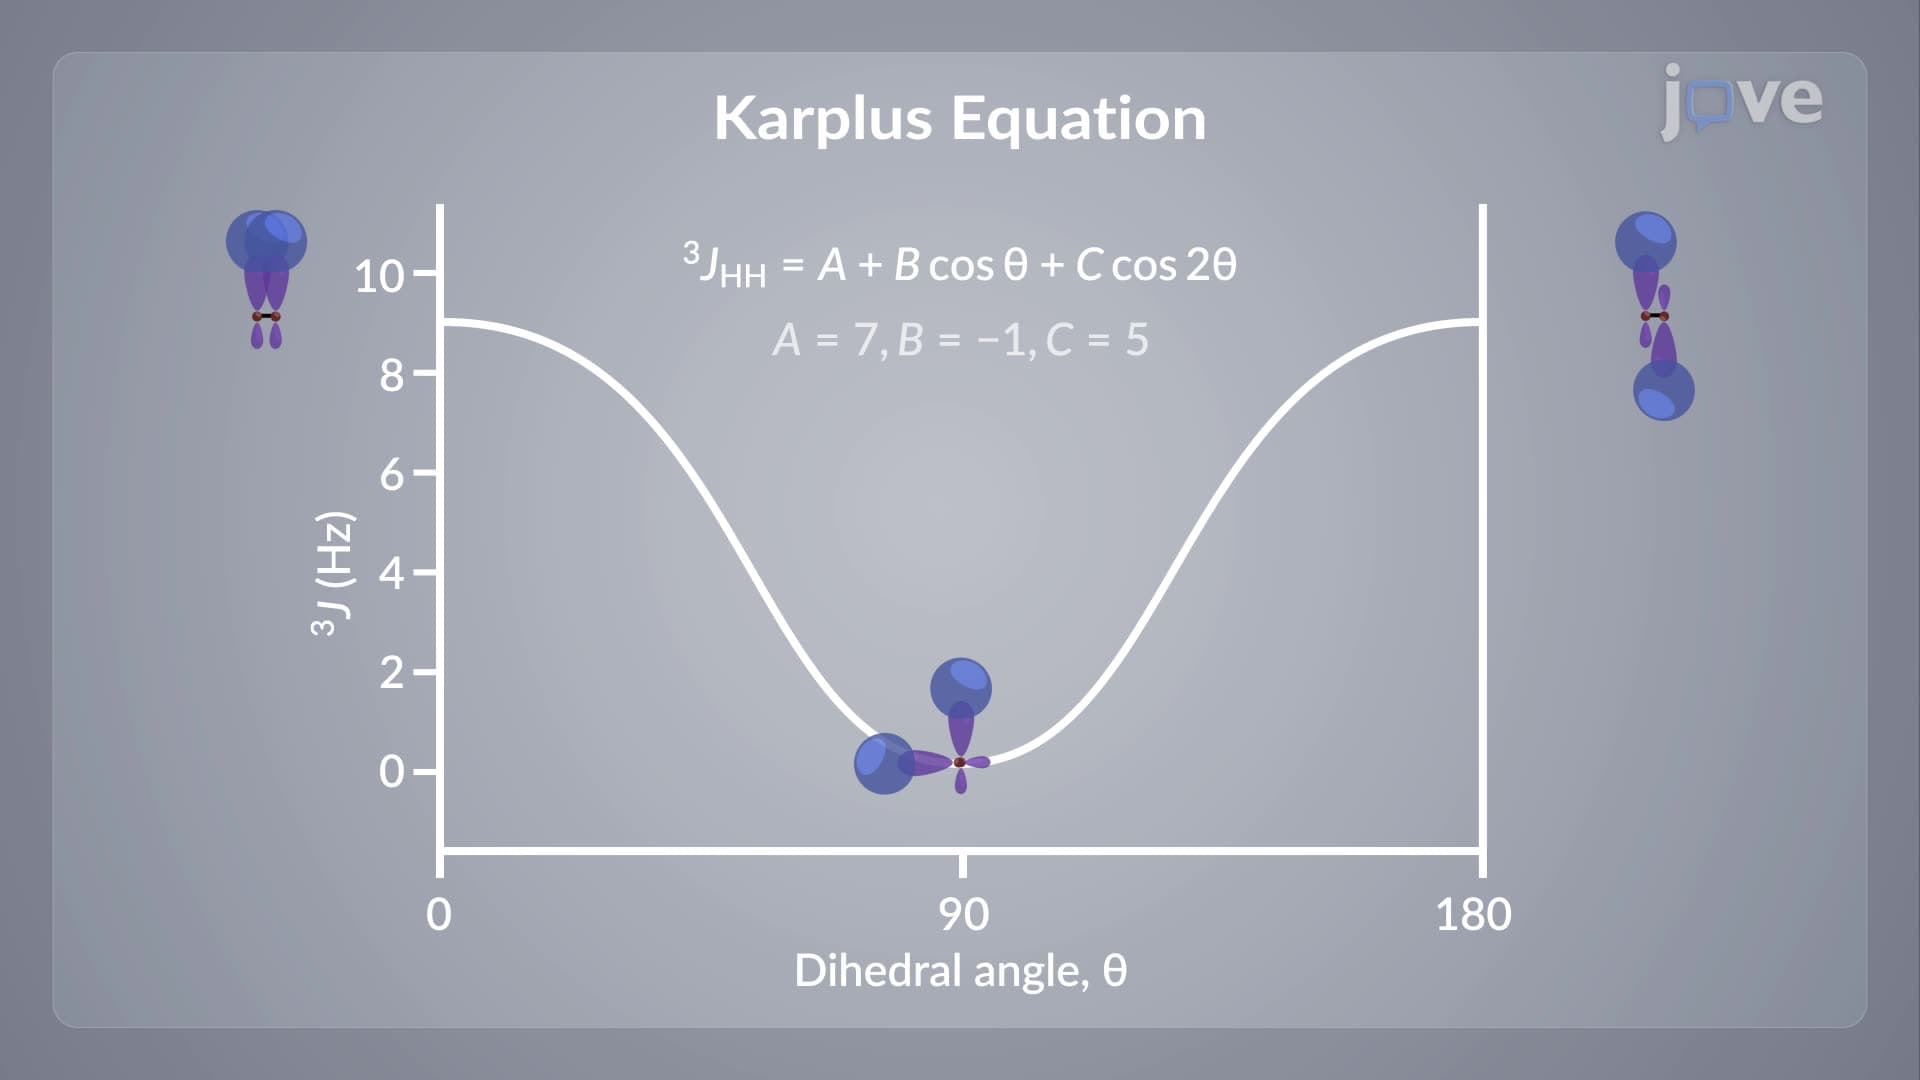
Spin–Spin Coupling: Three-Bond Coupling (Vicinal Coupling)
The extent of coupling depends on the C‑C bond length, the two H‑C‑C angles, any electron-withdrawing substituents, and the dihedral angle between the...