Related Experiment Video
Updated: Apr 18, 2026

In Vitro Analysis of PDZ-dependent CFTR Macromolecular Signaling Complexes
Published on: August 13, 2012
On the interactions between nucleotide binding domains and membrane spanning domains in cystic fibrosis transmembrane
1Istituto di Biofisica, CNR, Genova, Italy; Centre for Integrative Biology, CIBIO, University of Trento, Trento, Italy.

Abstract:
The Cystic Fibrosis Transmembrane Regulator (CFTR) is a membrane protein whose mutations cause cystic fibrosis, a lethal genetic disease. We performed a molecular dynamic (MD) study of the properties of the nucleotide binding domains (NBD) whose conformational changes, upon ATP binding, are the direct responsible of the gating mechanisms of CFTR. This study was done for the wild type (WT) CFTR and for the two most common mutations, ΔF508, that produces a traffic defect of the protein, and the mutation G551D, that causes a gating defect on CFTR. Using an homology model of the open channel conformation of the CFTR we thus introduced the mutations to the structure. Although the overall structures of the G551D and ΔF508 are quite well conserved, the NBD1-NBD2 interactions are severely modified in both mutants. NBD1 and NBD2 are indeed destabilized with a higher internal energy (Ei) in the ΔF508-CFTR. Differently, Ei does not change in the NBDs of G551D but, while the number of close contacts between NBD1 and NBD2 in ΔF508 is increased, a significant reduction of close contacts is found in the G551D mutated form. Hydrogen bonds formation between NBDs of the two mutated forms is also altered and it is slightly increased for the ΔF508, while are severely reduced in G551D. A consequent modification of the NBDs-ICLs interactions between residues involved in the transduction of the ATP binding and the channel gating is also registered. Indeed, while a major interaction is noticed between NBDs interface and ICL2 and ICL4 in the WT, this interaction is somehow altered in both mutated forms plausibly with effect on channel gating. Thus, single point mutations of the CFTR protein can reasonably results in channel gating defects due to alteration of the interaction mechanisms between the NBDs and NBDs-ICLs interfaces upon ATP-binding process.
Insights
Molecular dynamics simulations reveal how mutations in the Cystic Fibrosis Transmembrane Regulator (CFTR) disrupt nucleotide binding domain interactions, impacting CFTR protein function and leading to cystic fibrosis.
Area of Science:
- Biophysics
- Molecular Biology
- Genetics
Background:
- Cystic Fibrosis Transmembrane Regulator (CFTR) mutations cause cystic fibrosis.
- CFTR protein function relies on nucleotide binding domain (NBD) conformational changes upon ATP binding.
- Key CFTR mutations include ΔF508 (traffic defect) and G551D (gating defect).
Purpose of the Study:
- Investigate the impact of ΔF508 and G551D mutations on CFTR NBD properties using molecular dynamics.
- Analyze alterations in NBD1-NBD2 interactions, internal energy, and hydrogen bonding in mutated CFTR.
- Examine changes in NBDs-ICLs interactions and their potential effect on CFTR channel gating.
Main Methods:
- Molecular dynamics (MD) simulations.
- Utilized an homology model of the open channel conformation of CFTR.
- Introduced ΔF508 and G551D mutations into the CFTR model for analysis.
Main Results:
- While overall structures are conserved, NBD1-NBD2 interactions are significantly modified in both mutants.
- ΔF508-CFTR shows destabilized NBDs with higher internal energy and increased NBD1-NBD2 contacts.
- G551D-CFTR exhibits reduced NBD1-NBD2 contacts and severely reduced hydrogen bonds, with unchanged NBD internal energy.
- Interactions between NBDs and intracellular loops (ICLs) are altered in both mutants, affecting ATP binding signal transduction.
Conclusions:
- Single point mutations in CFTR can lead to channel gating defects.
- Altered interactions within NBDs and between NBDs-ICLs interfaces disrupt ATP-binding and gating mechanisms.
- These findings provide insights into the molecular basis of cystic fibrosis pathogenesis.
More Related Videos
 14:56
Expression and Purification of the Cystic Fibrosis Transmembrane Conductance Regulator Protein in Saccharomyces cerevisiae
Published on: March 10, 2012
 09:59
Functional Reconstitution and Channel Activity Measurements of Purified Wildtype and Mutant CFTR Protein
Published on: March 9, 2015
Related Concept Videos
 01:09
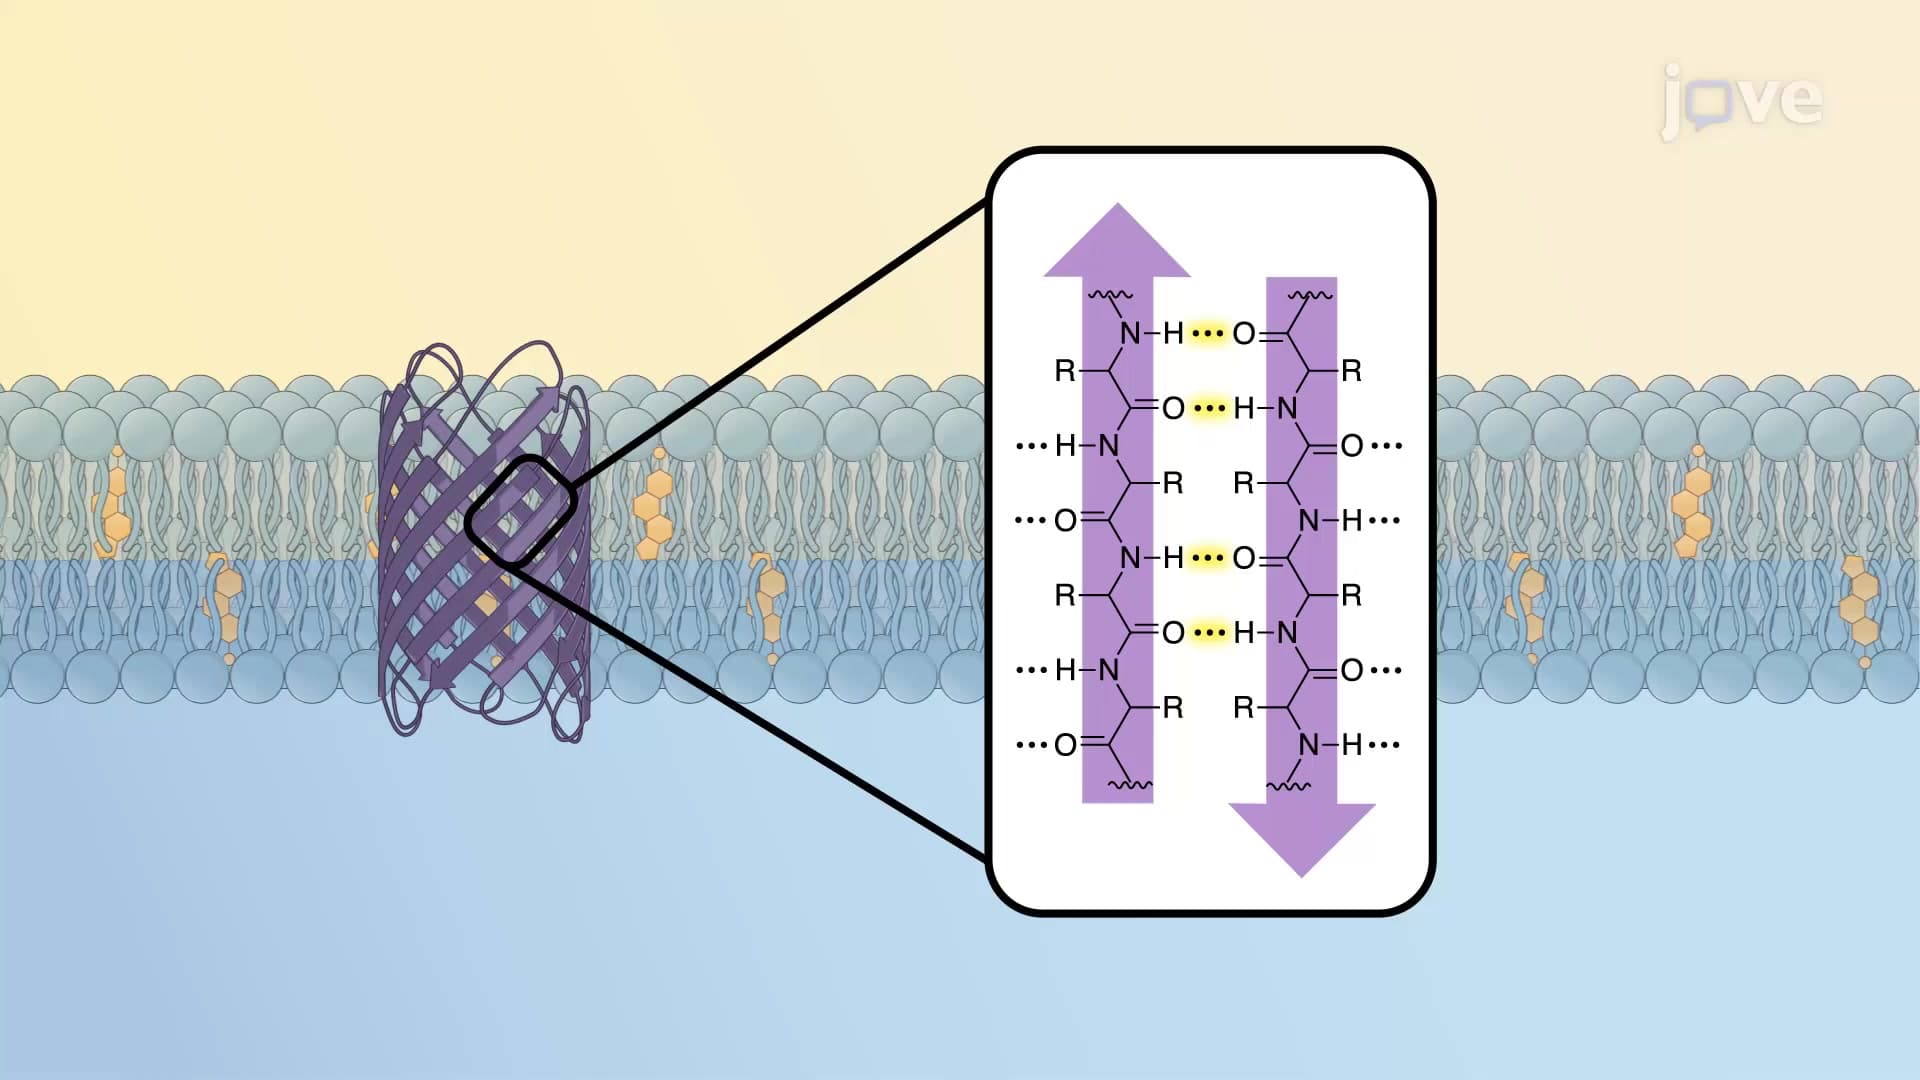
Multi-pass Transmembrane Proteins and β-barrels
α-Helix containing multi-pass transmembrane proteins
Multi-pass transmembrane proteins such as...
 01:23
Cystic Fibrosis: Pathogenesis
CF is primarily caused by a genetic mutation in a chromosome 7 gene coding for the cystic fibrosis transmembrane conductance regulator (CFTR) protein. The most common gene mutation leading to CF is the ΔF508 mutation,...
 00:59
Mechanisms of Membrane Domain Formation
Another mechanism for membrane domain formation involves membrane proteins interacting with...