De novo heterozygous mutations in SMC3 cause a range of Cornelia de Lange syndrome-overlapping phenotypes
Related Experiment Video
Updated: Apr 17, 2026
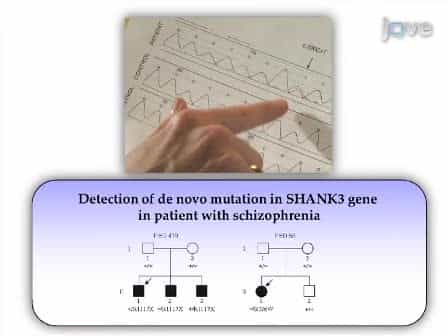
A Strategy to Identify de Novo Mutations in Common Disorders such as Autism and Schizophrenia
Published on: June 15, 2011
María Concepción Gil-Rodríguez1, Matthew A Deardorff, Morad Ansari
1Unit of Clinical Genetics and Functional Genomics, Departments of Pharmacology-Physiology and Pediatrics, Medical School, University of Zaragoza, CIBERER-GCV and ISS-Aragon, Zaragoza, Spain.

Abstract:
Cornelia de Lange syndrome (CdLS) is characterized by facial dysmorphism, growth failure, intellectual disability, limb malformations, and multiple organ involvement. Mutations in five genes, encoding subunits of the cohesin complex (SMC1A, SMC3, RAD21) and its regulators (NIPBL, HDAC8), account for at least 70% of patients with CdLS or CdLS-like phenotypes. To date, only the clinical features from a single CdLS patient with SMC3 mutation has been published. Here, we report the efforts of an international research and clinical collaboration to provide clinical comparison of 16 patients with CdLS-like features caused by mutations in SMC3. Modeling of the mutation effects on protein structure suggests a dominant-negative effect on the multimeric cohesin complex. When compared with typical CdLS, many SMC3-associated phenotypes are also characterized by postnatal microcephaly but with a less distinctive craniofacial appearance, a milder prenatal growth retardation that worsens in childhood, few congenital heart defects, and an absence of limb deficiencies. While most mutations are unique, two unrelated affected individuals shared the same mutation but presented with different phenotypes. This work confirms that de novo SMC3 mutations account for ∼ 1%-2% of CdLS-like phenotypes.
Insights
Mutations in the SMC3 gene cause Cornelia de Lange syndrome (CdLS)-like features, presenting distinct phenotypes compared to typical CdLS. This study analyzes 16 patients, revealing specific clinical characteristics associated with SMC3 mutations.
Area of Science:
- Genetics
- Molecular Biology
- Developmental Biology
Background:
- Cornelia de Lange syndrome (CdLS) is a genetic disorder with characteristic facial dysmorphism, growth failure, intellectual disability, limb malformations, and organ involvement.
- Mutations in cohesin complex genes (SMC1A, SMC3, RAD21) and regulators (NIPBL, HDAC8) explain over 70% of CdLS cases.
- Clinical data for CdLS patients with SMC3 mutations are scarce, with only one case previously published.
Purpose of the Study:
- To present a detailed clinical comparison of 16 patients with CdLS-like features caused by SMC3 mutations.
- To investigate the impact of SMC3 mutations on cohesin complex function and associated phenotypes.
- To delineate the specific clinical spectrum of SMC3-related CdLS.
Main Methods:
- International research and clinical collaboration involving 16 patients with CdLS-like features.
- Clinical data collection and comparative analysis of patient phenotypes.
- Protein structure modeling to understand the effect of mutations on the cohesin complex.
Main Results:
- SMC3 mutations account for approximately 1%-2% of CdLS-like phenotypes.
- SMC3-associated phenotypes include postnatal microcephaly, less distinct craniofacial features, milder prenatal growth retardation, fewer congenital heart defects, and absence of limb deficiencies compared to typical CdLS.
- While most mutations were unique, two unrelated patients shared the same mutation but exhibited different phenotypes, suggesting variable expressivity.
Conclusions:
- De novo SMC3 mutations are a cause of CdLS-like phenotypes with a distinct clinical profile.
- The identified SMC3 mutations likely exert a dominant-negative effect on the cohesin complex.
- Further research is needed to understand the variable expressivity observed in patients with identical SMC3 mutations.
More Related Videos
 06:41
In Vivo Functional Study of Disease-associated Rare Human Variants Using Drosophila
Published on: August 20, 2019
 08:22
A Novel Strategy Combining Array-CGH, Whole-exome Sequencing and In Utero Electroporation in Rodents to Identify Causative Genes for Brain Malformations
Published on: December 1, 2017
Related Concept Videos
 01:33
Pleiotropy
 00:49
Complementation Tests
Organisms heterozygous for different mutations are crossed pairwise in all combinations. If present on different genes, the mutations can complement each other by providing the missing...
 01:49
Multiple Allele Traits
01:49Multiple Allele Traits
 01:43
Incomplete Dominance
 02:20
Cohesins
Cohesin complexes in Meiotic Division
Meiosis involves two distinct rounds of chromosomal segregation and cell divisions— Meiosis I followed by Meiosis II – producing four daughter cells. Meiosis I includes the separation of...