Related Experiment Video
Updated: Apr 17, 2026

Assessment of Open Probability of the Mitochondrial Permeability Transition Pore in the Setting of Coenzyme Q Excess
Published on: June 1, 2022
COQ4 mutations cause a broad spectrum of mitochondrial disorders associated with CoQ10 deficiency
Gloria Brea-Calvo1, Tobias B Haack2, Daniela Karall3
1Centro Andaluz de Biología del Desarrollo, Universidad Pablo de Olavide - Consejo Superior de Investigaciones Científicas - Junta de Andalucía and Centro de Investigación Biomédica en Red de Enfermedades Raras, 41013 Sevilla, Spain.

Abstract:
Primary coenzyme Q10 (CoQ10) deficiencies are rare, clinically heterogeneous disorders caused by mutations in several genes encoding proteins involved in CoQ10 biosynthesis. CoQ10 is an essential component of the electron transport chain (ETC), where it shuttles electrons from complex I or II to complex III. By whole-exome sequencing, we identified five individuals carrying biallelic mutations in COQ4. The precise function of human COQ4 is not known, but it seems to play a structural role in stabilizing a multiheteromeric complex that contains most of the CoQ10 biosynthetic enzymes. The clinical phenotypes of the five subjects varied widely, but four had a prenatal or perinatal onset with early fatal outcome. Two unrelated individuals presented with severe hypotonia, bradycardia, respiratory insufficiency, and heart failure; two sisters showed antenatal cerebellar hypoplasia, neonatal respiratory-distress syndrome, and epileptic encephalopathy. The fifth subject had an early-onset but slowly progressive clinical course dominated by neurological deterioration with hardly any involvement of other organs. All available specimens from affected subjects showed reduced amounts of CoQ10 and often displayed a decrease in CoQ10-dependent ETC complex activities. The pathogenic role of all identified mutations was experimentally validated in a recombinant yeast model; oxidative growth, strongly impaired in strains lacking COQ4, was corrected by expression of human wild-type COQ4 cDNA but failed to be corrected by expression of COQ4 cDNAs with any of the mutations identified in affected subjects. COQ4 mutations are responsible for early-onset mitochondrial diseases with heterogeneous clinical presentations and associated with CoQ10 deficiency.
More Related Videos
 05:27
Inner Mitochondrial Membrane Sensitivity to Na+ Reveals Partially Segmented Functional CoQ Pools
Published on: July 20, 2022
 06:53
Visualization of Mitochondrial Respiratory Function using Cytochrome C Oxidase / Succinate Dehydrogenase COX/SDH Double-labeling Histochemistry
Published on: November 23, 2011
Related Concept Videos
 02:59
Animal Mitochondrial Genetics
 01:39
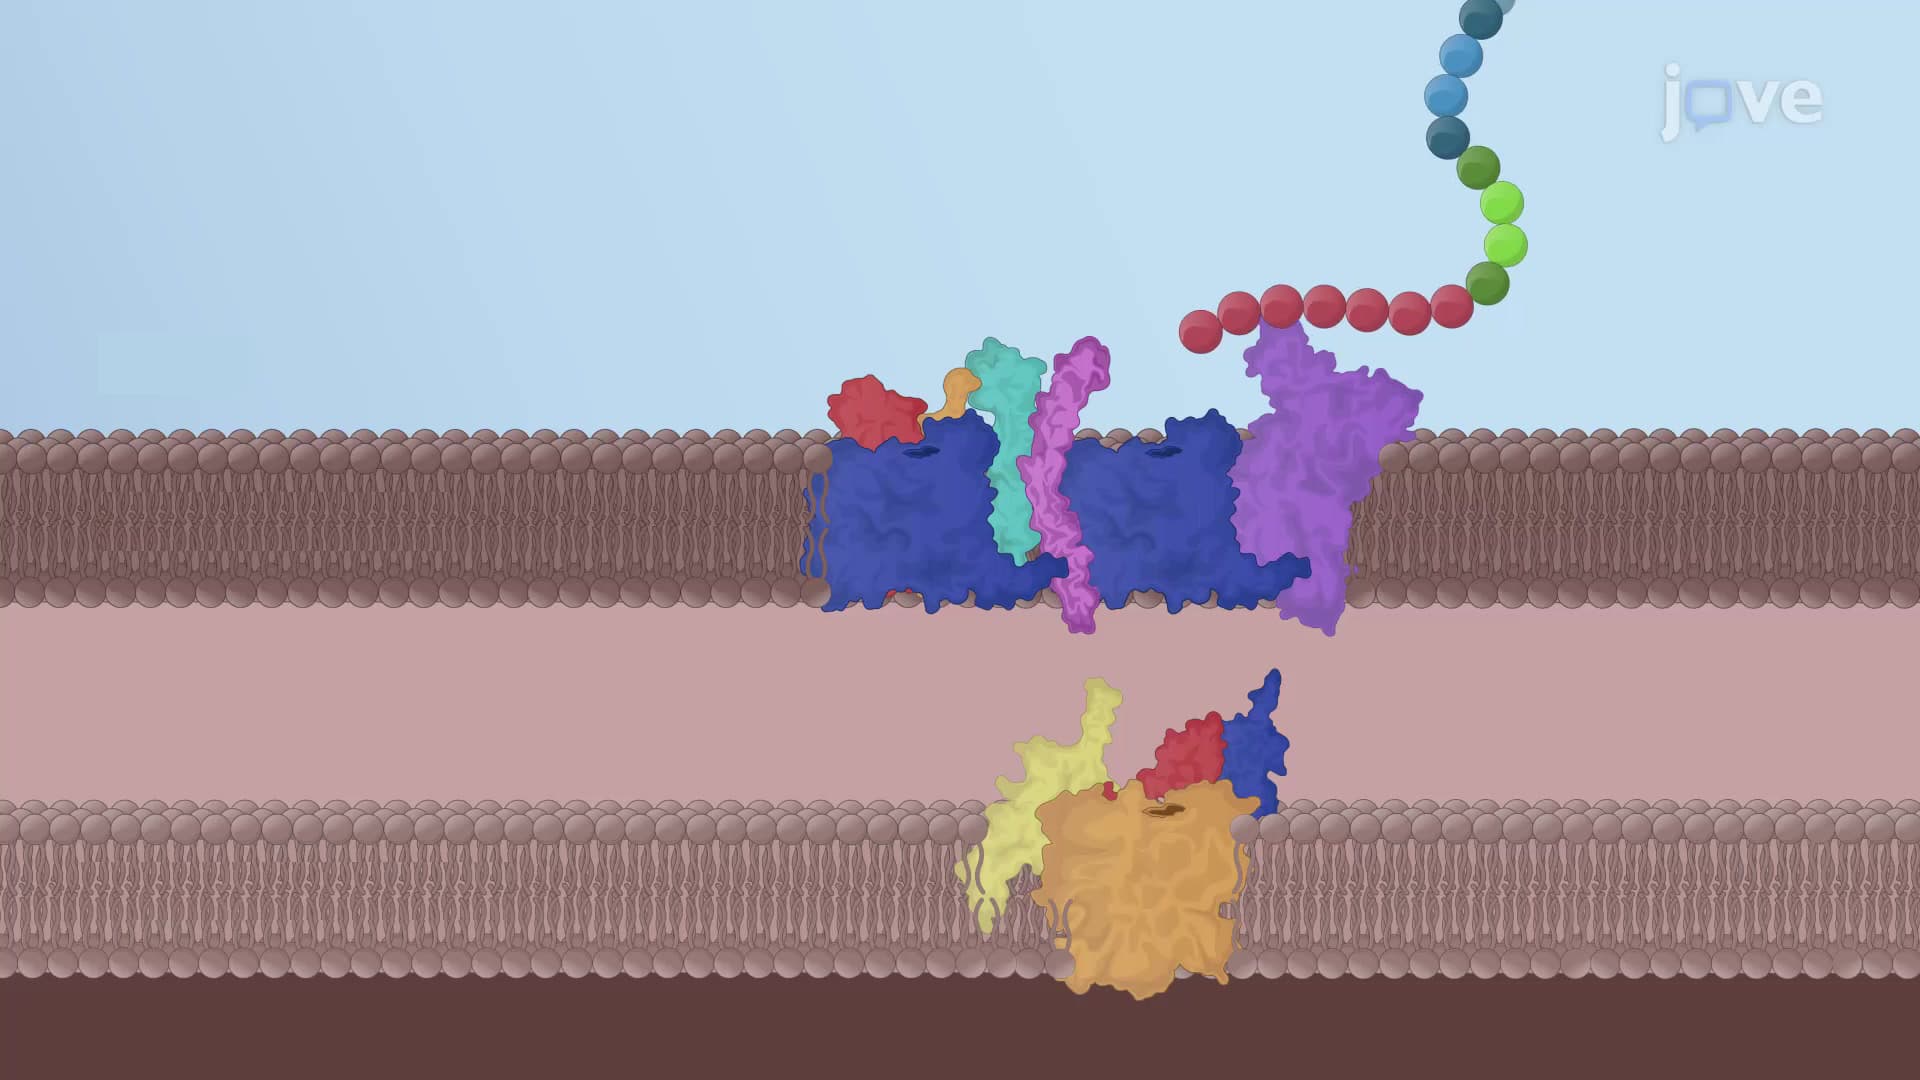
Mitochondrial Precursor Proteins
Most of the mitochondrial...
 01:39
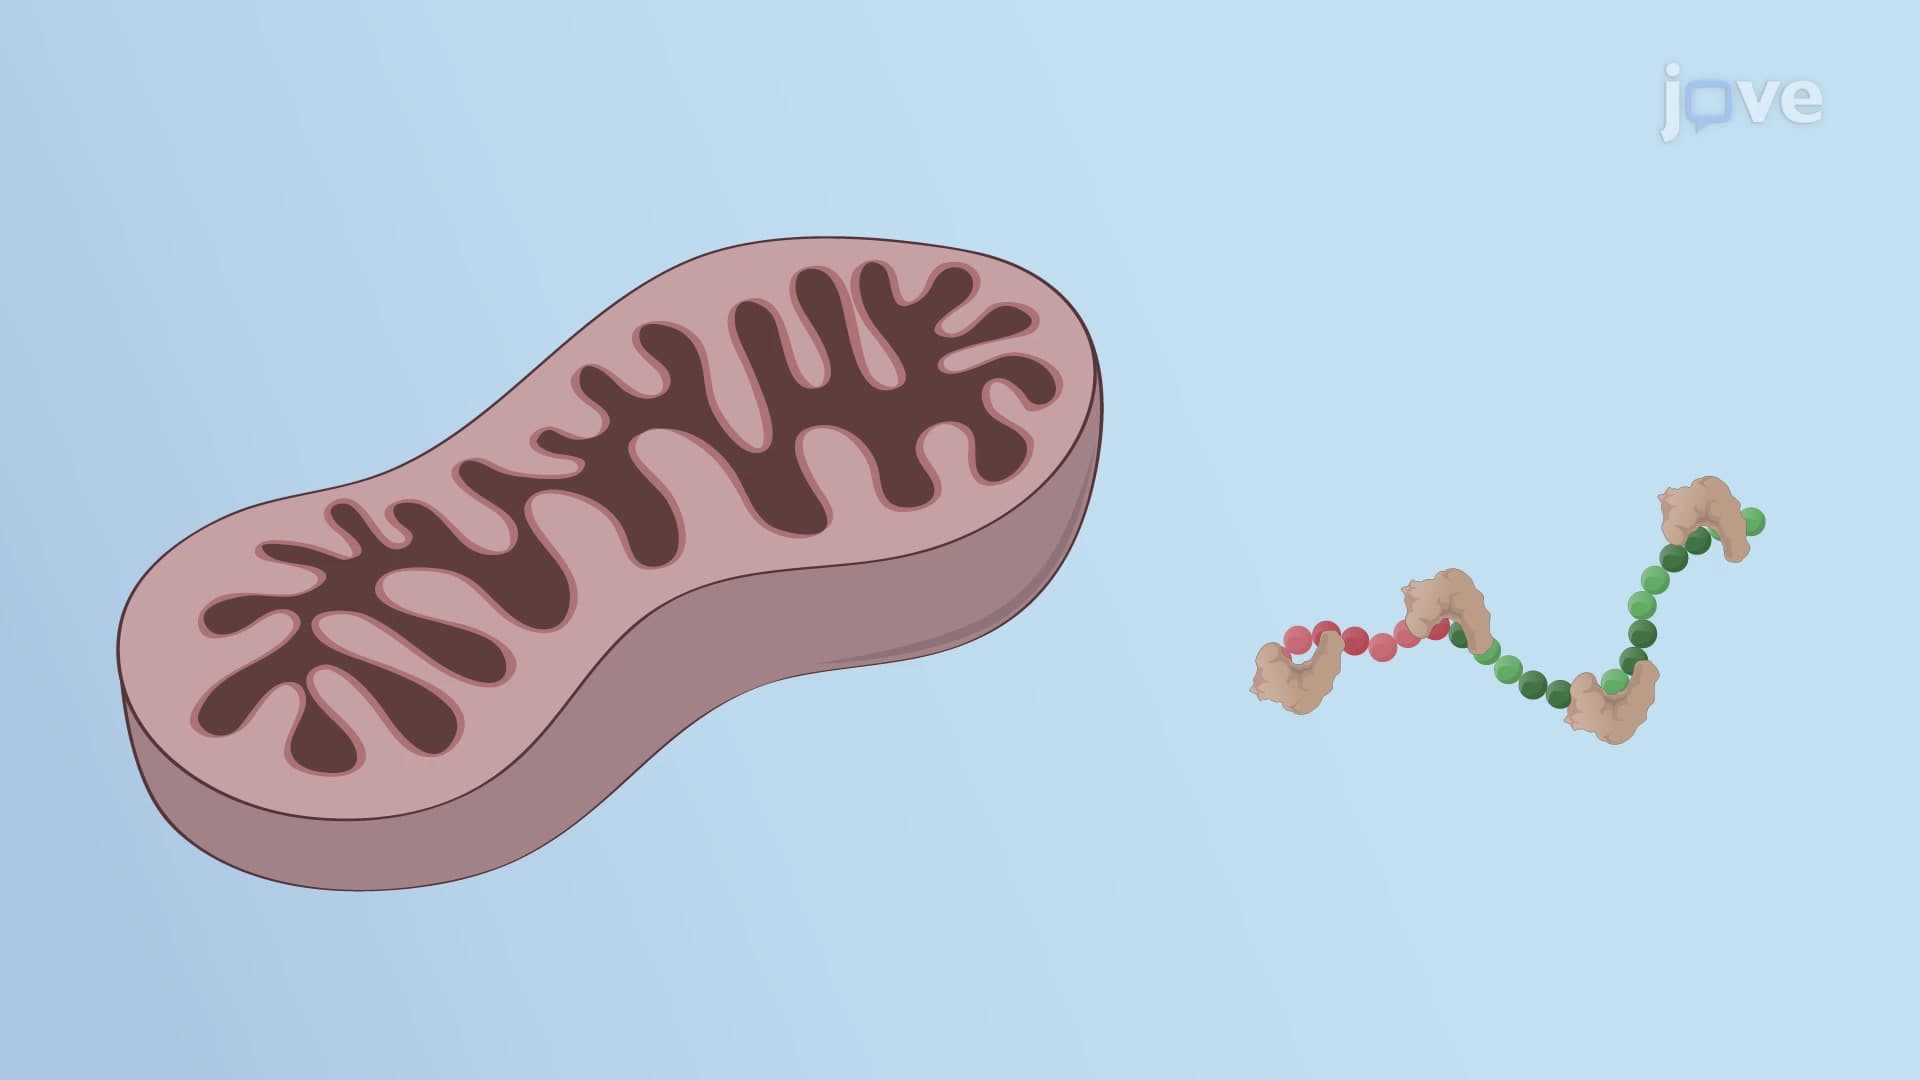
Mitochondrial Protein Sorting
Most of these mitochondrial proteins are encoded by the nucleus and imported to the mitochondria as unfolded or loosely folded precursors. Mitochondrial precursors...
 01:43
Electron Transport Chain: Complex III and IV
 01:45
Mitochondrial Membranes
01:45Mitochondrial Membranes