Related Experiment Video
Updated: Apr 17, 2026


Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Critical evaluation of the potential energy surface of the CH3 + HO2 reaction system
E P Faragó1, M Szőri1, M C Owen1
1Department of Chemical Informatics, Faculty of Education, University of Szeged, Boldogasszony sgt. 6, Szeged 6725, Hungary.

Abstract:
The CH3 + HO2 reaction system was studied theoretically by a newly developed, HEAT345-(Q) method based CHEAT1 protocol and includes the combined singlet and triplet potential energy surfaces. The main simplification is based on the CCSDT(Q)/cc-pVDZ calculation which is computationally inexpensive. Despite the economic and black-box treatment of higher excitations, the results are within 0.6 kcal/mol of the highly accurate literature values. Furthermore, the CHEAT1 surpassed the popular standard composite methods such as CBS-4M, CBS-QB3, CBS-APNO, G2, G3, G3MP2B3, G4, W1U, and W1BD mainly due to their poor performance in characterizing transition states (TS). For TS structures, various standard DFT and MP2 method have also been tested against the resulting CCSD/cc-pVTZ geometry of our protocol. A fairly good agreement was only found in the cases of the B2PLYP and BHandHLYP functionals, which were able to reproduce the structures of all TS studied within a maximum absolute deviation of 7%. The complex reaction mechanism was extended by three new low lying reaction channels. These are indirect water elimination from CH3OOH resulted formaldehyde, H2 elimination yielded methylene peroxide, and methanol and reactive triplet oxygen were formed via H-shift in the third channel. CHEAT1 protocol based on HEAT345-(Q) method is a robust, general, and cheap alternative for high accurate kinetic calculations.
More Related Videos
 05:51
Isotopic Effect in Double Proton Transfer Process of Porphycene Investigated by Enhanced QM/MM Method
Published on: July 19, 2019
 10:52
Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
Related Concept Videos
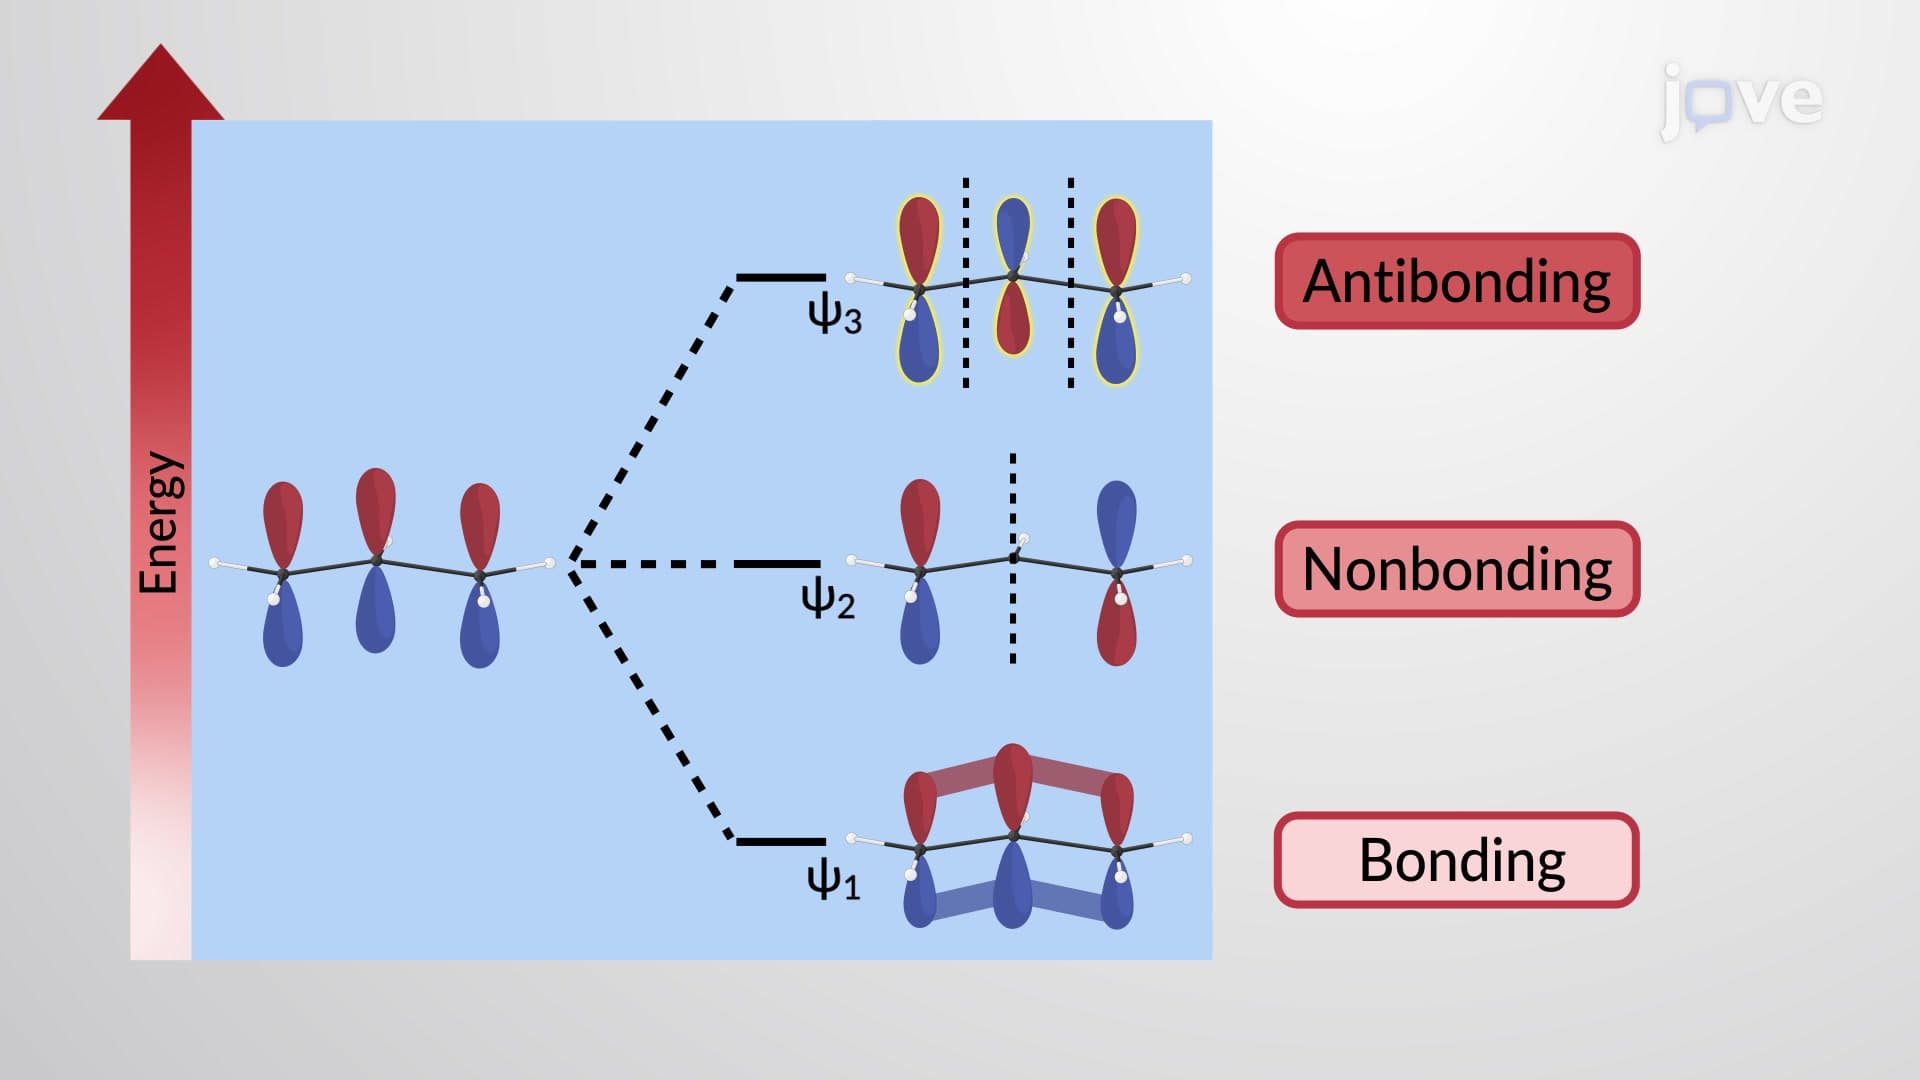 01:18
π Molecular Orbitals of the Allyl Cation and Anion
 02:36
Regioselectivity and Stereochemistry of Hydroboration
Hydroboration proceeds in a concerted fashion with the attack of borane on the π bond, giving a cyclic four-centered transition state. The –BH2 group is bonded to the less substituted carbon and –H to the more substituted carbon. The concerted nature requires the simultaneous addition of –H and –BH2 across the same face of the alkene giving syn...
 02:34
Regioselectivity and Stereochemistry of Acid-Catalyzed Hydration
 01:24
π Molecular Orbitals of 1,3-Butadiene
The simplest conjugated diene is 1,3-butadiene: a four-carbon system where each carbon is sp2-hybridized and has an unhybridized p orbital that contains an unpaired electron. According to molecular orbital theory, atomic orbitals combine to form molecular orbitals such that the...
 01:32
Radical Anti-Markovnikov Addition to Alkenes: Thermodynamics
 03:35
Hybridization of Atomic Orbitals II