Related Experiment Video
Updated: Apr 17, 2026


Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Analytic cubic and quartic force fields using density-functional theory
Magnus Ringholm1, Dan Jonsson1, Radovan Bast2
1Centre for Theoretical and Computational Chemistry (CTCC), Department of Chemistry, University of Tromsø-The Arctic University of Norway, 9037 Tromsø, Norway.

Abstract:
We present the first analytic implementation of cubic and quartic force constants at the level of Kohn-Sham density-functional theory. The implementation is based on an open-ended formalism for the evaluation of energy derivatives in an atomic-orbital basis. The implementation relies on the availability of open-ended codes for evaluation of one- and two-electron integrals differentiated with respect to nuclear displacements as well as automatic differentiation of the exchange-correlation kernels. We use generalized second-order vibrational perturbation theory to calculate the fundamental frequencies of methane, ethane, benzene, and aniline, comparing B3LYP, BLYP, and Hartree-Fock results. The Hartree-Fock anharmonic corrections agree well with the B3LYP corrections when calculated at the B3LYP geometry and from B3LYP normal coordinates, suggesting that the inclusion of electron correlation is not essential for the reliable calculation of cubic and quartic force constants.
More Related Videos
 10:52
Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
 08:04
Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
Related Concept Videos
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than...
 04:01
Real Gases: Effects of Intermolecular Forces and Molecular Volume Deriving Van der Waals Equation
 02:36
Molecular Geometry and Dipole Moments
 02:40
MO Theory and Covalent Bonding
 01:10
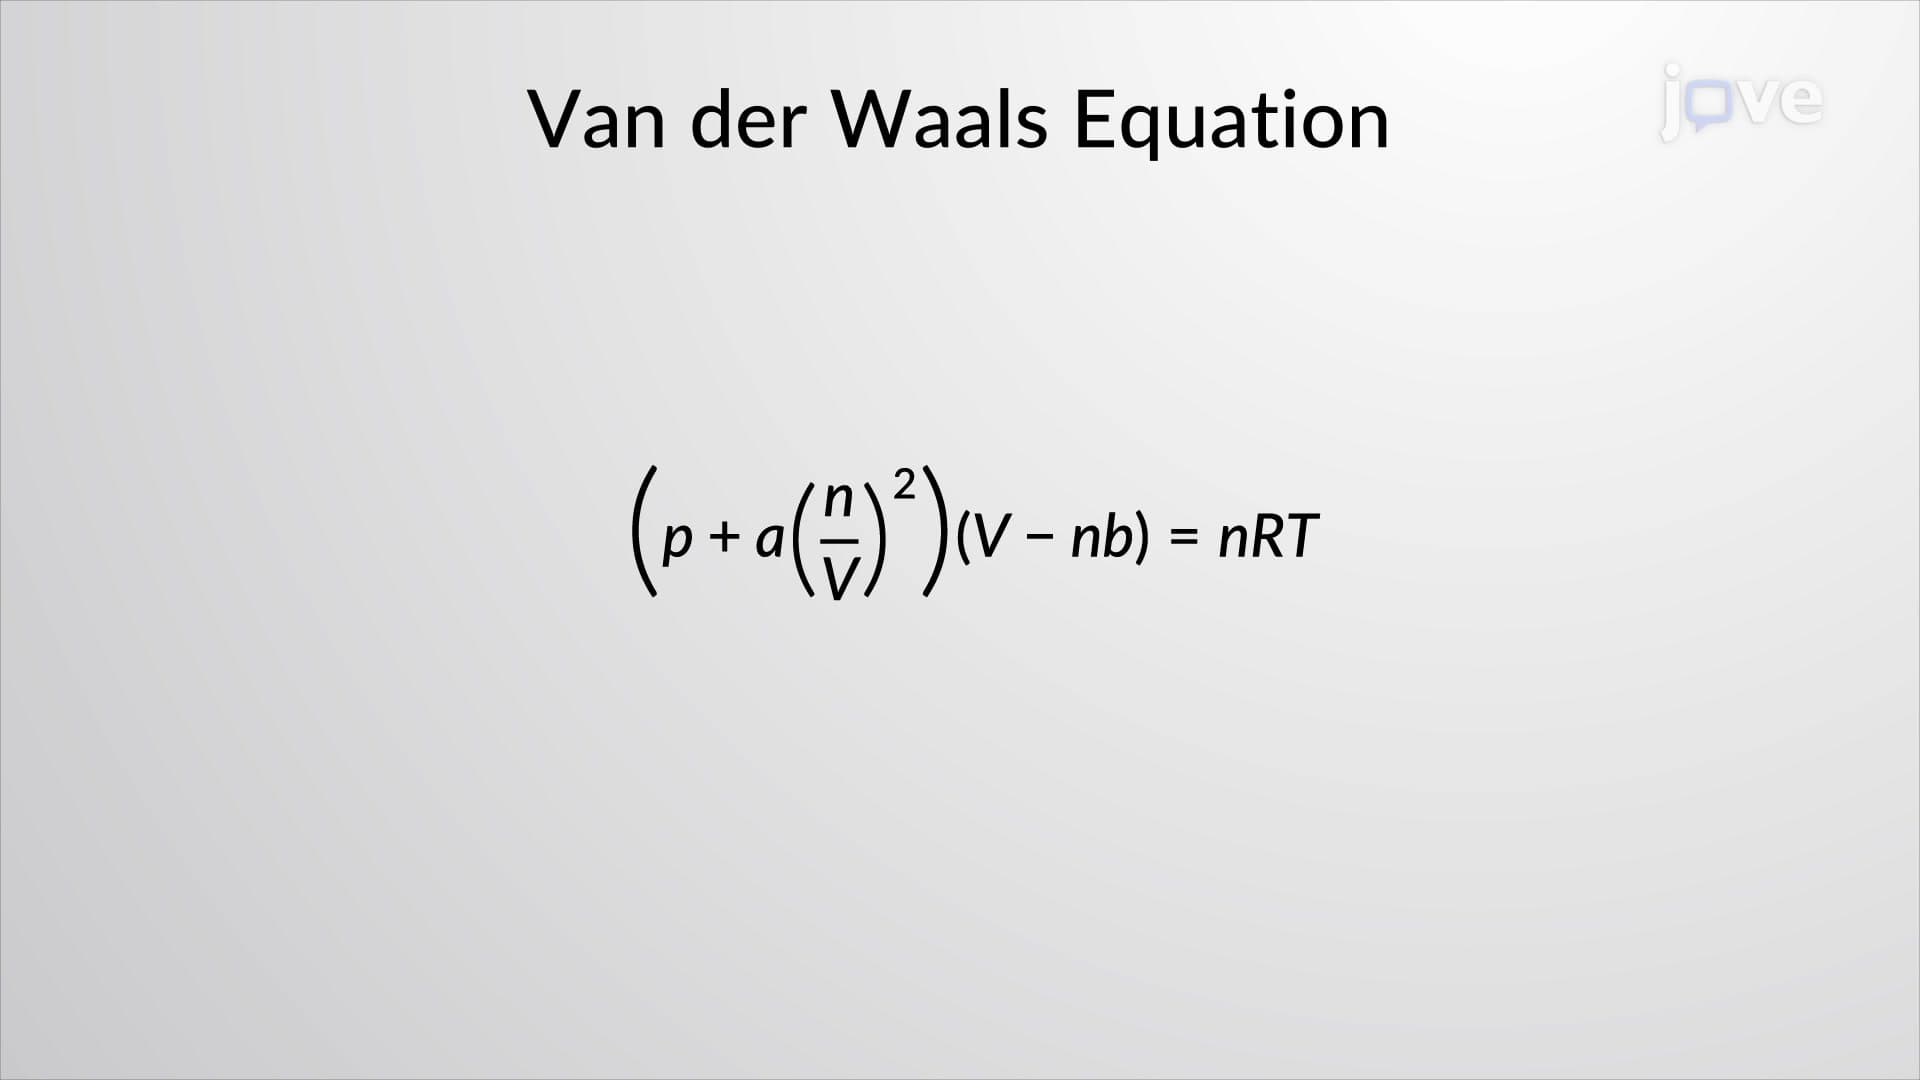
Van der Waals Equation
First, the attractive forces between molecules, which are stronger at higher densities and reduce the pressure, are considered by adding to the pressure a term equal to the square of the molar density multiplied by a positive coefficient a. Second, the...