Related Experiment Video
Updated: Apr 17, 2026

Using Microwave and Macroscopic Samples of Dielectric Solids to Study the Photonic Properties of Disordered Photonic Bandgap Materials
Published on: September 26, 2014
Theoretical estimation of the optical bandgap in a series of poly(aryl-ethynylene)s: a DFT study
José M Granadino-Roldán1, Andrés Garzón2, Mónica Moral1
1Departamento de Química Física y Analítica, Facultad de Ciencias Experimentales, Universidad de Jaén, Paraje las Lagunillas, s/n. 23071, Jaén, Spain.

Abstract:
Aimed to optimize the ratio accuracy/computational cost, in this work we study the performance of three different theoretical methodologies in the calculation of the optical bandgap for a test set made of a number of poly(aryl-ethynylene)s related polymers. Infinite, ideal polymer chains were first optimized by means of periodic calculations. Different length oligomers were afterward generated by direct replication of the corresponding periodic structure and their optical bandgaps were calculated by means of different time dependent-density functional theory (TD-DFT) methodologies. These results were fitted to an exponential function for each oligomer family in order to get a theoretical estimation of the optical bandgap for each polymer to be compared to the experimental reported values. The best result was obtained for TD-M06-2X yielding an average deviation of 3.4% with respect to the experimental values.

Related Concept Videos
 01:29
UV–Vis Spectroscopy: Woodward–Fieser Rules
 01:32
UV–Vis Spectroscopy of Conjugated Systems
One of the factors influencing λmax is...
 01:16
UV–Vis Spectroscopy: Molecular Electronic Transitions
 01:29
IR and UV–Vis Spectroscopy of Aldehydes and Ketones
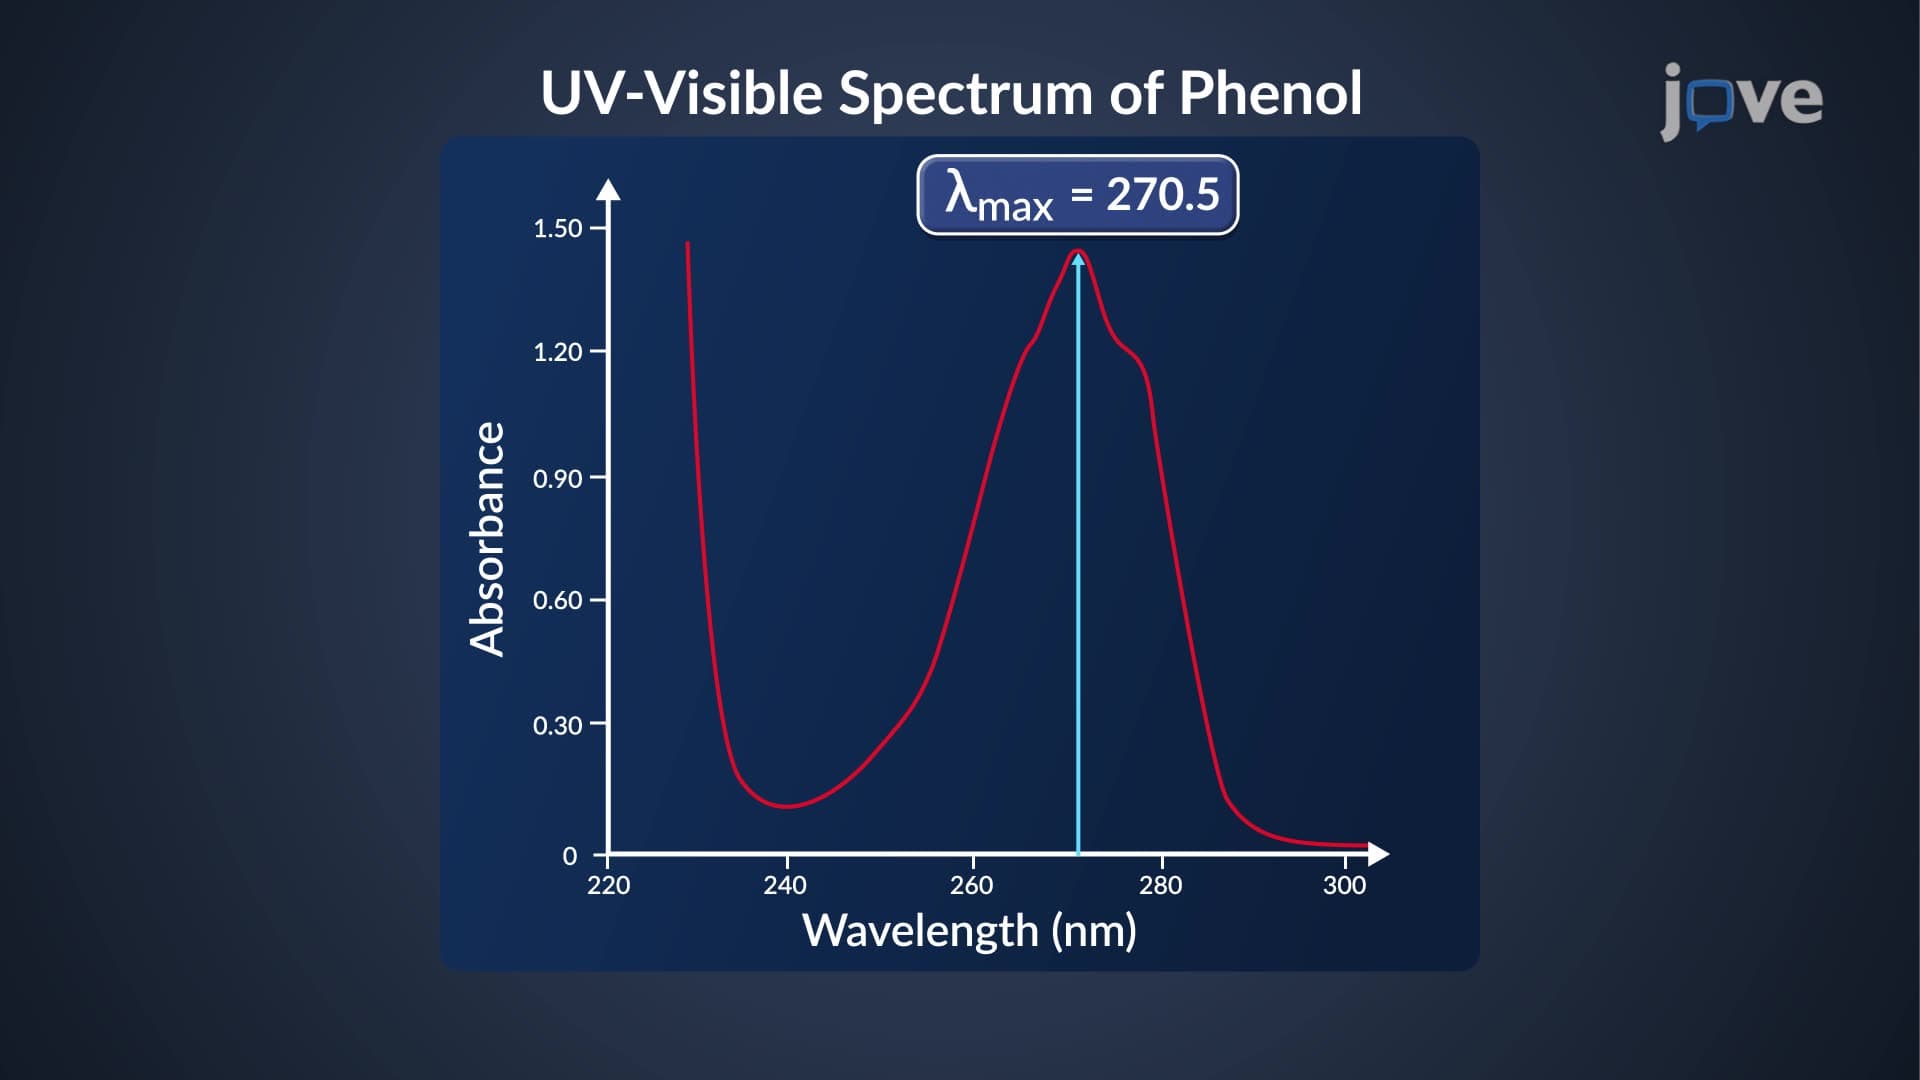 01:30
UV–Vis Spectrum
The UV–Vis spectrum of a molecule is the plot of its absorbance versus wavelength. The plot is drawn by taking molar...
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than...