Related Experiment Video
Updated: Apr 17, 2026
Nanomanipulation of Single RNA Molecules by Optical Tweezers
Published on: August 20, 2014
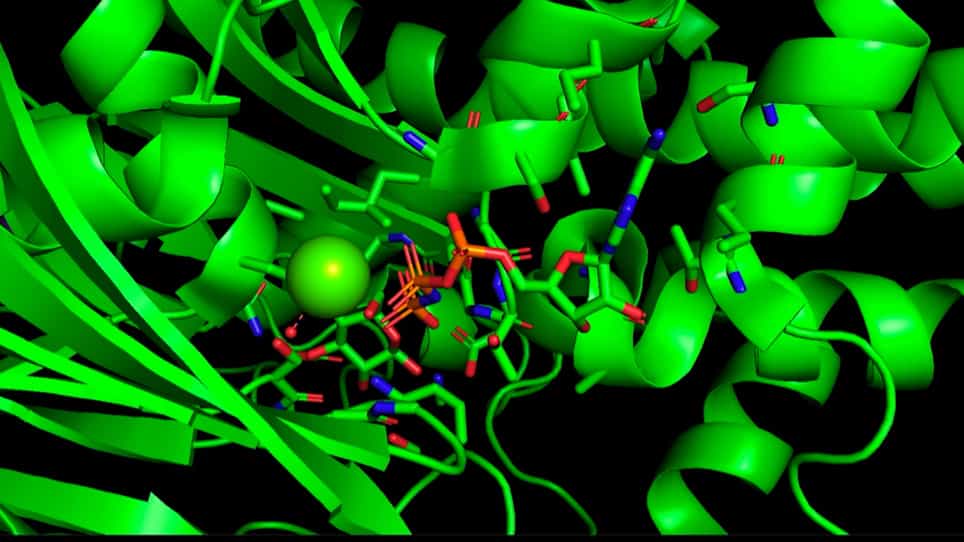
Reactive conformation of the active site in the hairpin ribozyme achieved by molecular dynamics simulations with ε/ζ
Vojtěch Mlýnský1, Petra Kührová1, Marie Zgarbová1
1†Regional Centre of Advanced Technologies and Materials, Department of Physical Chemistry, Faculty of Science, Palacky University, tr. 17 listopadu 12, 771 46 Olomouc, Czech Republic.

Abstract:
X-ray crystallography can provide important insights into the structure of RNA enzymes (ribozymes). However, the details of a ribozyme's active site architecture are often altered by the inactivating chemical modifications necessary to inhibit self-cleavage. Molecular dynamics (MD) simulations are able to complement crystallographic data and model the conformation of the ribozyme's active site in its native form. However, the performance of MD simulations is driven by the quality of the force field used. Force fields are primarily parametrized and tested for a description of canonical structures and thus may be less accurate for noncanonical RNA elements, including ribozyme catalytic cores. Here, we show that our recent reparametrization of ε/ζ torsions significantly improves the description of the hairpin ribozyme's scissile phosphate conformational behavior. In addition, we find that an imbalance in the force field description of the nonbonded interactions of the ribose 2'-OH contributes to the conformational behavior observed for the scissile phosphate in the presence of a deprotonated G8(-). On the basis of the new force field, we obtain a reactive conformation for the hairpin ribozyme active site that is consistent with the most recent mechanistic and structural data.
Related Concept Videos
 02:47
Ribozymes
Ribozymes can...
02:47Ribozymes
 01:13
Induced-fit Model
Enzymes exhibit substrate specificity, meaning that they can only bind to certain substrates. This is mainly determined by the shape and chemical...
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally...
 01:13
Introduction to Mechanisms of Enzyme Catalysis