Related Experiment Video
Updated: Apr 16, 2026


Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
"Plug and play" full-dimensional ab initio potential energy and dipole moment surfaces and anharmonic vibrational
Chen Qu1, Riccardo Conte, Paul L Houston
1Department of Chemistry and Cherry L. Emerson Centrer for Scientific Computations, Emory University, Atlanta, GA 30322, USA. jmbowma@emory.edu.

Abstract:
The potential energy surface of the methane-water dimer is represented as the sum of a new intrinsic two-body potential energy surface and pre-existing intramolecular potentials for the monomers. Different fits of the CH4-H2O intrinsic two-body energy are reported. All these fits are based on 30 467 ab initio interaction energies computed at CCSD(T)-F12b/haTZ (aug-cc-pVTZ for C and O, cc-pVTZ for H) level of theory. The benchmark fit is a full-dimensional, permutationally-invariant analytical representation with root-mean-square (rms) fitting error of 3.5 cm(-1). Two other computationally more efficient two-body potentials are also reported, albeit with larger rms fitting errors. Of these a compact permutationally invariant fit is shown to be the best one in combining precision and speed of evaluation. An intrinsic two-body dipole moment surface is also obtained, based on MP2/haTZ expectation values, with an rms fitting error of 0.002 au. As with the potential, this dipole moment surface is combined with existing monomer ones to obtain the full surface. The vibrational ground state of the dimer and dissociation energy, D0, are determined by diffusion Monte Carlo calculations, and MULTIMODE calculations are performed for the IR spectrum of the intramolecular modes. The relative accuracy of the different intrinsic two-body potentials is analyzed by comparing the energetics and the harmonic frequencies of the global minimum well, and the maximum impact parameter employed in a sample methane-water scattering calculation.
More Related Videos
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
 10:52
Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
Related Concept Videos
 02:36
Molecular Geometry and Dipole Moments
 03:24
Hybridization of Atomic Orbitals I
 01:24
π Molecular Orbitals of 1,3-Butadiene
The simplest conjugated diene is 1,3-butadiene: a four-carbon system where each carbon is sp2-hybridized and has an unhybridized p orbital that contains an unpaired electron. According to molecular orbital theory, atomic orbitals combine to form molecular orbitals such that the...
 01:18
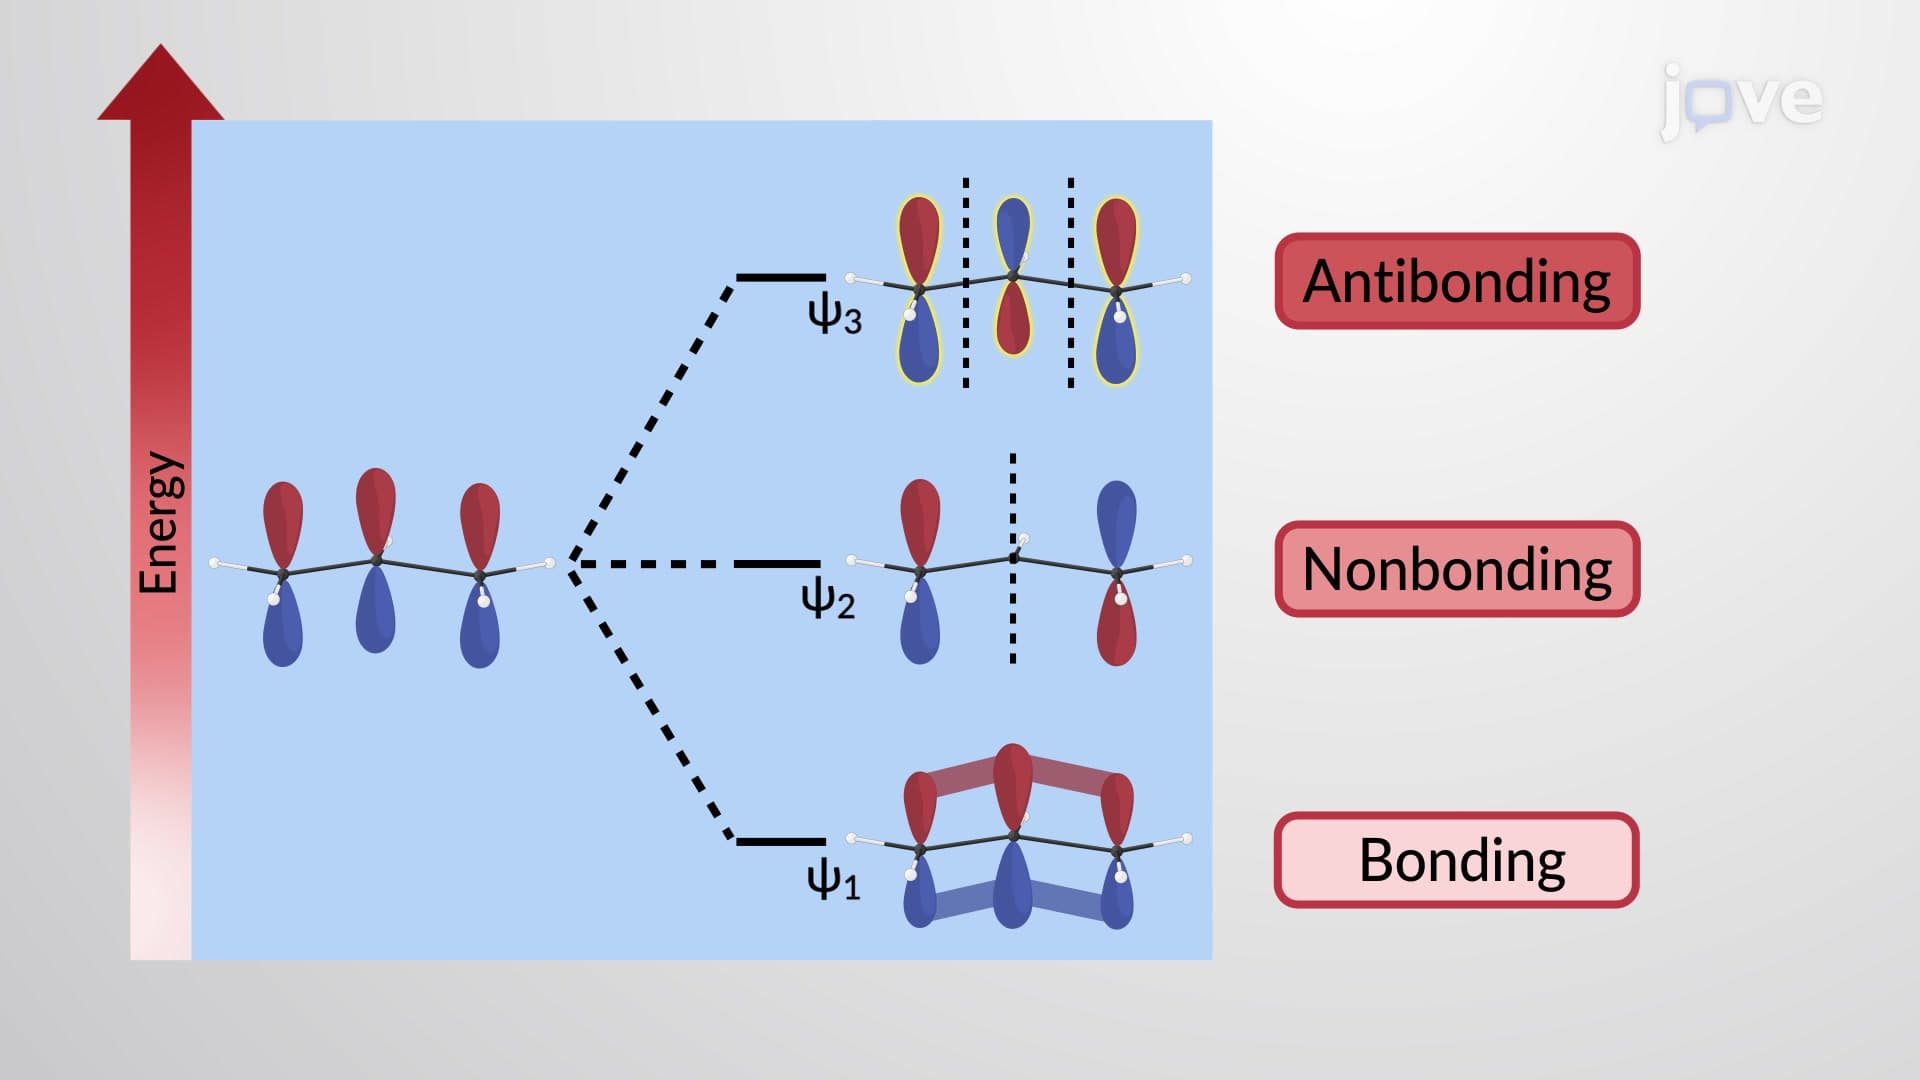
π Molecular Orbitals of the Allyl Cation and Anion
 02:52
VSEPR Theory and the Basic Shapes
 02:37
VSEPR Theory