Electronic spectra and DFT calculations of some pyrimido[1,2-a]benzimidazole derivatives

Mohamed E Elshakre1, H Moustafa1, Huwaida M E Hassaneen1

  • 1Chemistry Department, College of Science, Cairo University, University Ave., Dokki, 12613 Cairo, Egypt.

Related Concept Videos

NMR Spectroscopy of Benzene Derivatives01:37

NMR Spectroscopy of Benzene Derivatives

Simple unsubstituted benzene has six aromatic protons, all chemically equivalent. Therefore, benzene exhibits only a singlet peak at δ 7.3 ppm in the 1H NMR spectrum. The observed shift is far downfield because the aromatic ring current strongly deshields the protons. Any substitution on the benzene ring makes the aromatic protons nonequivalent, and the protons split each other. The peak is, therefore, no longer a singlet and the splitting pattern and their associated coupling...
12.4K
π Electron Effects on Chemical Shift: Aromatic and Antiaromatic Compounds01:14

π Electron Effects on Chemical Shift: Aromatic and Antiaromatic Compounds
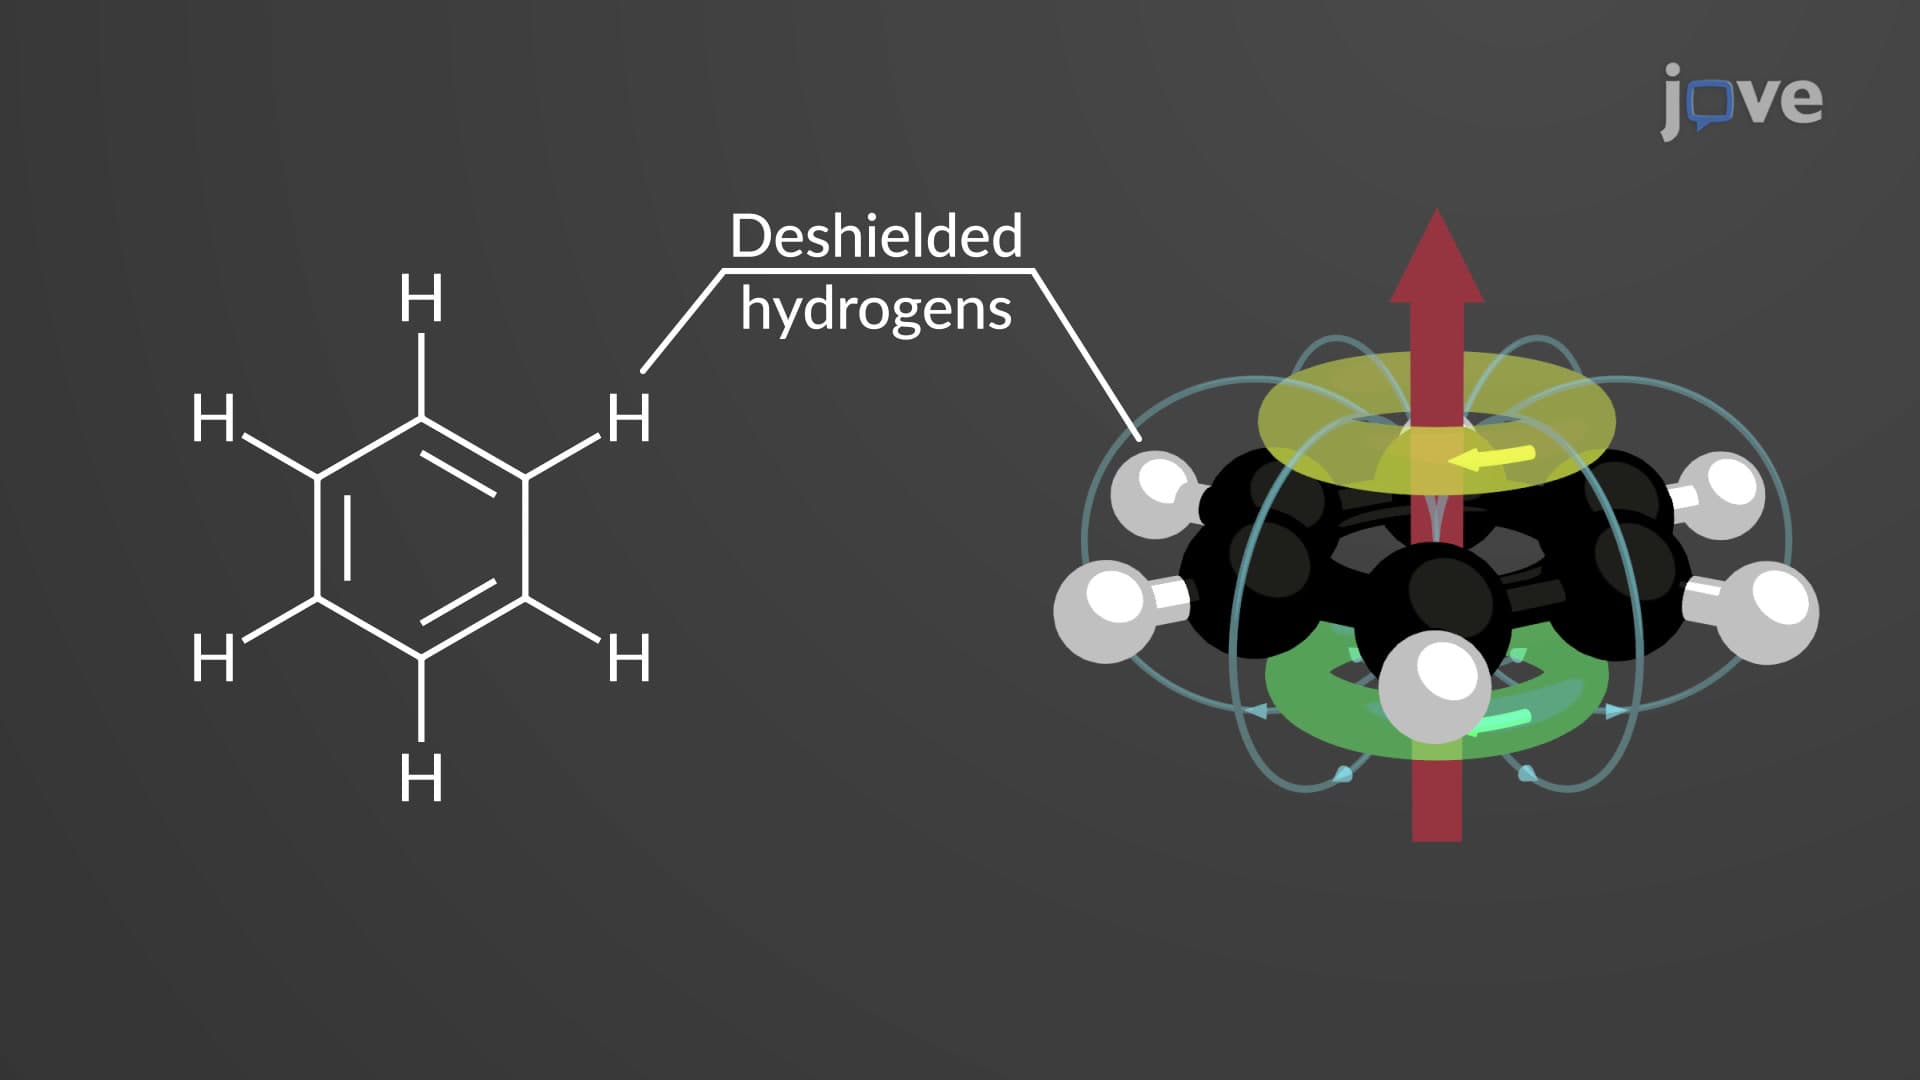
In aromatic compounds, such as benzene, the circulation of (4n + 2) π-electrons sets up a diamagnetic or diatropic ring current around the perimeter of the molecule. This current induces a magnetic field that opposes the external field inside the ring and reinforces it on the outside. The protons in benzene are deshielded and exhibit high chemical shifts in the range 6.5–8.5 ppm. The shielding effect at the center of the ring is evident in complex aromatic molecules, such as...
2.1K
UV–Vis Spectroscopy: Woodward–Fieser Rules01:29

UV–Vis Spectroscopy: Woodward–Fieser Rules
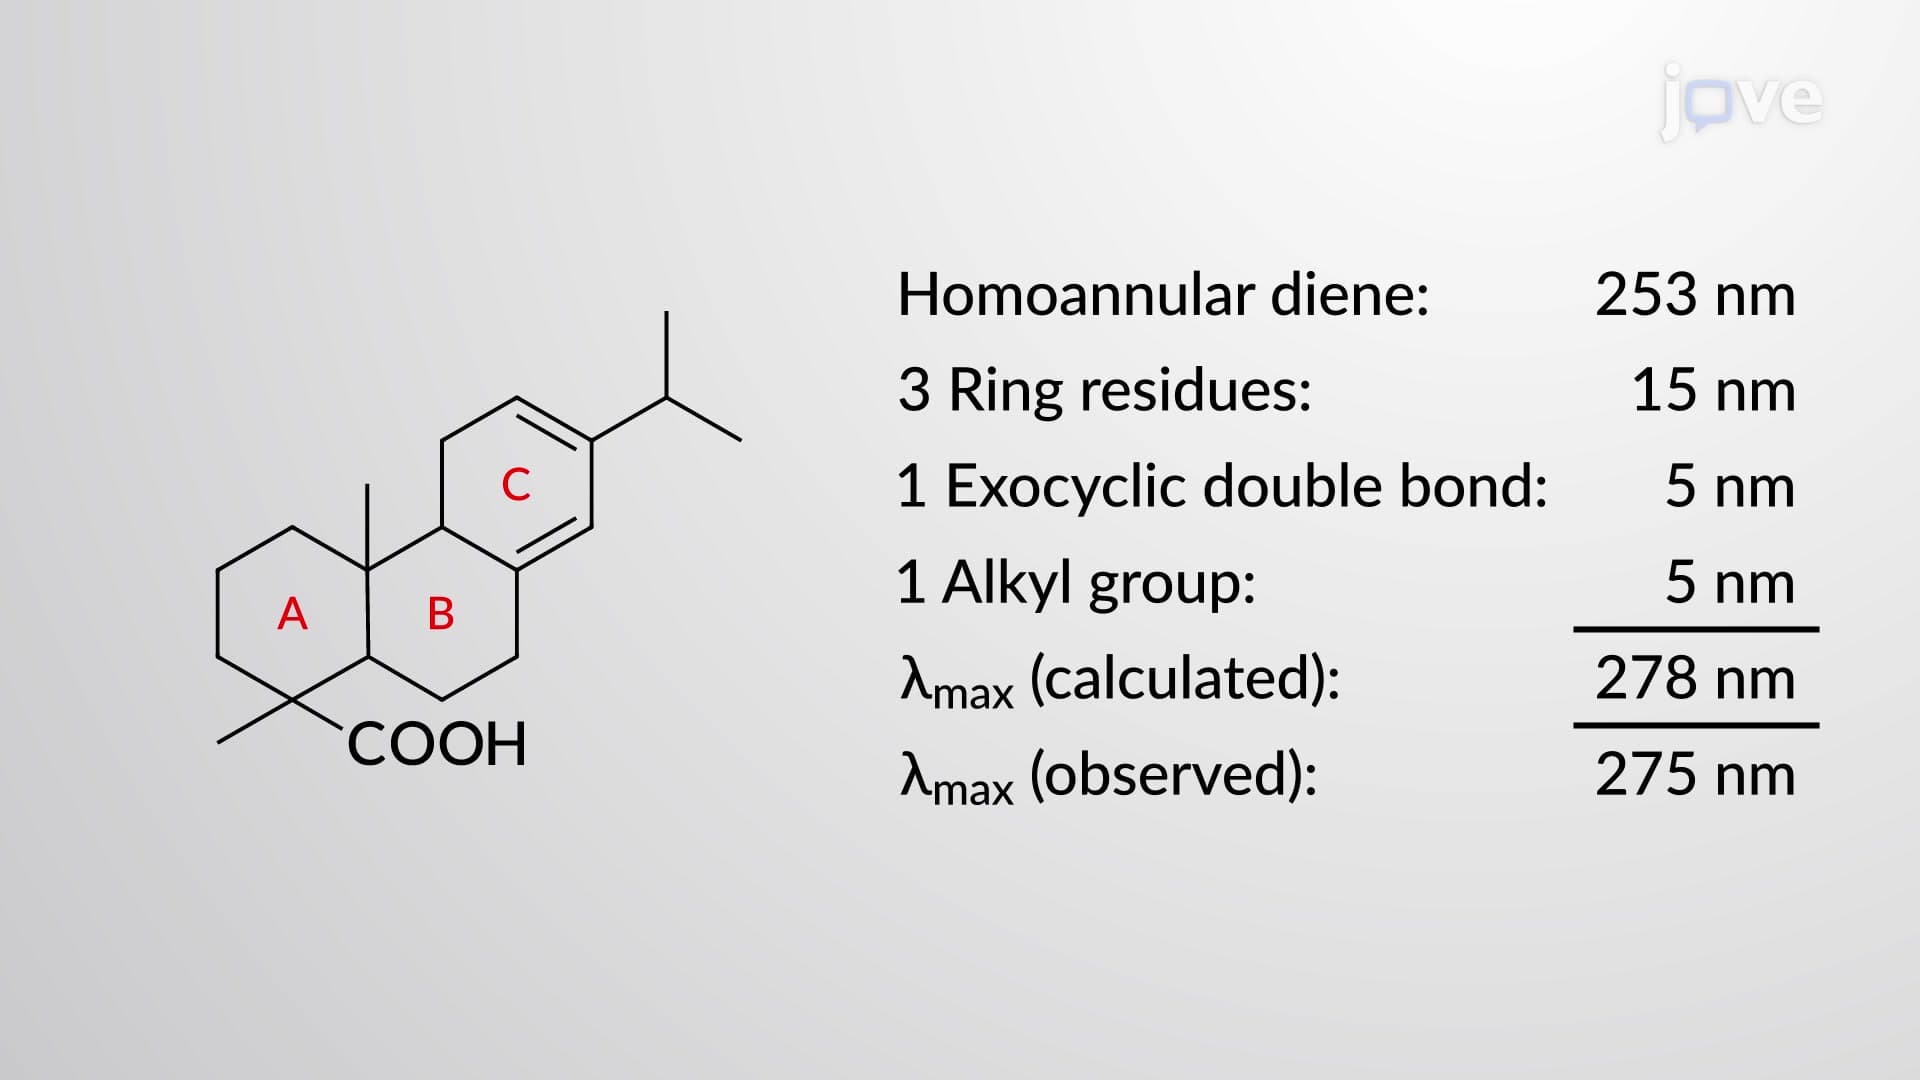
UV–Visible absorption spectra of conjugated dienes arise from the lowest energy π → π* transitions. The light-absorbing part of the molecule is called the chromophore, and the substituents directly attached to the chromophore are called auxochromes. A strong correlation exists between the absorption maxima, λmax, and the structure of a conjugated π system. The Woodward–Fieser rules predict the value of λmax for a...
30.0K
¹H NMR Signal Integration: Overview00:58

¹H NMR Signal Integration: Overview

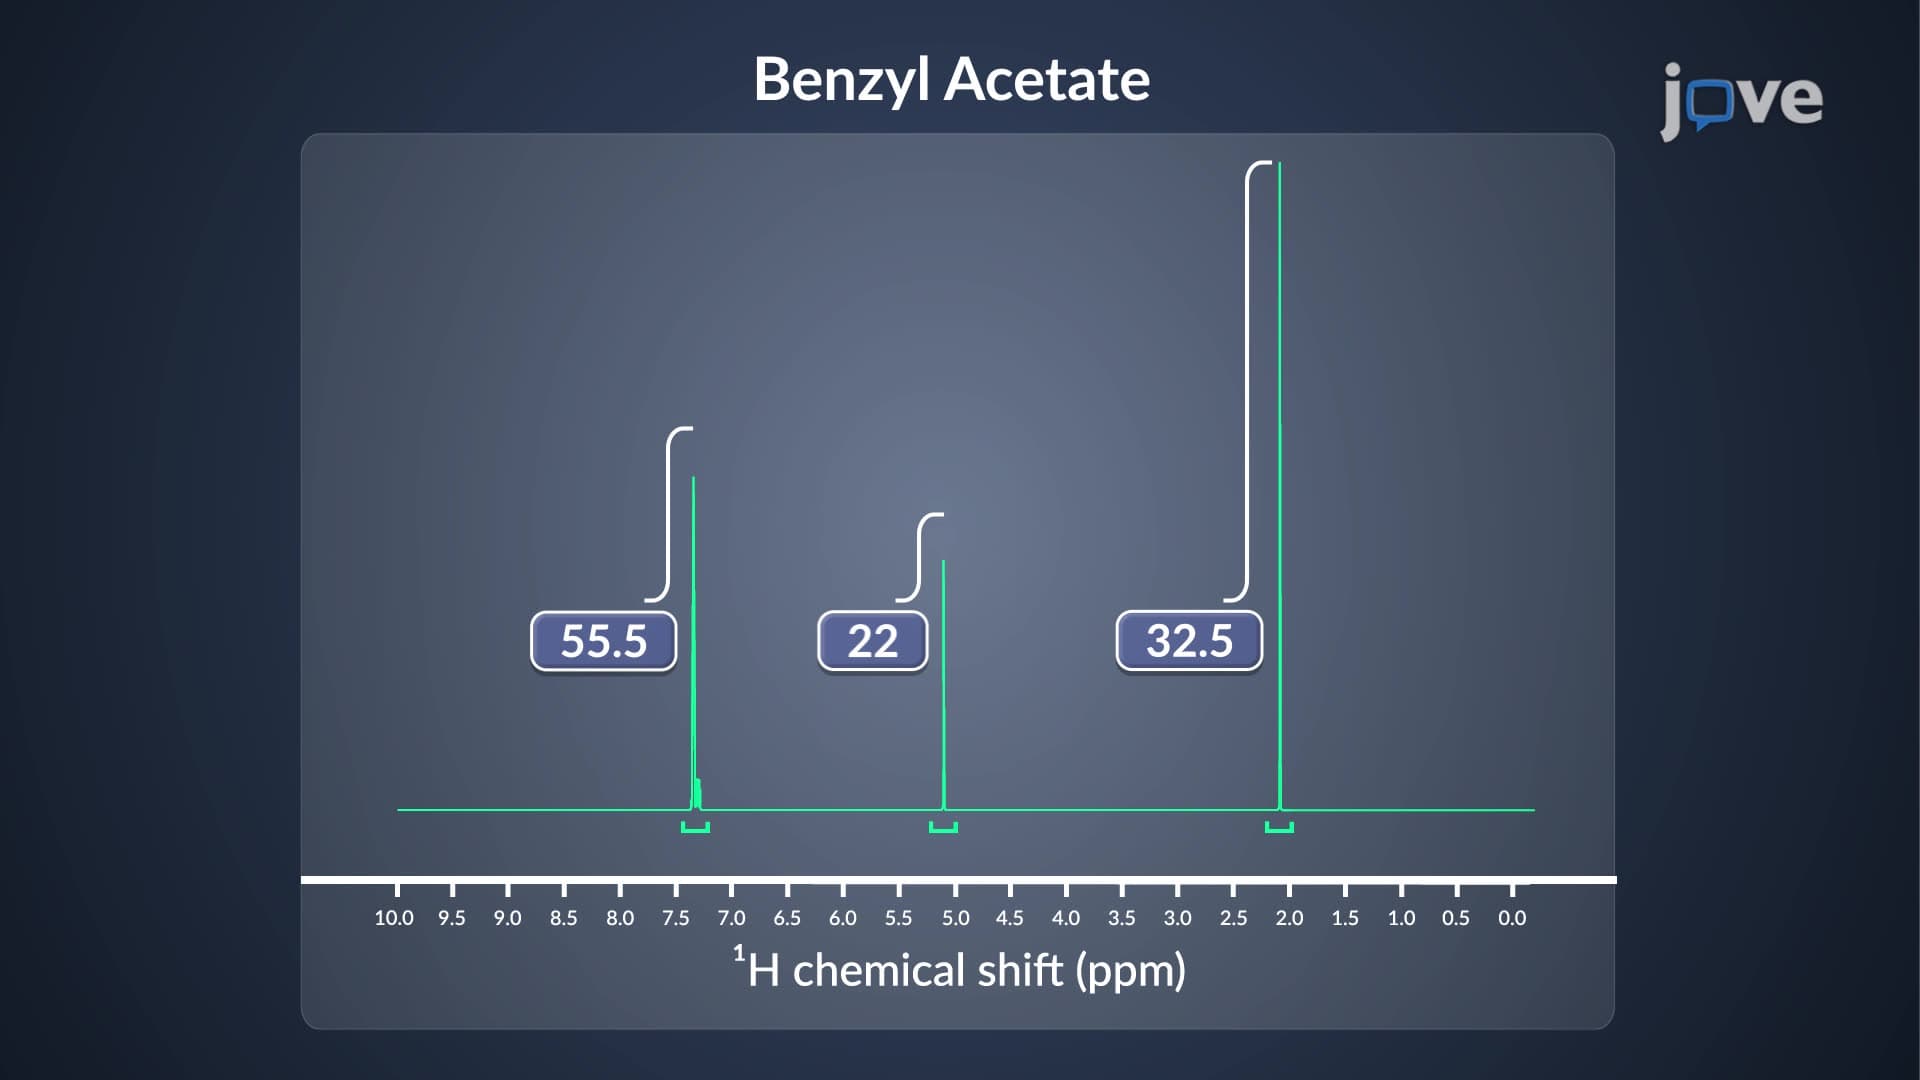
The intensity of a signal, which can be represented by the area under the peak, depends on the number of protons contributing to that signal. The area under each peak is shown as a vertical line called an integral, with the integral value listed under it, as seen in the proton NMR spectrum of benzyl acetate. Each integral value is divided by the smallest integral value to obtain the ratio of the number of protons producing each signal. The ratio reveals the relative number of protons and not...
4.5K
IR Spectrum Peak Splitting: Symmetric vs Asymmetric Vibrations01:08

IR Spectrum Peak Splitting: Symmetric vs Asymmetric Vibrations
Identical bonds within a polyatomic group can stretch symmetrically (in-phase) or asymmetrically (out-of-phase). Similar to hydrogen bonding, these vibrations also influence the shape of the IR peak. Generally, asymmetric stretching frequencies are higher than symmetric stretching frequencies. For example, primary amines exhibit two distinct IR peaks between 3300–3500 cm−1 corresponding to the symmetric and asymmetric N-H stretching, while secondary amines exhibit a single...
2.3K
NMR Spectroscopy of Aromatic Compounds01:14

NMR Spectroscopy of Aromatic Compounds

Aromatic compounds can be identified or analyzed using proton NMR and carbon‐13 NMR. Typically, aromatic hydrogens or hydrogens directly bonded to the aromatic rings are strongly deshielded by the aromatic ring current. Therefore, they absorb in the range of 6.5–8.0 ppm in proton NMR spectra. For instance, aromatic hydrogens directly bonded to the benzene ring absorb at 7.3 ppm. However, aromatic hydrogens of larger rings absorb farther upfield or downfield than the ideal range.
6.8K