Related Experiment Video
Updated: Apr 15, 2026

Use of Rabbit Eyes in Pharmacokinetic Studies of Intraocular Drugs
Published on: July 23, 2016
Population pharmacokinetic analysis for a single 1,200-milligram dose of oritavancin using data from two pivotal
C M Rubino1, S M Bhavnani2, G Moeck3
1Institute for Clinical Pharmacodynamics, Latham, New York, USA CRubino@ICPD.com.

Abstract:
Oritavancin is a lipoglycopeptide antibiotic with activity against Gram-positive bacteria. Here we describe oritavancin population pharmacokinetics and the impact of patient-specific covariates on drug exposure variability. Concentration-time data were analyzed from two phase 3 clinical trials, SOLO I and SOLO II, in which oritavancin was administered as a single 1,200-mg dose to patients with acute bacterial skin and skin structure infections. A total of 1,337 drug concentrations from 297 patients (90% of whom had 4 or 5 pharmacokinetic samples) were available for analysis. A previously derived population model based on data from 12 phase 1, 2, and 3 oritavancin studies was applied to the SOLO data set. Alterations to the structural model were made, as necessary, based on model fit. Analyses utilized Monte Carlo parametric expectation maximization (S-ADAPT 1.5.6). The previous population pharmacokinetic model fit the data well (r(2) = 0.972), and population pharmacokinetic parameters were estimated with acceptable precision and lack of bias. Covariate evaluations revealed statistically significant relationships between central compartment volume and age and between clearance and height; however, these relationships did not indicate a clinically relevant impact on oritavancin exposure over the range of age and height observed in the SOLO studies. The mean (coefficient of variation [CV]) area under the plasma concentration-time curve from time zero to 72 h (AUC0-72) and maximum plasma concentration (Cmax) were 1,530 (36.9%) μg · h/ml and 138 (23%) μg/ml, respectively. The mean (CV) half-life at alpha phase (t1/2α), t1/2β, and t1/2γ were 2.29 (49.8%), 13.4 (10.5%), and 245 (14.9%) hours, respectively. These analyses are the first to describe oritavancin pharmacokinetics following a single 1,200-mg dose. Covariate analyses suggested that no dose adjustments are required for renal impairment (creatinine clearance, >29 ml/min), mild or moderate hepatic impairment, age, weight, gender, or diabetes status.
More Related Videos
 06:18
A Novel Method to Determine the Longitudinal Antibacterial Activity of Drug-Eluting Materials
Published on: March 3, 2023
 09:17
A Robust Pneumonia Model in Immunocompetent Rodents to Evaluate Antibacterial Efficacy against S. pneumoniae, H. influenzae, K. pneumoniae, P. aeruginosa or A. baumannii
Published on: January 2, 2017
Related Concept Videos
 01:15
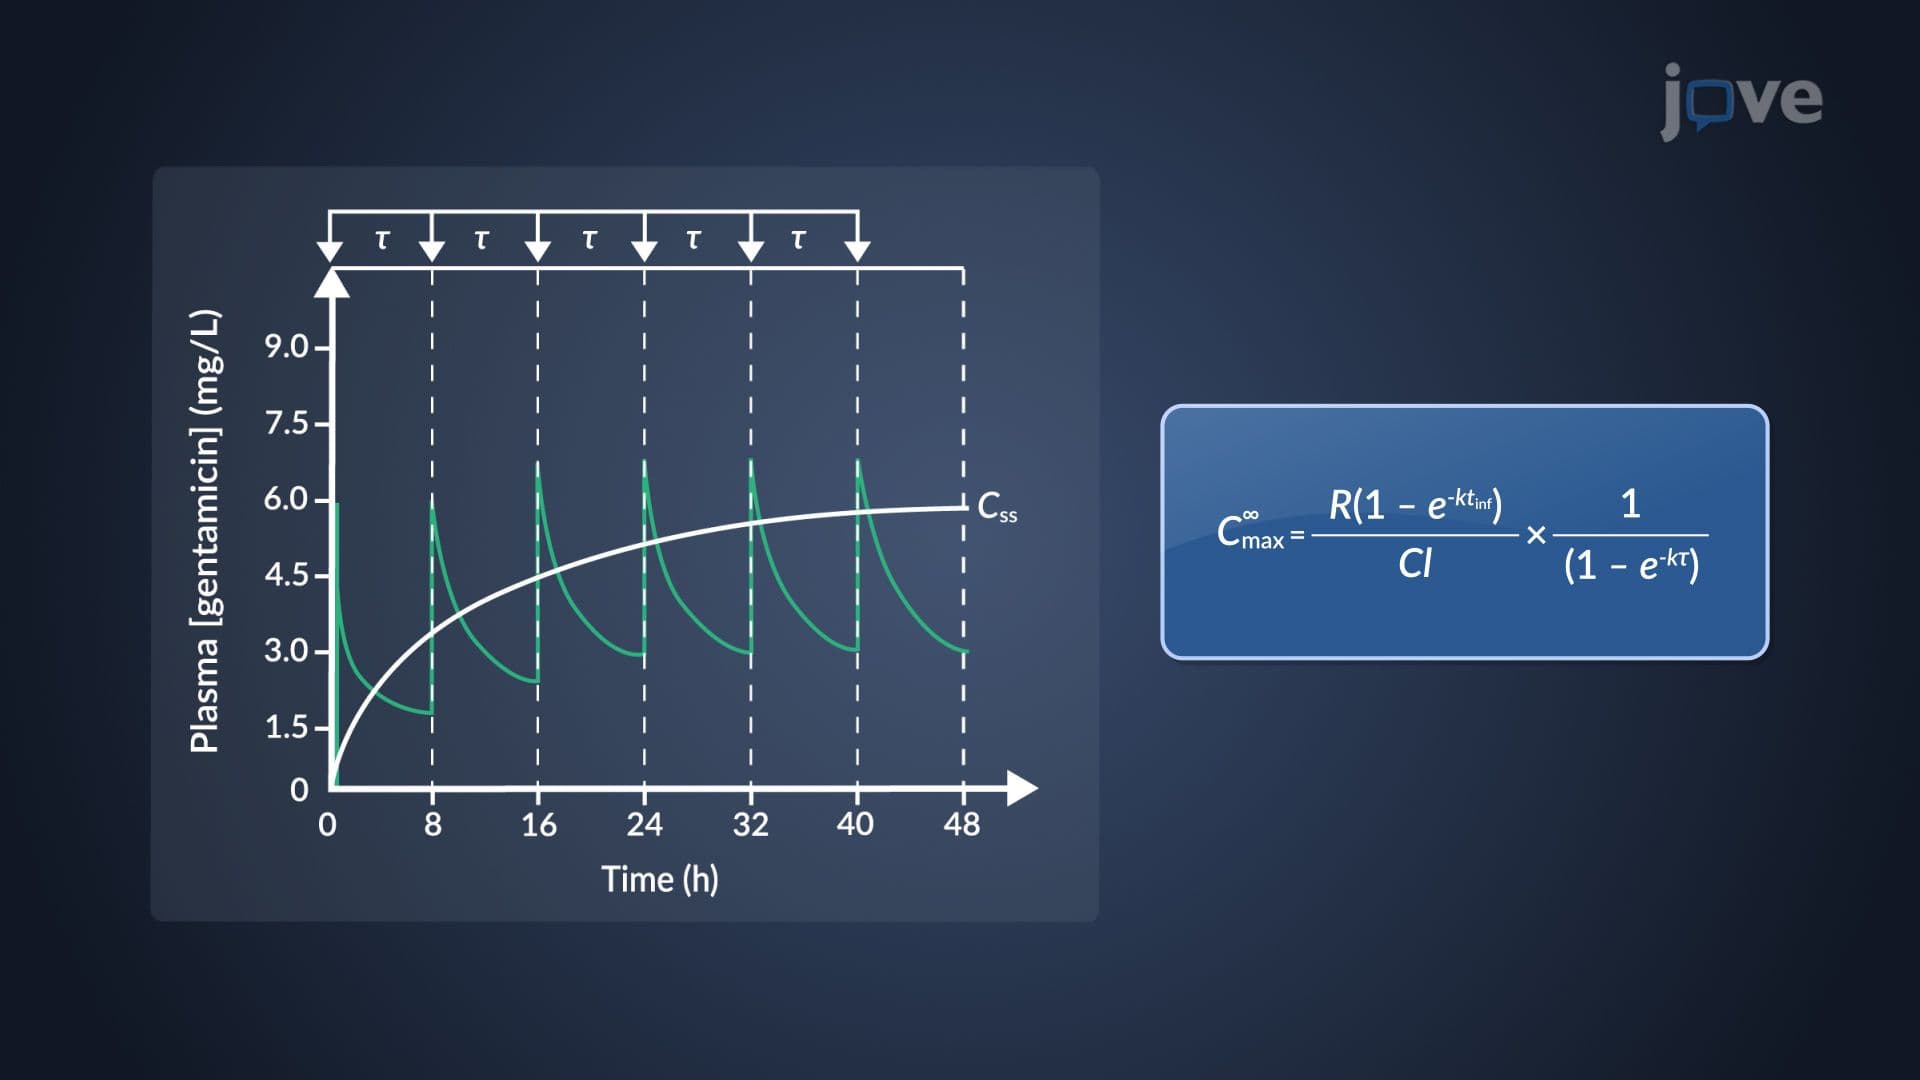
Determination of Multiple Dosing Parameters: Steady-State, Minimum and Maximum Concentrations
 00:54
Noncompartmental Analysis: Miscellaneous Pharmacokinetic Parameters
One key aspect of the noncompartmental approach is determining a drug's total clearance. This can be done by dividing the drug dose by the area under the concentration-time curve from zero to infinity. The area under the concentration-time curve represents the drug's...
 01:01
Dosage Regimens: Partial Pharmacokinetic Parameters
 01:19
One-Compartment Open Model for IV Bolus Administration: General Considerations
The drug's presence in the body is defined by an equation representing the difference between the rates of drug entry and exit. Key parameters—elimination rate constant,...
 01:14
Pharmacokinetic–Pharmacodynamic Relationship: Duration of Dose-Effect Relationship
 01:09
One-Compartment Open Model for IV Bolus Administration: Estimation of Elimination Rate Constant, Half-Life and Volume of Distribution