Related Experiment Video
Updated: Apr 15, 2026


Quantitative Structure-Activity Relationship, Activity Prediction, and Molecular Dynamics of Non-nucleotide Reverse Transcriptase Inhibitors
Published on: May 9, 2025
Quantitative structure-activity relationship and molecular docking of artemisinin derivatives to vascular endothelial
Mohamed E M Saeed1, Onat Kadioglu1, Ean-Jeong Seo1
1Department of Pharmaceutical Biology, Institute of Pharmacy and Biochemistry, Johannes Gutenberg University, Mainz, Germany.

Background/Aim:
The antimalarial drug artemisinin has been shown to exert anticancer activity through anti-angiogenic effects. For further drug development, it may be useful to have derivatives with improved anti-angiogenic properties.
Material And Methods:
We performed molecular docking of 52 artemisinin derivatives to vascular endothelial growth factor receptors (VEGFR1, VEGFR2), and VEGFA ligand using Autodock4 and AutodockTools-1.5.7.rc1 using the Lamarckian genetic algorithm. Quantitative structure-activity relationship (QSAR) analyses of the compounds prepared by Corina Molecular Networks were performed using the Molecular Operating Environment MOE 2012.10.
Results:
A statistically significant inverse relationship was obtained between in silico binding energies to VEGFR1 and anti-angiogenic activity in vivo of a test-set of artemisinin derivatives (R=-0.843; p=0.035). This served as a control experiment to validate molecular docking predicting anti-angiogenc effects. Furthermore, 52 artemisinin derivatives were docked to VEGFR1 and in selected examples also to VEGFR2 and VEGFA. Higher binding affinities were calculated for receptors than for the ligand. The best binding affinities to VEGFR1 were found for an artemisinin dimer, 10-dihydroartemisinyl-2-propylpentanoate, and dihydroartemisinin α-hemisuccinate sodium salt. QSAR analyses revealed significant relationships between VEGFR1 binding energies and defined molecular descriptors of 35 artemisinins assigned to the training set (R=0.0848, p<0.0001) and 17 derivatives assigned to the test set (R=0.761, p<0.001).
Conclusion:
Molecular docking and QSAR calculations can be used to identify novel artemisinin derivatives with anti-angiogenic effects.
More Related Videos
 13:18
Network Pharmacology Prediction and Experimental Validation of Trichosanthes-Fritillaria thunbergii Action Mechanism Against Lung Adenocarcinoma
Published on: March 3, 2023
 08:49
Incorporating Target Protein Structure Flexibility and Dynamics in Computational Drug Discovery Using Ensemble-Based Docking Analysis
Published on: June 20, 2025
Related Concept Videos
 01:28
Structure-Activity Relationships and Drug Design
SAR studies the intricate relationship between a drug's chemical structure and biological activity. It focuses on understanding how modifications to a drug's structure can influence...
 01:15
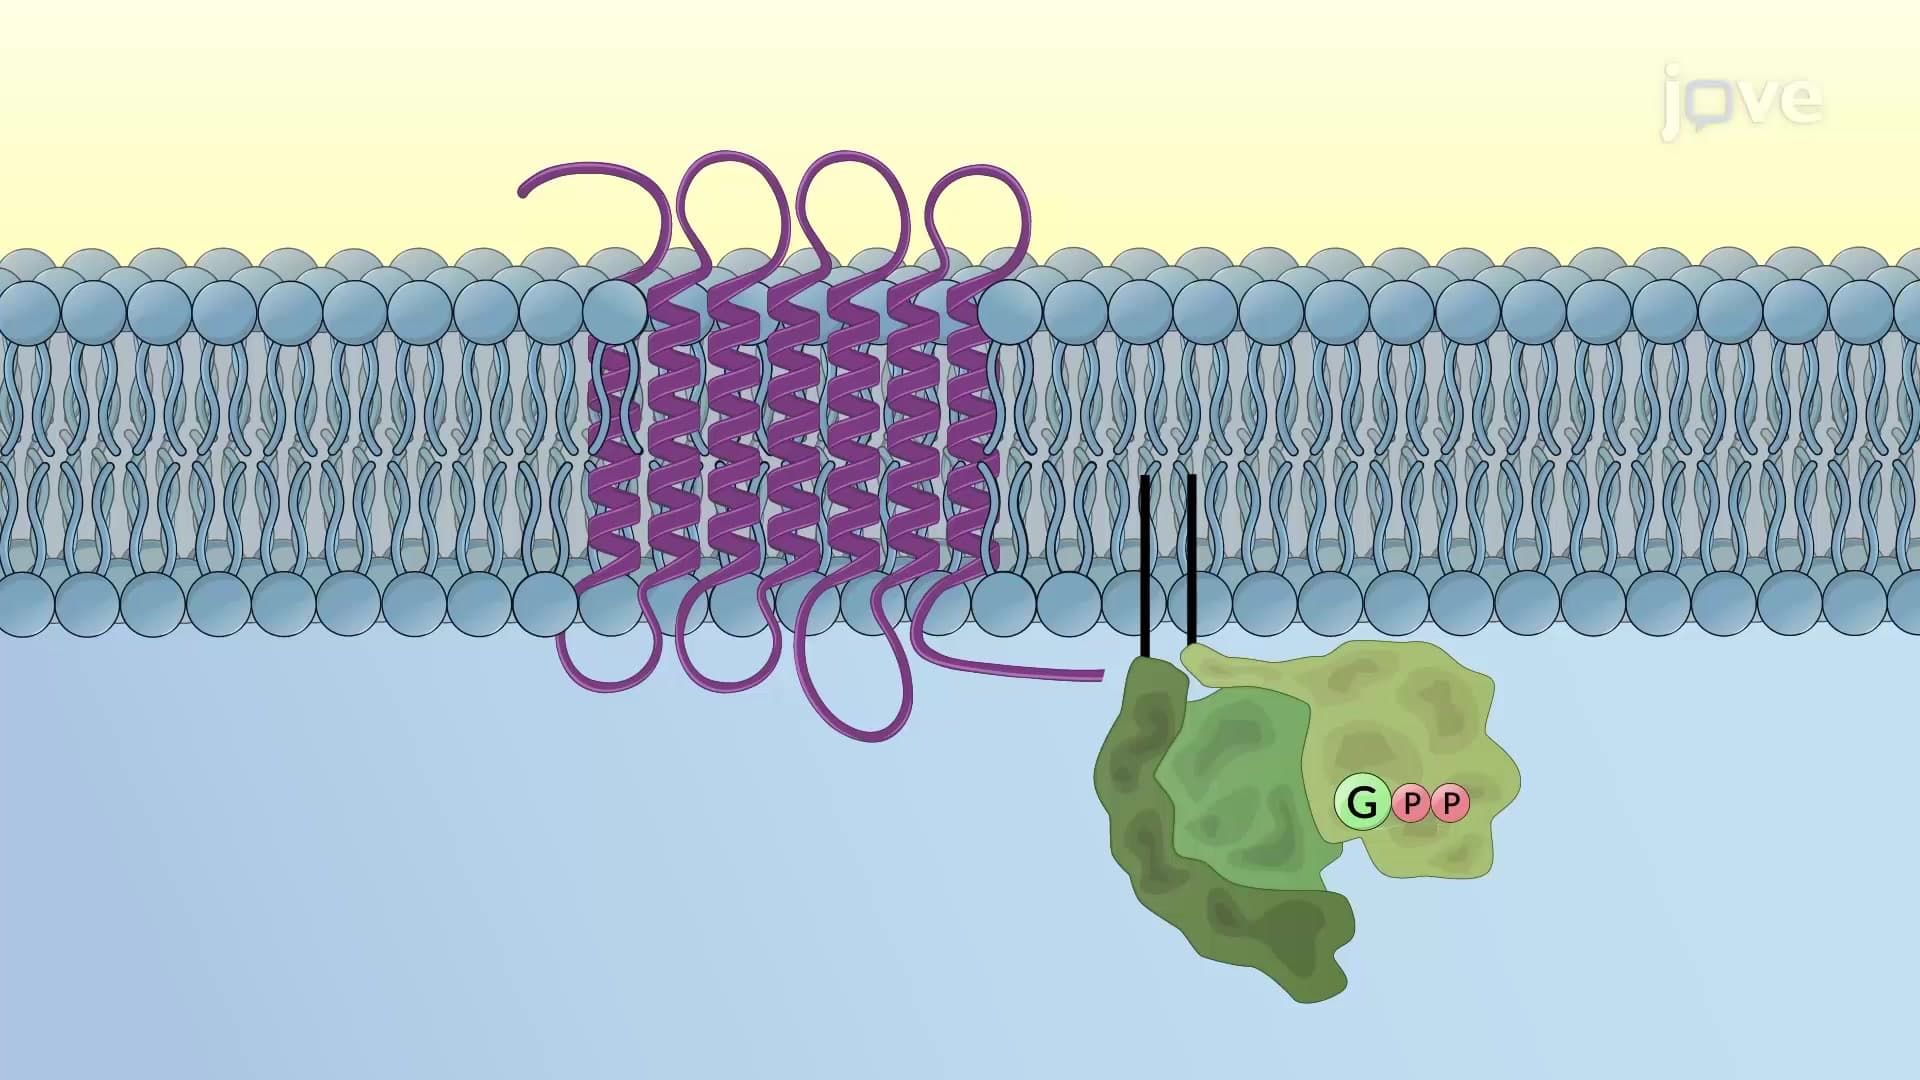
G Protein-coupled Receptors
GPCRs are also called heptahelical, 7TM, or serpentine receptors, and consist of seven (H1-H7) transmembrane alpha-helices that span the bilayer to form a cylindrical core. The transmembrane helices are connected by three extracellular loops and three...
 01:30
Quantitative Aspects of Drug-Receptor Interaction
 01:16
Adrenergic Agonists: Chemistry and Structure-Activity Relationship
Aromatic ring substitutions: Substituting the aromatic ring with –OH groups at positions 3 and 4 yields catecholamines (e.g., epinephrine), which have a high affinity for adrenoceptors. Hydrogen bonding between –OH groups and receptors enhances adrenergic activity.
Separation of...