Related Experiment Video
Updated: Apr 12, 2026

Preparation of SNS CobaltII Pincer Model Complexes of Liver Alcohol Dehydrogenase
Published on: March 19, 2020
Structure and Energetics of Complexes of B12N12 with Hydrogen Halides-SAPT(DFT) and MP2 Study
Sirous Yourdkhani1,2, Tatiana Korona2, Nasser L Hadipour1
1†Department of Chemistry, Tarbiat Modares University, P.O. Box 14115-175, Tehran, Iran.

Abstract:
Molecular complexes of a fullerene analogue B12N12 with hydrogen halides (HCl, HBr, and HI) were studied with symmetry-adapted perturbation theory with density-functional theory applied for a description of monomers (SAPT(DFT)), Møller-Plesset theory to the second order (MP2), and its spin-component-scaled variant (SCS-MP2) in a limit of a complete basis set. For each halide five symmetry-distinct minimum structures of the complex have been found on the potential energy hypersurface, with interaction energies ranging from -6 to -18 kJ/mol. The natural bond orbital and the atom-in-molecules analysis of noncovalent bonds resulted in a division of these configurations into three categories: hydrogen-bonded, halogen-bonded, and those of a mixed type, involving simultaneously a hydrogen bonding and a π-hole bonding between halogen and boron atoms. A comparison of various approaches for the calculation of interaction energies shows that the SCS-MP2 supermolecular method gives results which are in a close agreement with SAPT(DFT), while the MP2 interaction energies are systematically more negative than the SAPT values. The ability of the B12N12 nanocage to bind hydrogen halides through several active sites on its surface puts under question the selectivity of the binding necessary in crystal engineering, especially for the hydrogen bromide and hydrogen iodide cases, which show small differences in stabilization energies for their minimum structures. The directionality of noncovalent bonds is explained on grounds of the anisotropy of some SAPT components, like electrostatics and induction, as well as by the σ-hole and π-hole models.
More Related Videos
 10:52
Line Shape Analysis of Dynamic NMR Spectra for Characterizing Coordination Sphere Rearrangements at a Chiral Rhenium Polyhydride Complex
Published on: July 27, 2022
 06:35
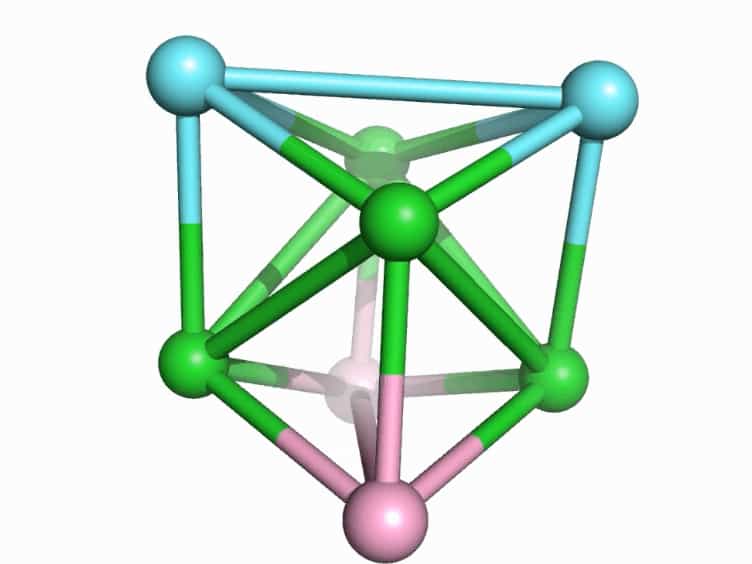
Construction and Systematical Symmetric Studies of a Series of Supramolecular Clusters with Binary or Ternary Ammonium Triphenylacetates
Published on: February 15, 2016
Related Concept Videos
 02:42
Valence Bond Theory
 02:45
Valence Bond Theory
 02:54
Coordination Compounds and Nomenclature
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 03:45
Formation of Complex Ions
 02:54
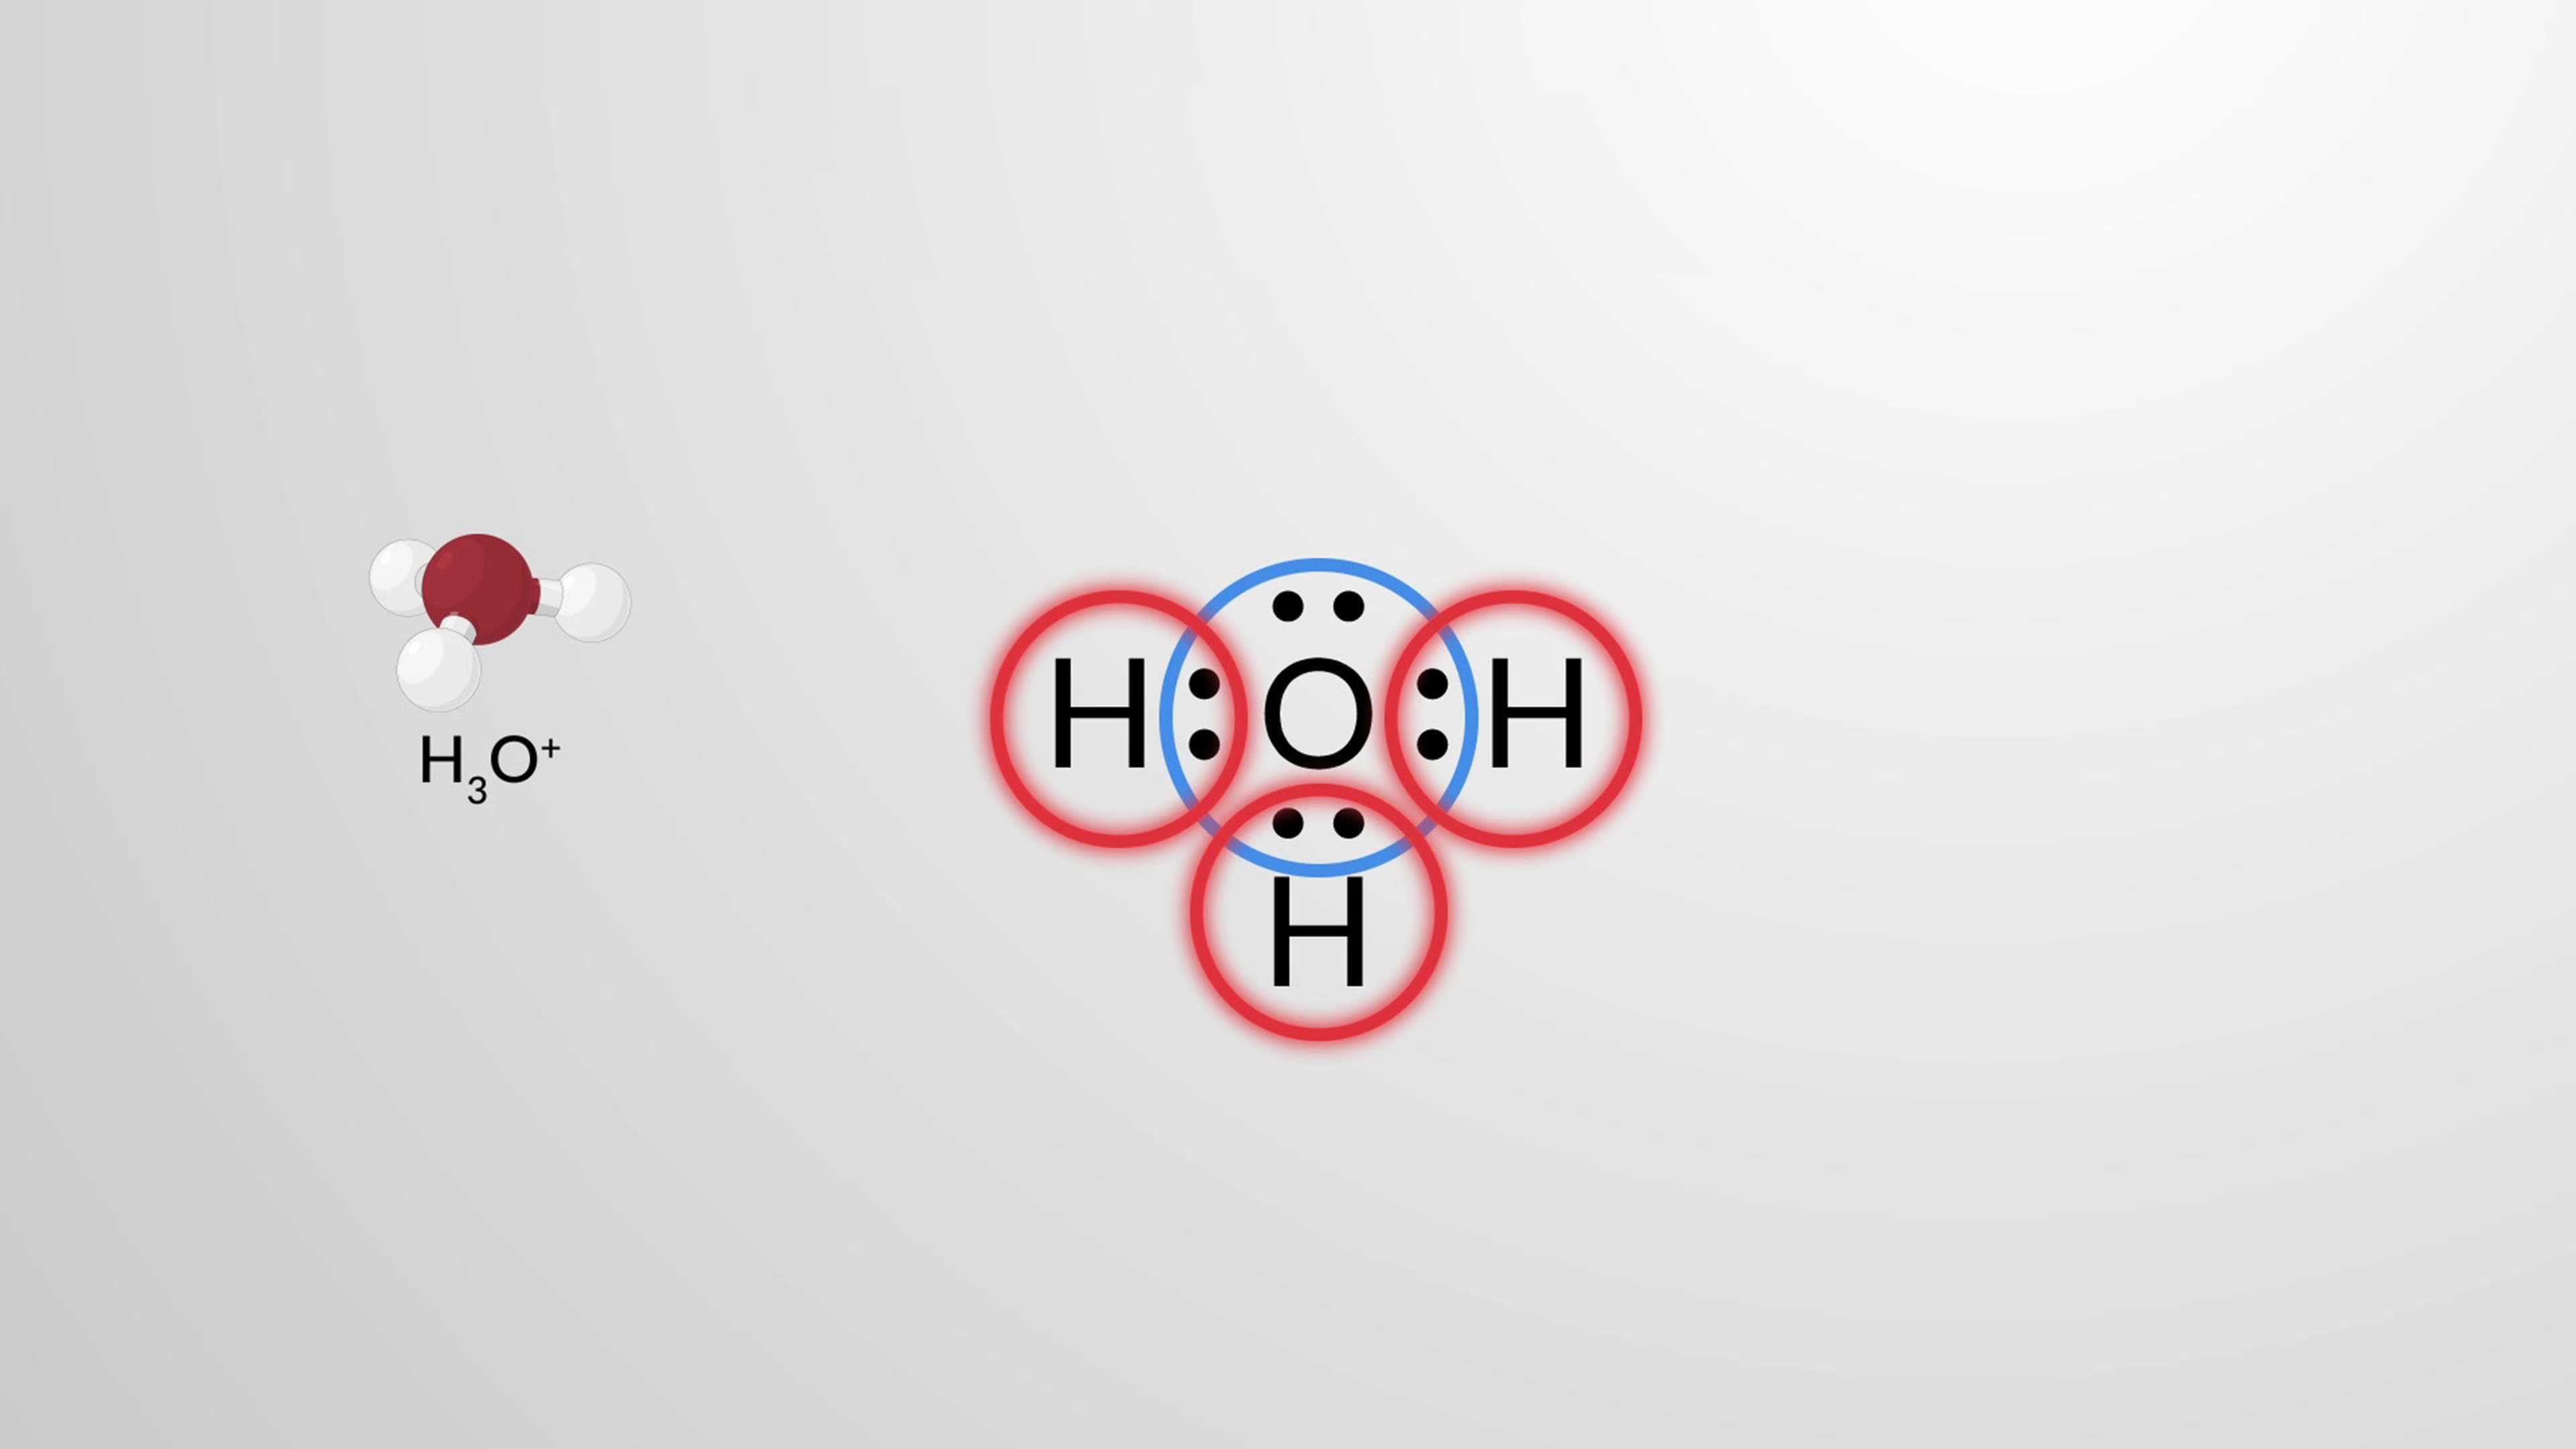
Lewis Structures of Molecular Compounds and Polyatomic Ions