Related Experiment Video
Updated: Apr 11, 2026
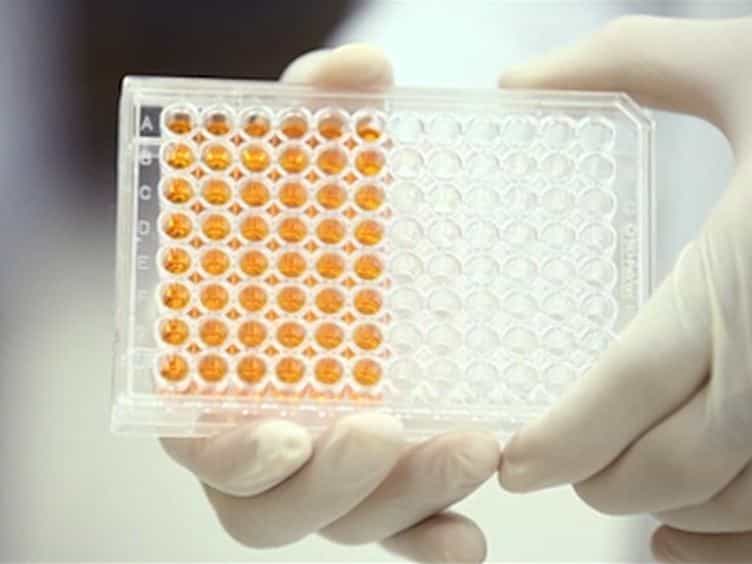

Development of an in vitro model system for studying the interaction of Equus caballus IgE with its high-affinity receptor FcεRI
Published on: November 1, 2014
Genome-Wide Association Study of Insect Bite Hypersensitivity in Swedish-Born Icelandic Horses
Merina Shrestha1, Susanne Eriksson1, Anouk Schurink1
1From the Department of Animal Breeding and Genetics, Swedish University of Agricultural Sciences, SE-750 07 Uppsala, Sweden (Shrestha, Eriksson, Bergström, and Lindgren); Animal Breeding and Genomic Centre, Wageningen University, 6700 AH Wageningen, The Netherlands (Shrestha, Schurink, and Ducro); Capilet Genetics AB, SE-725 93 Västerås, Sweden (Andersson); Östra Greda Research Group, SE-387 91 Borgholm, Sweden (Sundquist); Norsholms Animal Hospital, SE-602 37 Norrköping, Sweden (Frey); and Department of Clinical Sciences, Swedish University of Agricultural Sciences, SE-750 07 Uppsala, Sweden (Broström).

Abstract:
Insect bite hypersensitivity (IBH) is the most common allergic skin disease in horses and is caused by biting midges, mainly of the genus Culicoides. The disease predominantly comprises a type I hypersensitivity reaction, causing severe itching and discomfort that reduce the welfare and commercial value of the horse. It is a multifactorial disorder influenced by both genetic and environmental factors, with heritability ranging from 0.16 to 0.27 in various horse breeds. The worldwide prevalence in different horse breeds ranges from 3% to 60%; it is more than 50% in Icelandic horses exported to the European continent and approximately 8% in Swedish-born Icelandic horses. To minimize the influence of environmental effects, we analyzed Swedish-born Icelandic horses to identify genomic regions that regulate susceptibility to IBH. We performed a genome-wide association (GWA) study on 104 affected and 105 unaffected Icelandic horses genotyped using Illumina® EquineSNP50 Genotyping BeadChip. Quality control and population stratification analyses were performed with the GenABEL package in R (λ = 0.81). The association analysis was performed using the Bayesian variable selection method, Bayes C, implemented in GenSel software. The highest percentage of genetic variance was explained by the windows on X chromosomes (0.51% and 0.36% by 73 and 74 mb), 17 (0.34% by 77 mb), and 18 (0.34% by 26 mb). Overlapping regions with previous GWA studies were observed on chromosomes 7, 9, and 17. The windows identified in our study on chromosomes 7, 10, and 17 harbored immune system genes and are priorities for further investigation.
More Related Videos
 07:01
Zebrafish Animal Model for the Study of Allergic Reactions in Response to Tick Saliva Biomolecules
Published on: September 16, 2022
 10:33
Multi-locus Variable-number Tandem-repeat Analysis of the Fish-pathogenic Bacterium Yersinia ruckeri by Multiplex PCR and Capillary Electrophoresis
Published on: June 17, 2019
