Related Experiment Video
Updated: Apr 10, 2026
Determination of the Photoisomerization Quantum Yield of a Hydrazone Photoswitch
Published on: February 7, 2022
Probing the π→π* photoisomerization mechanism of cis-azobenzene by multi-state ab initio on-the-fly trajectory
1Institute of Molecular Science, Department of Applied Chemistry, and Center for Interdisciplinary Molecular Science, National Chiao-Tung University, Hsinchu 300, Taiwan. cyzhu@mail.nctu.edu.tw.

Abstract:
Based on a newly developed algorithm to compute global nonadiabatic switching probability by using only electronic adiabatic potential energy surfaces and gradients, we performed on-the-fly, trajectory-surface hopping simulations at the 5SA-CASSCF(6,6)/6-31G quantum level to probe the π→π* photoisomerization mechanism of the azobenzene within four singlet low-lying electronic states (S0, S1, S2, and S3) coupled with a complicated conical intersection network. We found that four conical intersections between the S1 and S2 states (one is near the cis-isomer region, another near the trans-isomer region, and two others between cis and trans) play the most important roles for understanding the photoisomerization mechanism of azobenzene upon S2 and S3ππ* excitation. We studied six cases to demonstrate the photoisomerization mechanism in detail by choosing eight (six) typical reactive (nonreactive) trajectories, namely, two-step fast-fast processes having lifetimes of several tenths to one hundred femtoseconds and two-step, fast-slow and slow-slow processes having lifetimes of several hundred to one thousand femtoseconds. We found for the first time from simulation that once a trajectory visits the conical intersection near the trans-isomer after ππ* excitation, it could rapidly go through the inversion pathway to trans-azobenzene, and confirms the most recent experimental observations. We performed 536 sampling trajectories (336 from S2 and 200 from S3), initially starting from the Franck-Condon region of cis-azobenzene, and obtained a total reactive quantum yield of 0.3-0.45 in very good agreement with recent experimental results of 0.24-0.50. Moreover, the current method can estimate overall nonadiabatic transition probability for each sampling trajectory from beginning to end. This can greatly accelerate convergence of nonadiabatic molecular dynamic simulation, and, for instance, results in a quantum yield of 0.53 estimated from only eight typical reactive trajectories.
Related Concept Videos
 01:26
Photochemical Electrocyclic Reactions: Stereochemistry
Selection Rules: Photochemical Activation
 01:26
Thermal and Photochemical Electrocyclic Reactions: Overview
 01:24
π Molecular Orbitals of 1,3-Butadiene
The simplest conjugated diene is 1,3-butadiene: a four-carbon system where each carbon is sp2-hybridized and has an unhybridized p orbital that contains an unpaired electron. According to molecular orbital theory, atomic orbitals combine to form molecular orbitals such that the number...
 01:18
Structure of Benzene: Molecular Orbital Model
 01:16
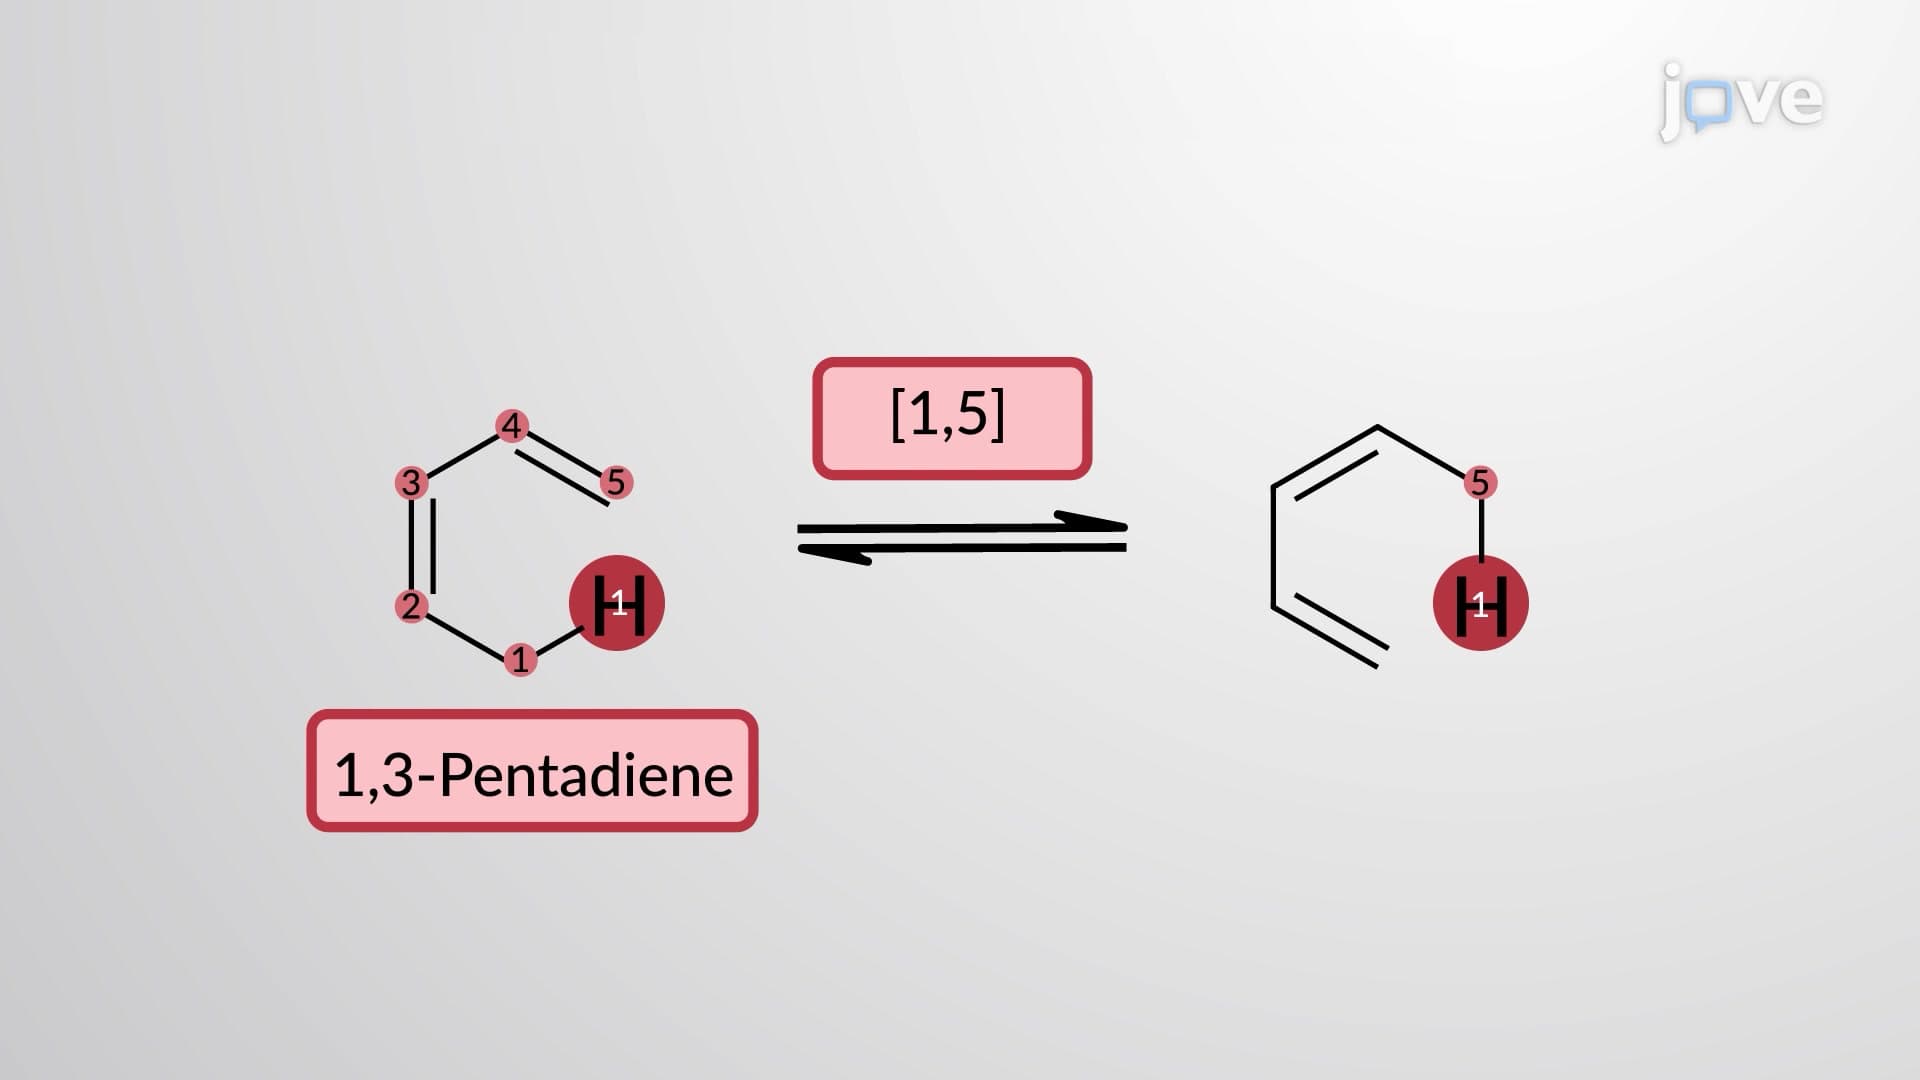
Thermal Sigmatropic Reactions: Overview
Sigmatropic shifts are classified based on an order term [i, j ], where i and j indicate the number of atoms across which each end of the σ bond migrates. Below are examples of a [3,3] sigmatropic shift in 1,5-hexadiene, referred...
 01:21
[3,3] Sigmatropic Rearrangement of 1,5-Dienes: Cope Rearrangement