Related Experiment Video
Updated: Apr 9, 2026
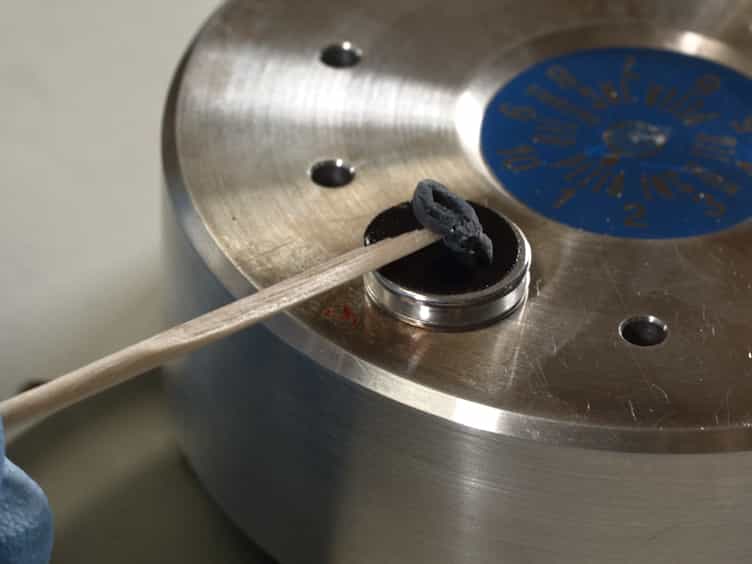

Scanning Electron Microscopy of Macerated Tissue to Visualize the Extracellular Matrix
Published on: June 14, 2016
Chronic sustained inflammation links to left ventricular hypertrophy and aortic valve sclerosis: a new link between
Ling Yan1, Marion A Hofmann Bowman1
1Department of Medicine, Section of Cardiology, The University of Chicago, Chicago, IL 60637 USA.

Insights
Human S100/calgranulin accelerates cardiovascular disease in chronic kidney disease (CKD) mice. This S100/RAGE-mediated inflammation upregulates FGF23, promoting cardiac hypertrophy and calcification.
Area of Science:
- Nephrology
- Cardiology
- Molecular Biology
Background:
- Cardiovascular disease (CVD) is prevalent in chronic kidney disease (CKD), with S100A12 and FGF23 as key biomarkers.
- Left ventricular hypertrophy (LVH), diastolic dysfunction, and valvular calcification are common CVD manifestations in CKD patients.
Purpose of the Study:
- To investigate the role of human S100/calgranulin in accelerating cardiovascular disease in a mouse model of CKD.
- To elucidate the underlying mechanisms involving S100 proteins, RAGE, and FGF23 in CKD-associated cardiac pathology.
Main Methods:
- Developed transgenic hBAC-S100 mice expressing human S100A12 and S100A8/9 proteins.
- Induced CKD in hBAC-S100 and wild-type (WT) mice via ureteral ligation.
- Assessed cardiac and valvular pathology, FGF23 levels, and the role of RAGE.
Main Results:
- hBAC-S100 mice with CKD exhibited increased cardiac FGF23, LVH, diastolic dysfunction, and valvular calcification.
- This phenotype was dependent on S100/RAGE signaling, as it was absent in WT mice or hBAC-S100 mice lacking RAGE.
- In vitro studies showed inflammatory stimuli upregulate FGF23 in cardiac fibroblasts.
Conclusions:
- Myeloid-derived human S100/calgranulin promotes cardiac hypertrophy and ectopic calcification in CKD via a RAGE-dependent pathway.
- FGF23, upregulated by cardiac fibroblasts in response to S100/RAGE-mediated inflammation, may act paracrinely to exacerbate cardiac dysfunction.
- This study provides a mechanistic link between CKD, systemic inflammation, and cardiovascular complications like LVH and diastolic dysfunction.
Background:
Cardiovascular disease including left ventricular hypertrophy, diastolic dysfunction and ectopic valvular calcification are common in patients with chronic kidney disease (CKD). Both S100A12 and fibroblast growth factor 23 (FGF23) have been identified as biomarkers of cardiovascular morbidity and mortality in patients with CKD. We tested the hypothesis that human S100/calgranulin would accelerate cardiovascular disease in mice subjected to CKD.
Methods:
This review paper focuses on S100 proteins and their receptor for advanced glycation end products (RAGE) and summarizes recent findings obtained in novel developed transgenic hBAC-S100 mice that express S100A12 and S100A8/9 proteins. A bacterial artificial chromosome of the human S100/calgranulin gene cluster containing the genes and regulatory elements for S100A8, S100A9 and S100A12 was expressed in C57BL/6J mice (hBAC-S100). CKD was induced by ureteral ligation, and hBAC-S100 mice and WT mice were studied after 10 weeks of chronic uremia.
Results:
hBAC-S100 mice with CKD showed increased FGF23 in the heart, left ventricular hypertrophy (LVH), diastolic dysfunction, focal cartilaginous metaplasia and calcification of the mitral and aortic valve annulus together with aortic valve sclerosis. This phenotype was not observed in WT mice with CKD or in hBAC-S100 mice lacking RAGE with CKD, suggesting that the inflammatory milieu mediated by S100/RAGE promotes pathological cardiac hypertrophy in CKD. In vitro, inflammatory stimuli including IL-6, TNFα, LPS, or serum from hBAC-S100 mice up regulated FGF23 mRNA and protein in primary murine neonatal and adult cardiac fibroblasts.
Conclusions:
Taken together, our study shows that myeloid-derived human S100/calgranulin is associated with the development of cardiac hypertrophy and ectopic cardiac calcification in a RAGE dependent manner in a mouse model of CKD. We speculate that FGF23 produced by cardiac fibroblasts in response to cytokines may act in a paracrine manner to accelerate LVH and diastolic dysfunction in hBAC-S100 mice with CKD. We suggest that S100/RAGE-mediated chronic sustained systemic inflammation is linked to pathological cardiac remodeling via direct up regulation of FGF23 in cardiac fibroblasts, thereby providing a new mechanistic understanding for the common association between CKD, diabetes, metabolic syndrome, or hypertension with left ventricular hypertrophy with diastolic dysfunction.
Related Concept Videos
 01:22
Mitral Stenosis I: Introduction
 01:20
Mitral Regurgitation I: Introduction
 01:15
Aortic Regurgitation I: Introduction
 01:23
Rheumatic Heart Disease I: Introduction
 01:29
Heart Failure II: Pathophysiology
 01:22
Aortic Regurgitation II: Clinical Features and Diagnostic Tests