Related Experiment Video
Updated: Apr 7, 2026
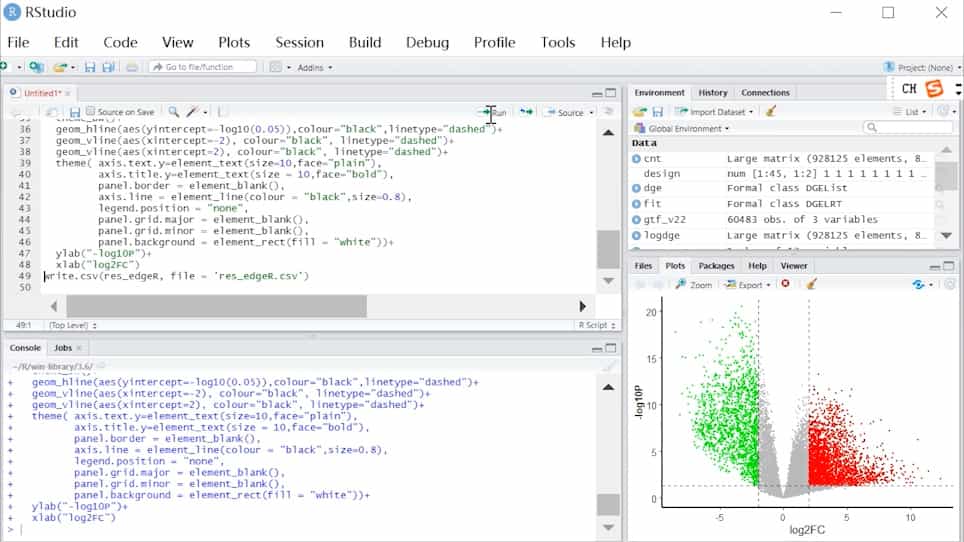

Three Differential Expression Analysis Methods for RNA Sequencing: limma, EdgeR, DESeq2
Published on: September 18, 2021
DINGO: differential network analysis in genomics
Min Jin Ha1, Veerabhadran Baladandayuthapani1, Kim-Anh Do1
1Department of Biostatistics, The University of Texas MD Anderson Cancer Center, Houston, TX 77030, USA.

Motivation:
Cancer progression and development are initiated by aberrations in various molecular networks through coordinated changes across multiple genes and pathways. It is important to understand how these networks change under different stress conditions and/or patient-specific groups to infer differential patterns of activation and inhibition. Existing methods are limited to correlation networks that are independently estimated from separate group-specific data and without due consideration of relationships that are conserved across multiple groups.
Method:
We propose a pathway-based differential network analysis in genomics (DINGO) model for estimating group-specific networks and making inference on the differential networks. DINGO jointly estimates the group-specific conditional dependencies by decomposing them into global and group-specific components. The delineation of these components allows for a more refined picture of the major driver and passenger events in the elucidation of cancer progression and development.
Results:
Simulation studies demonstrate that DINGO provides more accurate group-specific conditional dependencies than achieved by using separate estimation approaches. We apply DINGO to key signaling pathways in glioblastoma to build differential networks for long-term survivors and short-term survivors in The Cancer Genome Atlas. The hub genes found by mRNA expression, DNA copy number, methylation and microRNA expression reveal several important roles in glioblastoma progression.
Availability And Implementation:
R Package at: odin.mdacc.tmc.edu/∼vbaladan.
Contact:
veera@mdanderson.org
Supplementary Information:
Supplementary data are available at Bioinformatics online.
Insights
This study introduces DINGO, a novel method for analyzing differential gene networks in cancer. DINGO accurately identifies group-specific molecular changes, improving our understanding of cancer progression and patient stratification.
Area of Science:
- Genomics
- Bioinformatics
- Computational Biology
Background:
- Cancer development involves complex molecular network aberrations across multiple genes and pathways.
- Understanding differential network patterns under various conditions is crucial for cancer research.
- Current methods for network analysis often overlook conserved relationships across patient groups.
Purpose of the Study:
- To develop a novel pathway-based differential network analysis model named DINGO.
- To improve the estimation of group-specific and conserved molecular network components.
- To provide a refined understanding of driver and passenger events in cancer progression.
Main Methods:
- DINGO jointly estimates group-specific conditional dependencies by decomposing them into global and group-specific components.
- The model facilitates inference on differential networks, highlighting key molecular interactions.
- An R package is available for implementing the DINGO model.
Main Results:
- Simulation studies confirm DINGO's superior accuracy in estimating group-specific dependencies compared to separate approaches.
- Application to glioblastoma data reveals differential networks for long-term and short-term survivors.
- Key hub genes identified through multi-omics data (mRNA, DNA copy number, methylation, microRNA) are implicated in glioblastoma progression.
Conclusions:
- DINGO offers a robust framework for pathway-based differential network analysis in cancer genomics.
- The method enhances the identification of molecular mechanisms driving cancer progression and patient outcomes.
- The findings provide insights into glioblastoma pathogenesis and potential therapeutic targets.
More Related Videos
 10:44
Inherent Dynamics Visualizer, an Interactive Application for Evaluating and Visualizing Outputs from a Gene Regulatory Network Inference Pipeline
Published on: December 7, 2021
 11:13
RNA-Seq Analysis of Differential Gene Expression in Electroporated Chick Embryonic Spinal Cord
Published on: November 1, 2014
Related Concept Videos
 02:26
Comparing Copy Number Variations and SNPs
Copy number variations or CNVs are the structural variations that cover more than 1kb of DNA sequence. The single nucleotide polymorphism (SNP), on the other hand, is a single nucleotide change or a point mutation that is found in more than 1%...
 02:54
Evolutionary Relationships through Genome Comparisons
 01:09
Epistasis Analysis