Related Experiment Video
Updated: Apr 6, 2026


Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
Hellmann-Feynman forces within the DFT + U in Wannier functions basis
D Novoselov1, Dm M Korotin, V I Anisimov
1Institute of Metal Physics, S.Kovalevskoy St. 18, 620137 Yekaterinburg, Russia.

Abstract:
The most general way to describe localized atomic-like electronic states in strongly correlated compounds is to use Wannier functions. In the present paper we continue development of widely-used DFT + U method with the Wannier function basis set and propose a technique to calculate Hubbard contribution to atomic forces. The technique was implemented as a part of plane-waves pseudopotential code Quantum-ESPRESSO and tested on two compounds: charge transfer insulator NiO with cubic crystal structure and correlated metal SrVO3 with perovskite structure.
More Related Videos
 08:44
Measurements of Long-range Electronic Correlations During Femtosecond Diffraction Experiments Performed on Nanocrystals of Buckminsterfullerene
Published on: August 22, 2017
 09:43
Interfacial Molecular-level Structures of Polymers and Biomacromolecules Revealed via Sum Frequency Generation Vibrational Spectroscopy
Published on: August 13, 2019
Related Concept Videos
 01:04
Force and Potential Energy in Three Dimensions
 02:45
The Quantum-Mechanical Model of an Atom
 01:10
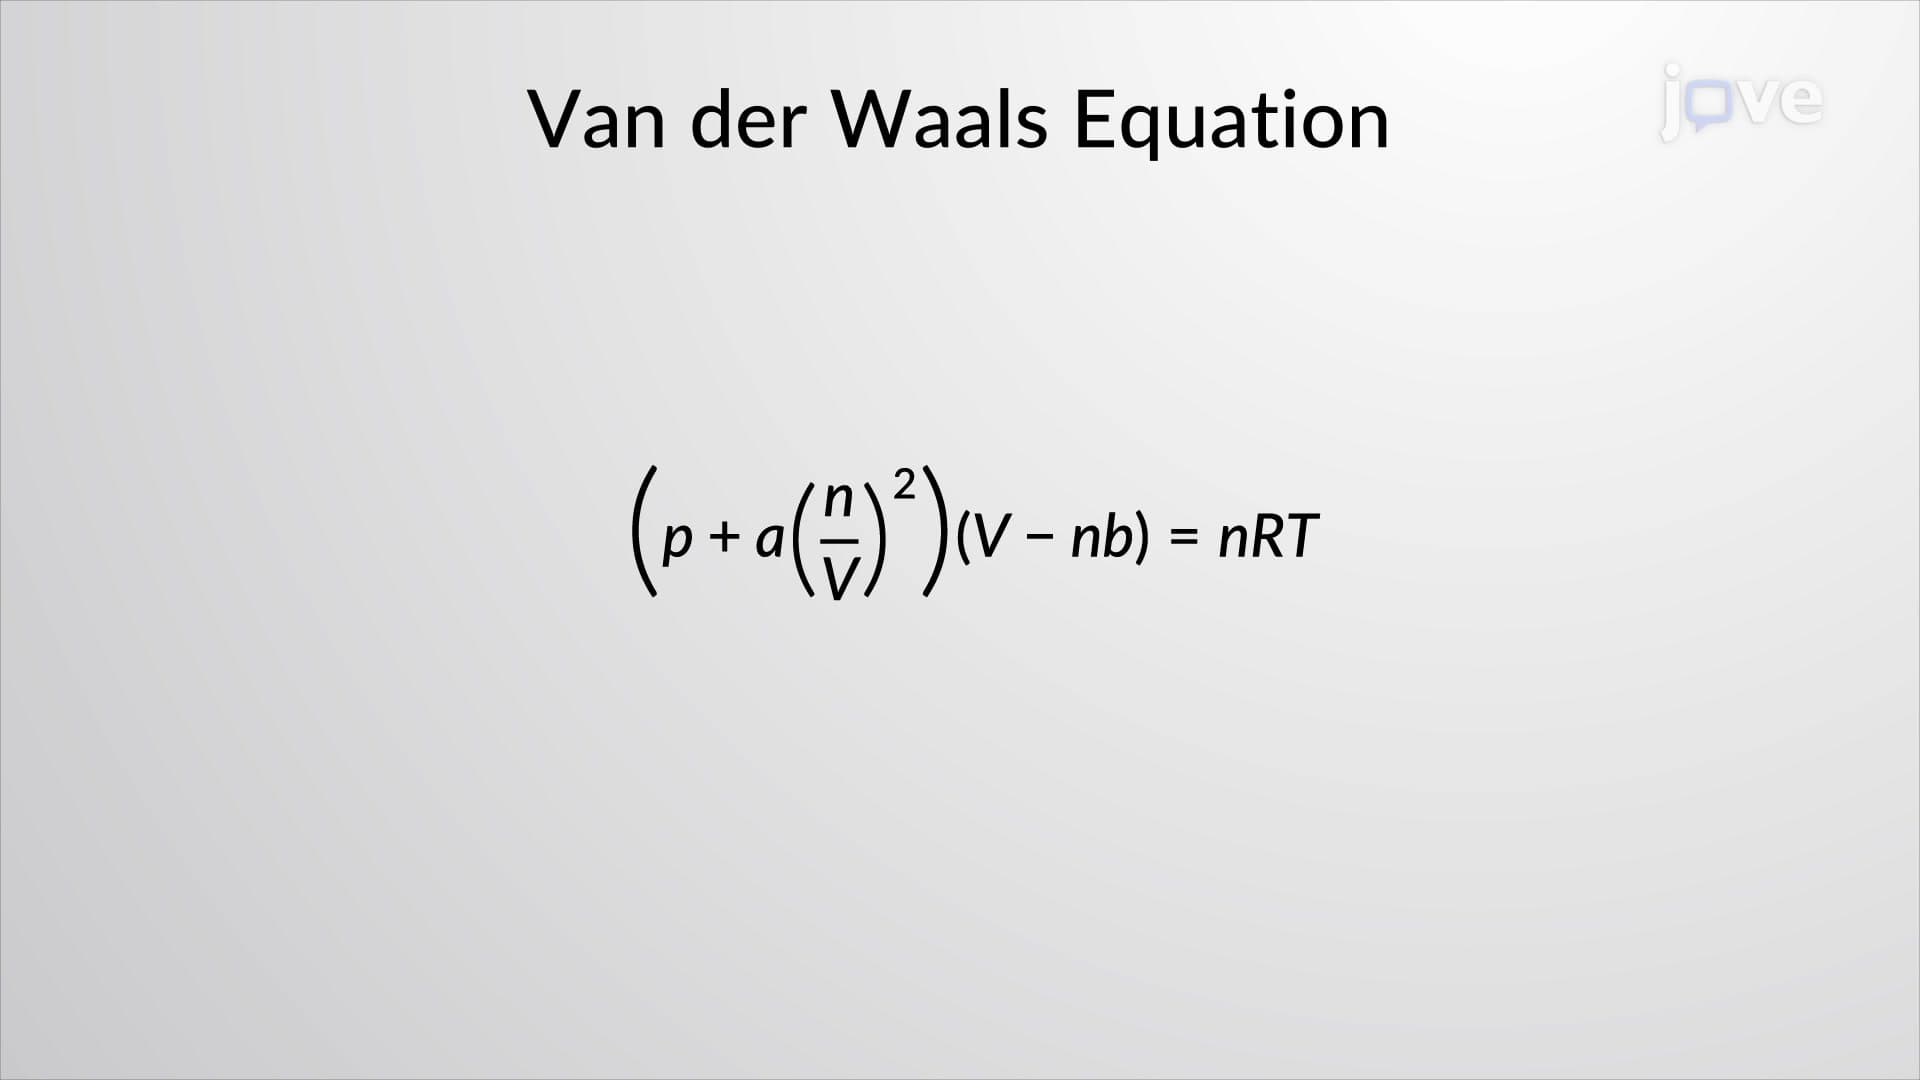
Van der Waals Equation
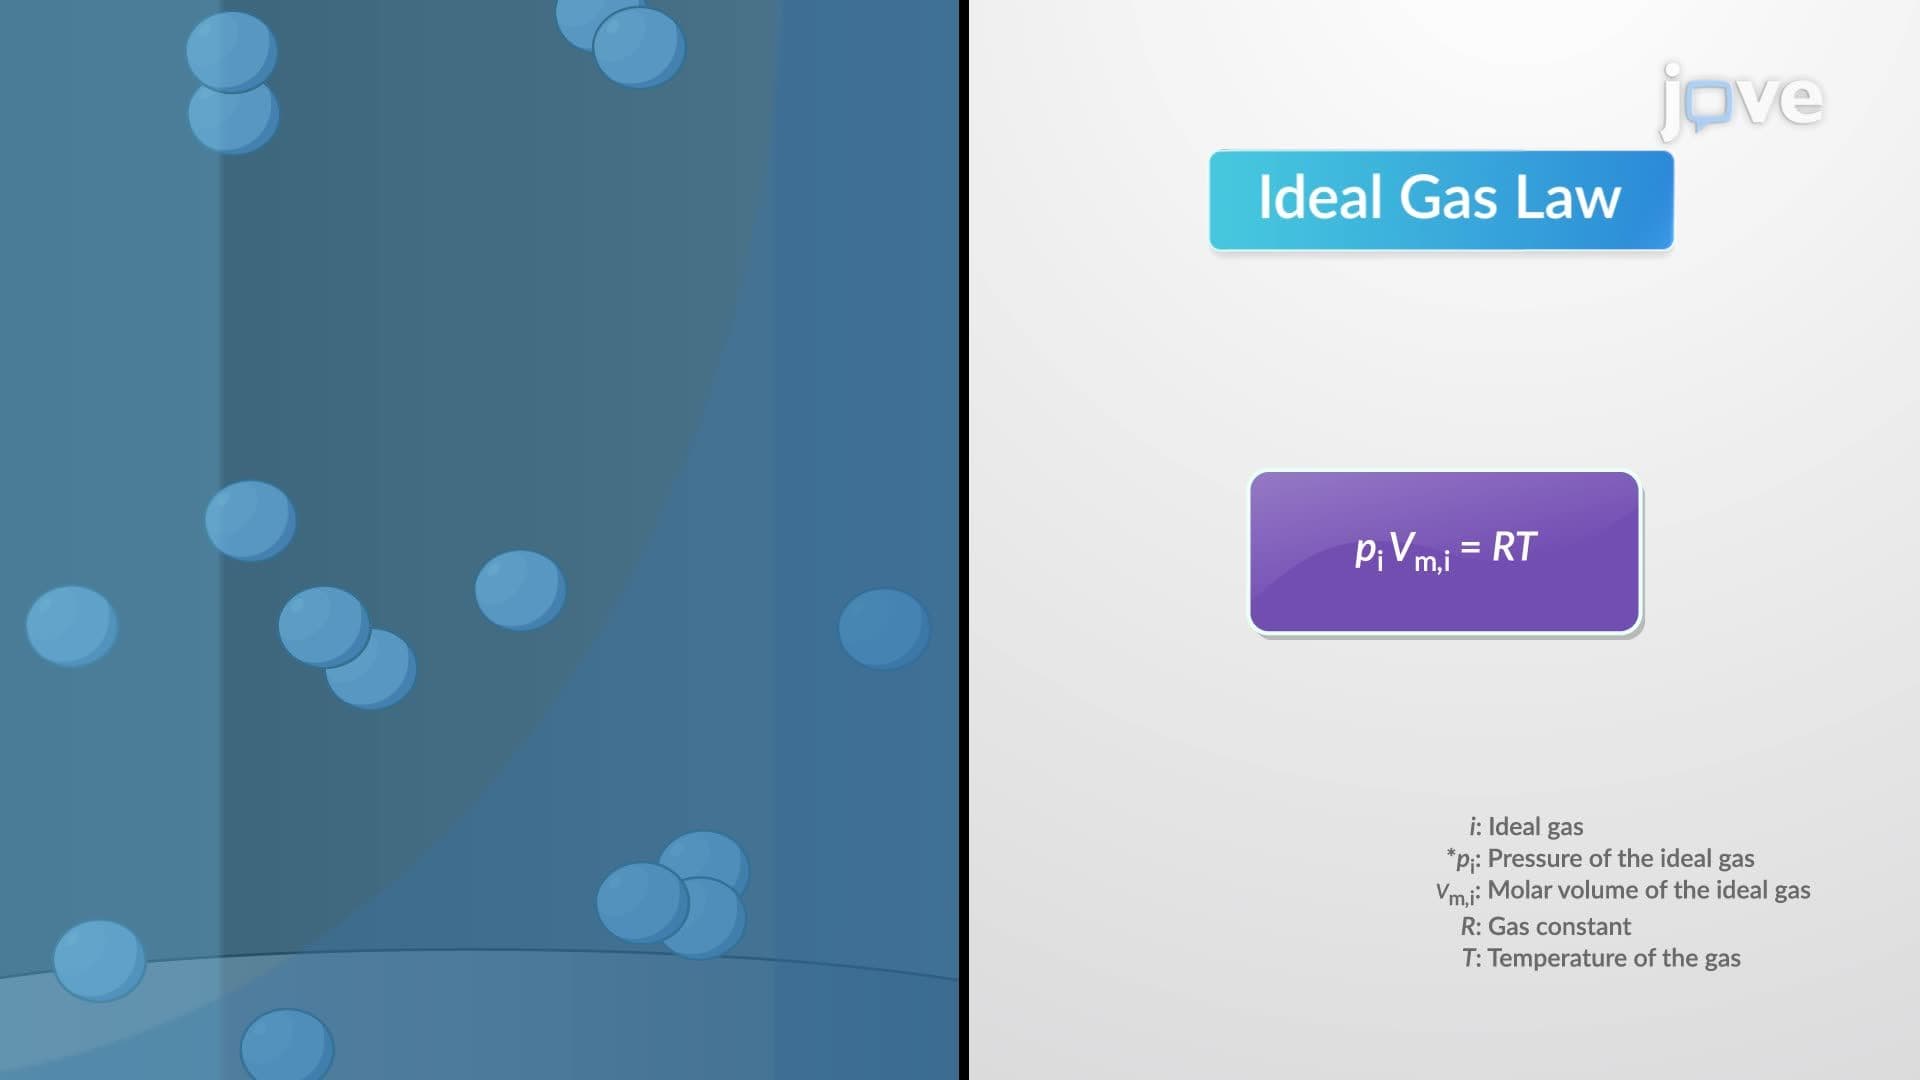
First, the attractive forces between molecules, which are stronger at higher densities and reduce the pressure, are considered by adding to the pressure a term equal to the square of the molar density multiplied by a positive coefficient a. Second, the volume...
 01:26
The Van der Waals Equation
 01:24
Van der Waals Interactions
 01:13
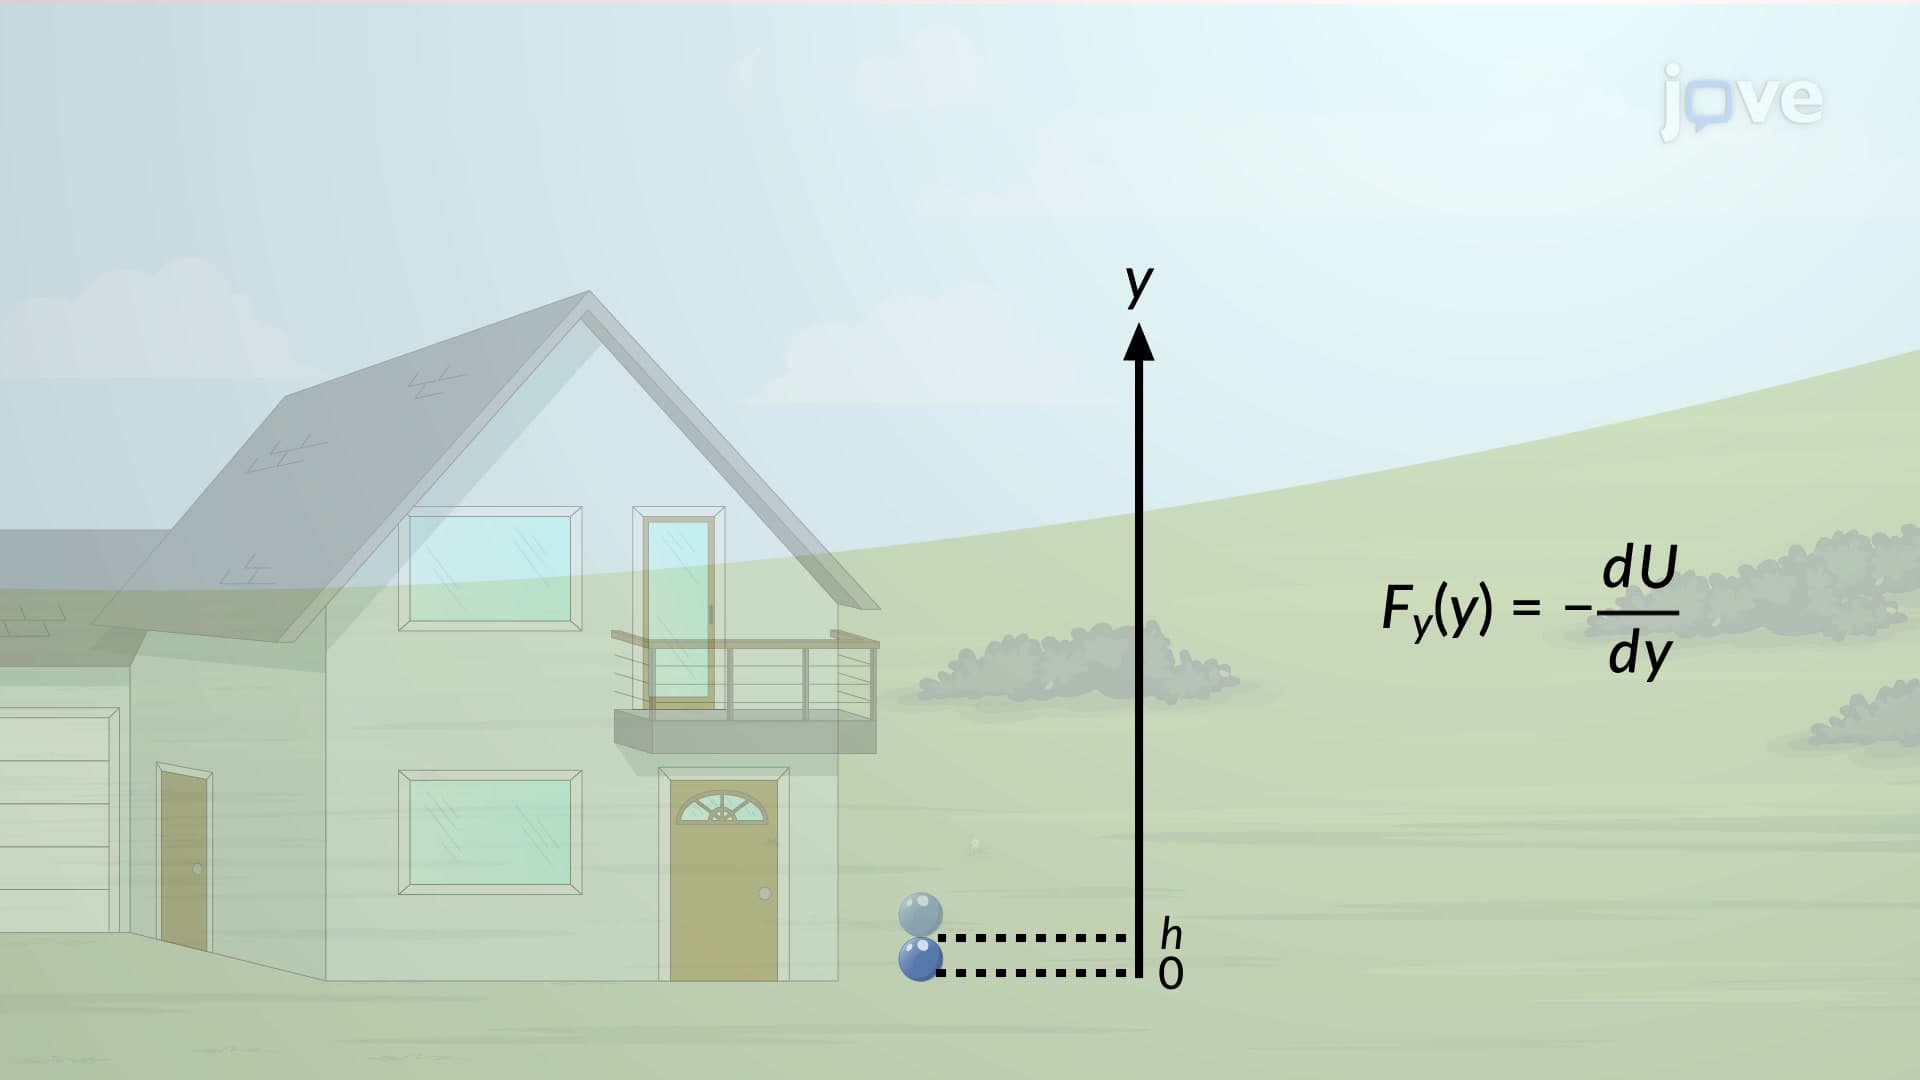
Force and Potential Energy in One Dimension