Related Experiment Video
Updated: Apr 5, 2026

Quantitative Atomic-Site Analysis of Functional Dopants/Point Defects in Crystalline Materials by Electron-Channeling-Enhanced Microanalysis
Published on: May 10, 2021
A DFT+U study of A-site and B-site substitution in BaFeO3-δ
Zarah Medina Baiyee1, Chi Chen, Francesco Ciucci
1Department of Mechanical and Aerospace Engineering, The Hong Kong University of Science and Technology, Hong Kong, China. francesco.ciucci@ust.hk.

Abstract:
BaFeO3-δ (BFO)-based perovskites have emerged as cheap and effective oxygen electrocatalysts for oxygen reduction reaction at high temperatures. The BFO cubic phase facilitates a high oxygen deficiency and is commonly stabilised by partial substitution. Understanding the electronic mechanisms of substitution and oxygen deficiency is key to rational material design, and can be realised through DFT analysis. In this work an in-depth first principle DFT+U study is undertaken to determine site distinctive characteristics for 12.5%, Y, La and Ce substitutions in BFO. In particular, it is shown that B-site doped structures exhibit a lower energy cost for oxygen vacancy formation relative to A site doping and pristine BFO. This is attributed to the stabilisation of holes in the oxygen sub-lattice and increased covalency of the Fe-O bonds of the FeO6 octahedra in B-site-substituted BFO. Charge analysis shows that A-site substitution amounts to donor doping and consequently impedes the accommodation of other donors (i.e. oxygen vacancies). However, A-site substitution may also exhibit a higher electronic conductivity due to less lattice distortion for oxygen deficiency compared to B-site doped structures. Furthermore, analysis of the local structural effects provides physical insight into stoichiometric expansions observed for this material.
More Related Videos
Related Concept Videos
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 02:42
Valence Bond Theory
 00:49
Ligand Binding and Linkage
 01:08
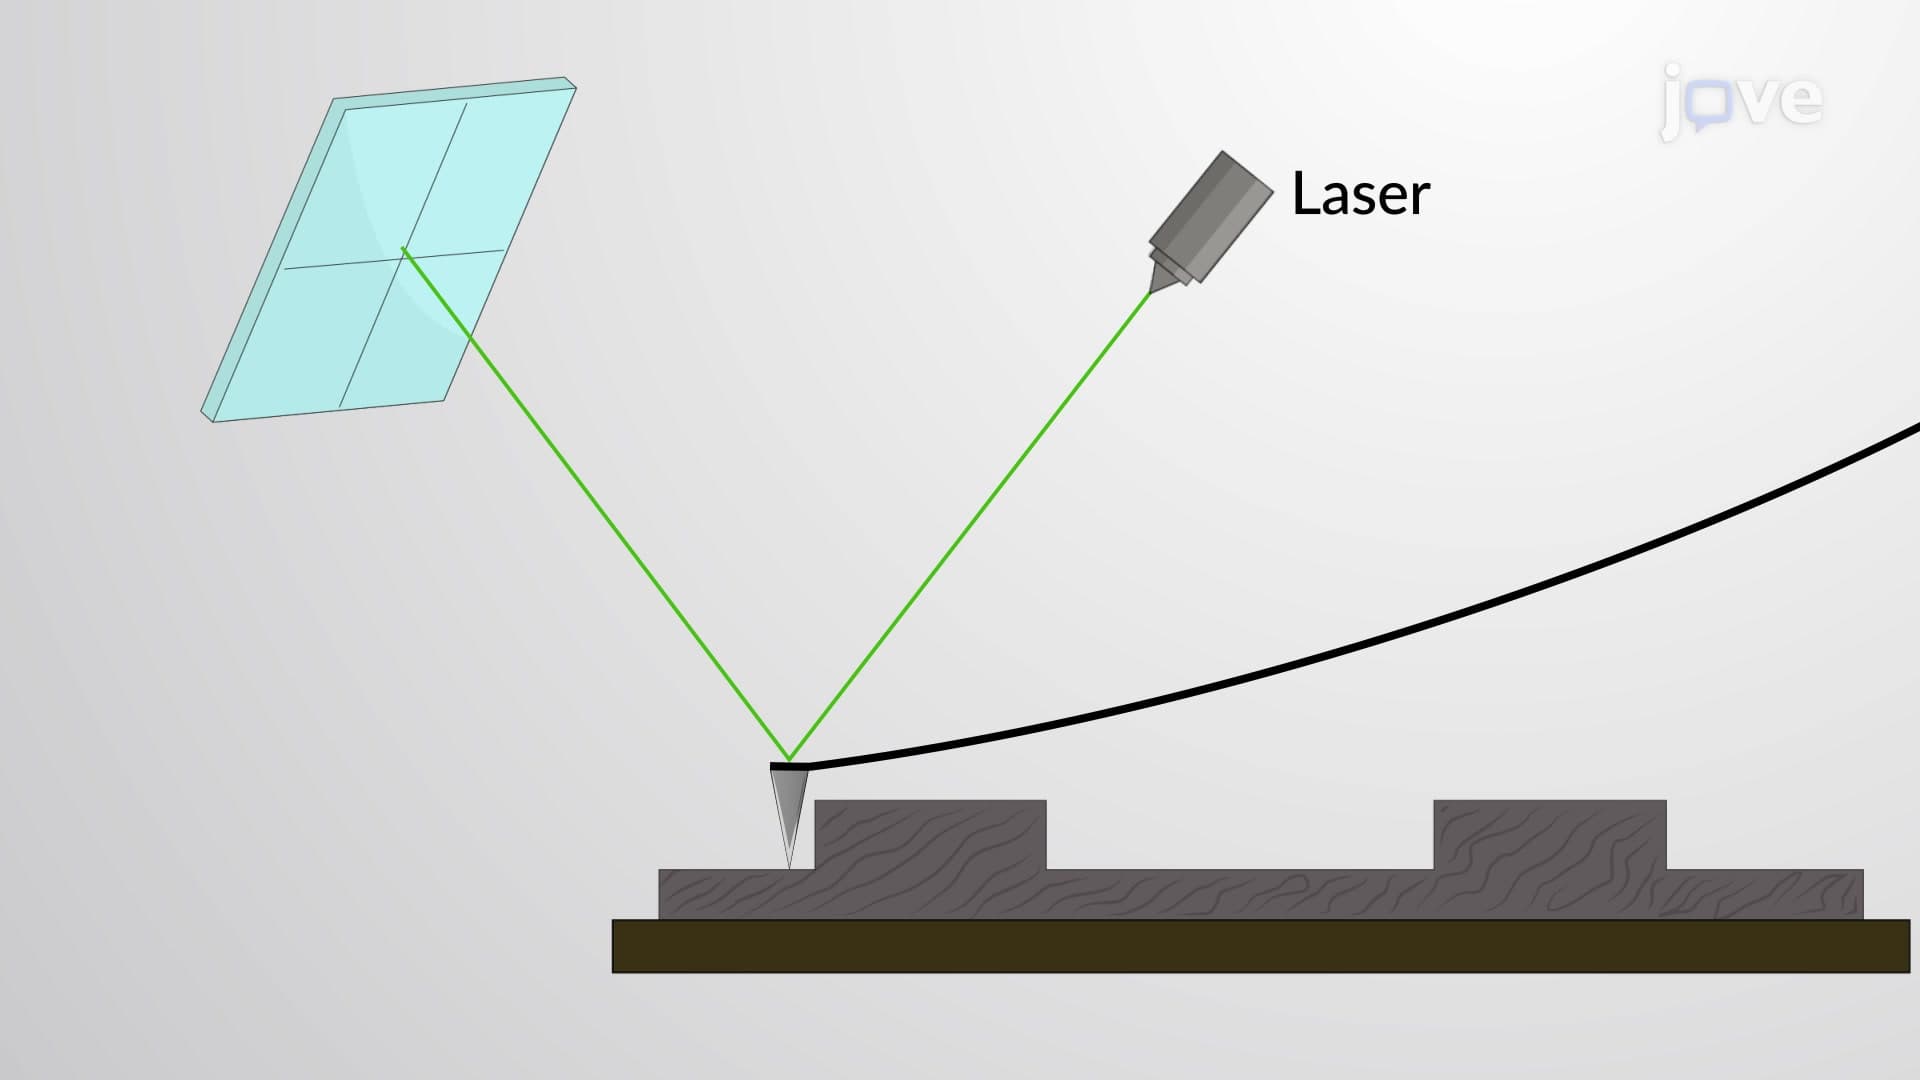
Atomic Force Microscopy
The AFM Probe
The probe is regarded as the heart of any AFM setup and comprises the...
 03:24
Hybridization of Atomic Orbitals I