
Molecular pathomechanisms and cell-type-specific disease phenotypes of MELAS caused by mutant mitochondrial tRNA(Trp)
Hideyuki Hatakeyama1,2, Ayako Katayama3, Hirofumi Komaki4,3
1Department of Mental Retardation and Birth Defect Research, National Institute of Neuroscience, National Center of Neurology and Psychiatry, Kodaira, Tokyo, 187-8502, Japan. hideyuki@ncnp.go.jp.

Introduction:
Numerous pathogenic mutations responsible for mitochondrial diseases have been identified in mitochondrial DNA (mtDNA)-encoded tRNA genes. In most cases, however, the detailed molecular pathomechanisms and cellular pathophysiology of these mtDNA mutations -how such genetic defects determine the variation and the severity of clinical symptoms in affected individuals- remain unclear. To investigate the molecular pathomechanisms and to realize in vitro recapitulation of mitochondrial diseases, intracellular mutant mtDNA proportions must always be considered.
Results:
We found a disease-causative mutation, m.5541C>T heteroplasmy in MT-TW gene, in a patient exhibiting mitochondrial myopathy, encephalopathy, lactic acidosis, and stroke-like episodes (MELAS) with multiple organ involvement. We identified the intrinsic molecular pathomechanisms of m.5541C>T. This mutation firstly disturbed the translation machinery of mitochondrial tRNA(Trp) and induced mitochondrial respiratory dysfunction, followed by severely injured mitochondrial homeostasis. We also demonstrated cell-type-specific disease phenotypes using patient-derived induced pluripotent stem cells (iPSCs) carrying ~100 % mutant m.5541C>T. Significant loss of terminally differentiated iPSC-derived neurons, but not their stem/progenitor cells, was detected most likely due to serious mitochondrial dysfunction triggered by m.5541C>T; in contrast, m.5541C>T did not apparently affect skeletal muscle development.
Conclusions:
Our iPSC-based disease models would be widely available for understanding the "definite" genotype-phenotype relationship of affected tissues and organs in various mitochondrial diseases caused by heteroplasmic mtDNA mutations, as well as for further drug discovery applications.
Insights
A novel mitochondrial DNA mutation, m.5541C>T, causes MELAS by disrupting tRNA(Trp) translation and mitochondrial function. Patient-derived iPSC models reveal cell-type-specific neuronal loss, aiding genotype-phenotype understanding.
Area of Science:
- Mitochondrial genetics and disease
- Cellular pathophysiology
- Stem cell biology
Background:
- Mitochondrial diseases often stem from mutations in mitochondrial DNA (mtDNA)-encoded tRNA genes.
- The precise molecular mechanisms linking these genetic defects to varied clinical symptoms remain largely unknown.
- Understanding intracellular mutant mtDNA levels is crucial for studying disease pathomechanisms and in vitro modeling.
Observation:
- A pathogenic mutation, m.5541C>T heteroplasmy in the MT-TW gene, was identified in a patient with MELAS (mitochondrial myopathy, encephalopathy, lactic acidosis, and stroke-like episodes).
- This mutation was found to disrupt the mitochondrial tRNA(Trp) translation machinery, leading to mitochondrial respiratory dysfunction and impaired homeostasis.
- Patient-derived induced pluripotent stem cells (iPSCs) with near-homoplasmic m.5541C>T mutation exhibited cell-type-specific phenotypes.
Findings:
- The m.5541C>T mutation specifically impacted terminally differentiated neurons derived from iPSCs, causing significant cell loss due to mitochondrial dysfunction.
- In contrast, the mutation did not appear to affect neuronal stem/progenitor cells or skeletal muscle development.
- This highlights cell-type-specific vulnerability to mitochondrial dysfunction caused by the m.5541C>T mutation.
Implications:
- Induced pluripotent stem cell (iPSC)-based disease models offer a platform for elucidating genotype-phenotype correlations in mitochondrial diseases.
- These models are valuable for understanding tissue-specific effects of heteroplasmic mtDNA mutations.
- The developed models can facilitate drug discovery and therapeutic development for mitochondrial disorders.
Related Concept Videos
 02:59
Animal Mitochondrial Genetics
 01:19
Translocation of Proteins into the Mitochondria
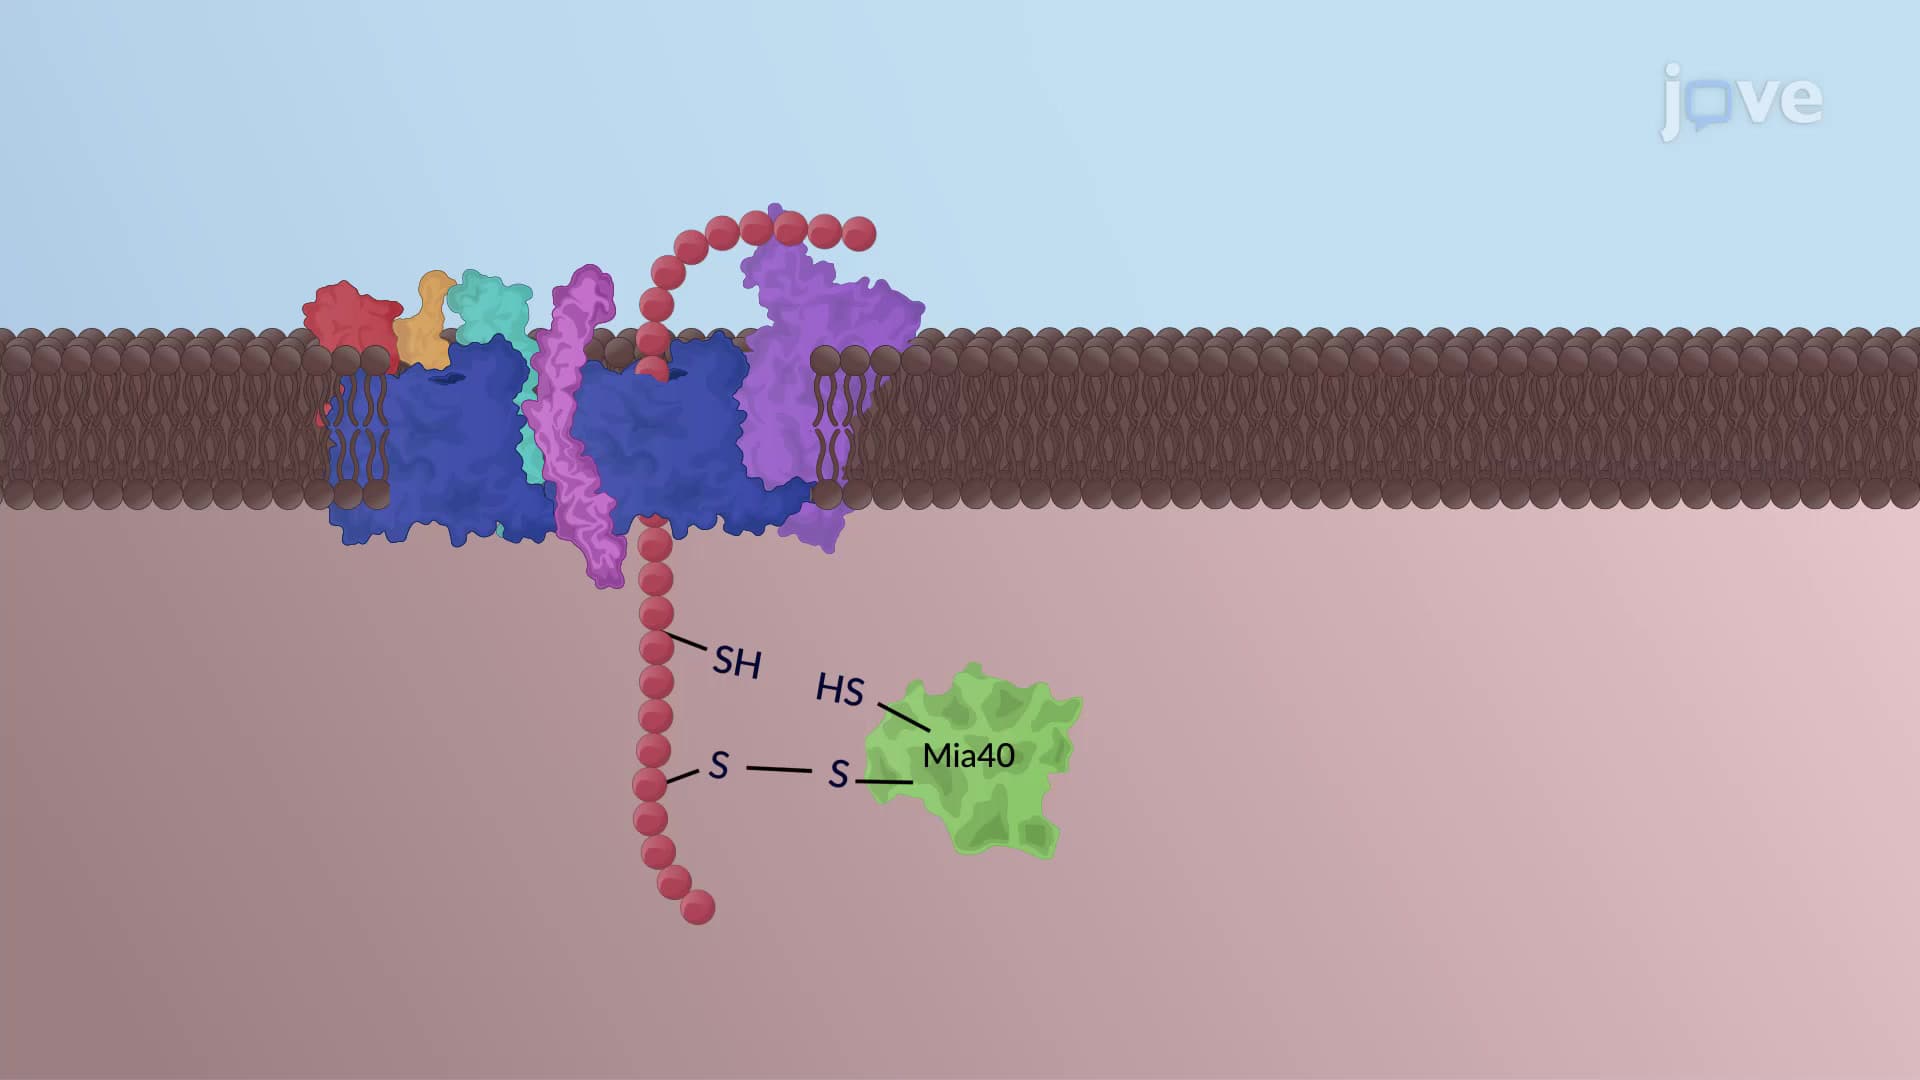
Sorting of outer membrane proteins:
Mitochondrial outer membrane proteins are of two types: the transmembrane, beta-barrel porins, and the membrane-anchored, alpha-helical proteins. Beta-barrel porin precursors are translocated by the TOM complex and inserted into the outer mitochondrial membrane by the SAM complex. In contrast,...
 01:31
Translation
Translation is the process of synthesizing proteins from the genetic information carried by messenger RNA (mRNA). Following transcription, it constitutes the final step in the expression of genes. This process is carried out by ribosomes, complexes of protein and specialized RNA molecules. Ribosomes, transfer RNA (tRNA), and other proteins produce a chain of amino acids—the polypeptide—as the end product of translation.
Translation Produces the Building Blocks of...
01:31Translation
Translation Produces the Building Blocks of Life
Proteins are...
 01:48
ATP Synthase: Mechanism
 01:46
Electron Transport Chain: Complex I and II
ROS generation is regulated and maintained at moderate levels necessary...