Related Experiment Video
Updated: Apr 4, 2026


Isotopic Effect in Double Proton Transfer Process of Porphycene Investigated by Enhanced QM/MM Method
Published on: July 19, 2019
A global full-dimensional potential energy surface and quasiclassical trajectory study of the O((1)D) + CH4
Kejie Shao1, Bina Fu, Dong H Zhang
1State Key Laboratory of Molecular Reaction Dynamics and Center for Theoretical and Computational Chemistry, Dalian Institute of Chemical Physics, Chinese Academy of Sciences, Dalian, P. R. China 116023. bina@dicp.ac.cn zhangdh@dicp.ac.cn.

Abstract:
We report a new global, full-dimensional ground-state potential energy surface (PES) of the O((1)D) + CH4 multichannel reaction, based on high-level ab initio calculations and fitting procedures. The PES is a permutationally invariant fit to roughly 340 000 electronic energies calculated by the MRCI + Q/aug-cc-pVTZ level of theory. Extensive quasiclassical trajectory calculations were carried out on the new PES at the collision energy of relevance to the previously universal crossed molecular beam experiments. The product branching ratios, translational energy distributions and angular distributions of OH + CH3, H + CH2OH/CH3O and H2 + HCOH/H2CO product channels were calculated and compared with the available experimental results. Very good agreement between theory and experiment has been achieved. The O((1)D) + CH4 reaction mainly proceeds through the CH3OH intermediate via a trapped abstraction mechanism, starting with the abstraction of the hydrogen atom, rather than the direct insertion pathway with the O((1)D) atom directly inserting into the C-H bond of CH4. The process with a very short lifetime behaves like an abstraction reaction, producing a pronounced forward scattering peak as found in the OH + CH3 channel, while the process with a relatively long lifetime produces reaction products with nearly forward and backward scattering symmetry, similar to an insertion reaction, as found in other reaction channels.
More Related Videos
 10:52
Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
Related Concept Videos
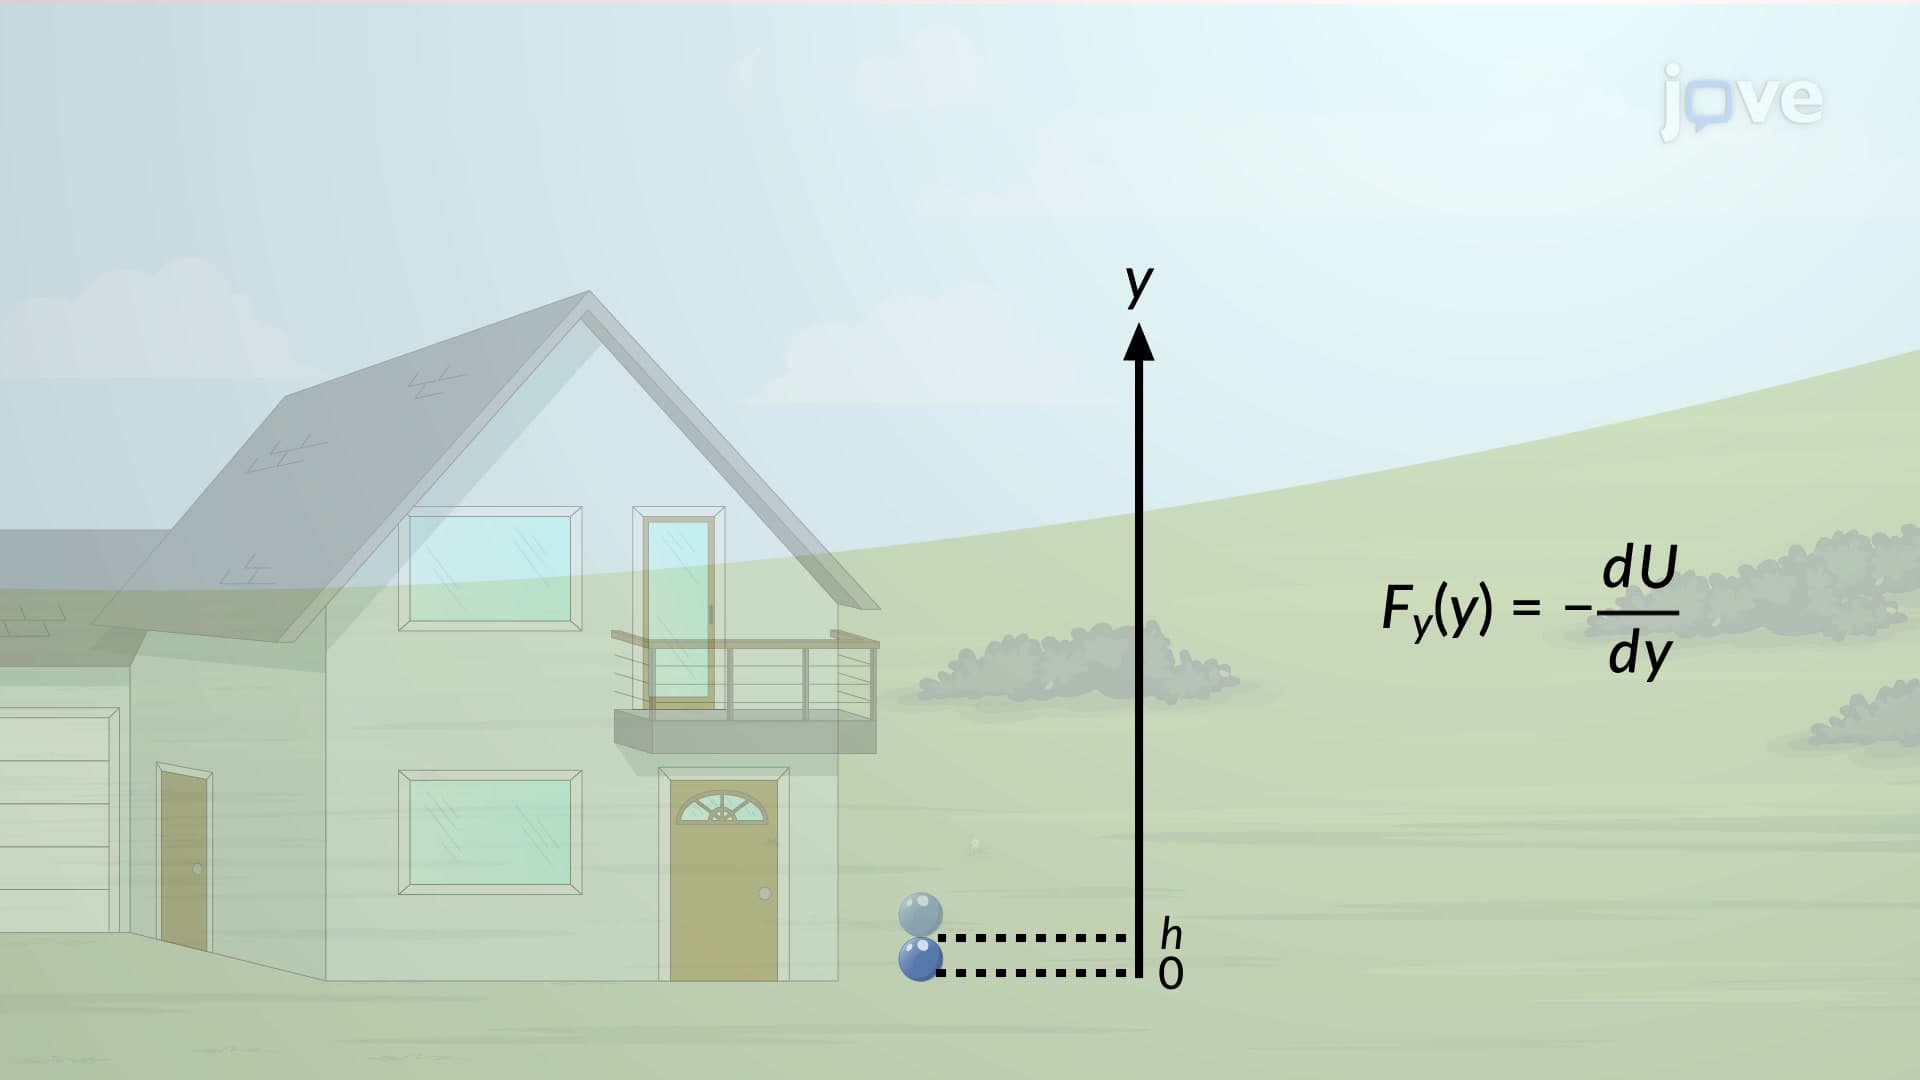 01:13
Force and Potential Energy in One Dimension
 02:13
Energy Diagrams, Transition States, and Intermediates
 02:36
Molecular Geometry and Dipole Moments
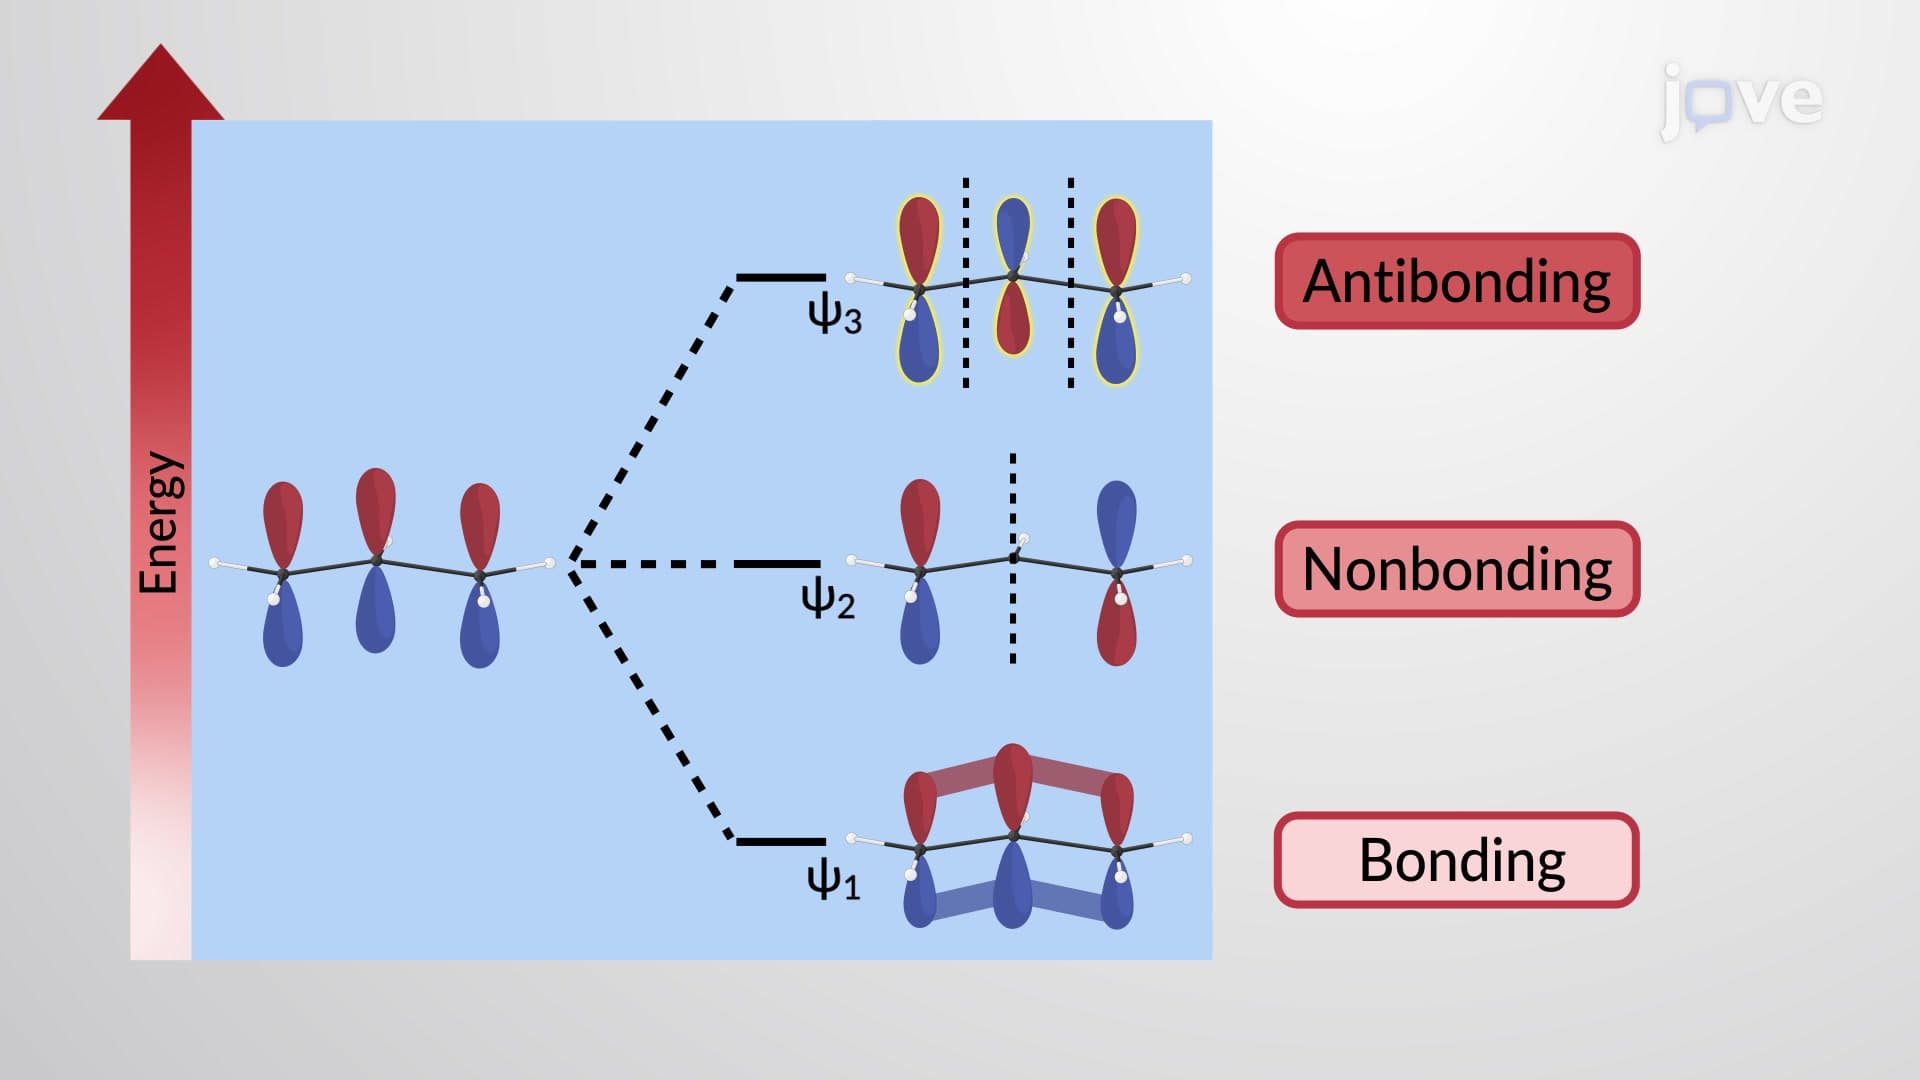 01:18
π Molecular Orbitals of the Allyl Cation and Anion
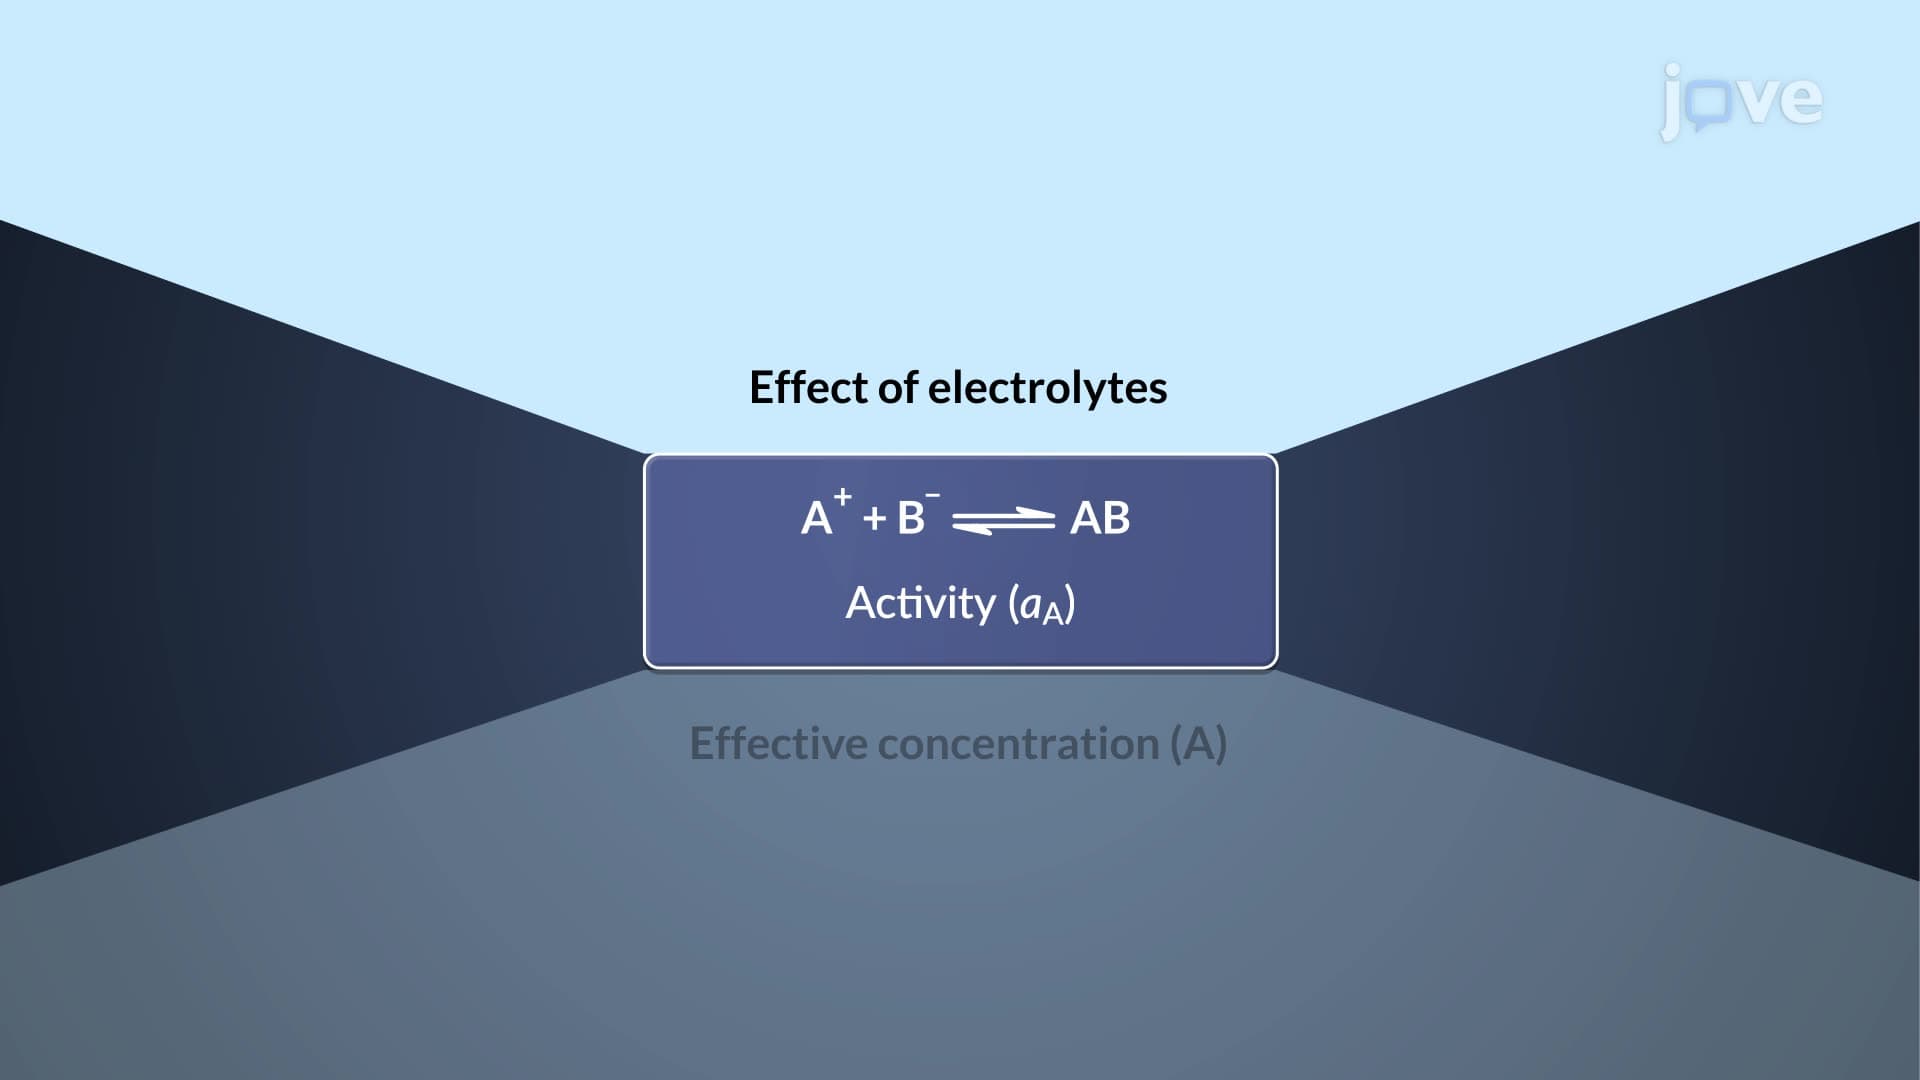 01:10
Thermodynamics: Chemical Potential and Activity
The thermodynamic equilibrium constant is more accurately defined in terms of activity rather than concentration.
 02:30
Stability of Substituted Cyclohexanes
The two chair conformations of cyclohexanes undergo rapid interconversion at room temperature. Both forms have identical energies and stabilities, each comprising equal amounts of the equilibrium mixture. Replacing a hydrogen atom with a functional group makes the two conformations energetically non-equivalent.
For example, in...