Related Experiment Video
Updated: Aug 12, 2026
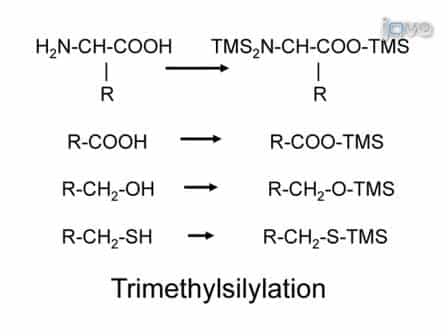

One-step Metabolomics: Carbohydrates, Organic and Amino Acids Quantified in a Single Procedure
Published on: June 25, 2010
[Clinical and biochemical diagnosis of galactosemia among our cases]

Insights
This study identifies key clinical and biochemical markers for diagnosing galactosemia in infants, emphasizing early detection through newborn screening and family history. Prompt diagnosis and a galactose-free diet are crucial for managing this genetic disorder.
Area of Science:
- Biochemistry
- Genetics
- Pediatrics
Context:
- Galactosemia is a rare genetic disorder affecting infants.
- Early diagnosis is critical for preventing severe health complications.
- Newborn screening and family history aid in identifying affected individuals.
Purpose:
- To analyze clinical and biochemical findings in 17 galactosemia cases.
- To evaluate diagnostic methods including enzymatic assays and metabolite levels.
- To highlight the importance of early detection and management.
Summary:
- Seventeen cases of galactosemia from 15 families were diagnosed between 1 day and 11 months of age.
- Common symptoms include hepatomegaly, jaundice, splenomegaly, vomiting, diarrhea, and cataracts.
- Biochemical diagnosis confirmed absent galactose-1-phosphate uridyl transferase activity and elevated galactose-1-phosphate levels in red blood cells.
Impact:
- Establishes characteristic clinical and biochemical profiles for galactosemia diagnosis.
- Reinforces the value of newborn screening and family history in early detection.
- Underscores the necessity of a galactose-free diet for managing galactosemia.
Abstract:
Clinical and biochemical diagnostic studies concerned 17 cases of galactosemia coming from 15 not consauguineous families. Galactosemia was diagnosed between 1-st day and 11-th month of life. Tentative diagnosis based on clinical picture was made in 12 infants, others were detected through family history of galactosemia and/or biochemical newborn screening carried out at the National Research Institute of Mother and Child since 1969. Clinical symptoms of galactosemia occurred in most patients in the first week of life. They were the following (tab. II): hepatomegaly (in 94%), jaundice (81%), splenomegaly (79%), vomitus (62%) and diarrhoea in 56% of patients. Cataract was found in 6 infants (38%). Biochemical diagnosis was based on the results of enzymatic estimation of galactose-1-phosphate uridyl transferase activity in blood, galactose-1-phosphate in red blood cells and galactose in blood and urine. No activity of galactose-1-phosphate uridyl transferase was found in all patients, and the concentration of galactose-1-phosphate was higher than 25 mg/100 ml of red blood cells. High galactose level was observed in blood and urine in all patients with typical clinical course of galactosemia. In 2 patients however without clinical symptoms of the disease only trace amounts of galactose was detected in blood and urine. All these patients were treated with galactose free diet.
Related Concept Videos
 01:27
Glucose Transporters
Facilitated diffusion-glucose transporters (GLUTs) are encoded by the solute-linked carrier (SLC) family 2, subfamily A gene family, or SLC2A. The 14 GLUT protein members are distributed into three classes:
 01:20
Inborn Errors of Metabolism
 01:30
Serum Laboratory Studies, Stool Test, Breath Test
 01:19
Type I Diabetes III: Clinical Manifestations
 01:25
Type II Diabetes Mellitus III: Clinical Manifestations and Diagnosis
 01:25
Diabetic Ketoacidosis l: Introduction