Related Experiment Video
Updated: Apr 3, 2026

Author Spotlight: Decoding Mitochondrial Aging
Published on: June 30, 2023
Mitochondrial dynamics and quality control in Huntington's disease
Pedro Guedes-Dias1, Brígida R Pinho2, Tânia R Soares2
1REQUIMTE/LAQV, Department of Drug Sciences, Faculty of Pharmacy, University of Porto, Porto, Portugal; Department of Cell and Developmental Biology, University College London, London WC1E 6BT, UK.

Insights
Huntington's disease (HD) involves mutant huntingtin (mHtt) disrupting mitochondrial function, especially in the striatum. This review details mHtt's impact on mitochondrial dynamics and quality control, crucial for understanding HD neurodegeneration.
Area of Science:
- Neuroscience
- Genetics
- Cell Biology
Background:
- Huntington's disease (HD) is an inherited neurodegenerative disorder.
- It stems from polyglutamine expansion mutations in the huntingtin protein.
- Mutant huntingtin (mHtt) particularly affects medium spiny neurons in the striatum.
Purpose of the Study:
- To review evidence linking mHtt to mitochondrial dysfunction in HD.
- To focus on brain and neuronal data, especially striatal vulnerability.
- To explore mHtt's effects on mitochondrial dynamics and quality control.
Main Methods:
- Literature review of current evidence.
- Focus on studies examining mitochondrial biogenesis, protein import, and complex assembly.
- Analysis of data on mitochondrial fission, fusion, transport, and mitophagy.
Main Results:
- mHtt induces abnormalities in mitochondrial dynamics and quality control.
- Specific effects include impaired mitochondrial biogenesis, protein import, and complex assembly.
- Dysfunctional mitochondrial transport and mitophagy are also observed.
Conclusions:
- Mitochondrial dysfunction is a key pathogenic mechanism in HD.
- mHtt disrupts multiple aspects of mitochondrial health and function.
- Further research into these mechanisms may reveal therapeutic targets for HD.
Abstract:
Huntington's disease (HD) is an inherited neurodegenerative disorder caused by polyglutamine expansion mutations in the huntingtin protein. Despite its ubiquitous distribution, expression of mutant huntingtin (mHtt) is particularly detrimental to medium spiny neurons within the striatum. Mitochondrial dysfunction has been associated with HD pathogenesis. Here we review the current evidence for mHtt-induced abnormalities in mitochondrial dynamics and quality control, with a particular focus on brain and neuronal data pertaining to striatal vulnerability. We address mHtt effects on mitochondrial biogenesis, protein import, complex assembly, fission and fusion, mitochondrial transport, and on the degradation of damaged mitochondria via autophagy (mitophagy). For an integrated perspective on potentially converging pathogenic mechanisms, we also address impaired autophagosomal transport and abnormal mHtt proteostasis in HD.
More Related Videos
Related Concept Videos
 01:45
Mitochondrial Membranes
 01:19
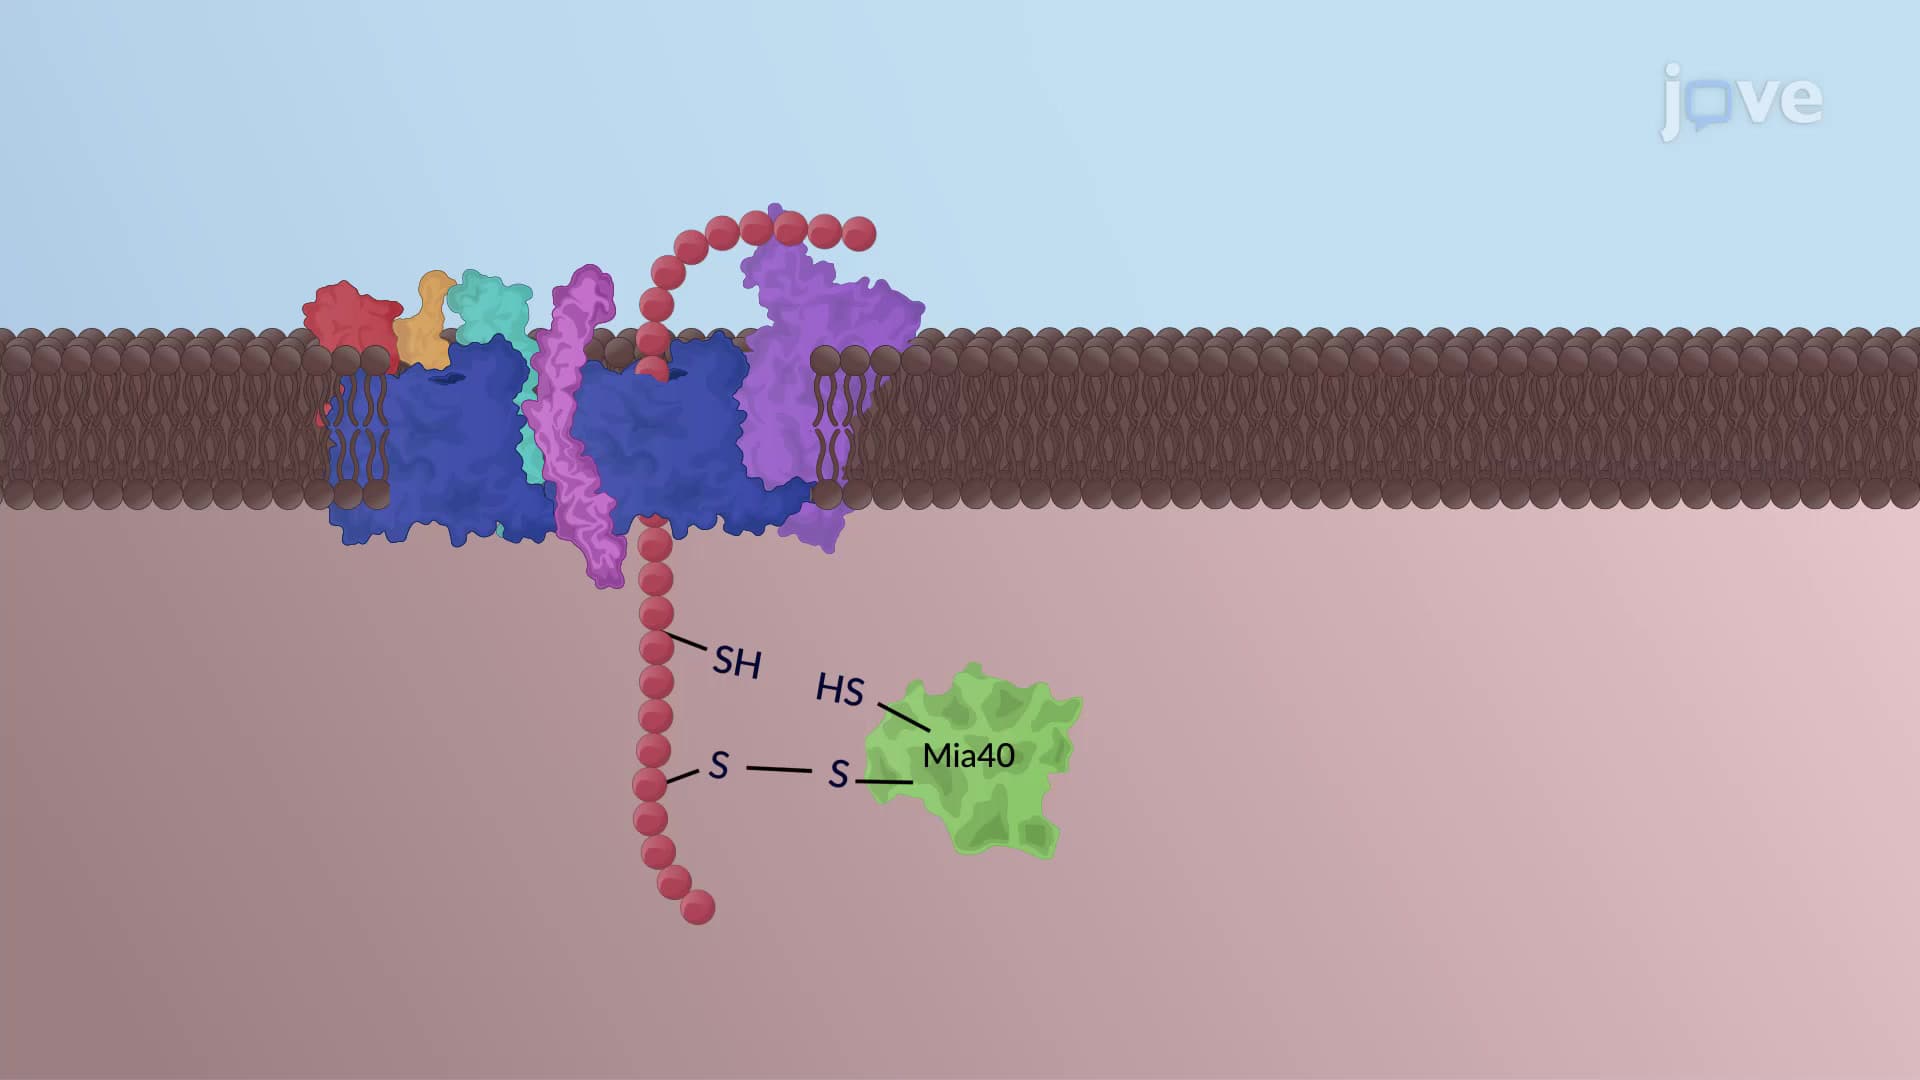
Translocation of Proteins into the Mitochondria
Sorting of outer membrane proteins:
Mitochondrial outer membrane proteins are of two types: the transmembrane, beta-barrel porins, and the membrane-anchored, alpha-helical proteins. Beta-barrel porin precursors are translocated by the TOM complex and inserted into the outer mitochondrial membrane by the SAM complex. In contrast,...
 01:37
Mitochondria
 01:39
Mitochondrial Protein Sorting
Most of these mitochondrial proteins are encoded by the nucleus and imported to the mitochondria as unfolded or loosely folded precursors. Mitochondrial precursors...
 02:59
Animal Mitochondrial Genetics