Free Energy Calculations using a Swarm-Enhanced Sampling Molecular Dynamics Approach
Related Experiment Video
Updated: Apr 1, 2026

Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Kepa K Burusco1, Neil J Bruce1,2, Irfan Alibay1
1Manchester Pharmacy School, University of Manchester, Oxford Road, Manchester, M13 9PT, UK.

Abstract:
Free energy simulations are an established computational tool in modelling chemical change in the condensed phase. However, sampling of kinetically distinct substates remains a challenge to these approaches. As a route to addressing this, we link the methods of thermodynamic integration (TI) and swarm-enhanced sampling molecular dynamics (sesMD), where simulation replicas interact cooperatively to aid transitions over energy barriers. We illustrate the approach by using alchemical alkane transformations in solution, comparing them with the multiple independent trajectory TI (IT-TI) method. Free energy changes for transitions computed by using IT-TI grew increasingly inaccurate as the intramolecular barrier was heightened. By contrast, swarm-enhanced sampling TI (sesTI) calculations showed clear improvements in sampling efficiency, leading to more accurate computed free energy differences, even in the case of the highest barrier height. The sesTI approach, therefore, has potential in addressing chemical change in systems where conformations exist in slow exchange.
More Related Videos
 10:52
Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
 05:51
Isotopic Effect in Double Proton Transfer Process of Porphycene Investigated by Enhanced QM/MM Method
Published on: July 19, 2019
Related Concept Videos
 02:49
Calculating Standard Free Energy Changes
 02:39
Gibbs Free Energy
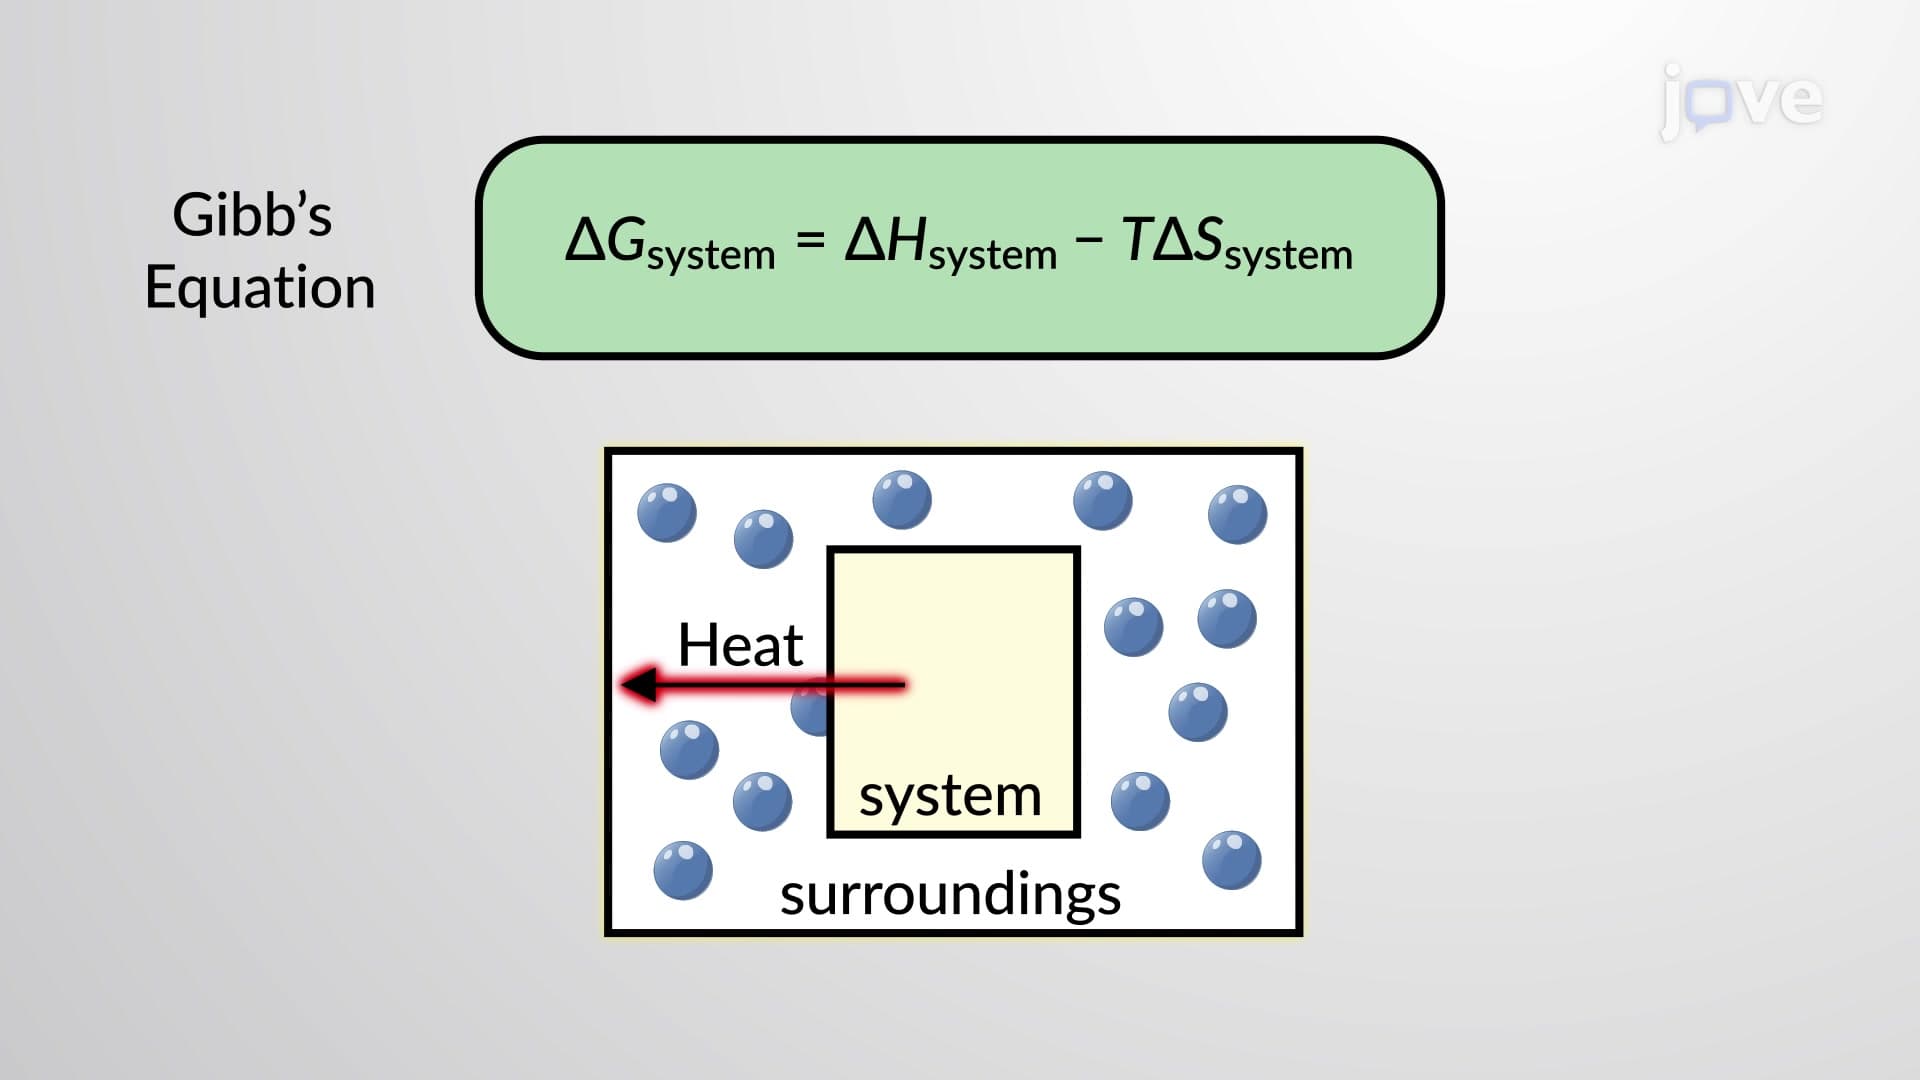 01:05
An Introduction to Free Energy
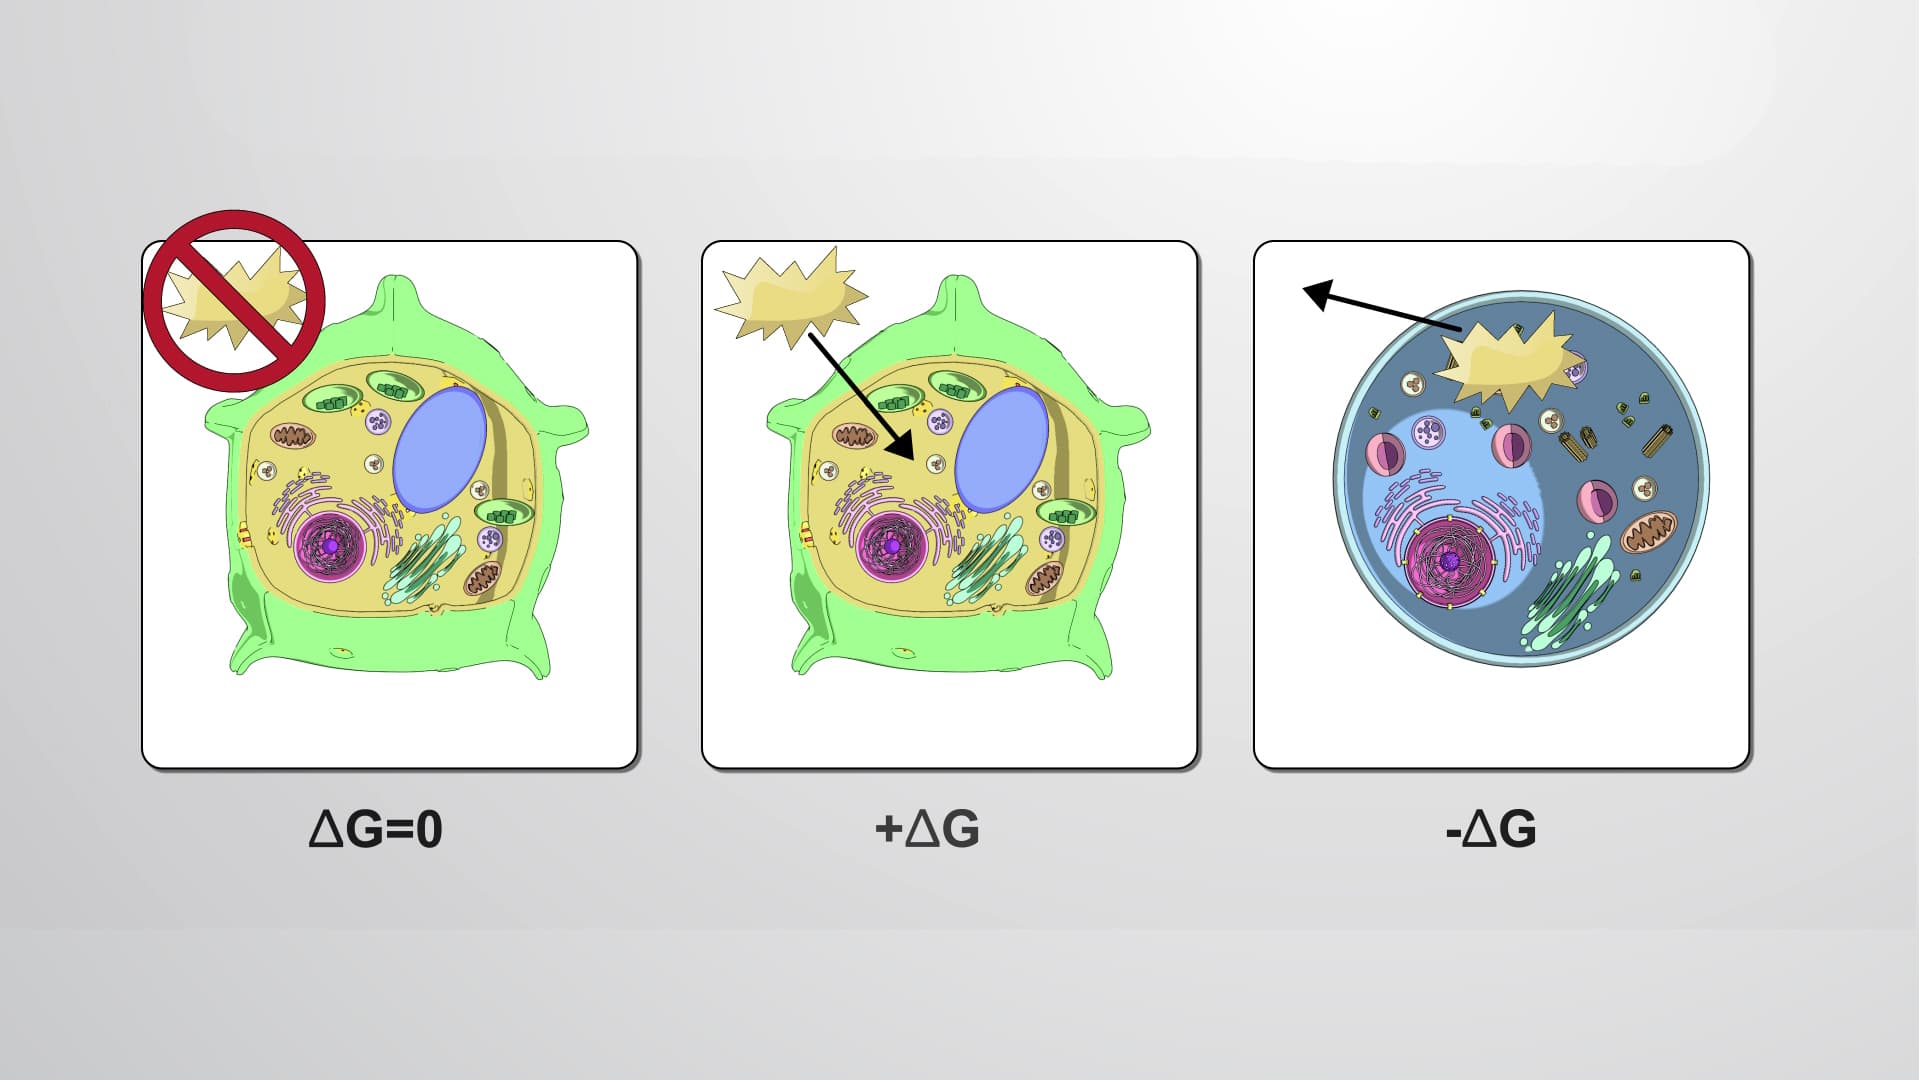 01:21
Free Energy
 03:25
Free Energy Changes for Nonstandard States
 01:21
Molecular Kinetic Energy