Kinetics and specificity of serine proteases in peptide synthesis catalyzed in organic solvents
1Centre de Biochimie et de Biologie Moléculaire du Centre National de la Recherche Scientifique, Marseille, France.

Abstract:
Initial rates of peptide-bond synthesis catalyzed by poly(ethylene glycol)-modified chymotrypsin in benzene were determined using high-performance liquid chromatography. Enzymatic synthesis of N-benzoyl-L-tyrosyl-L-phenylalanine amide from N-benzoyl-L-tyrosine ethyl ester and L-phenylalanine amide was found to obey Michaelis-Menten kinetics an to be consistent with a ping-pong mechanism modified by a hydrolytic branch. The catalytic activity of modified chymotrypsin was dependent on both water concentration and type of organic solvent, the highest synthesis rate being obtained in toluene. Since the chymotrypsin specificity in the organic phase was actually altered, the enzyme's apparent kinetic parameters were determined for different substrates and compared to those obtained with other serine proteases in benzene. Both N-benzoyl-L-tyrosine ethyl ester and N-alpha-benzoyl-L-lysine methyl ester were comparable acyl donors in benzene and the (kcat/Km)app value of modified chymotrypsin was only 10-fold smaller than that obtained with poly(ethylene glycol)-modified trypsin in the synthesis of N-alpha-benzoyl-L-lysyl-L-phenylalanine amide. The change in chymotrypsin specificity was also confirmed through the binding of trypsin inhibitors in benzene. The overall results suggest that hydrophobic bonding between the enzyme and its substrate should not be taken into account during catalysis in the organic phase. In general, if hydrophobic interactions are involved in the binding of substrates to the active site in aqueous media, the replacement of water by hydrophobic solvents will induce some change in enzyme specificity. Moreover, secondary residues of enzyme-binding sites may also exert a significant influence on specificity since, as observed in this study, chymotrypsin exhibited high affinity for cationic substrates and cationic inhibitors as well in apolar solvents.
More Related Videos
 09:47
The Determination of Protease Specificity in Mouse Tissue Extracts by MALDI-TOF Mass Spectrometry: Manipulating PH to Cause Specificity Changes
Published on: May 25, 2018
 06:19
An Inexpensive Adaptation of a Commercial Microwave Reactor for Solid Phase Peptide Synthesis
Published on: November 22, 2024
Related Concept Videos
 02:34
Nucleophilic Substitution Reactions
In 1896, the German chemist Paul Walden discovered that he could interconvert pure enantiomeric (+) and (-) malic acids through a series of reactions. This conversion suggested the involvement of optical inversion during the substitution reaction. Further, in 1930, Sir Christopher Ingold described for the first time two different forms of nucleophilic substitution reactions, which are known as SN1 (nucleophilic substitution unimolecular) and SN2 (nucleophilic substitution...
 02:05
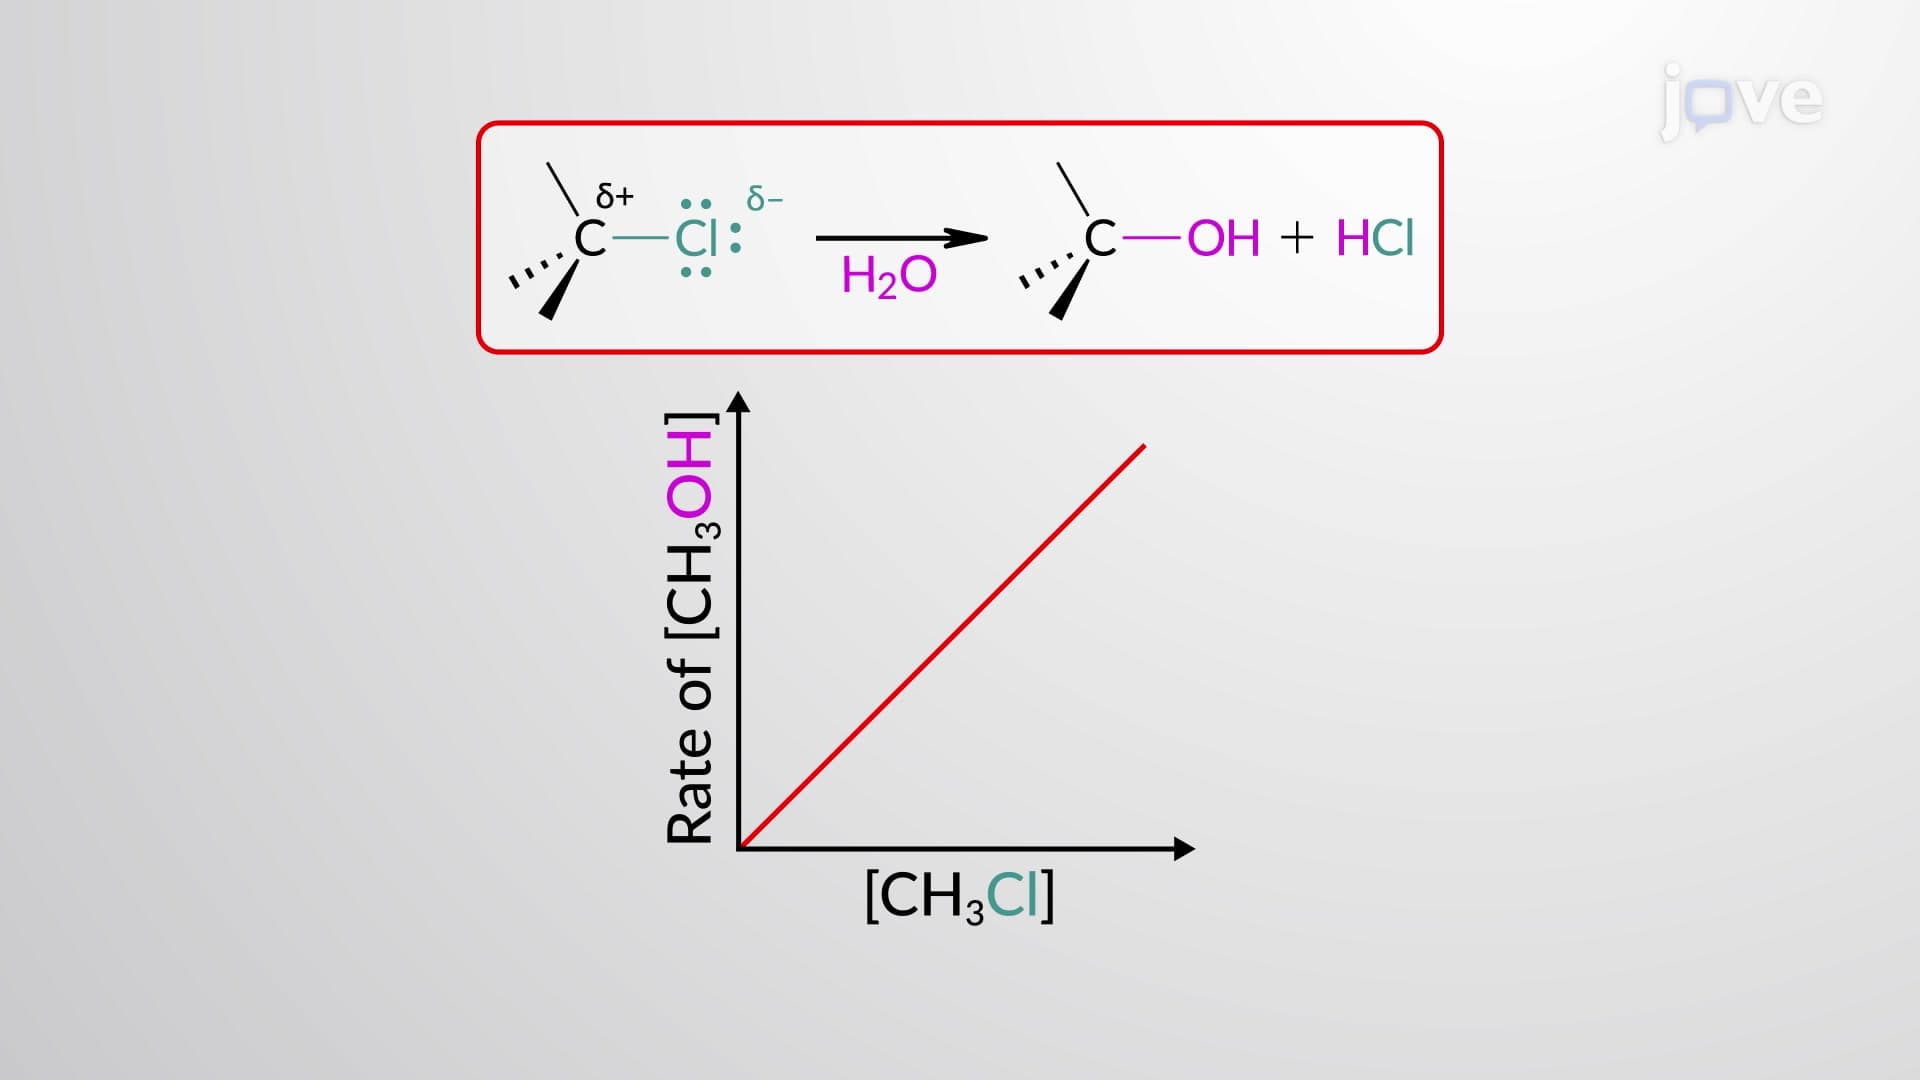
SN1 Reaction: Kinetics
However, Sir Christopher Ingold and Edward D. Hughes, who studied the kinetics of various nucleophilic substitution reactions, noticed that a tertiary alkyl halide does undergo a nucleophilic substitution reaction in the presence of a weak nucleophile. While studying the substitution...
 02:25
SN1 Reaction: Mechanism
Firstly, the haloalkane ionizes to generate a carbocation intermediate and a halide ion. This heterolytic cleavage is highly endothermic with large activation energy. The ionization of the substrate, facilitated by a polar...
 02:27
Predicting Products: SN1 vs. SN2
With increased substitution on the alkyl halide,...
 02:45
E2 Reaction: Kinetics and Mechanism
 01:13
Esters to Carboxylic Acids: Acid-Catalyzed Hydrolysis
During hydrolysis, the ester is first activated towards nucleophilic attack through the protonation of the carboxyl oxygen atom by the acid catalyst. The protonation makes the ester carbonyl carbon more electrophilic. In the next step, water acts as a nucleophile and adds to the...