Related Experiment Video
Updated: Mar 30, 2026


Analyzing Melts and Fluids from Ab Initio Molecular Dynamics Simulations with the UMD Package
Published on: September 17, 2021
Very fast averaging of thermal properties of crystals by molecular simulation
Sabry G Moustafa1, Andrew J Schultz1, David A Kofke1
1Department of Chemical and Biological Engineering, University at Buffalo, The State University of New York, Buffalo, New York 14260-4200, USA.

Abstract:
Knowledge of approximate harmonic behavior of crystals is introduced into a new "mapped averaging" framework to yield alternative expressions for the thermodynamic properties of crystalline systems. The expressions separate the known harmonic behavior from residual averages, which thus encapsulate anharmonic contributions to the properties. With harmonic contributions removed, direct measurement of these anharmonic contributions by molecular simulation can be accomplished without contamination by noise produced by the already-known harmonic behavior. We show with application to the Lennard-Jones model that first-derivative properties (pressure, energy) can be obtained to a given precision via this harmonically mapped averaging at least 10 times faster than by using conventional averaging, and second-derivative properties (e.g., heat capacity) are obtained at least 100 times faster; in more favorable cases, the speedup exceeds a millionfold. Free-energy calculations are accelerated by 50 to 1000 times. Data obtained using these formulations are rigorous and not subject to any added approximation, and in fact are less sensitive to inaccuracies relating to finite-size effects, potential truncation, equilibration, and similar considerations. Moreover, the approach does not require any alteration in how sampling is performed during the simulation, so it may be used with standard Monte Carlo or molecular dynamics methods. However, the mapped averages do require evaluation of first and second derivatives of the intermolecular potential, for evaluation of first and second thermodynamic-derivative properties, respectively. Apart from its usefulness to simulation, the formalism developed here may constitute a basis for new theoretical treatments of crystals.
More Related Videos
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
 05:56
Exploring Caspase Mutations and Post-Translational Modification by Molecular Modeling Approaches
Published on: October 13, 2022
Related Concept Videos
 02:54
Molecular and Ionic Solids
Molecular Solids
Molecular crystalline solids, such as ice, sucrose (table sugar), and iodine, are solids that are composed of neutral molecules as their constituent units. These molecules are held together by weak intermolecular forces such as London dispersion forces, dipole-dipole interactions, or hydrogen bonds, which...
 01:29
Determination of Crystal Structures
 01:21
Polymer Classification: Crystallinity
Crystalline domains are the regions where polymer chains are aligned in an orderly manner and held together in proximity by intermolecular forces. For example, chains in the crystalline domains of polyethylene and nylon are bound together by van der Waals...
 02:39
Phase Transitions: Melting and Freezing
 01:25
Crystal Growth: Principles of Crystallization
Initiating crystallization involves manipulating the concentration of the solute and the temperature of the solution. Since crystal growth occurs when the ratio of concentration and solubility of the solute in the solvent...
 01:16
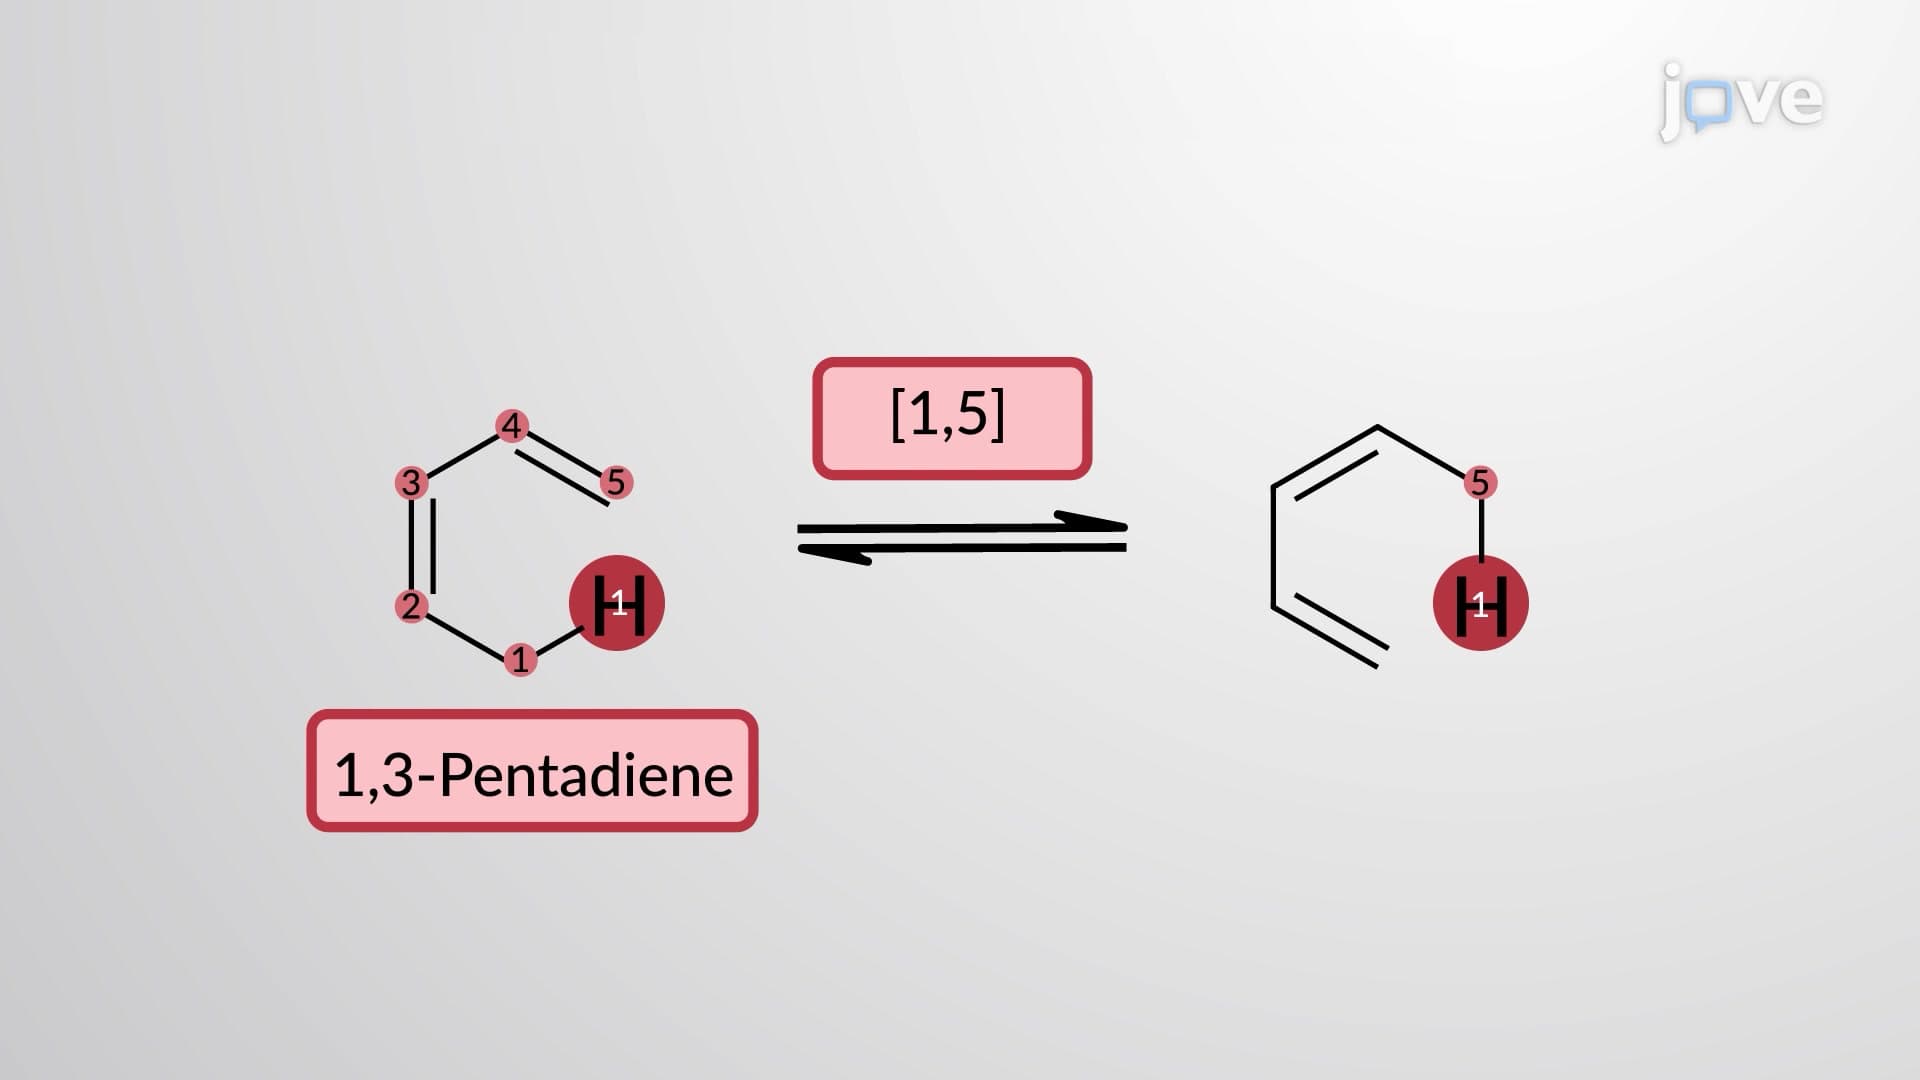
Thermal Sigmatropic Reactions: Overview
Sigmatropic shifts are classified based on an order term [i, j ], where i and j indicate the number of atoms across which each end of the σ bond migrates. Below are examples of a [3,3] sigmatropic shift in 1,5-hexadiene, referred...