A Metabolic Signature of Mitochondrial Dysfunction Revealed through a Monogenic Form of Leigh Syndrome
Julie Thompson Legault1, Laura Strittmatter2, Jessica Tardif3
1Department of Nutrition, Université de Montréal, Montreal, QC H3C 3J7, Canada; Research Centre, Montreal Heart Institute, Montreal, QC H1T 1C8, Canada.

Abstract:
A decline in mitochondrial respiration represents the root cause of a large number of inborn errors of metabolism. It is also associated with common age-associated diseases and the aging process. To gain insight into the systemic, biochemical consequences of respiratory chain dysfunction, we performed a case-control, prospective metabolic profiling study in a genetically homogenous cohort of patients with Leigh syndrome French Canadian variant, a mitochondrial respiratory chain disease due to loss-of-function mutations in LRPPRC. We discovered 45 plasma and urinary analytes discriminating patients from controls, including classic markers of mitochondrial metabolic dysfunction (lactate and acylcarnitines), as well as unexpected markers of cardiometabolic risk (insulin and adiponectin), amino acid catabolism linked to NADH status (α-hydroxybutyrate), and NAD(+) biosynthesis (kynurenine and 3-hydroxyanthranilic acid). Our study identifies systemic, metabolic pathway derangements that can lie downstream of primary mitochondrial lesions, with implications for understanding how the organelle contributes to rare and common diseases.
Related Concept Videos
 01:18
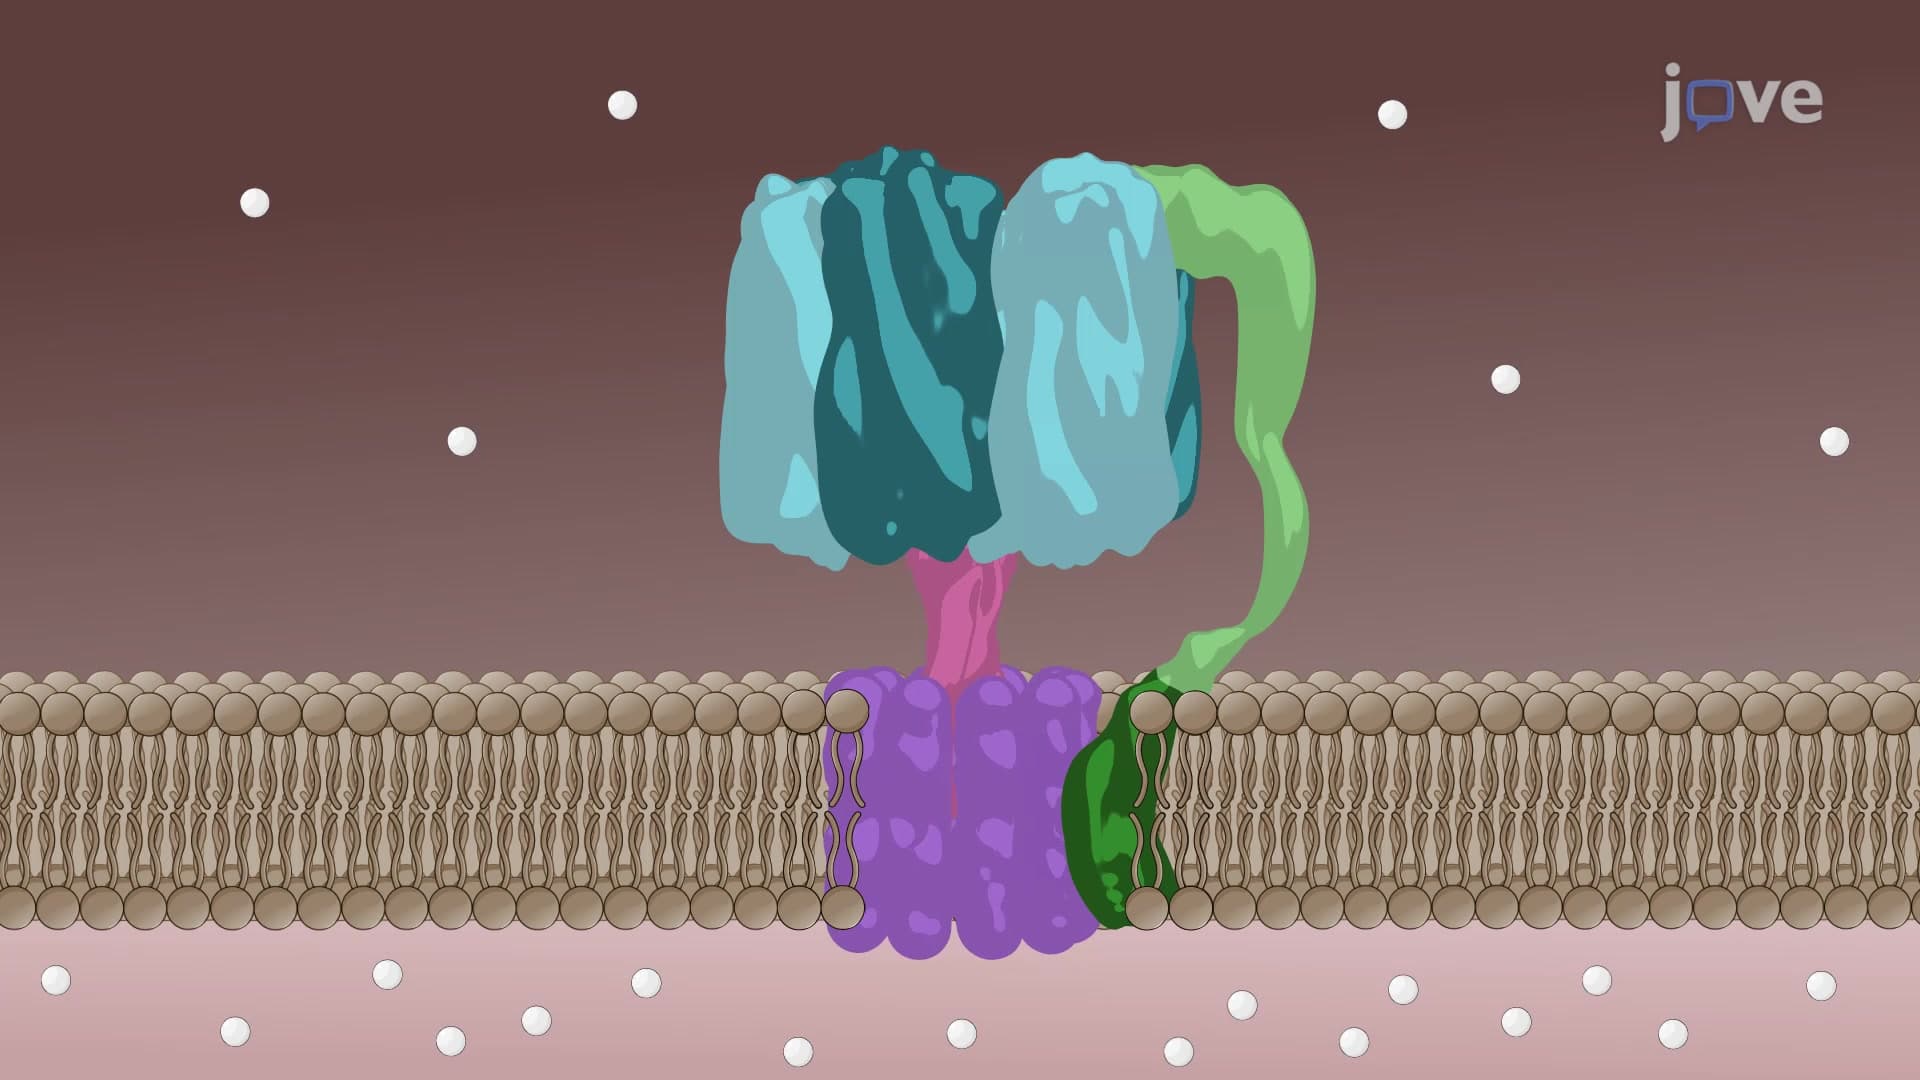
ATP Synthase: Structure
 01:45
Mitochondrial Membranes