Related Experiment Video
Updated: Dec 6, 2025

Author Spotlight: Functionalizing Metal-Organic Frameworks: Advancements, Challenges, and the Power of Post-Synthetic Ligand Exchange
Published on: June 23, 2023
Performance of Density Functionals for Activation Energies of Zr-Mediated Reactions
1Beijing National Laboratory for Molecular Sciences (BNLMS), CAS Key Laboratory of Photochemistry, Institute of Chemistry, Chinese Academy of Sciences , Beijing, 100190, China.

Abstract:
Coupled cluster CCSD(T) calculations with core-valence correlation and complete basis set (CBS) limit extrapolation are used to benchmark the performance of commonly used density functionals in computing energy barriers for Zr-mediated reactions involving zirconocene species. These reactions include (a) insertions of the Zr-H bond of Cp2Zr(H)Cl into C═C, C≡C, and C═O bonds and (b) C-H activations by Zr═N bond in Cp2Zr═NH. The best performing functionals are M06-L, M06, and M06-2X in the M06 series, all having mean unsigned deviations (MUD) less than 2 kcal/mol. The worst performing functional is OLYP, with a distinctly large MUD of more than 10 kcal/mol. Considering also the trends in barrier heights and the systematic barrier height deviation, our best recommended functional is M06-2X. In this work, DFT empirical dispersion correction (DFT-D3) is found to improve the performance of barrier height values for most functionals (especially of OLYP and B3LYP). With DFT empirical dispersion correction, we also recommend M06-2X for reaction barrier calculations of Zr-mediated reactions.
More Related Videos
 16:11
Thermochemical Studies of NiII and ZnII Ternary Complexes Using Ion Mobility-Mass Spectrometry
Published on: June 8, 2022
 10:52
Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
Related Concept Videos
 02:34
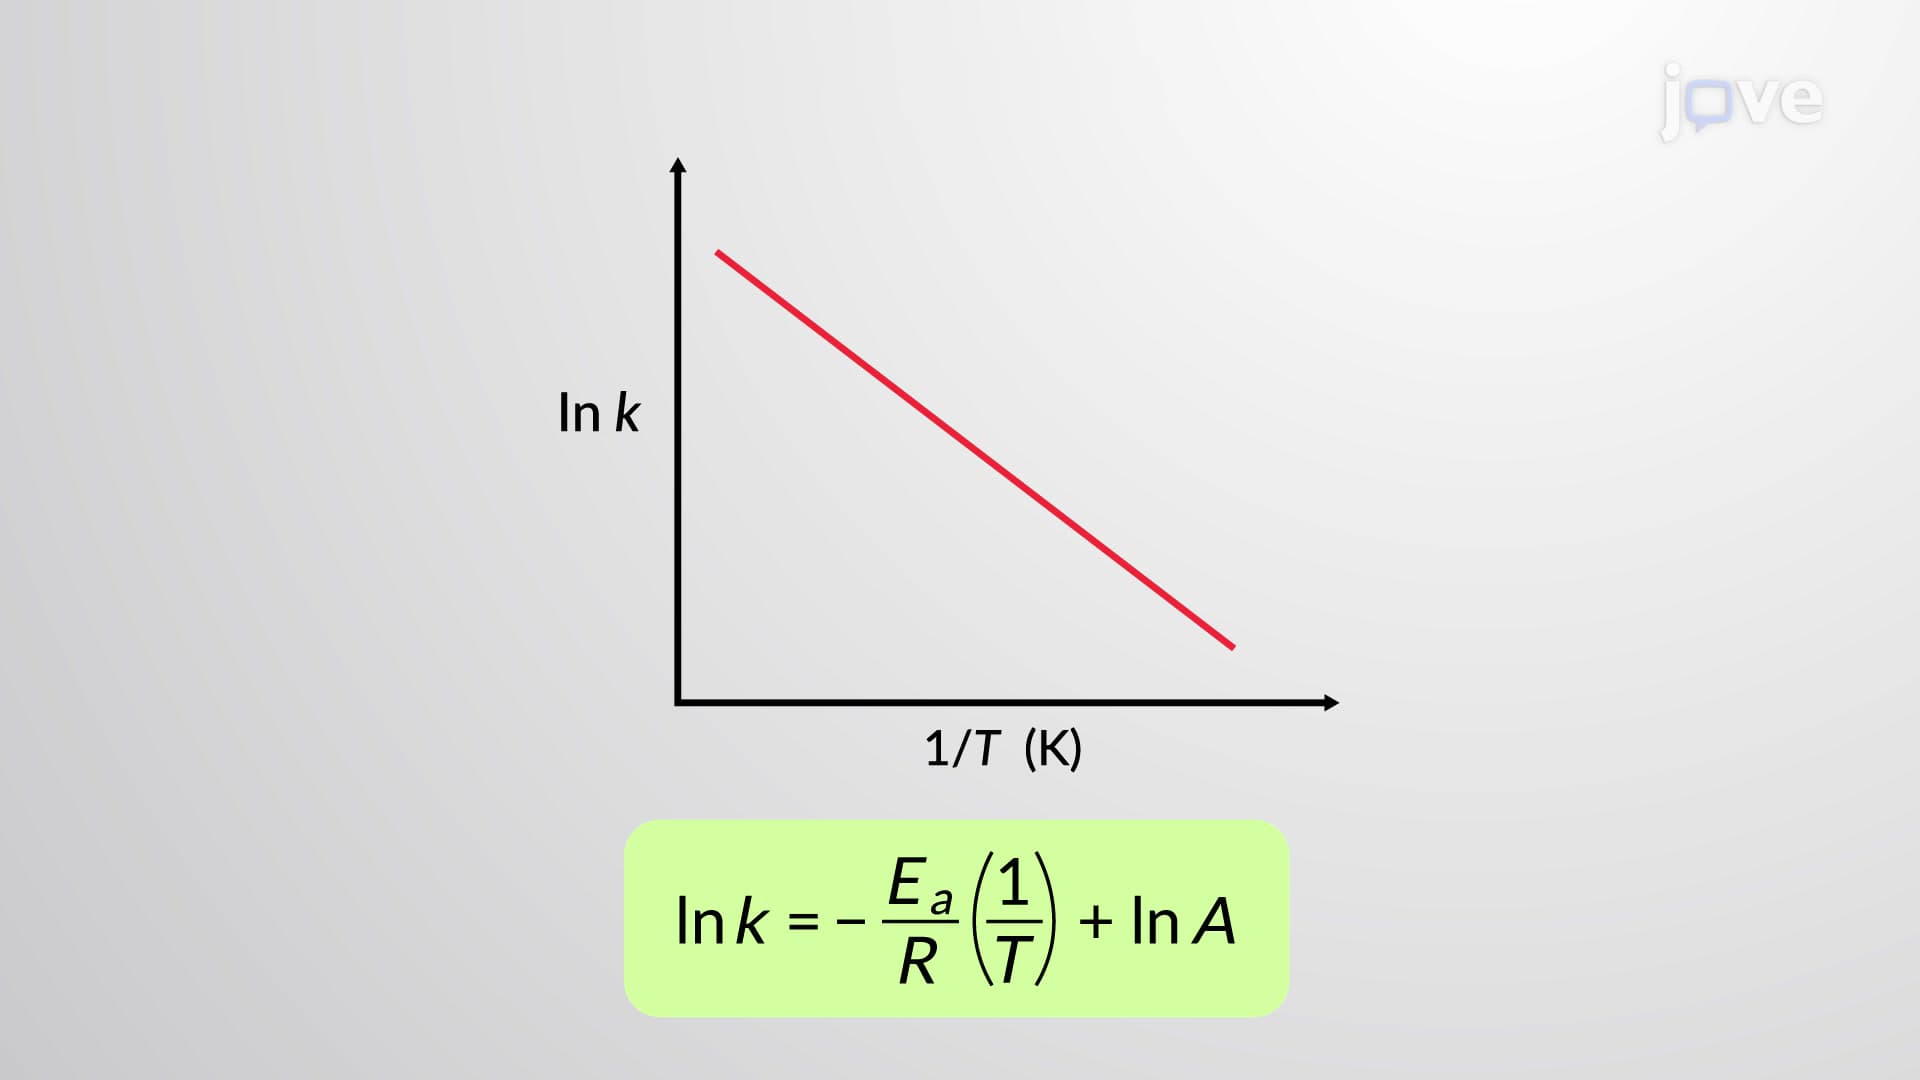
Arrhenius Plots
The Arrhenius equation can be used...
 02:13
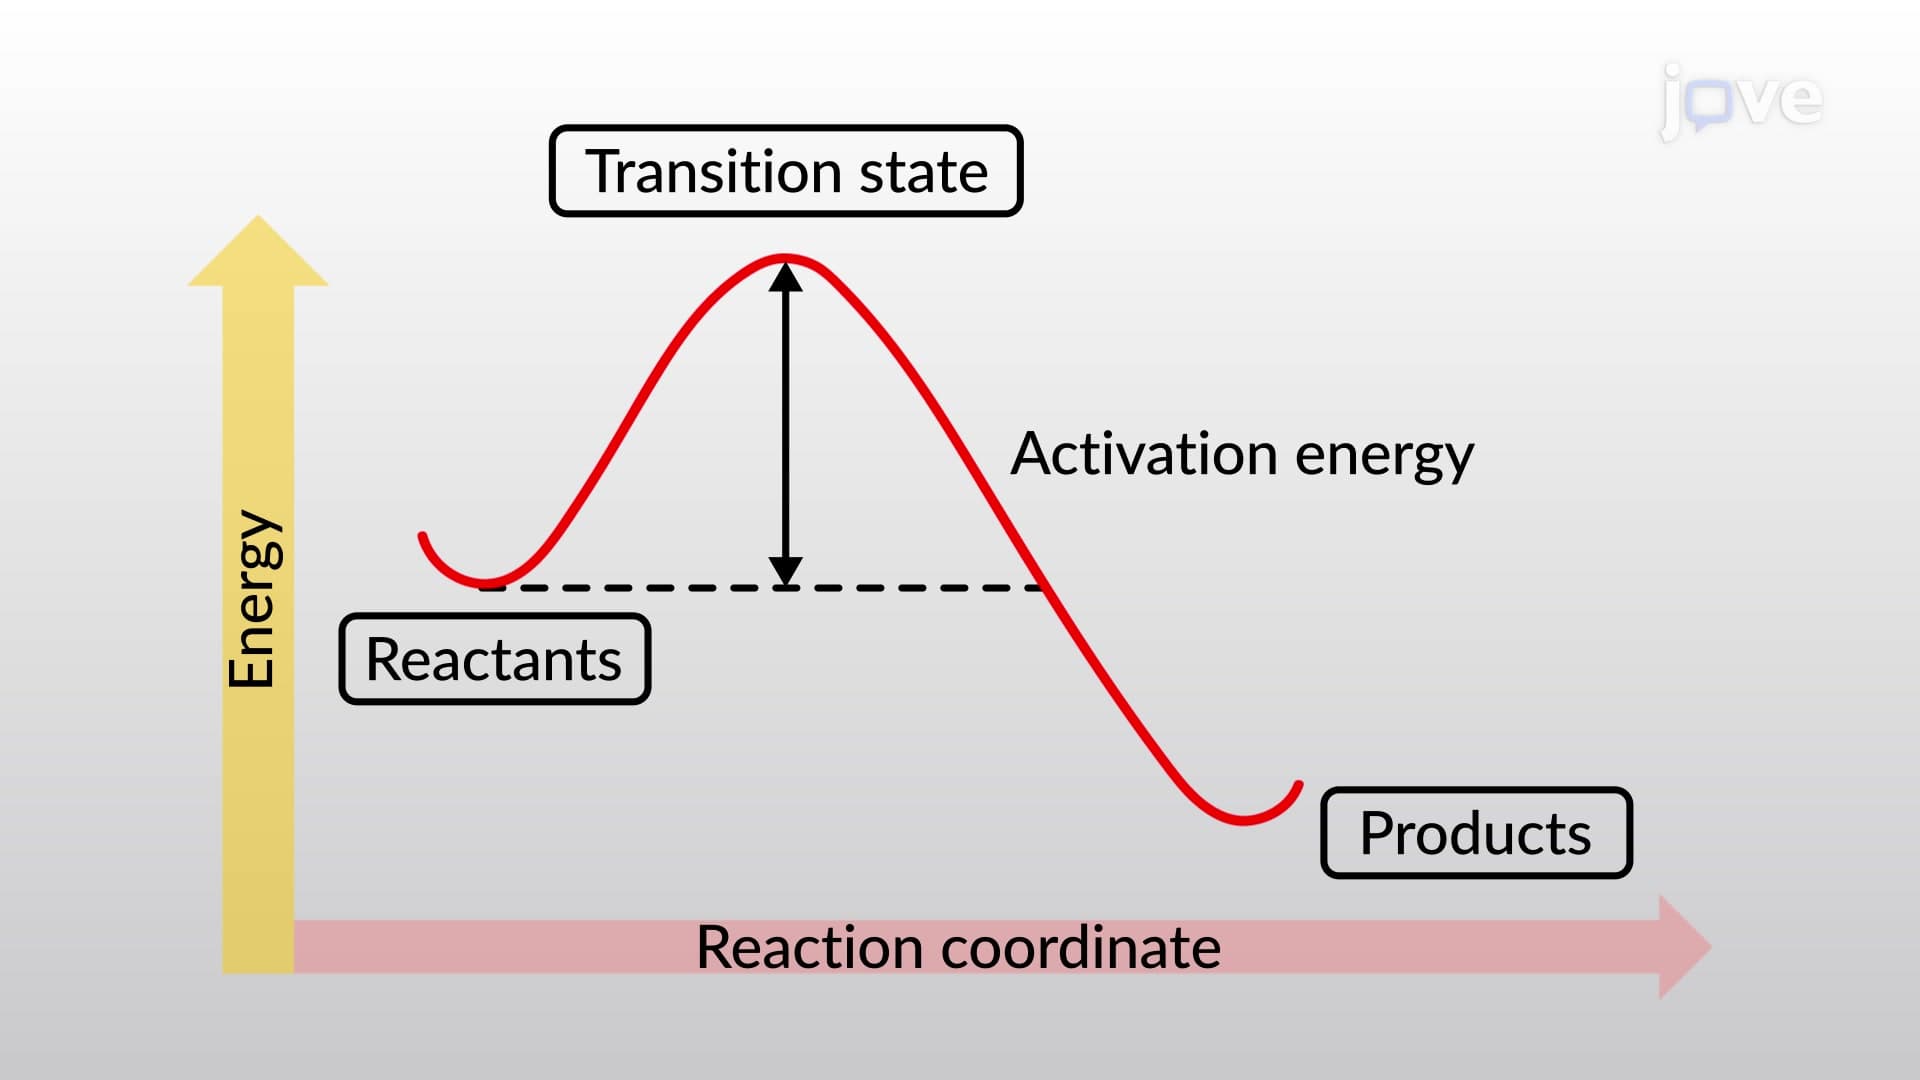
Bond Dissociation Energy and Activation Energy
 01:13
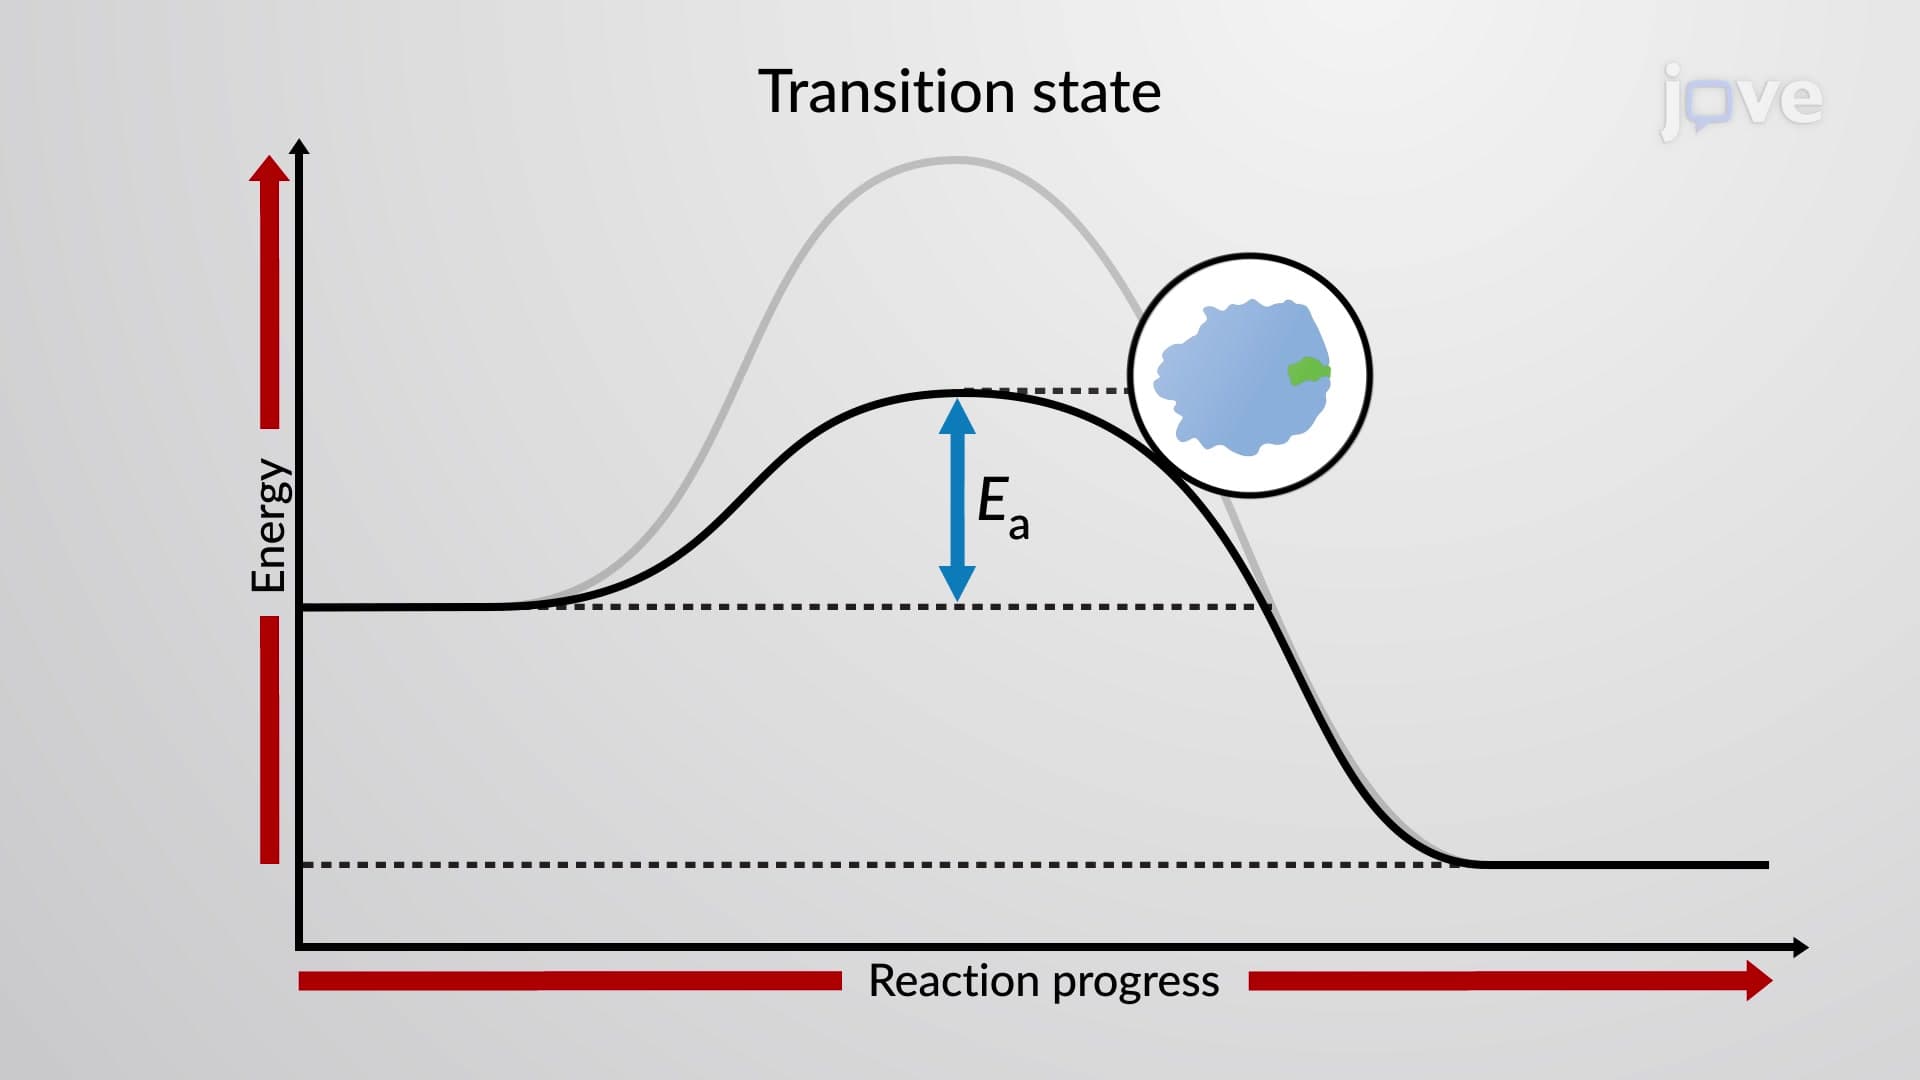
Enzymes and Activation Energy
 02:13
Energy Diagrams, Transition States, and Intermediates
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 02:44
The Born-Haber Cycle