Related Experiment Video
Updated: Mar 30, 2026

Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
First-Principles Elucidation of Atomic Size Effects Using DFT-Chemical Pressure Analysis: Origins of Ca36Sn23's
Joshua Engelkemier1, Veronica M Berns1, Daniel C Fredrickson1
1Department of Chemistry, University of Wisconsin-Madison , 1101 University Avenue, Madison, Wisconsin 53706, United States.

Abstract:
The space requirements of atoms are empirically known to play key roles in determining structure and reactivity across compounds ranging from simple molecules to extended solid state phases. Despite the importance of this concept, the effects of atomic size on stability remain difficult to extract from quantum mechanical calculations. Recently, we outlined a quantitative yet visual and intuitive approach to the theoretical analysis of atomic size in periodic structures: the DFT-Chemical Pressure (DFT-CP) analysis. In this Article, we describe the methodological details of this DFT-CP procedure, with a particular emphasis on refinements of the method to make it useful for a wider variety of systems. A central improvement is a new integration scheme with broader applicability than our earlier Voronoi cell method: contact volume space-partitioning. In this approach, we make explicit our assumption that the pressure at each voxel is most strongly influenced by its two closest atoms. The unit cell is divided into regions corresponding to individual interatomic contacts, with each region containing all points that share the same two closest atoms. The voxel pressures within each contact region are then averaged, resulting in effective interatomic pressures. The method is illustrated through the verification of the role of Ca-Ca repulsion (deduced earlier from empirical considerations by Corbett and co-workers) in the long-period superstructure of the W5Si3 type exhibited by Ca36Sn23.
More Related Videos
 08:42
High-Sensitivity Nuclear Magnetic Resonance at Giga-Pascal Pressures: A New Tool for Probing Electronic and Chemical Properties of Condensed Matter under Extreme Conditions
Published on: October 10, 2014
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
Related Concept Videos
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 03:08
Atomic Radii and Effective Nuclear Charge
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 01:24
Van der Waals Interactions
 01:10
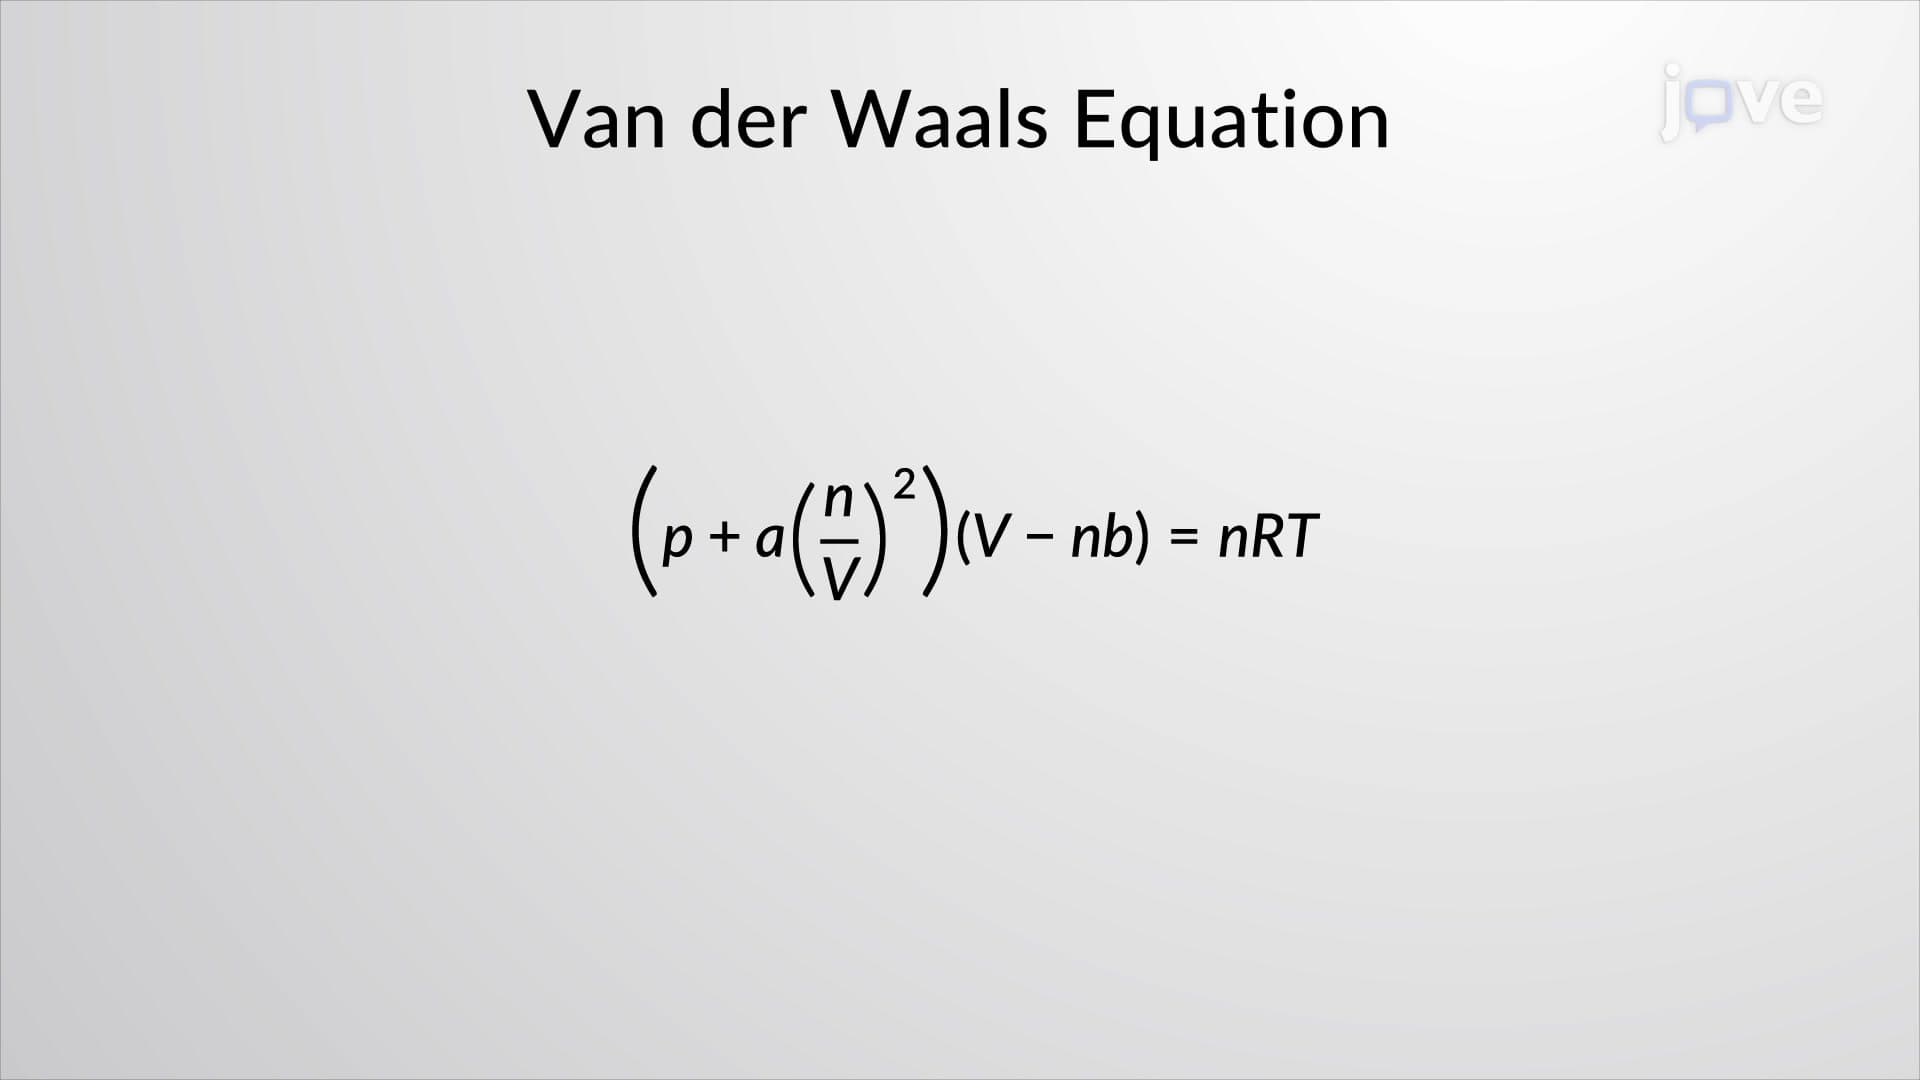
Van der Waals Equation
First, the attractive forces between molecules, which are stronger at higher densities and reduce the pressure, are considered by adding to the pressure a term equal to the square of the molar density multiplied by a positive coefficient a. Second, the volume...
 01:16
IR Spectroscopy: Hooke's Law Approximation of Molecular Vibration
According to Hooke's law, the vibrational frequency is directly proportional to...