Related Experiment Video
Updated: Mar 29, 2026


Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Performance of Density Functional Theory Procedures for the Calculation of Proton-Exchange Barriers: Unusual Behavior
Bun Chan1, Andrew T B Gilbert, Peter M W Gill
1School of Chemistry, University of Sydney , Sydney, NSW 2006, Australia.

Abstract:
We have examined the performance of a variety of density functional theory procedures for the calculation of complexation energies and proton-exchange barriers, with a focus on the Minnesota-class of functionals that are generally highly robust and generally show good accuracy. A curious observation is that M05-type and M06-type methods show an atypical decrease in calculated barriers with increasing proportion of Hartree-Fock exchange. To obtain a clearer picture of the performance of the underlying components of M05-type and M06-type functionals, we have investigated the combination of MPW-type and PBE-type exchange and B95-type and PBE-type correlation procedures. We find that, for the extensive E3 test set, the general performance of the various hybrid-DFT procedures improves in the following order: PBE1-B95 → PBE1-PBE → MPW1-PBE → PW6-B95. As M05-type and M06-type procedures are related to PBE1-B95, it would be of interest to formulate and examine the general performance of an alternative Minnesota DFT method related to PW6-B95.
More Related Videos
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
 16:11
Thermochemical Studies of NiII and ZnII Ternary Complexes Using Ion Mobility-Mass Spectrometry
Published on: June 8, 2022
Related Concept Videos
 01:20
¹³C NMR: Distortionless Enhancement by Polarization Transfer (DEPT)
 01:28
Debye–Huckel–Onsager Conductance Equation
 01:27
Calculations of Electric Potential II
Consider a...
 01:30
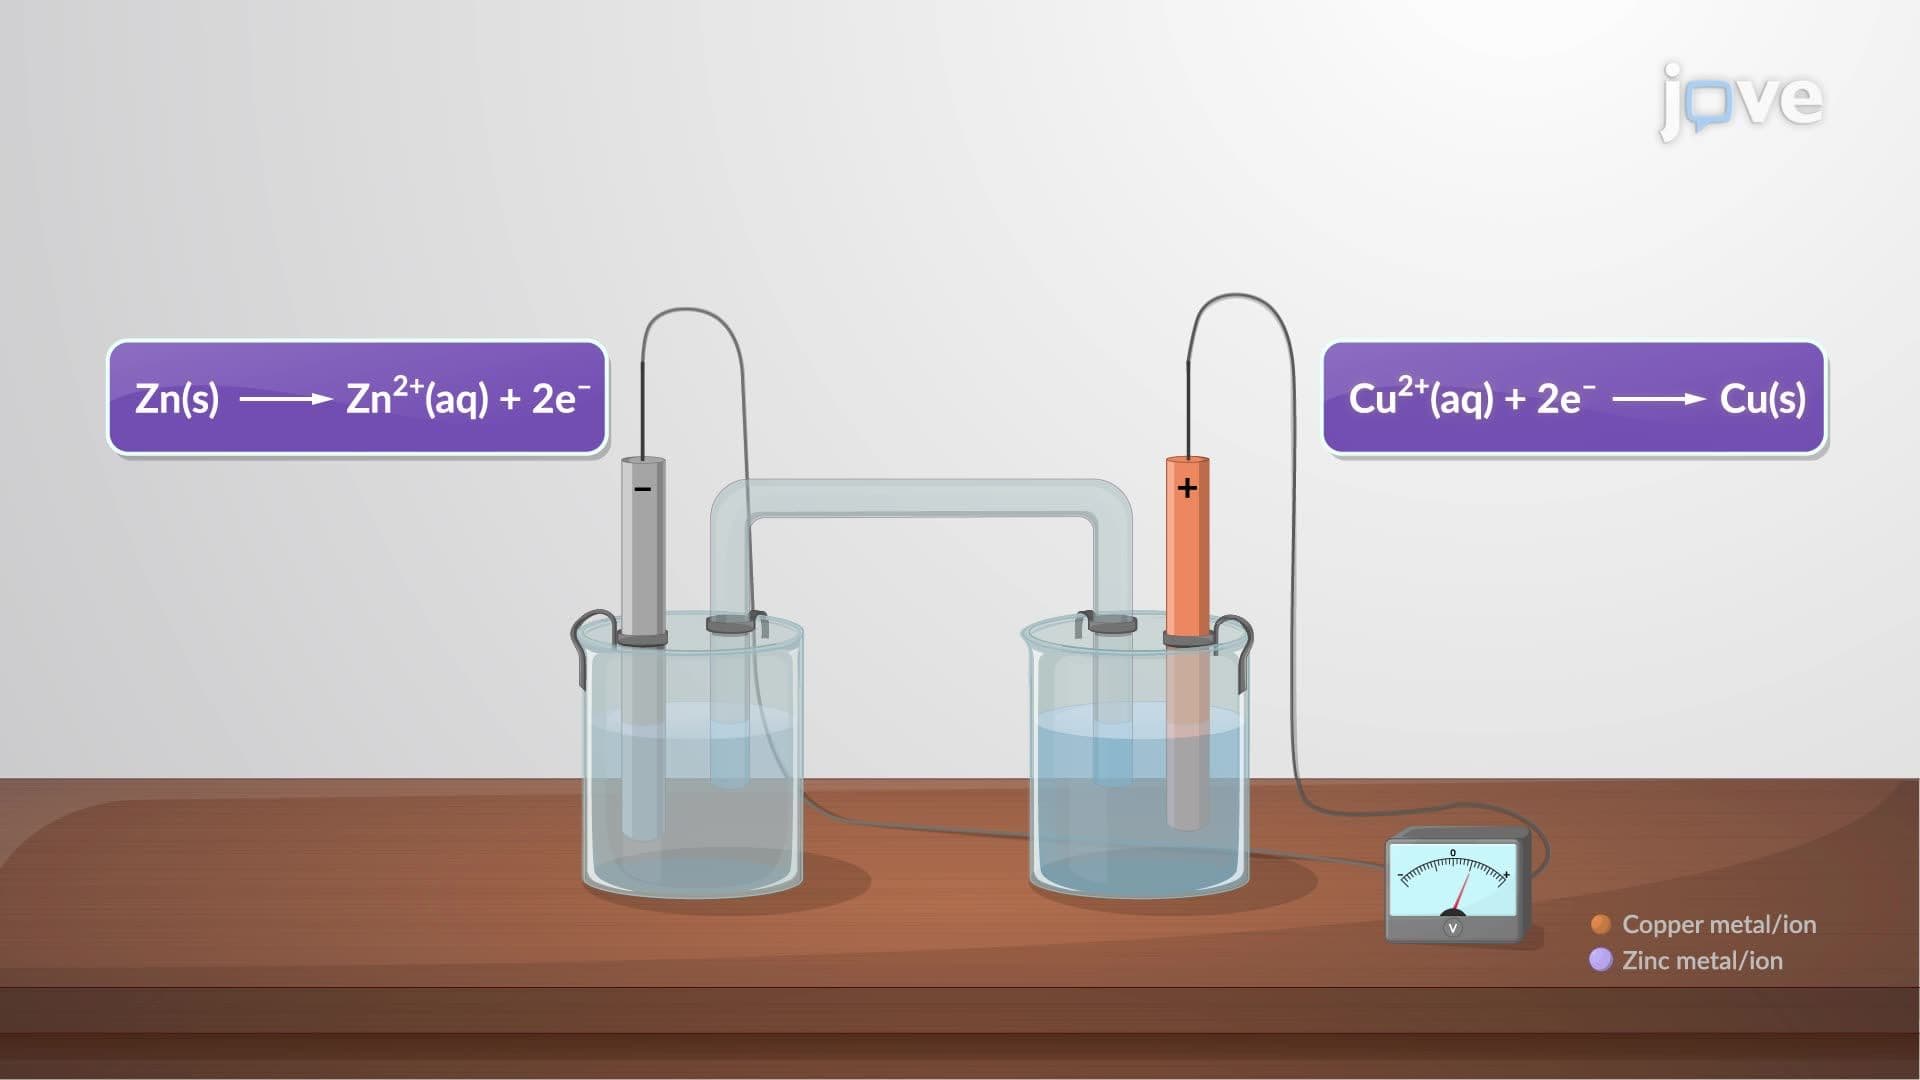
Processes at Electrodes
 00:48
¹H NMR of Labile Protons: Deuterium (²H) Substitution
 03:51
Molecular Orbital Theory II