Related Experiment Video
Updated: Mar 29, 2026
Analyzing Melts and Fluids from Ab Initio Molecular Dynamics Simulations with the UMD Package
Published on: September 17, 2021
Liquid Methanol from DFT and DFT/MM Molecular Dynamics Simulations
Nicolas Sieffert1, Michael Bühl2, Marie-Pierre Gaigeot3,4
1Université Joseph Fourier Grenoble I, CNRS UMR-5250 Département de Chimie Moléculaire, 301 rue de la Chimie, 38041 Grenoble Cedex 9, France.

Abstract:
We present a comparative study of computational protocols for the description of liquid methanol from ab initio molecular dynamics simulations, in view of further applications directed at the modeling of chemical reactivity of organic and organometallic molecules in (explicit) methanol solution. We tested density functional theory molecular dynamics (DFT-MD) in its Car-Parrinello Molecular Dynamics (CPMD) and Quickstep/Born-Oppenheimer MD (CP2K) implementations, employing six popular density functionals with and without corrections for dispersion interactions (namely BLYP, BLYP-D2, BLYP-D3, BP86, BP86-D2, and B97-D2). Selected functionals were also tested within the two QM/MM frameworks implemented in CPMD and CP2K, considering one DFT molecule in a MM environment (described by the OPLS model of methanol). The accuracy of each of these methods at describing the bulk liquid phase under ambient conditions was evaluated by analyzing their ability to reproduce (i) the average structure of the liquid, (ii) the mean squared displacement of methanol molecules, (iii) the average molecular dipole moments, and (iv) the gas-to-liquid red-shift observed in their infrared spectra. We show that it is difficult to find a DFT functional that describes these four properties equally well within full DFT-MD simulations, despite a good overall performance of B97-D2. On the other hand, DFT/MM-MD provides a satisfactory description of the solvent-solute polarization effects with all functionals and thus represents a good alternative for the modeling of methanol solutions in the context of chemical reactivity in an explicit environment.
More Related Videos
 10:52
Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
Related Concept Videos
 02:34
Comparing Intermolecular Forces: Melting Point, Boiling Point, and Miscibility
Temporary attractive forces like dispersion are present in all molecules, whether they are polar or nonpolar. They...
 02:56
Intermolecular Forces and Physical Properties
 02:40
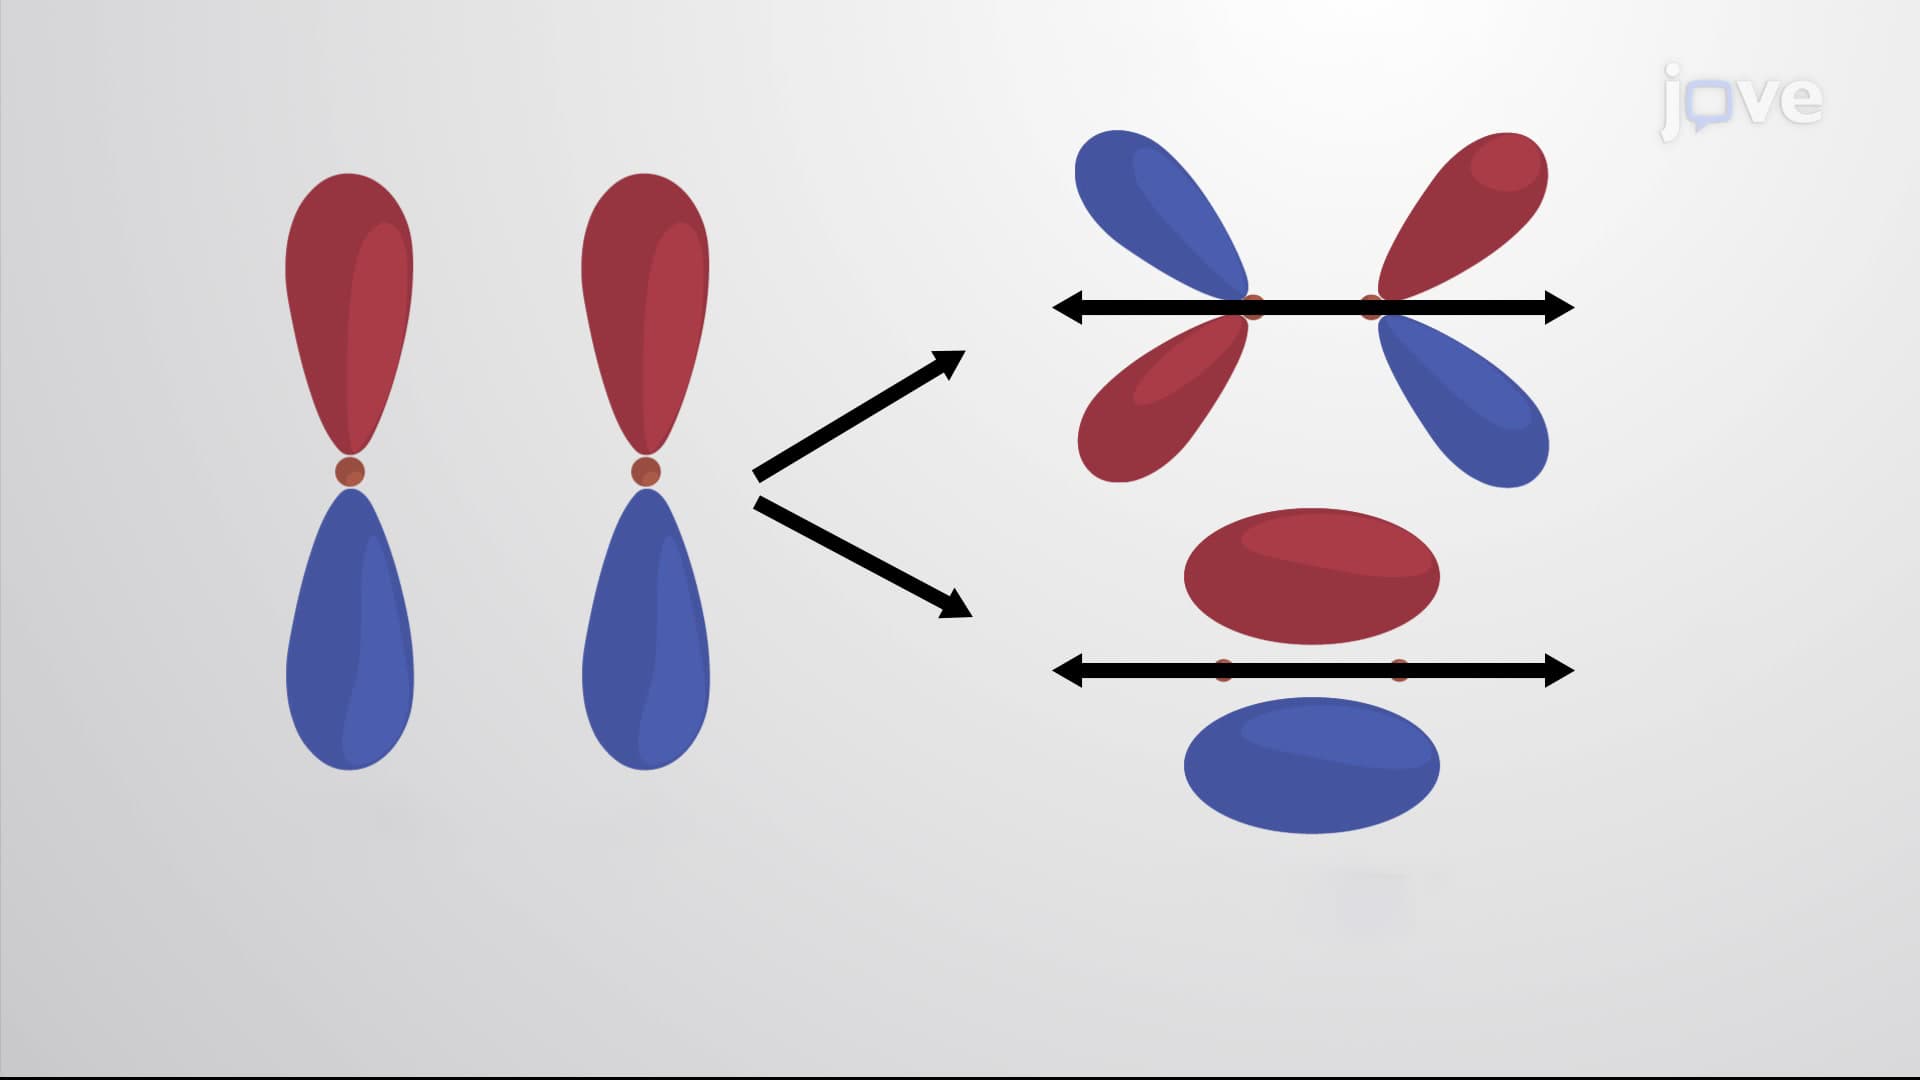
MO Theory and Covalent Bonding
 02:00
Molecular Models
 02:26
Molecular Comparison of Gases, Liquids, and Solids
 04:01
Real Gases: Effects of Intermolecular Forces and Molecular Volume Deriving Van der Waals Equation