Related Experiment Video
Updated: Mar 29, 2026

Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Accurate Prediction of Noncovalent Interaction Energies with the Effective Fragment Potential Method: Comparison of
Joanna C Flick1, Dmytro Kosenkov1, Edward G Hohenstein2
1Department of Chemistry, Purdue University , West Lafayette, Indiana 47907, United States.

Abstract:
Noncovalent interactions play an important role in the stabilization of biological molecules. The effective fragment potential (EFP) is a computationally inexpensive ab initio-based method for modeling intermolecular interactions in noncovalently bound systems. The accuracy of EFP is benchmarked against the S22 and S66 data sets for noncovalent interactions [Jurečka, P.; Šponer, J.; Černý, J.; Hobza, P. Phys. Chem. Chem. Phys.2006, 8, 1985; Řezáč, J.; Riley, K. E.; Hobza, P. J. Chem. Theory Comput.2011, 7, 2427]. The mean unsigned error (MUE) of EFP interaction energies with respect to coupled-cluster singles, doubles, and perturbative triples in the complete basis set limit [CCSD(T)/CBS] is 0.9 and 0.6 kcal/mol for S22 and S66, respectively, which is similar to the MUE of MP2 and SCS-MP2 for the same data sets, but with a greatly reduced computational expense. Moreover, EFP outperforms classical force fields and popular DFT functionals such as B3LYP and PBE, while newer dispersion-corrected functionals provide a more accurate description of noncovalent interactions. Comparison of EFP energy components with the symmetry-adapted perturbation theory (SAPT) energies for the S22 data set shows that the main source of errors in EFP comes from Coulomb and polarization terms and provides a valuable benchmark for further improvements in the accuracy of EFP and force fields in general.
More Related Videos
Related Concept Videos
 02:27
Predicting Molecular Geometry
 01:13
Force and Potential Energy in One Dimension
 02:13
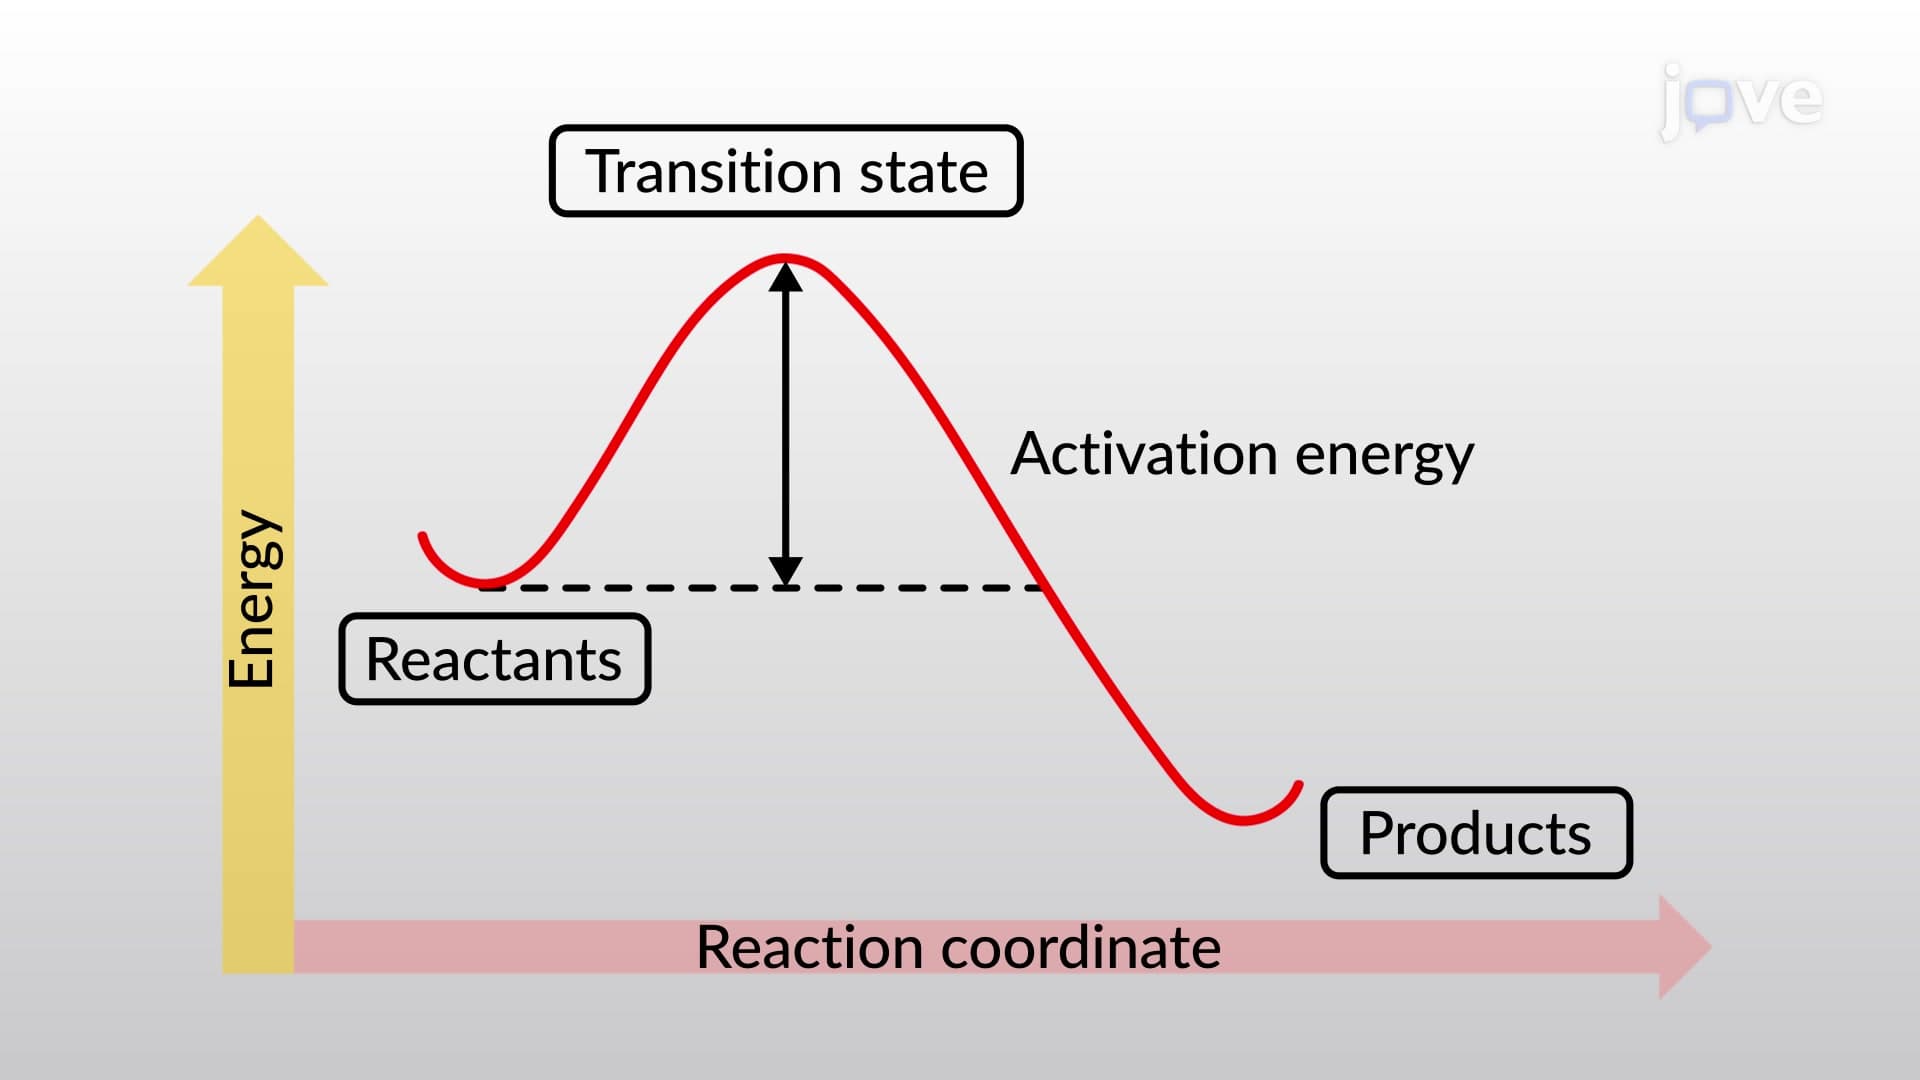
Bond Dissociation Energy and Activation Energy
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 02:35
Noncovalent Attractions in Biomolecules
02:35Noncovalent Attractions in Biomolecules
Four types of noncovalent interactions are hydrogen bonds, van der Waals forces, ionic bonds, and hydrophobic interactions.
Hydrogen bonding results from the electrostatic attraction of a hydrogen atom covalently bonded to a strong-electronegative atom like oxygen,...