

Eigensystem Representation of the Electronic Susceptibility Tensor for Intermolecular Interactions within Density
A Scherrer1, V Verschinin1, D Sebastiani1
1Dahlem Center for Complex Quantum Systems, Physics Department, Free University Berlin , Arnimallee 14, 14195 Berlin, Germany.

Abstract:
We present an efficient implementation of the electronic susceptibility tensor within density functional theory. The susceptibility is represented by means of its eigensystem, which is computed using an iterative Lanczos diagonalization technique for the susceptibility tensor within density functional perturbation theory. We show that a representation in a finite basis of eigenstates is sufficiently accurate to compute the linear response of the electronic density to external potentials. Once the eigensystem representation is computed, the actual response computation can be done at very low computational cost. The method is applied to the water molecule in a dipole field as a benchmark system. The results illustrate the potential of the approach for the first-principles calculation of supramolecular interactions in complex disordered systems in the condensed phase.
More Related Videos
 08:04
Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
 06:53
Author Spotlight: Magnetometric Characterization of Intermediates in the Solid-State Electrochemistry of Redox-Active Metal-Organic Frameworks
Published on: June 9, 2023
Related Concept Videos
 02:42
Valence Bond Theory
 02:45
Valence Bond Theory
 01:28
Debye–Huckel–Onsager Conductance Equation
 01:24
Van der Waals Interactions
 02:36
Molecular Geometry and Dipole Moments
 01:10
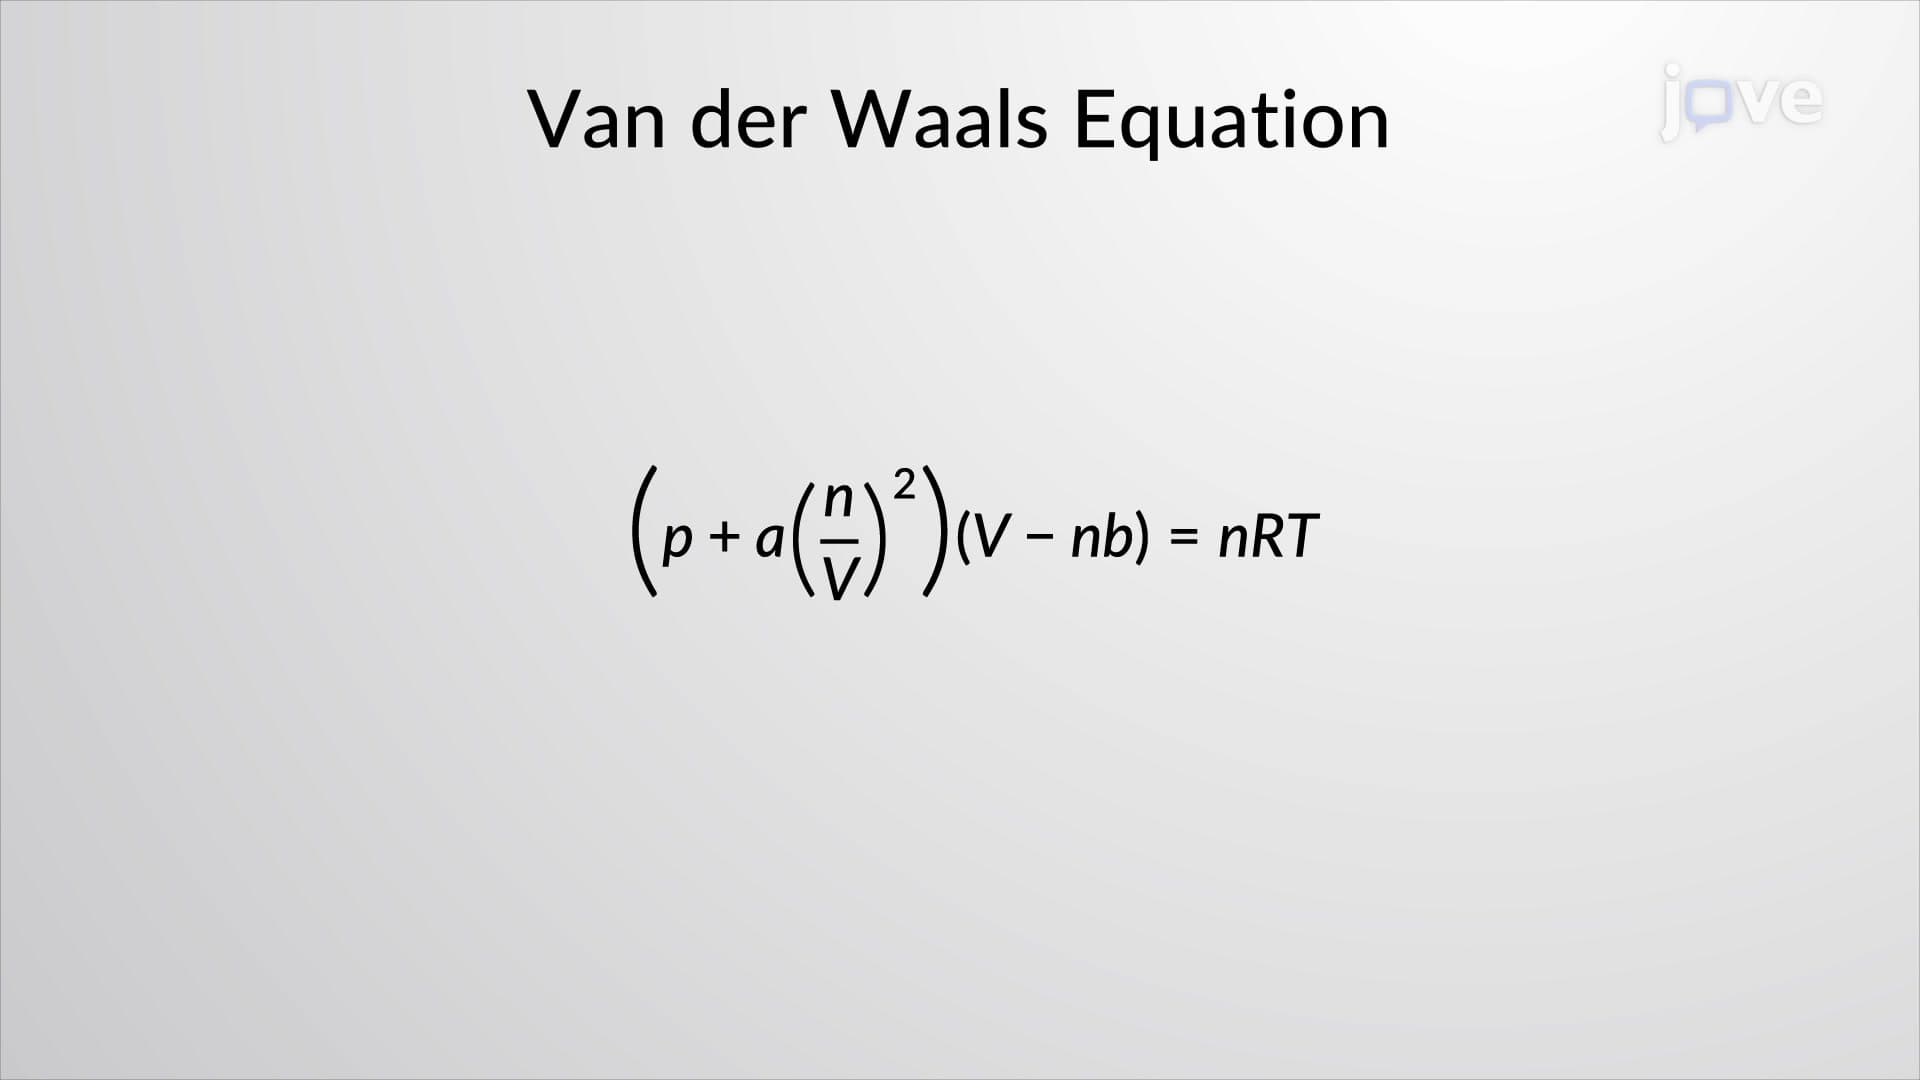
Van der Waals Equation
First, the attractive forces between molecules, which are stronger at higher densities and reduce the pressure, are considered by adding to the pressure a term equal to the square of the molar density multiplied by a positive coefficient a. Second, the volume...