Related Experiment Video
Updated: Mar 29, 2026


Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Accuracy of Calculated Chemical Shifts in Carbon 1s Ionization Energies from Single-Reference ab Initio Methods and
Alf Holme1, Knut J Børve1, Leif J Sæthre1
1Department of Chemistry, University of Bergen , NO-5007 Bergen, Norway.

Abstract:
A database of 77 adiabatic carbon 1s ionization energies has been prepared, covering linear and cyclic alkanes and alkenes, linear alkynes, and methyl- or fluoro-substituted benzenes. Individual entries are believed to carry uncertainties of less than 30 meV in ionization energies and less than 20 meV for shifts in ionization energies. The database provides an unprecedented opportunity for assessing the accuracy of theoretical schemes for computing inner-shell ionization energies and their corresponding chemical shifts. Chemical shifts in carbon 1s ionization energies have been computed for all molecules in the database using Hartree-Fock, Møller-Plesset (MP) many-body perturbation theory of order 2 and 3 as well as various approximations to full MP4, and the coupled-cluster approximation with single- and double-excitation operators (CCSD) and also including a perturbational estimate of the energy effect of triple-excitation operators (CCSD(T)). Moreover, a wide range of contemporary density functional theory (DFT) methods are also evaluated with respect to computing experimental shifts in C1s ionization energies. Whereas the top ab initio methods reproduce the observed shifts almost to within the experimental uncertainty, even the best-performing DFT approaches meet with twice the root-mean-squared error and thrice the maximum error compared to CCSD(T). However, a number of different density energy functionals still afford sufficient accuracy to become tools in the analysis of complex C1s photoelectron spectra.
More Related Videos
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
 08:04
Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
Related Concept Videos
 01:17
Chemical Shift: Internal References and Solvent Effects
The internal reference compound generally used in NMR spectroscopy is tetramethylsilane (TMS). TMS is preferred because it is chemically inert, soluble in NMR solvents, and easily removable. Also, the highly shielded methyl protons in TMS yield an intense...
 01:15
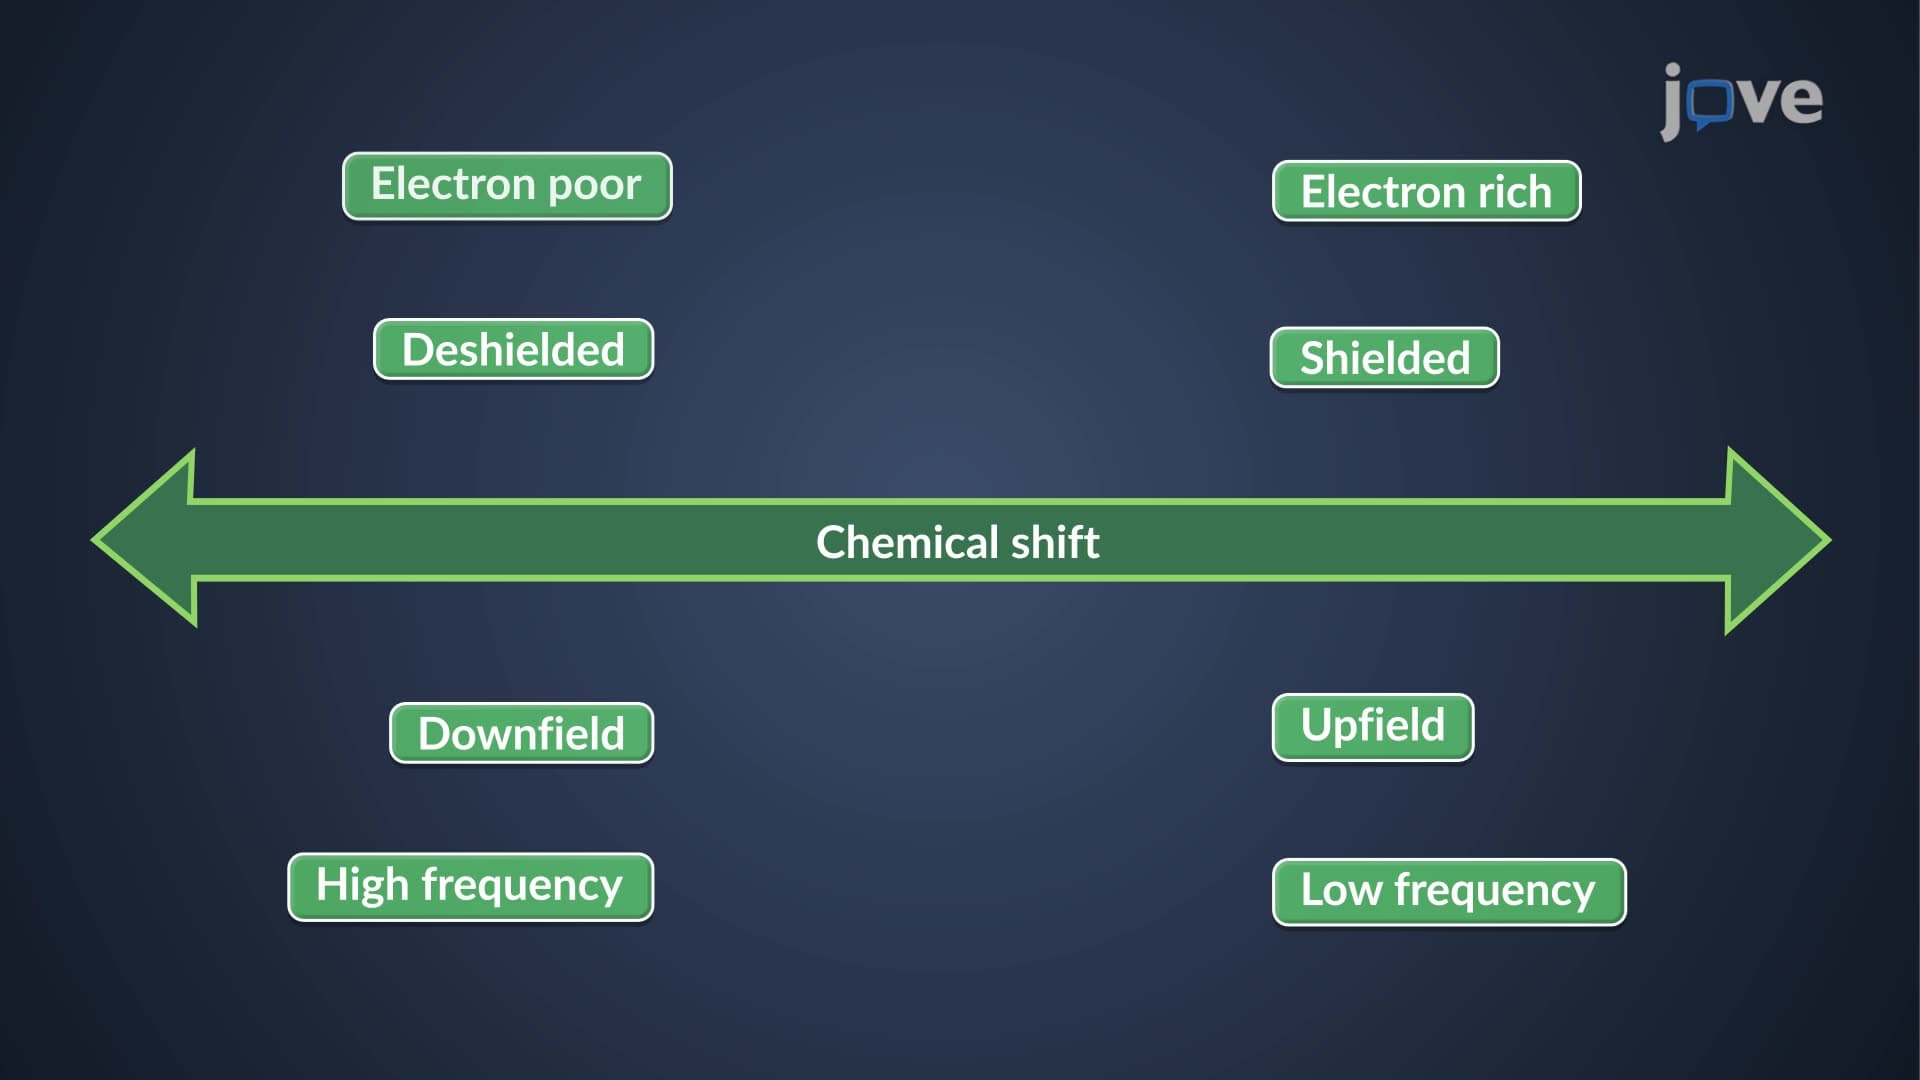
NMR Spectroscopy: Chemical Shift Overview
For instance, the proton...
 01:10
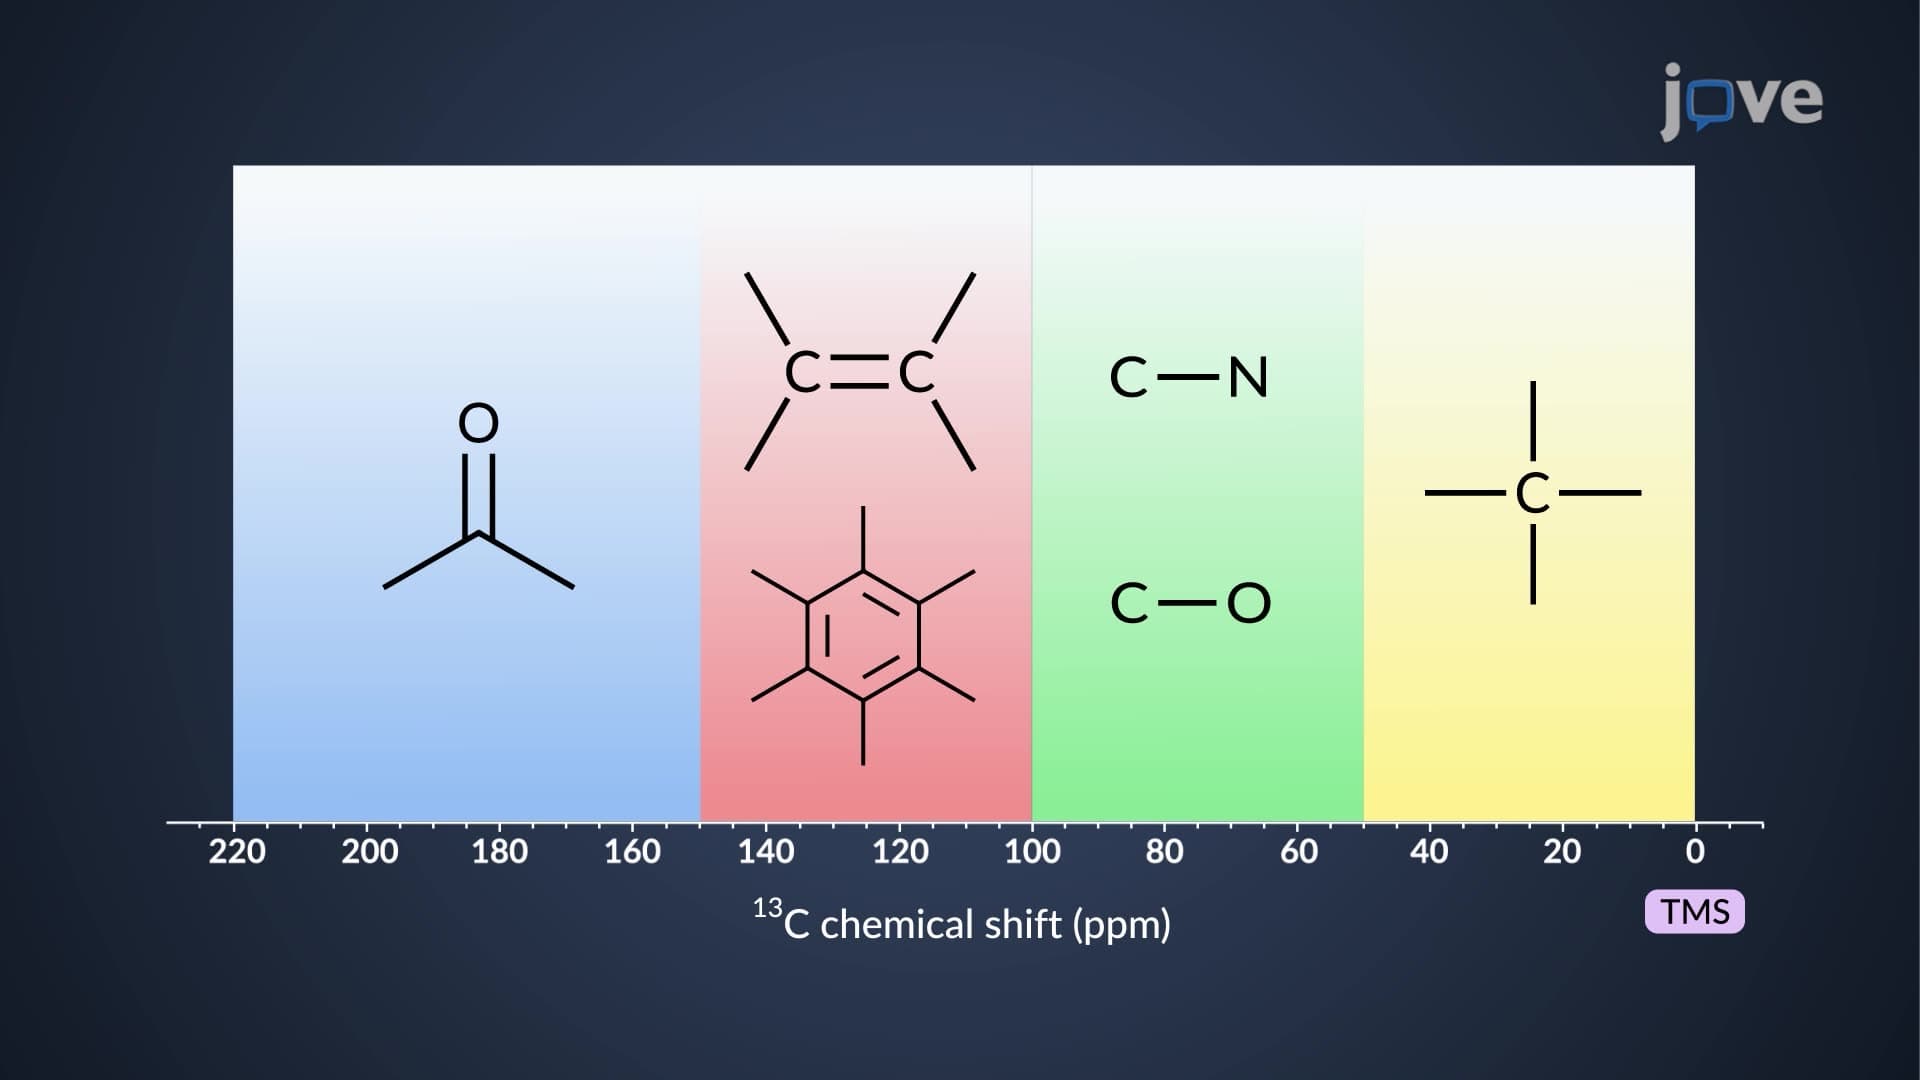
Carbon-13 (¹³C) NMR: Overview
 01:27
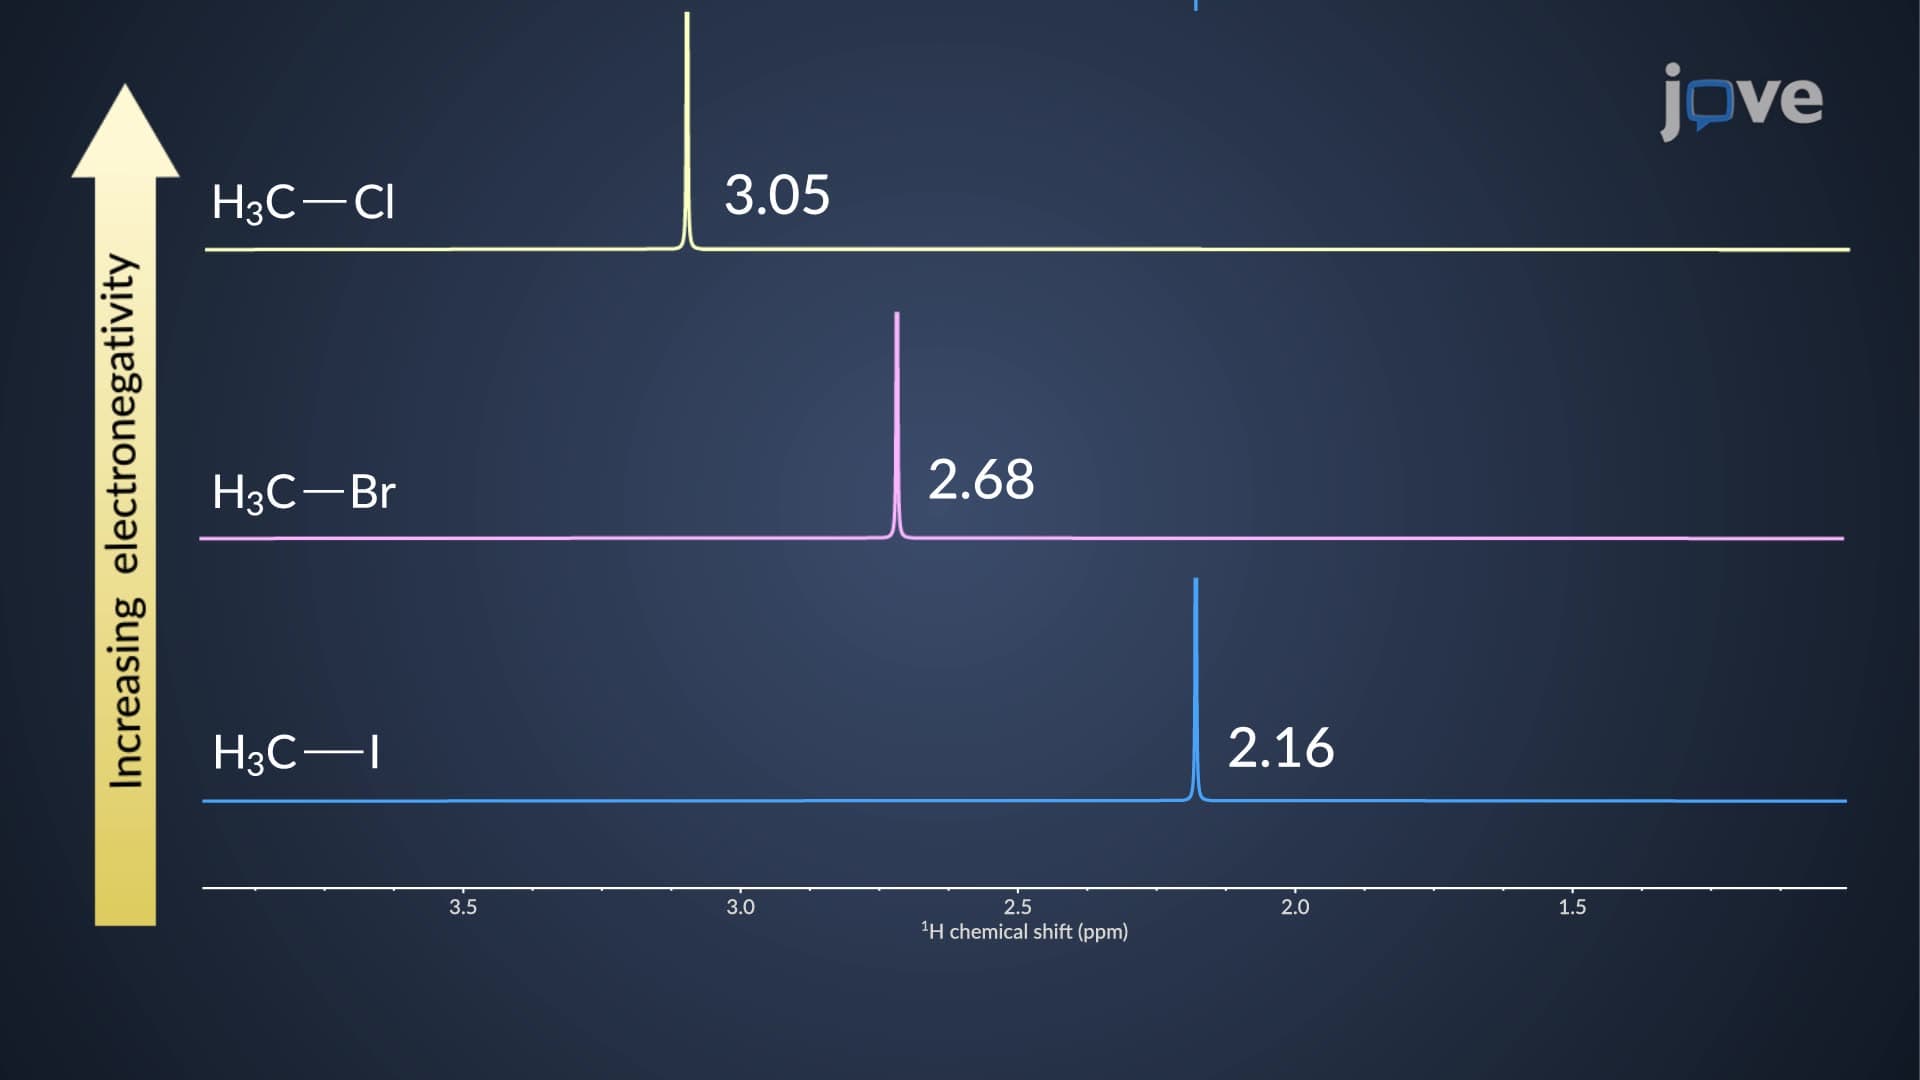
Inductive Effects on Chemical Shift: Overview
 01:27
π Electron Effects on Chemical Shift: Overview
 01:02
¹H NMR: Interpreting Distorted and Overlapping Signals
As Δν decreases and the signals move closer, the doublets appear increasingly distorted. The intensities of the inner lines increase at the cost of those of the outer lines as the signals are...