Related Experiment Video
Updated: Mar 29, 2026

Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Accuracy of Effective Core Potentials and Basis Sets for Density Functional Calculations, Including Relativistic
1Department of Chemistry and Supercomputing Institute, University of Minnesota , Minneapolis, Minnesota 55455-0431, United States.

Abstract:
For molecules containing the fourth-period element arsenic, we test (i, ii) the accuracy of all-electron (AE) basis sets from the def2-xZVP and ma-xZVP series (where xZ is S, TZ, or QZ), (iii) the accuracy of the 6-311G series of AE basis sets with additional polarization and diffuse functions, and (iv) the performance of effective core potentials (ECPs). The first set of tests involves basis-set convergence studies with eleven density functionals for five cases: equilibrium dissociation energy (De) of As2, vertical ionization potential (VIP) of As2, IP of As, acid dissociation of H3AsO4, and De of FeAs. A second set of tests involves the same kinds of basis-set convergence studies for the VIP and De values of As3 and As4 clusters. Both relativistic and nonrelativistic calculations are considered, including in each case both AE calculations and calculations with ECPs. Convergence and accuracy are assessed by comparing to relativistic AE calculations with the cc-pV5Z-DK or ma-cc-pV5Z-DK basis and to nonrelativistic AE calculations with the cc-pV5Z or ma-cc-pV5Z basis. The primary objective of this study is to evaluate the abilities of ECPs with both their recommended basis sets and other basis sets to reproduce the results of all-electron relativistic calculations. The performance of the def2 and ma series basis sets is consistent with their sizes, and quadruple-ζ basis sets are the best. The def2-TZVP basis set performs better than most of the 6-311G series basis sets, which are the most commonly used basis sets in the previous studies of arsenic compounds. However, relativistic def2-TZVP calculations are not recommended. The large-core ECPs, which are the only available ECPs for arsenic in the popular Gaussian program, have average errors of 9-12 kcal/mol for the arsenic systems studied; therefore, these ECPs are not recommended. The triple-ζ small-core relativistic ECP (RECP) basis set cc-pVTZ-PP is found to have performance better than that of the def2-TZVP basis set, and it is highly recommended for arsenic-containing systems. The double-ζ RECP basis set ma-sc-SVP is recommended for large arsenic systems for which the def2-TZVP and cc-pVTZ-PP basis sets are unaffordable, if a basis-set error of ∼2 kcal/mol can be tolerated.
More Related Videos
 08:04
Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
Related Concept Videos
 03:08
Atomic Radii and Effective Nuclear Charge
 02:28
Electronic Structure of Atoms
An atom comprises protons and neutrons, which are contained inside the dense, central core called the nucleus, with electrons present around the nucleus. Taking into account the wave–particle duality of electrons and the uncertainty in position around the nucleus, quantum mechanics provides a more accurate model for the atomic structure. It describes atomic orbitals as the regions around the nucleus where electrons of discrete energy exist, characterized by four quantum...
 01:26
Thermodynamic Potentials
 02:35
Molecular Orbital Theory I
 01:27
π Electron Effects on Chemical Shift: Overview
 02:38
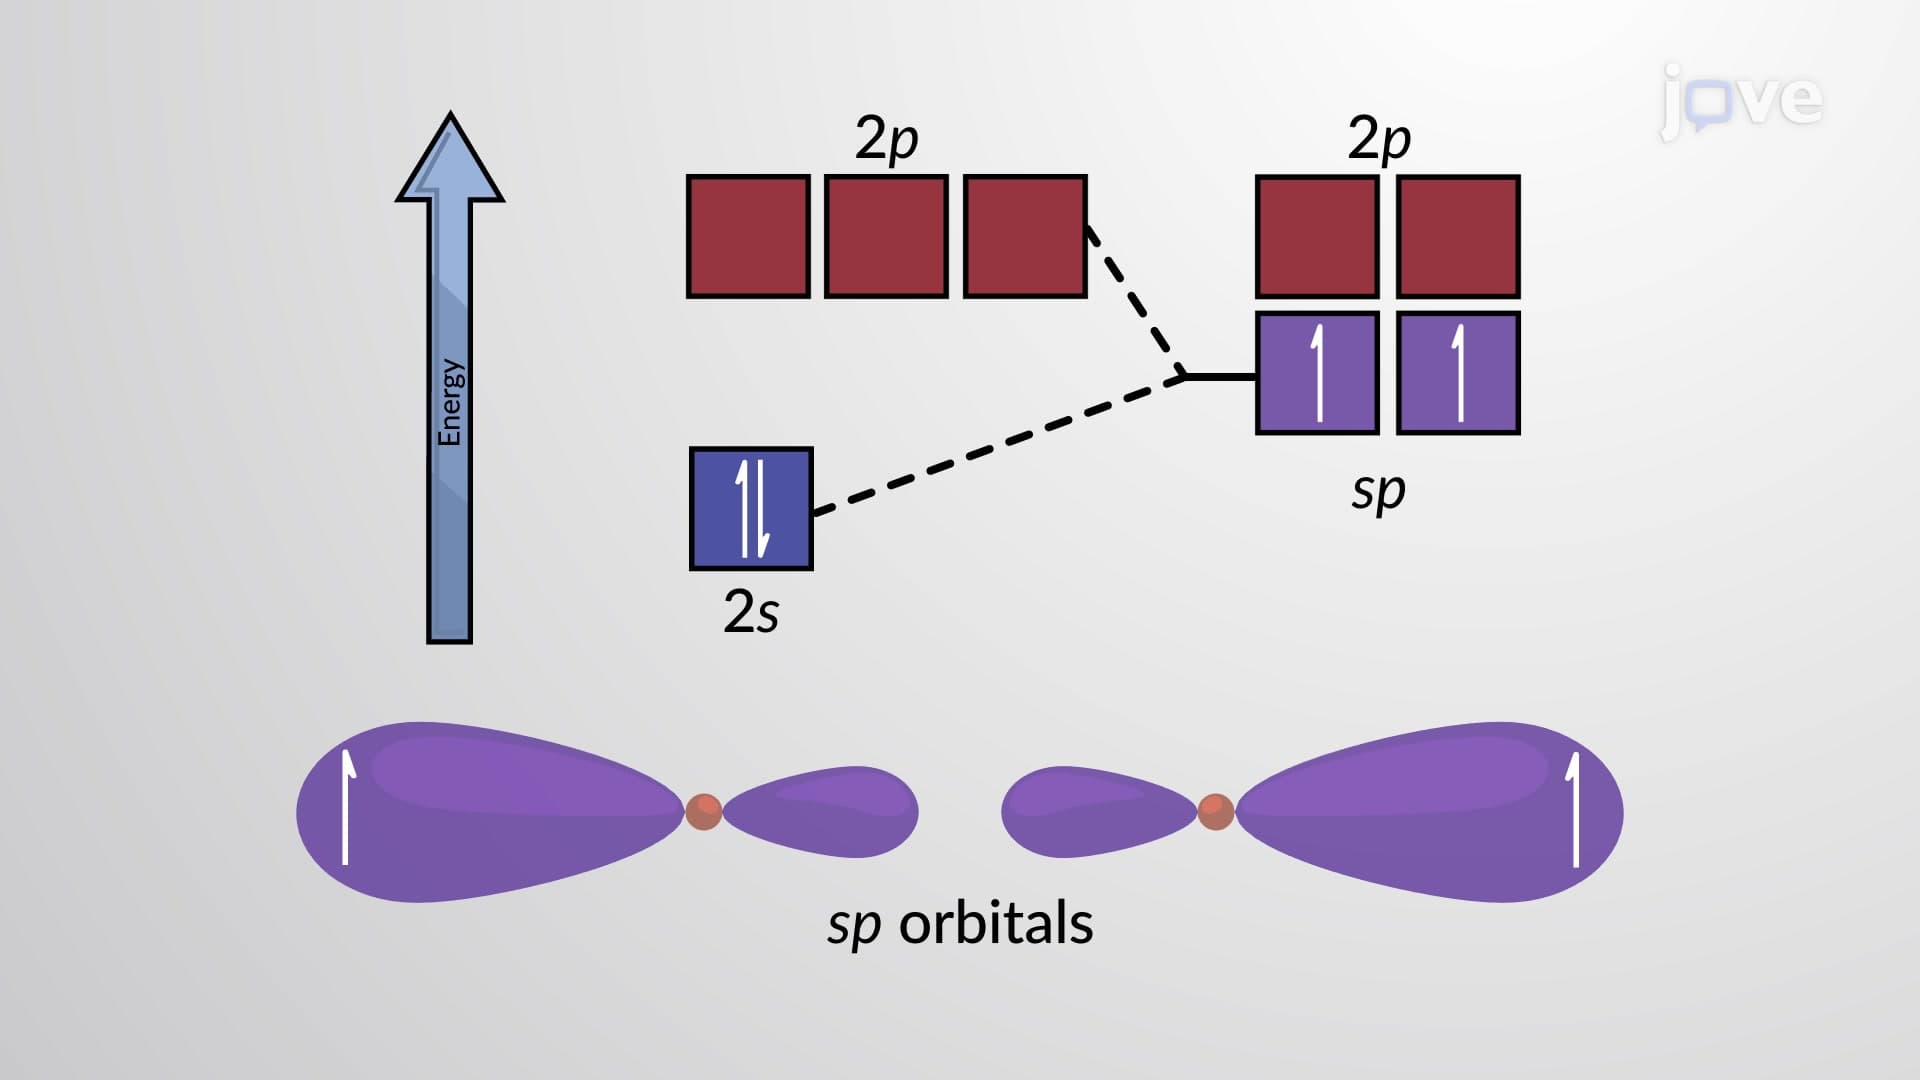
Valence Bond Theory and Hybridized Orbitals
A σ bond (single bond in a Lewis structure) is a covalent bond in which the electron density is...