Related Experiment Video
Updated: Mar 29, 2026


Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
Exploring the Limits of Density Functional Approximations for Interaction Energies of Molecular Precursors to Organic
Stephan N Steinmann1, Clemence Corminboeuf1
1Laboratory for Computational Molecular Design, Institut des Sciences et Ingénierie Chimiques, Ecole Polytechnique Fédérale de Lausanne , CH-1015 Lausanne, Switzerland.

Abstract:
Neutral and charged assemblies of π-conjugated molecules span the field of organic electronics. Electronic structure computations can provide valuable information regarding the nature of the intermolecular interactions within molecular precursors to organic electronics. Here, we introduce a database of neutral (Pi29n) and radical (Orel26rad) dimer complexes that represent binding energies between organic functional units. The new benchmarks are used to test approximate electronic structure methods. Achieving accurate interaction energies for neutral complexes (Pi29n) is straightforward, so long as dispersion interactions are properly taken into account. However, π-dimer radical cations (Orel26rad) are examples of highly challenging situations for density functional approximations. The role of dispersion corrections is crucial, yet simultaneously long-range corrected exchange schemes are necessary to provide the proper dimer dissociation behavior. Nevertheless, long-range corrected functionals seriously underestimate the binding energy of Orel26rad at equilibrium geometries. In fact, only ωB97X-D, an empirical exchange-correlation functional fitted together with an empirical "classical" dispersion correction, leads to suitable results. Valuable alternatives are the more demanding MP2/6-31G*(0.25) level, as well as the most cost-effective combination involving a dispersion corrected long-range functional together with a smaller practical size basis set (e.g., LC-ωPBEB95-dDsC/6-31G*). The Orel26rad test set should serve as an ideal benchmark for assessing the performance of improved schemes.
More Related Videos
 11:44
Using Cyclic Voltammetry, UV-Vis-NIR, and EPR Spectroelectrochemistry to Analyze Organic Compounds
Published on: October 18, 2018
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
Related Concept Videos
 02:13
Bond Dissociation Energy and Activation Energy
 01:24
Van der Waals Interactions
 02:13
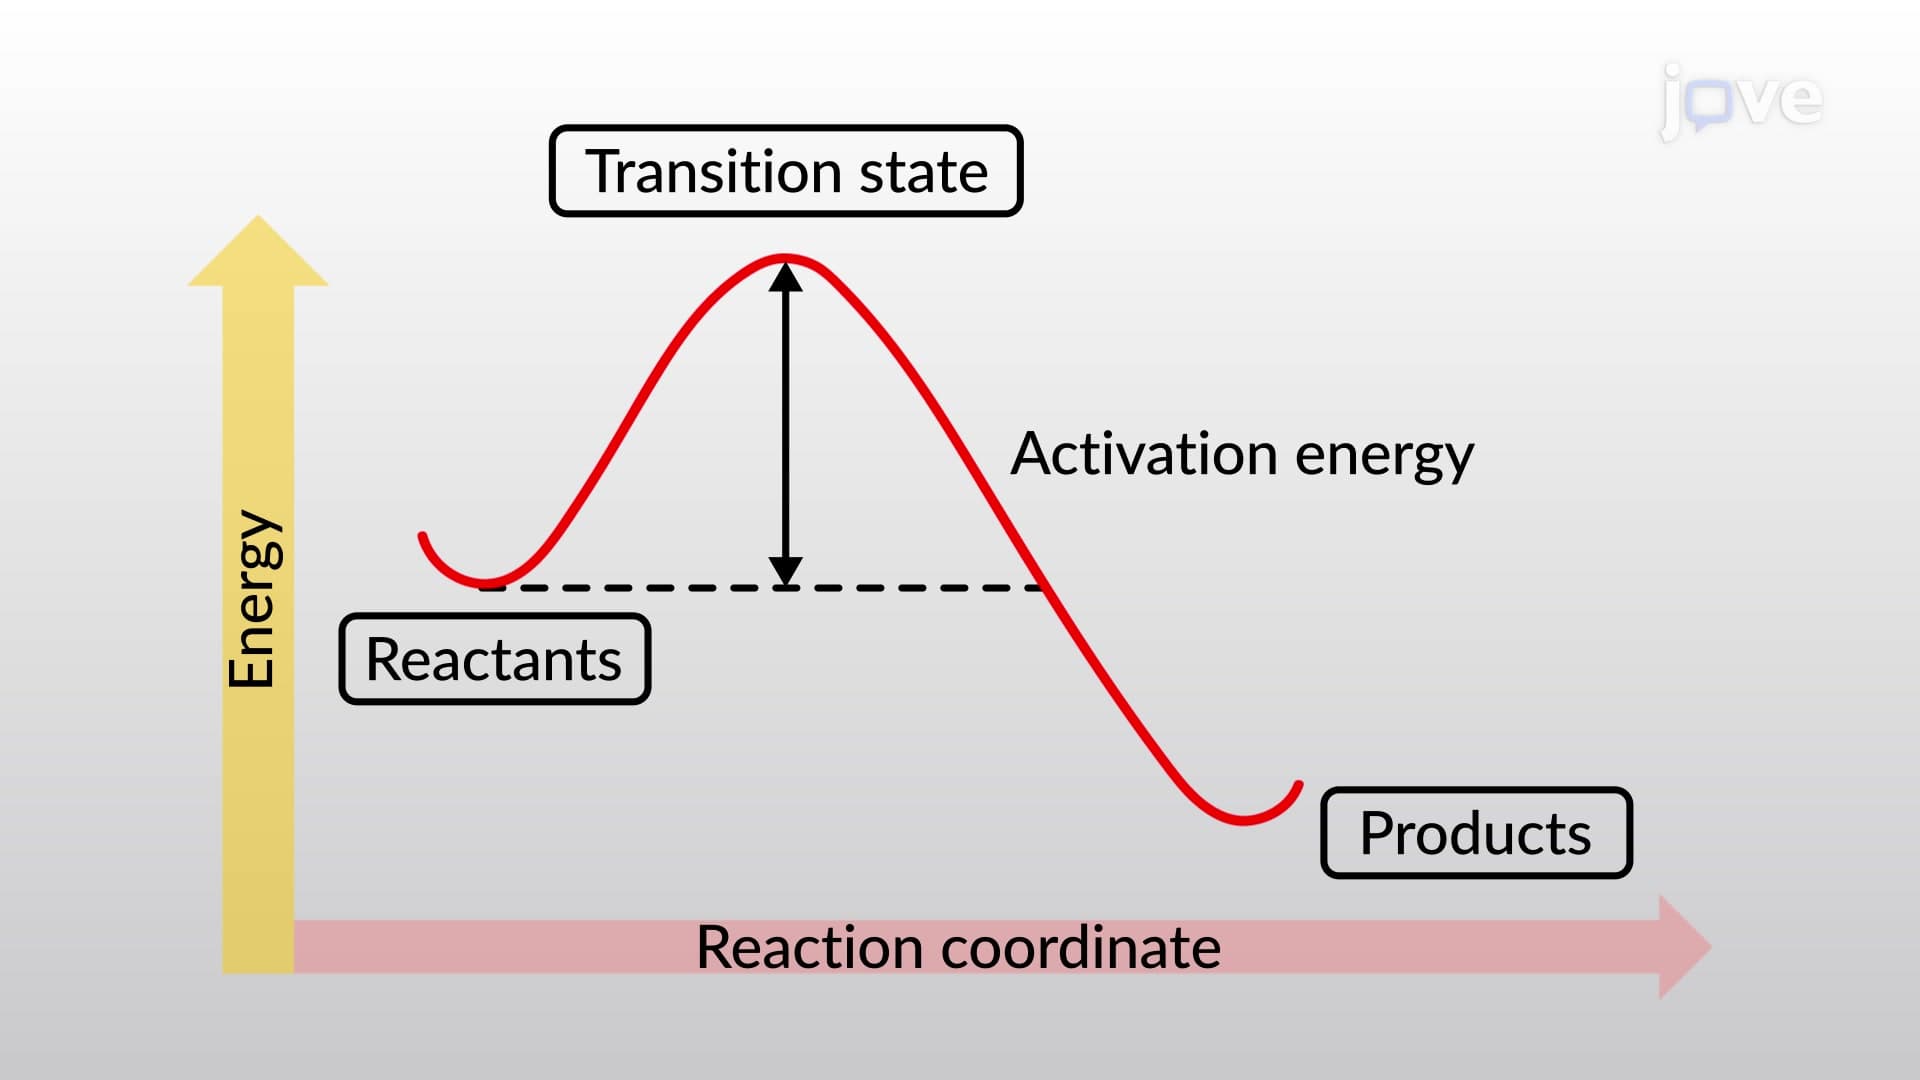
Energy Diagrams, Transition States, and Intermediates
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 03:51
Molecular Orbital Theory II
 01:28
Debye–Huckel–Onsager Conductance Equation