

Molecular Dynamics Simulations Predict a Favorable and Unique Mode of Interaction between Lithium (Li(+)) Ions and
Andrew S Thomas1, Adrian H Elcock1
1Department of Biochemistry, University of Iowa , Iowa City, Iowa 52242, United States.

Abstract:
We report the results of molecular dynamics simulations of the interaction thermodynamics of group I cations with hydrophobic molecules in water using a variety of nonpolarizable force fields. Surprisingly, we find that the Li(+) ion is predicted to form thermodynamically favorable interactions with methane and neopentane using a new mode of recognition that is intermediate between a direct contact and a solvent-separated complex. Further simulations show that this favorable interaction is only predicted by ion parameter sets that correctly reproduce Li(+)'s experimental hydration number.
More Related Videos
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
 05:37
Rapid in-silico Battery Electrolyte Electrochemical Reaction Generation using 3T-VASP Multi-Scale Energy Minimization
Published on: August 22, 2025
Related Concept Videos
 03:13
Intermolecular Forces
 02:42
Aqueous Solutions and Heats of Hydration
When ionic compounds dissolve in water, the ions in the solid separate and disperse uniformly throughout the solution because water molecules surround and solvate the ions, reducing the strong electrostatic forces between them. This process...
 03:51
Molecular Orbital Theory II
 01:23
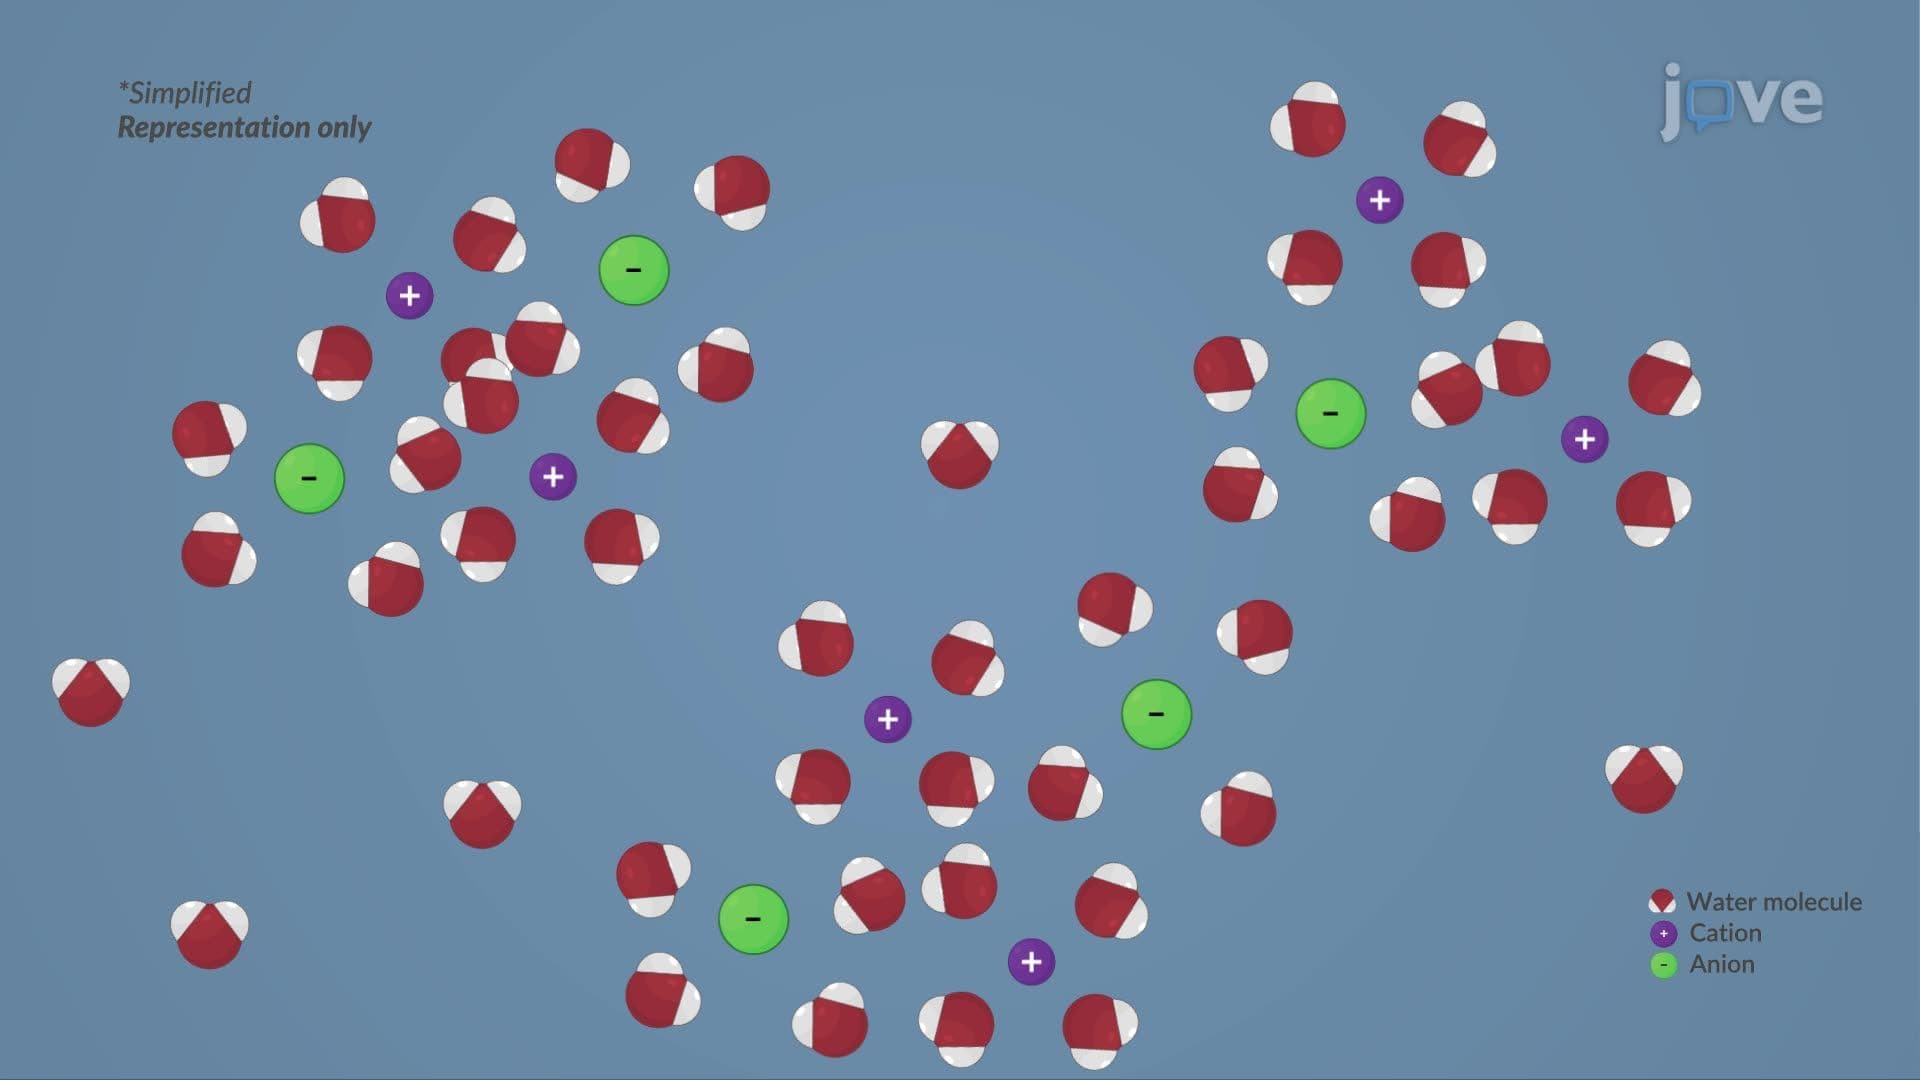
Theory of Strong Electrolytes
 01:27
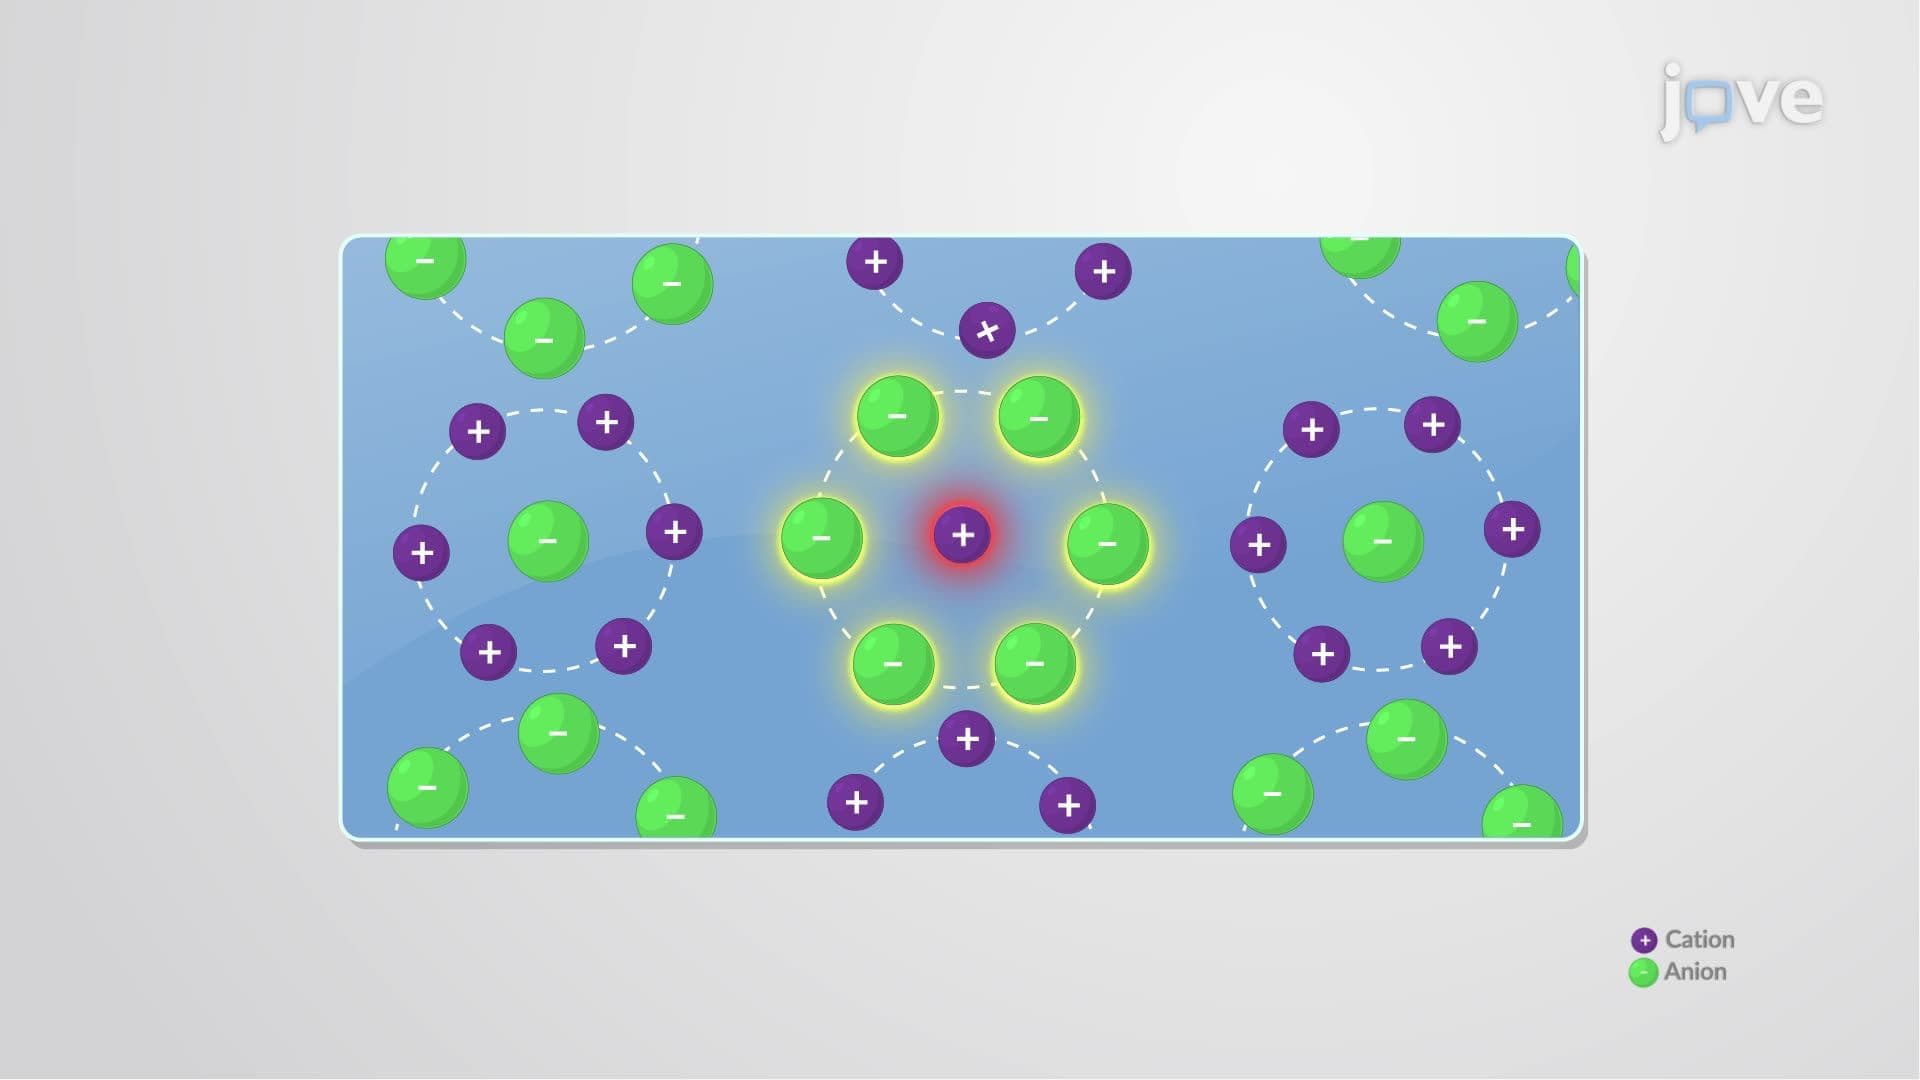
The Debye–Hückel Theory of Electrolyte Solutions
 01:28
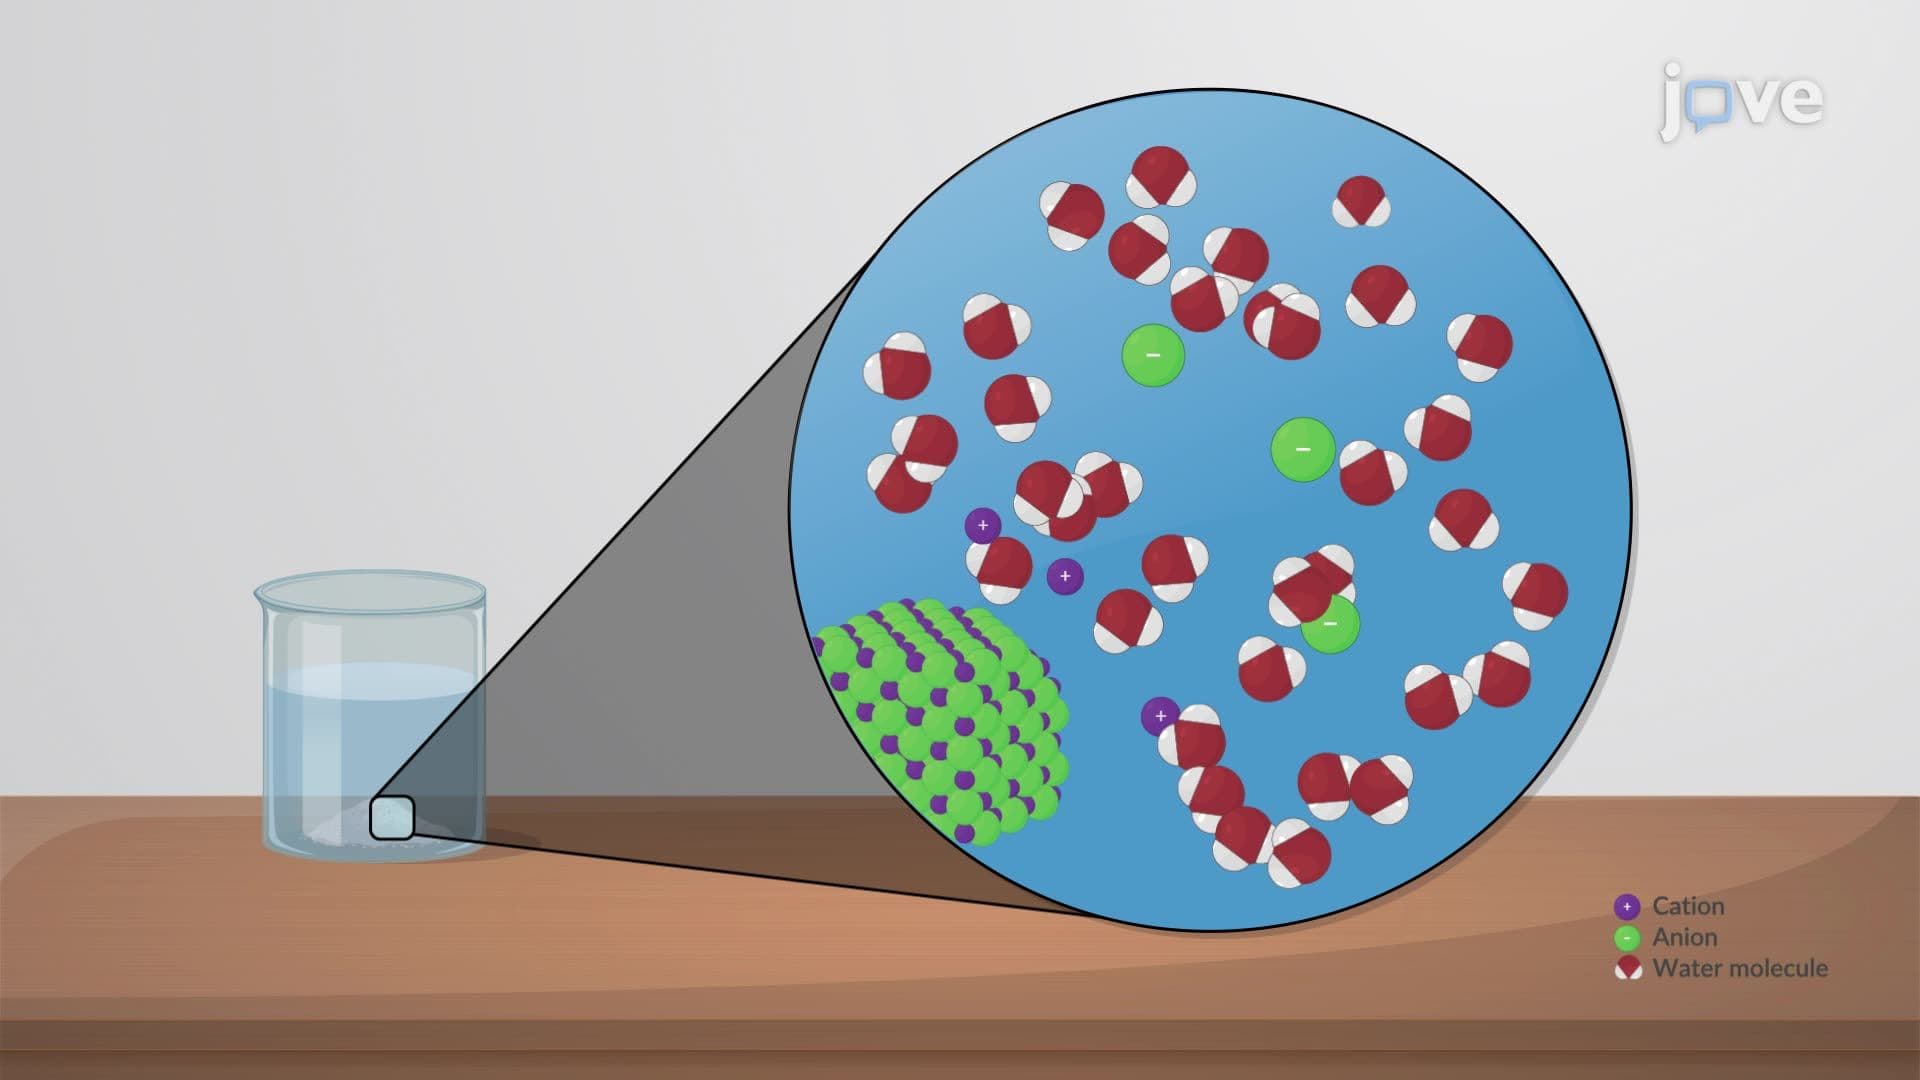
Ionic Association