Related Experiment Video
Updated: Mar 29, 2026

Analyzing and Building Nucleic Acid Structures with 3DNA
Published on: April 26, 2013
Lennard-Jones Parameters for B3LYP/CHARMM27 QM/MM Modeling of Nucleic Acid Bases
Ulla Pentikäinen1, Katherine E Shaw1, Kittusamy Senthilkumar1
1Centre for Computational Chemistry, School of Chemistry, University of Bristol, Cantock's Close, Bristol BS8 1TS, United Kingdom, and Department of Biological and Environmental Science and NanoScience Center, University of Jyväskylä, P.O. Box 35, 40014 Jyväskylä, Finland.

Abstract:
Combined quantum mechanics/molecular mechanics (QM/MM) methods allow computations on chemical events in large molecular systems. Here, we have tested the suitability of the standard CHARMM27 forcefield Lennard-Jones van der Waals (vdW) parameters for the treatment of nucleic acid bases in QM/MM calculations at the B3LYP/6-311+G(d,p)-CHARMM27 level. Alternative parameters were also tested by comparing the QM/MM hydrogen bond lengths and interaction energies with full QM [B3LYP/6-311+G(d,p)] results. The optimization of vdW parameters for nucleic acid bases is challenging because of the likelihood of multiple hydrogen bonds between the nucleic acid base and a water molecule. Two sets of optimized atomic vdW parameters for polar hydrogen, carbonyl carbon, and aromatic nitrogen atoms for nucleic acid bases are reported: base-dependent and base-independent. The results indicate that, for QM/MM investigations of nucleic acids, the standard forcefield vdW parameters may not be appropriate for atoms treated by QM. QM/MM interaction energies calculated with standard CHARMM27 parameters are found to be too large, by around 3 kcal/mol. This is because of overestimation of electrostatic interactions. Interaction energies closer to the full QM results are found using the optimized vdW parameters developed here. The optimized vdW parameters [developed by reference to B3LYP/6-311+G(d,p) results] were also tested at the B3LYP/6-31G(d) QM/MM level and were found to be transferable to the lower level. The optimized parameters also model the interaction energies of charged nucleic acid bases and deprotonation energies reasonably well.
More Related Videos
 07:31
Author Spotlight: Advancing Cell Membrane Biophysics - Exploring Interactions and Challenges Through Experimental and Computational Approaches
Published on: September 1, 2023
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
Related Concept Videos
 02:36
Molecular Geometry and Dipole Moments
 04:01
VSEPR Theory and the Effect of Lone Pairs
 02:37
VSEPR Theory
 01:22
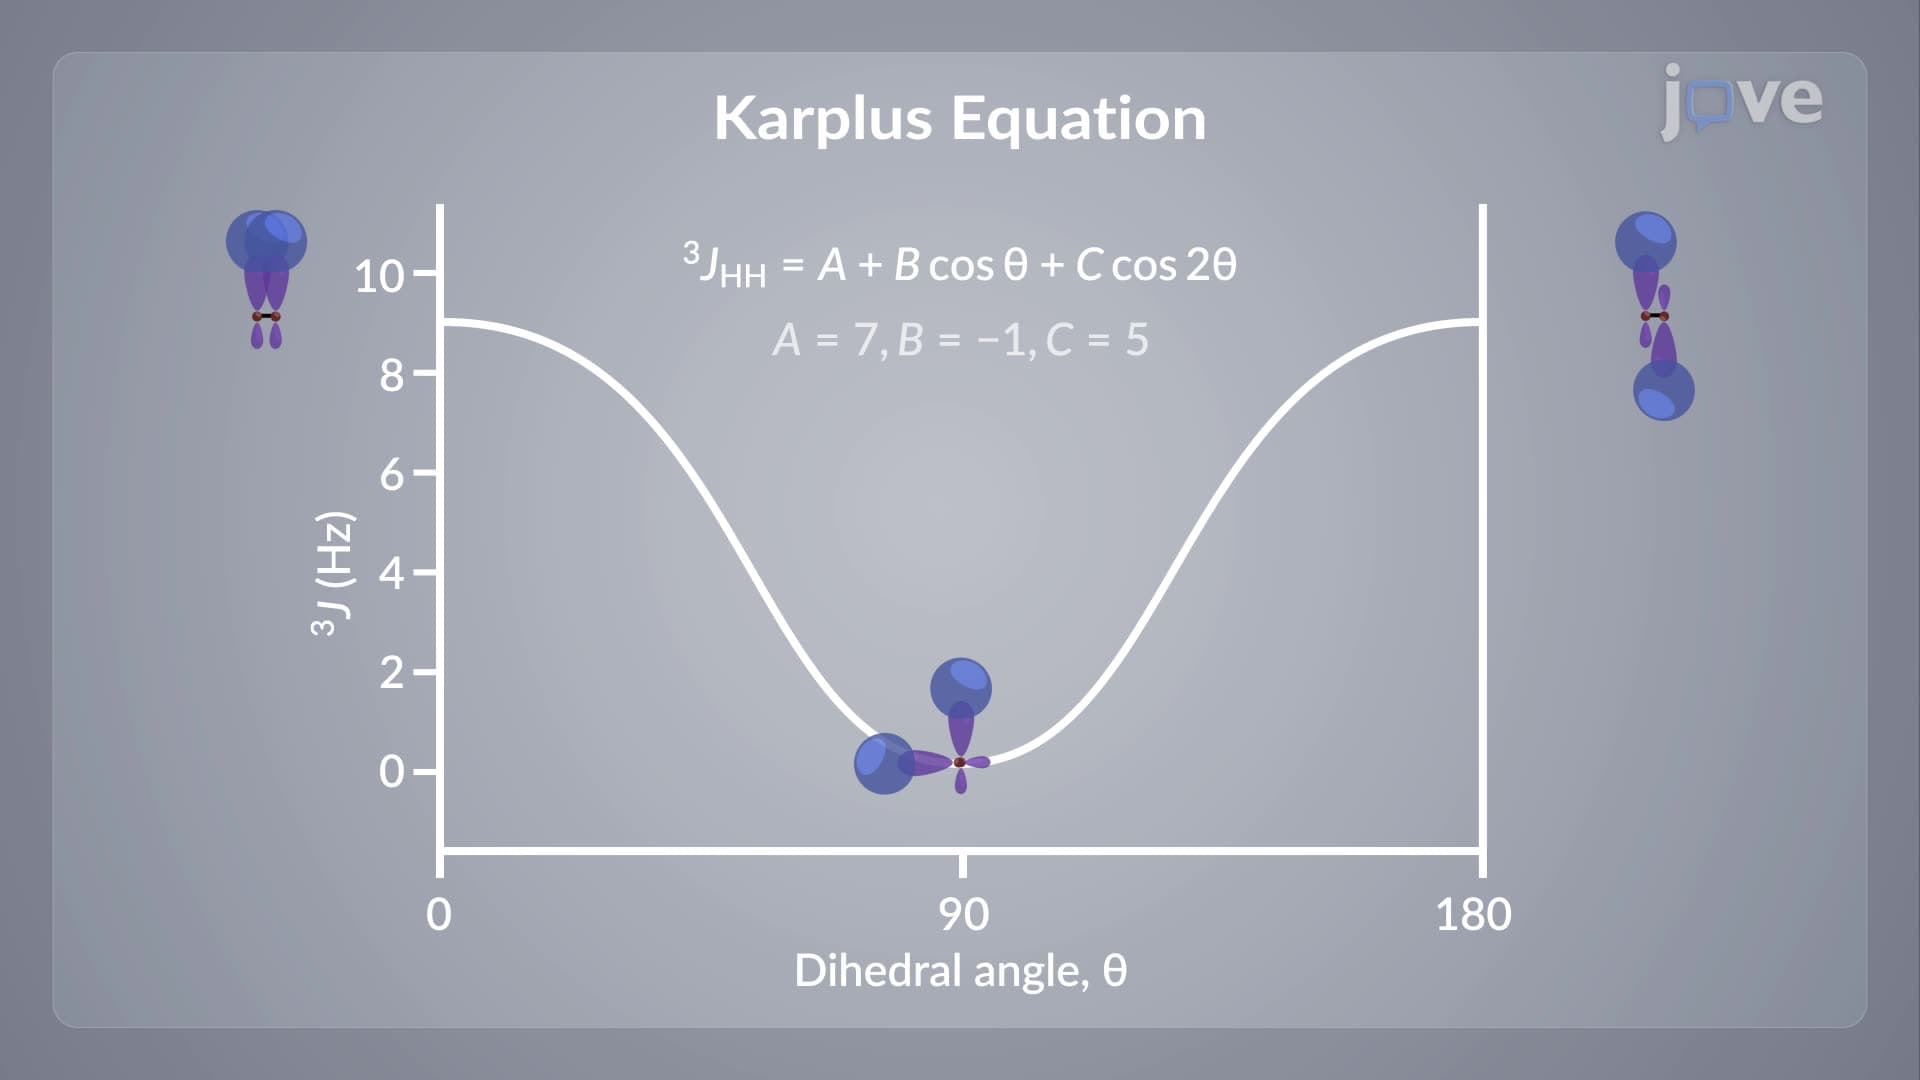
Spin–Spin Coupling: Three-Bond Coupling (Vicinal Coupling)
The extent of coupling depends on the C‑C bond length, the two H‑C‑C angles, any electron-withdrawing substituents, and the dihedral angle between the involved orbitals. The...
 01:25
Nucleic Acid Structure
DNA Structure
DNA...
 01:24
π Molecular Orbitals of 1,3-Butadiene
The simplest conjugated diene is 1,3-butadiene: a four-carbon system where each carbon is sp2-hybridized and has an unhybridized p orbital that contains an unpaired electron. According to molecular orbital theory, atomic orbitals combine to form molecular orbitals such that the number...