

Implementation and Performance of DFT-D with Respect to Basis Set and Functional for Study of Dispersion Interactions
Journal of Chemical Theory and Computation
|December 1, 2015

Abstract
No abstract available in PubMed .
More Related Videos
 13:56
Probe Type II Band Alignment in One-Dimensional Van Der Waals Heterostructures Using First-Principles Calculations
Published on: October 12, 2019
8.5K
 10:52
Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
13.5K
Related Concept Videos
 01:20
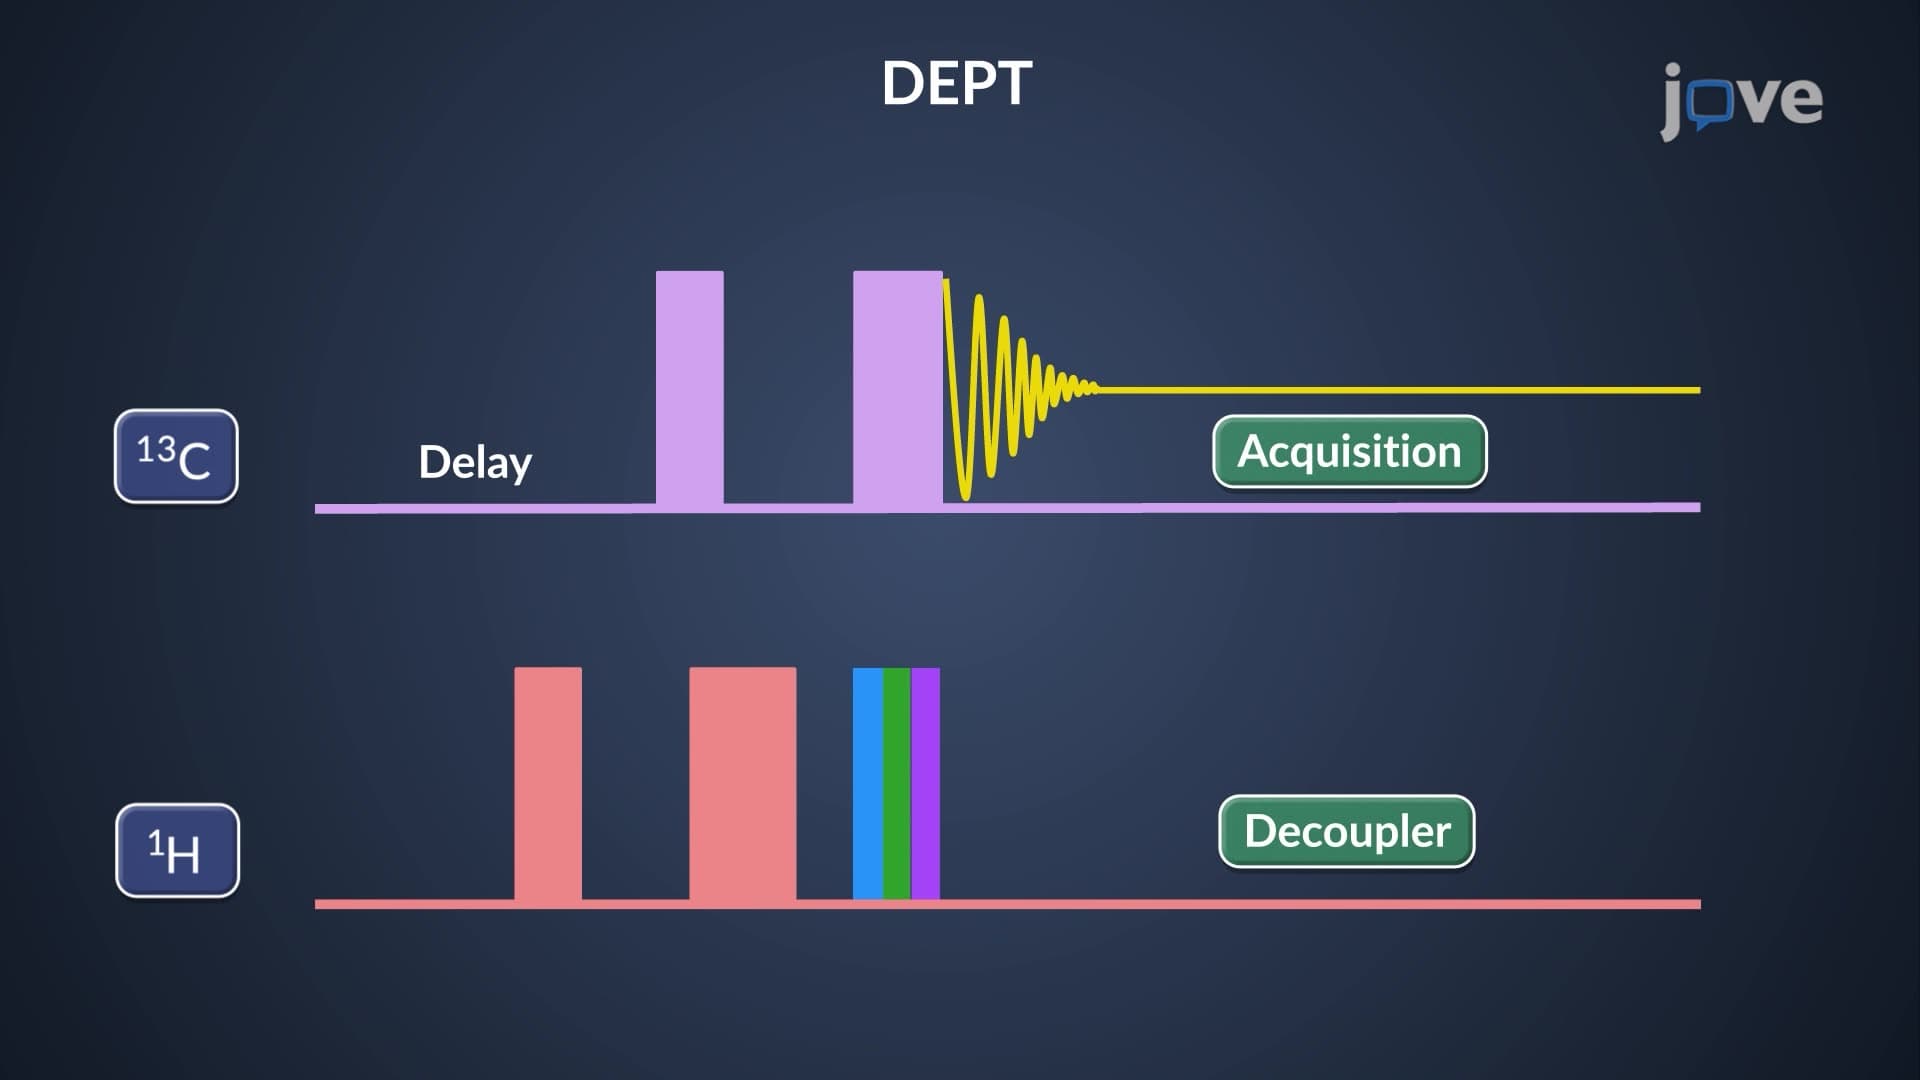
¹³C NMR: Distortionless Enhancement by Polarization Transfer (DEPT)
1.8K
When proton-coupled carbon-13 spectra are simplified by a broadband proton decoupling technique, structural information about the coupled protons is lost. Distortionless enhancement by polarization transfer (DEPT) is a technique that provides information on the number of hydrogens attached to each carbon in a molecule. While the DEPT experiment utilizes complex pulse sequences, the pulse delay and flip angle are specifically manipulated. The resulting signals have different phases depending on...
1.8K
 01:24
Van der Waals Interactions
73.2K
Atoms and molecules interact with each other through intermolecular forces. These electrostatic forces arise from attractive or repulsive interactions between particles with permanent, partial, or temporary charges. The intermolecular forces between neutral atoms and molecules are ion–dipole, dipole–dipole, and dispersion forces, collectively known as van der Waals forces.
73.2K
 01:29
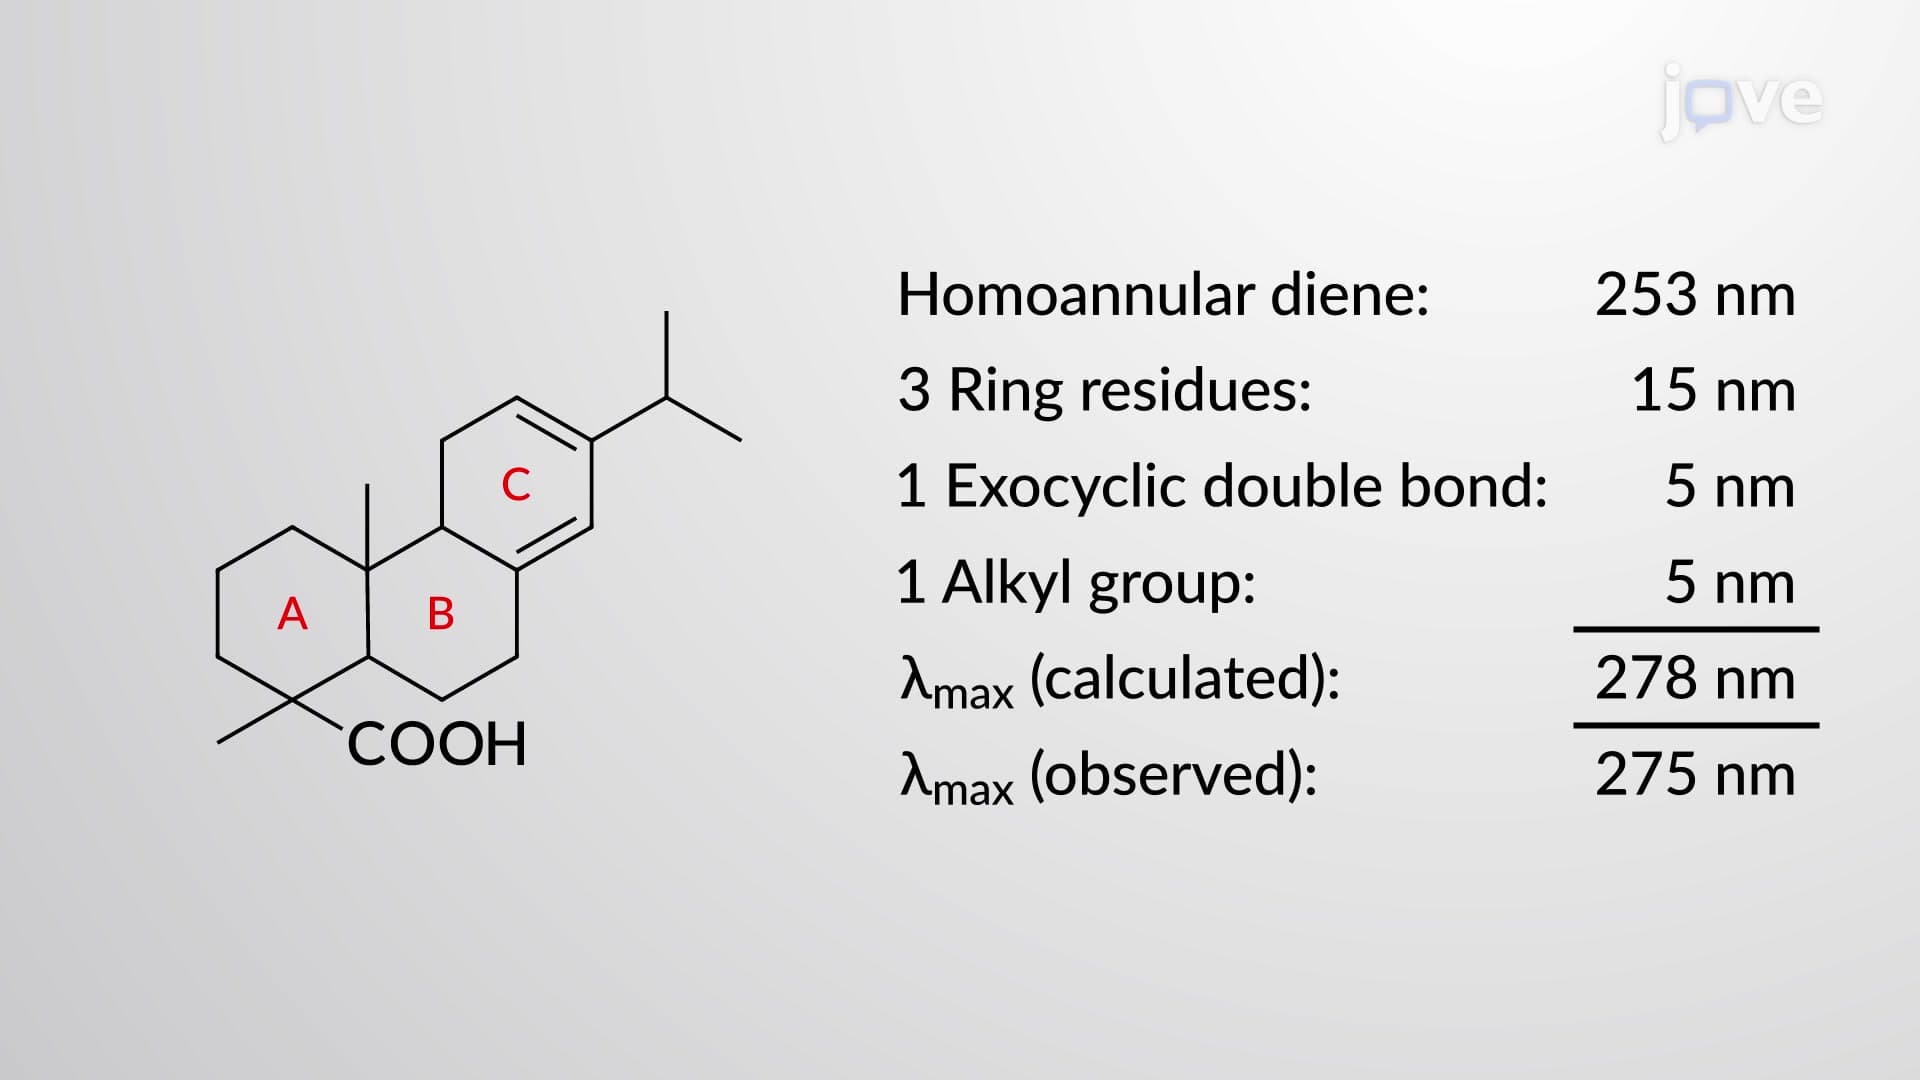
UV–Vis Spectroscopy: Woodward–Fieser Rules
29.6K
UV–Visible absorption spectra of conjugated dienes arise from the lowest energy π → π* transitions. The light-absorbing part of the molecule is called the chromophore, and the substituents directly attached to the chromophore are called auxochromes. A strong correlation exists between the absorption maxima, λmax, and the structure of a conjugated π system. The Woodward–Fieser rules predict the value of λmax for a given structure by adding the...
29.6K
 01:24
π Molecular Orbitals of 1,3-Butadiene
12.7K
Conjugated dienes have lower heats of hydrogenation than cumulated and isolated dienes, making them more stable. The enhanced stabilization of conjugated systems can be understood from their π molecular orbitals.
The simplest conjugated diene is 1,3-butadiene: a four-carbon system where each carbon is sp2-hybridized and has an unhybridized p orbital that contains an unpaired electron. According to molecular orbital theory, atomic orbitals combine to form molecular orbitals such that the number...
The simplest conjugated diene is 1,3-butadiene: a four-carbon system where each carbon is sp2-hybridized and has an unhybridized p orbital that contains an unpaired electron. According to molecular orbital theory, atomic orbitals combine to form molecular orbitals such that the number...
12.7K
 01:14
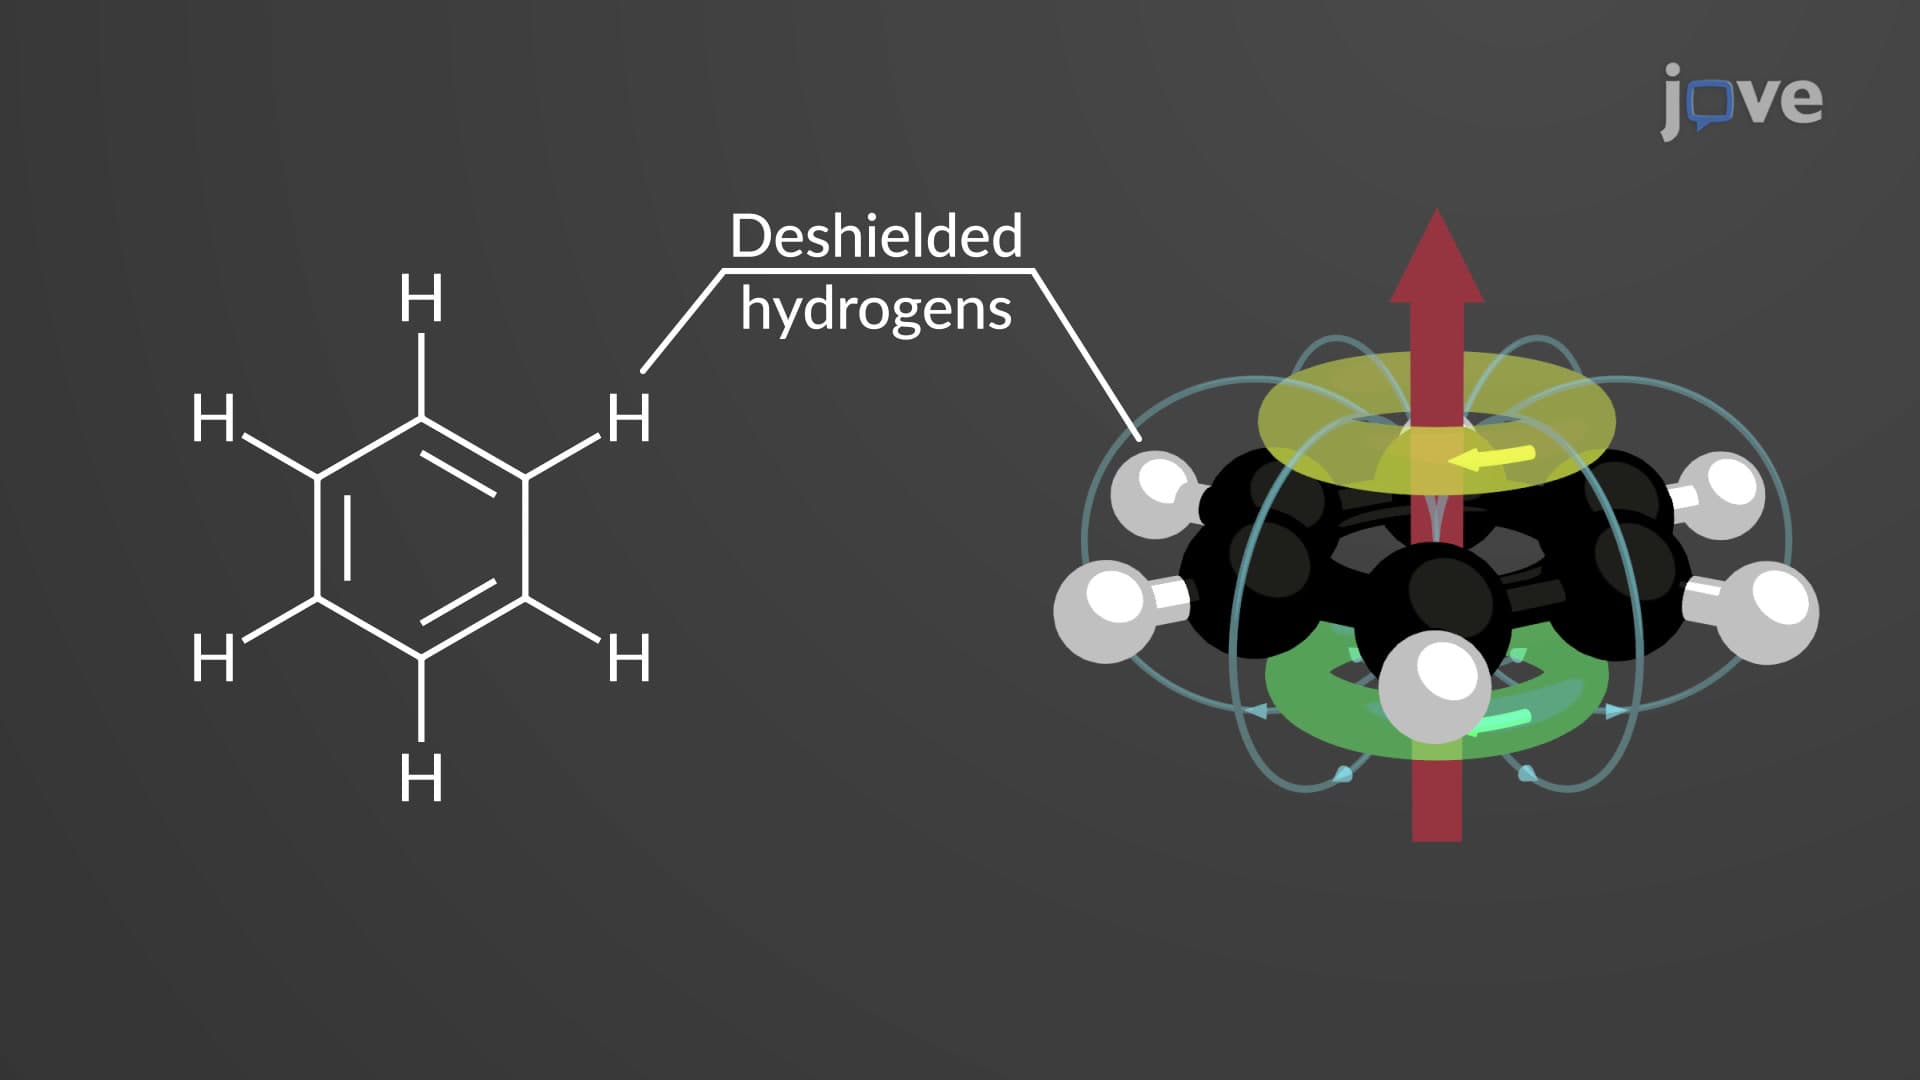
NMR Spectroscopy of Aromatic Compounds
6.7K
Aromatic compounds can be identified or analyzed using proton NMR and carbon‐13 NMR. Typically, aromatic hydrogens or hydrogens directly bonded to the aromatic rings are strongly deshielded by the aromatic ring current. Therefore, they absorb in the range of 6.5–8.0 ppm in proton NMR spectra. For instance, aromatic hydrogens directly bonded to the benzene ring absorb at 7.3 ppm. However, aromatic hydrogens of larger rings absorb farther upfield or downfield than the ideal range.
6.7K
 01:14
π Electron Effects on Chemical Shift: Aromatic and Antiaromatic Compounds
2.0K
In aromatic compounds, such as benzene, the circulation of (4n + 2) π-electrons sets up a diamagnetic or diatropic ring current around the perimeter of the molecule. This current induces a magnetic field that opposes the external field inside the ring and reinforces it on the outside. The protons in benzene are deshielded and exhibit high chemical shifts in the range 6.5–8.5 ppm. The shielding effect at the center of the ring is evident in complex aromatic molecules, such as...
2.0K