
Anharmonic Vibrational Analysis for the Propadienylidene Molecule (H2C═C═C:)
Qunyan Wu1, Qiang Hao1, Jeremiah J Wilke1
1Institute of Chemical Physics, Beijing Institute of Technology, Beijing, P. R. China 100081, Center for Computational Quantum Chemistry, University of Georgia, Athens, Georgia 30602, College of Chemistry, Beijing Normal University, Beijing, P. R. China 100875, and Center for Computational Quantum Chemistry, South China Normal University, Guangzhou, P. R. China 510631.

Abstract:
Maier et al. found that photolysis of singlet cyclopropenylidene (1S) in a matrix yields triplet propargylene (2T), which upon further irradiation is converted to singlet propadienylidene (vinylidenecarbene, 3S). Their discovery was followed by interstellar identification of 3S by Cernicharo et al. An accurate quartic force field for propadienylidene (3S) has been determined employing the ab initio coupled-cluster (CC) with single and double excitations and perturbative triple excitations [CCSD(T)] method and the correlation-consistent core-valence quadruple-ζ (cc-pCVQZ) basis set. Utilizing vibrational second-order perturbation theory (VPT2), vibration-rotation coupling constants, rotational constants, centrifugal distortion constants, vibrational anharmonic constants, and fundamental vibrational frequencies are determined. The predicted fundamental frequencies for 3S as well as its (13)C and deuterium isotopologues are in good agreement with experimental values. The theoretical zero-point vibration corrected rotational constants B0 are consistent with experimental values within 0.3% of errors. The isotopic shifts of B0 are in close to exact agreement with experimental observations. The mean absolute deviation between theoretical anharmonic and experimental fundamental vibrational frequencies for 24 modes (excluding CH2 s-str.) is only 2.6 cm(-1). The isotopic shifts of the vibrational frequencies are also in excellent agreement with the available experimental values. However, a large discrepancy is observed for the CH2 symmetric stretch, casting doubt on the experimental assignment for this mode.
More Related Videos
 09:35
Preparation of a Corannulene-functionalized Hexahelicene by CopperI-catalyzed Alkyne-azide Cycloaddition of Nonplanar Polyaromatic Units
Published on: September 18, 2016
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
Related Concept Videos
 01:24
π Molecular Orbitals of 1,3-Butadiene
The simplest conjugated diene is 1,3-butadiene: a four-carbon system where each carbon is sp2-hybridized and has an unhybridized p orbital that contains an unpaired electron. According to molecular orbital theory, atomic orbitals combine to form molecular orbitals such that the number...
 01:18
Aromatic Hydrocarbon Anions: Structural Overview
Due to the absence of continuous...
 01:18
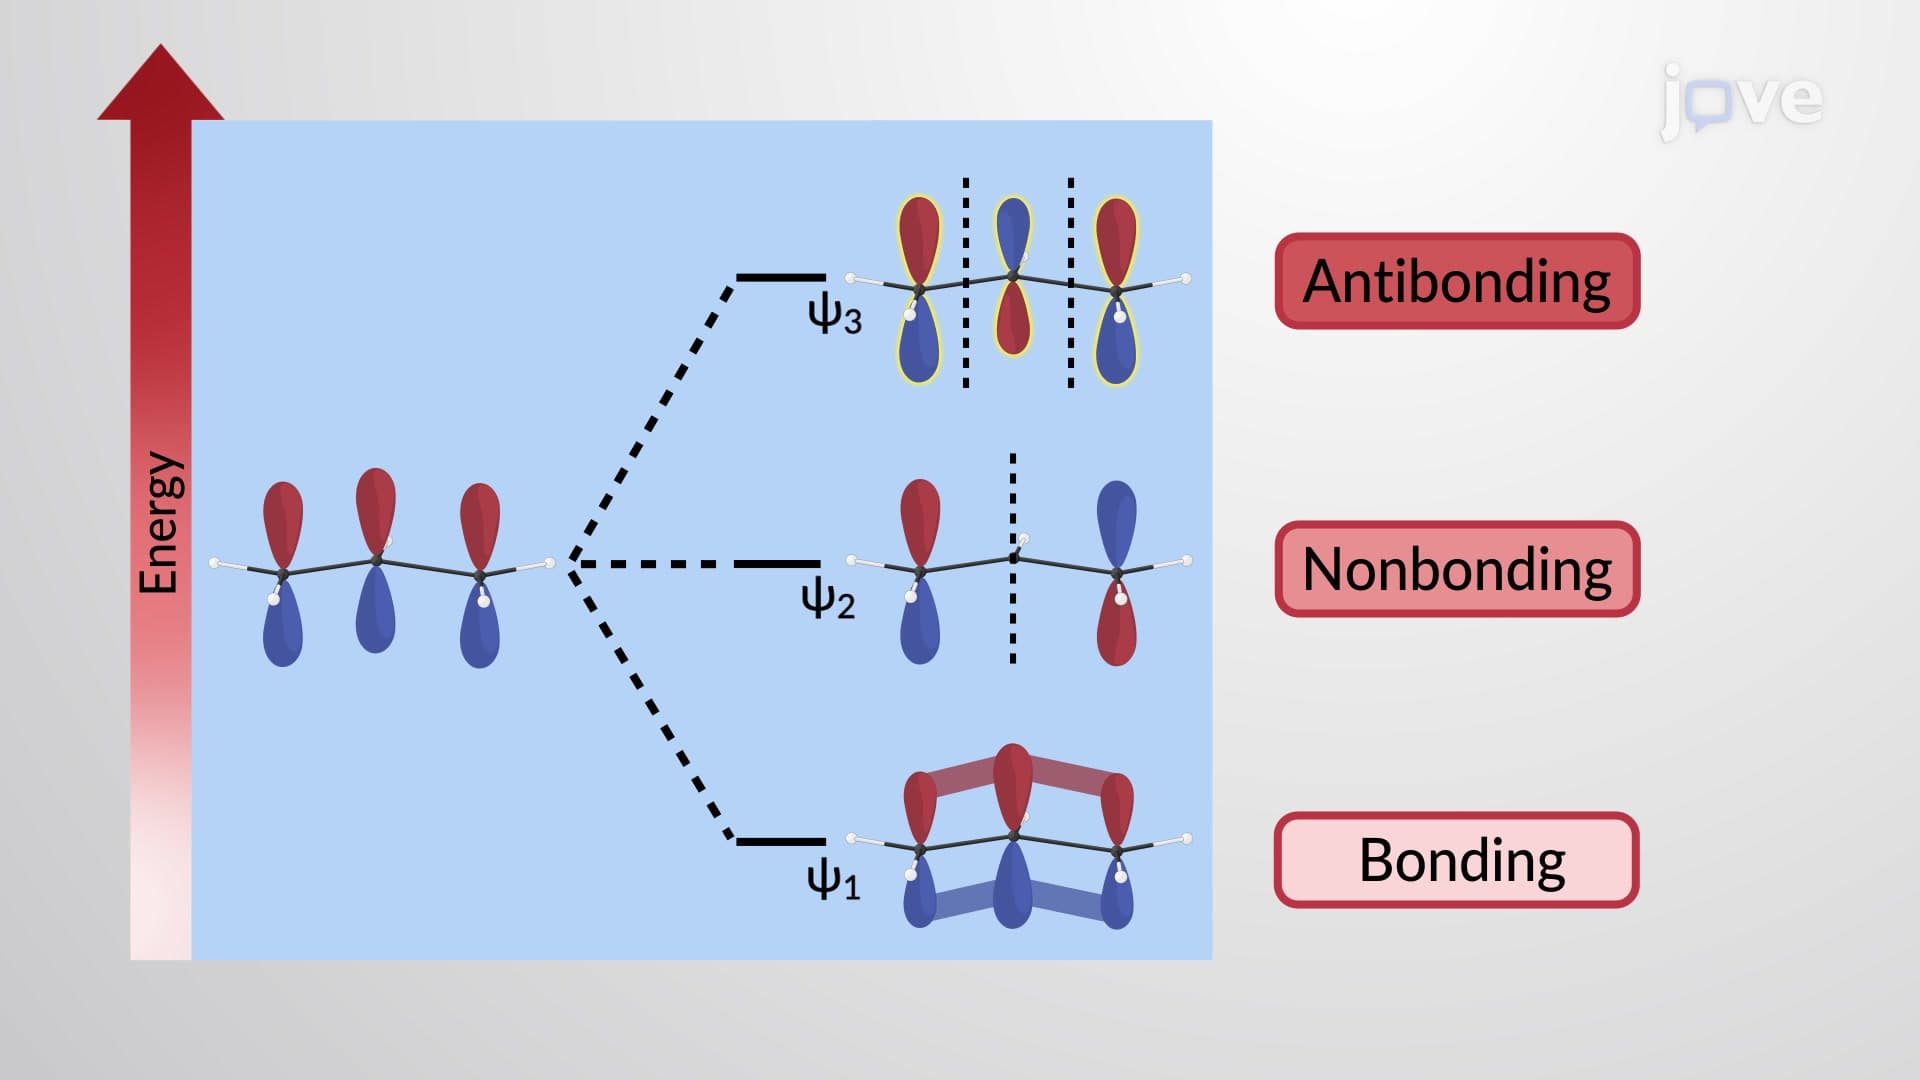
π Molecular Orbitals of the Allyl Cation and Anion
 01:20
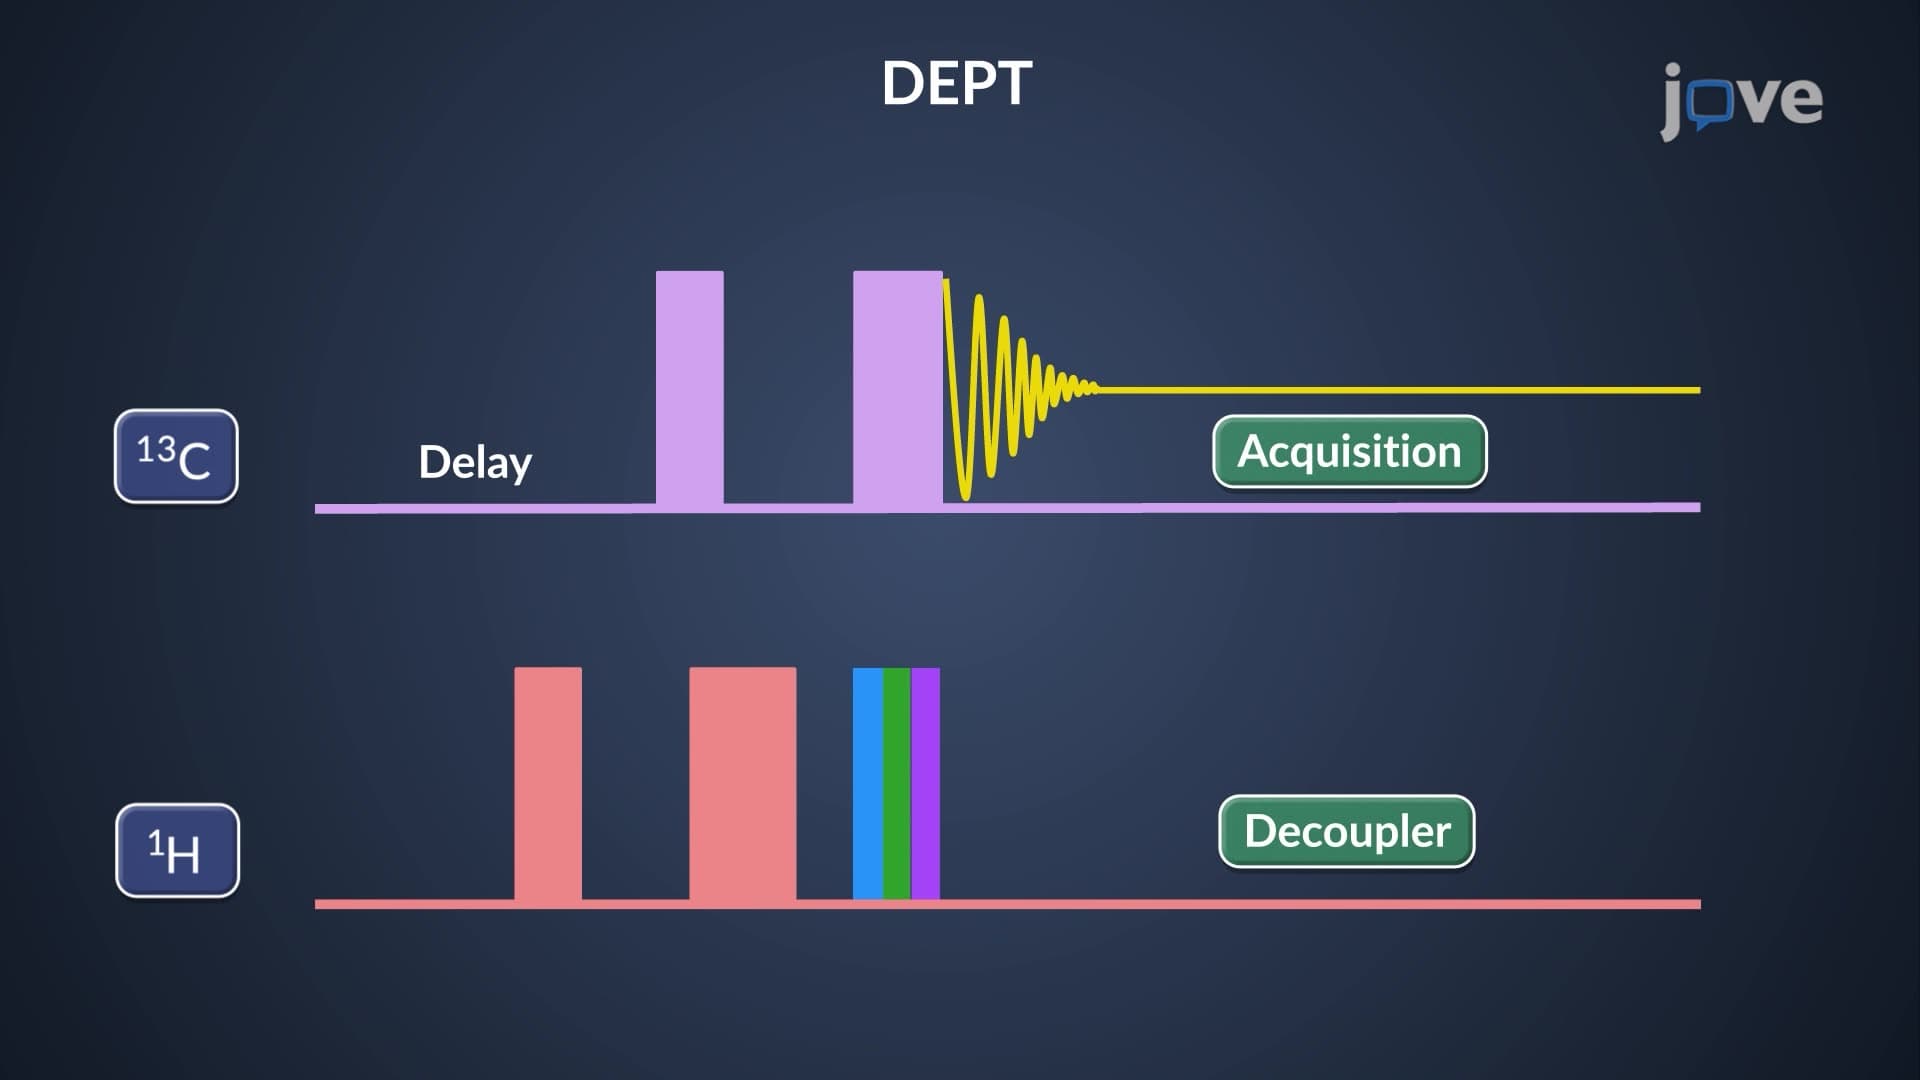
¹³C NMR: Distortionless Enhancement by Polarization Transfer (DEPT)
 01:24
IR Spectroscopy: Molecular Vibration Overview
Stretching vibrations are vibrational motions that occur along the bond line, changing the bond length or distance between two bonded atoms. They are further distinguished as symmetric or asymmetric. In symmetric stretching, the...
 01:14
π Electron Effects on Chemical Shift: Aromatic and Antiaromatic Compounds