Related Experiment Video
Updated: Mar 29, 2026
Curation of Computational Chemical Libraries Demonstrated with Alpha-Amino Acids
Published on: April 13, 2022
Molecular Polarization Effects on the Relative Energies of the Real and Putative Crystal Structures of Valine
Timothy G Cooper1, Katarzyna E Hejczyk1, William Jones1
1The Pfizer Institute for Pharmaceutical Materials Science, Department of Chemistry, University of Cambridge, Lensfield Road, Cambridge CB2 1EW, United Kingdom.

Abstract:
The computer-generation of the crystal structures of the α-amino acid valine is used as a challenging test of lattice energy modeling methods for crystal structure prediction of flexible polar organic molecules and, specifically, to examine the importance of molecular polarization on calculated relative energies. Total calculated crystal energies, which combine atom-atom model potential calculations of intermolecular interactions with density functional theory intramolecular energies, do not effectively distinguish the real (known) crystal structures from the rest of the low energy computer-generated alternatives when the molecular electrostatic models are derived from isolated molecule calculations. However, we find that introducing a simple model for the bulk crystalline environment when calculating the molecular energy and electron density distribution leads to important changes in relative total crystal energies and correctly distinguishes the observed crystal structures from the set of computer-generated possibilities. This study highlights the importance of polarization of the molecular charge distribution in crystal structure prediction calculations, especially for polar flexible molecules, and suggests a computationally inexpensive approach to include its effect in lattice energy calculations.
More Related Videos
 14:55
Atomic Scale Structural Studies of Macromolecular Assemblies by Solid-state Nuclear Magnetic Resonance Spectroscopy
Published on: September 17, 2017
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
Related Concept Videos
 02:37
VSEPR Theory
 02:36
Molecular Geometry and Dipole Moments
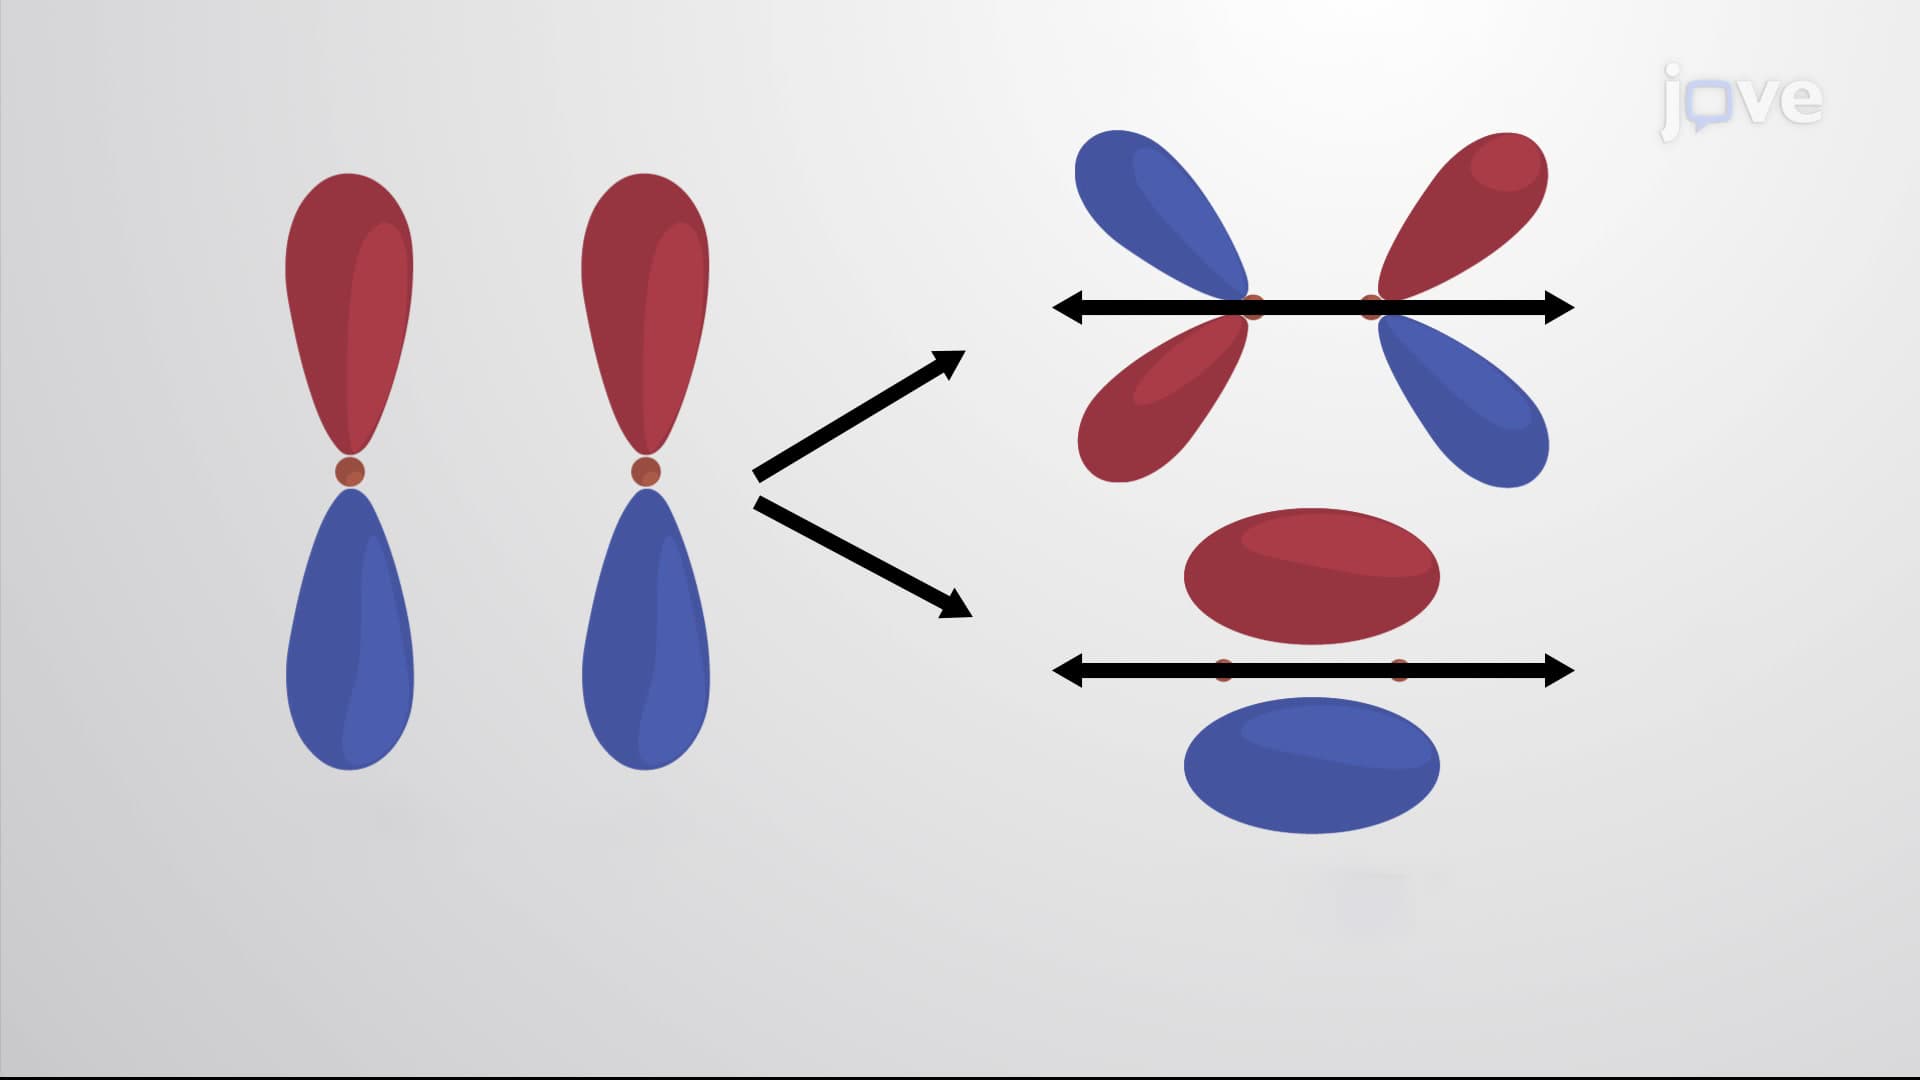 02:40
MO Theory and Covalent Bonding
 03:51
Molecular Orbital Theory II
 04:01
VSEPR Theory and the Effect of Lone Pairs
 03:37
Molecular Shape and Polarity