The MARTINI Coarse-Grained Force Field: Extension to Proteins
Luca Monticelli1, Senthil K Kandasamy1, Xavier Periole1
1Dept of Biological Sciences, University of Calgary, 2500 University Dr NW, Calgary, AB, T2N 1N4, Canada, Chemical Engineering Department, The University of Michigan, 2300 Hayward Street, Ann Arbor, Michigan 48109, and Groningen Biomolecular Sciences and Biotechnology Institute & Zernike Institute for Advanced Materials, University of Groningen, Nijenborgh 4, 9747 AG Groningen, The Netherlands.

Abstract:
Many biologically interesting phenomena occur on a time scale that is too long to be studied by atomistic simulations. These phenomena include the dynamics of large proteins and self-assembly of biological materials. Coarse-grained (CG) molecular modeling allows computer simulations to be run on length and time scales that are 2-3 orders of magnitude larger compared to atomistic simulations, providing a bridge between the atomistic and the mesoscopic scale. We developed a new CG model for proteins as an extension of the MARTINI force field. Here, we validate the model for its use in peptide-bilayer systems. In order to validate the model, we calculated the potential of mean force for each amino acid as a function of its distance from the center of a dioleoylphosphatidylcholine (DOPC) lipid bilayer. We then compared amino acid association constants, the partitioning of a series of model pentapeptides, the partitioning and orientation of WALP23 in DOPC lipid bilayers and a series of KALP peptides in dimyristoylphosphatidylcholine and dipalmitoylphosphatidylcholine (DPPC) bilayers. A comparison with results obtained from atomistic models shows good agreement in all of the tests performed. We also performed a systematic investigation of the partitioning of five series of polyalanine-leucine peptides (with different lengths and compositions) in DPPC bilayers. As expected, the fraction of peptides partitioned at the interface increased with decreasing peptide length and decreasing leucine content, demonstrating that the CG model is capable of discriminating partitioning behavior arising from subtle differences in the amino acid composition. Finally, we simulated the concentration-dependent formation of transmembrane pores by magainin, an antimicrobial peptide. In line with atomistic simulation studies, disordered toroidal pores are formed. In conclusion, the model is computationally efficient and effectively reproduces peptide-lipid interactions and the partitioning of amino acids and peptides in lipid bilayers.
More Related Videos
 09:17
Structure-Based Simulation and Sampling of Transcription Factor Protein Movements along DNA from Atomic-Scale Stepping to Coarse-Grained Diffusion
Published on: March 1, 2022
 06:45
Force Spectroscopy of Single Protein Molecules Using an Atomic Force Microscope
Published on: February 28, 2019
Related Concept Videos
 02:00
Molecular Models
 02:04
Protein-Protein Interfaces
02:04Protein-protein Interfaces
 01:19
Protein Dynamics in Living Cells
Fluorescent recovery after photobleaching (FRAP) is a fluorescent-protein-based detection technique used to quantify protein movement rates within the cell. This method exposes a small portion of the cell to an intense laser beam. The laser beam causes permanent photobleaching of the fluorophore-tagged proteins in the exposed region. As the bleached...
 02:15
Protein and Protein Structures
02:15Protein and Protein Structure
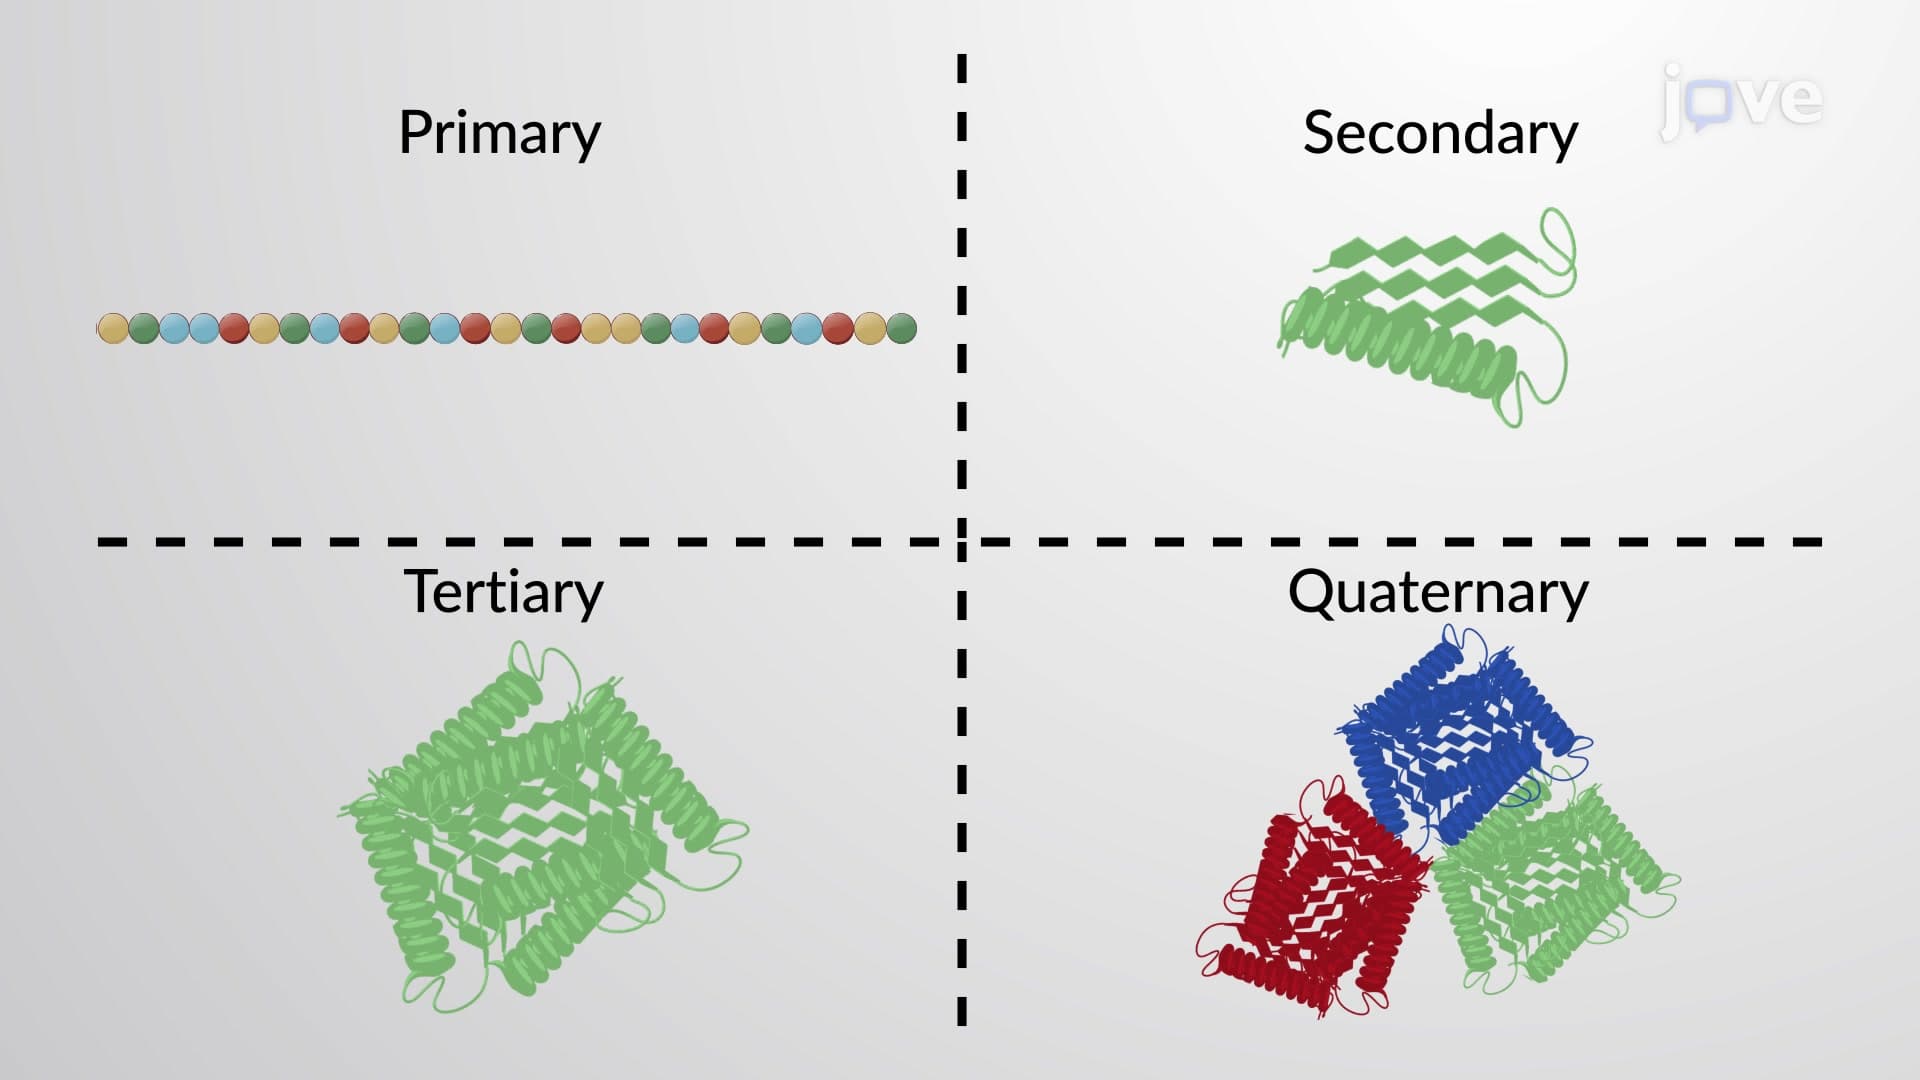
A protein's shape is critical to its function. For example, an enzyme...