Related Experiment Video
Updated: Mar 29, 2026
Author Spotlight: A Computational Approach to Decipher Amino Acid Preferences in Multispecific Protein-Protein Interactions
Published on: January 26, 2024
Benchmarking pKa Prediction Methods for Residues in Proteins
Courtney L Stanton1, Kendall N Houk1
1Department of Chemistry and Biochemistry, University of California Los Angeles, 607 Charles E. Young Drive East, Los Angeles, California 90095.

Abstract:
Methods for estimation of pKa values of residues in proteins were tested on a set of benchmark proteins with experimentally known pKa values. The benchmark set includes 80 different residues (20 each for Asp, Glu, Lys, and His), half of which consists of significantly variant cases (ΔpKa ≥ 1 pKa unit from the amino acid in solution). The method introduced by Case and co-workers [J. Am. Chem. Soc. 2004, 126, 4167-4180], referred to as the molecular dynamics/generalized-Born/thermodynamic integration (MD/GB/TI) technique, gives a root-mean-square deviation (rmsd) of 1.4 pKa units on the benchmark set. The use of explicit waters in the immediate region surrounding the residue was shown to generally reduce high errors for this method. Longer simulation time was also shown to increase the accuracy of this method. The empirical approach developed by Jensen and co-workers [Proteins 2005, 61, 704-721], PROPKA, also gives an overall rmsd of 1.4 pKa units and is more or less accurate based on residue type-the method does very well for Lys and Glu, but less so for Asp and His. Likewise, the absolute deviation is quite similar for the two methods-5.2 for PROPKA and 5.1 for MD/GB/TI. A comparison of these results with several prediction methods from the literature is presented. The error in pKa prediction is analyzed as a function of variation of the pKa from that in water and the solvent accessible surface area (SASA) of the residue. A case study of the catalytic lysine residue in 2-deoxyribose-5-phosphate aldolase (DERA) is also presented.


Related Concept Videos
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally...
 02:35
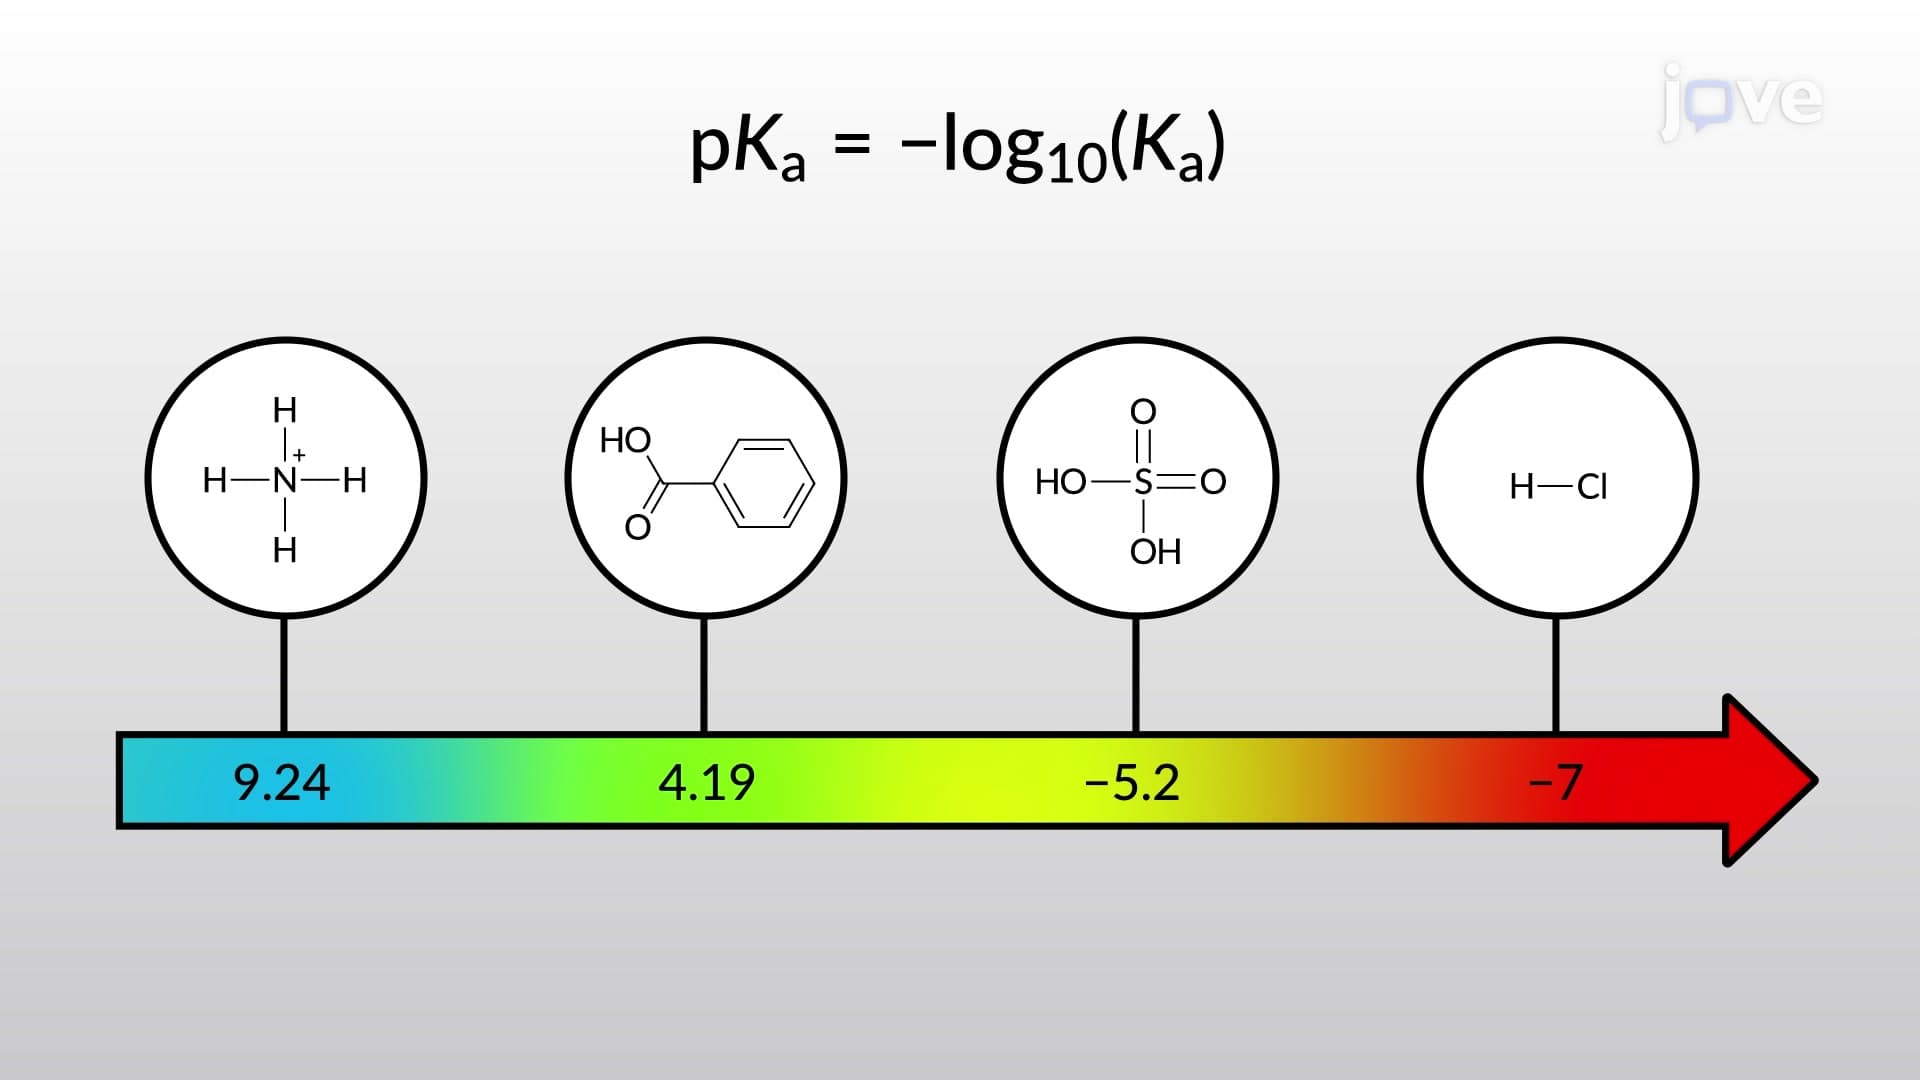
Acid and Bases: Ka, pKa, and Relative Strengths
 02:18
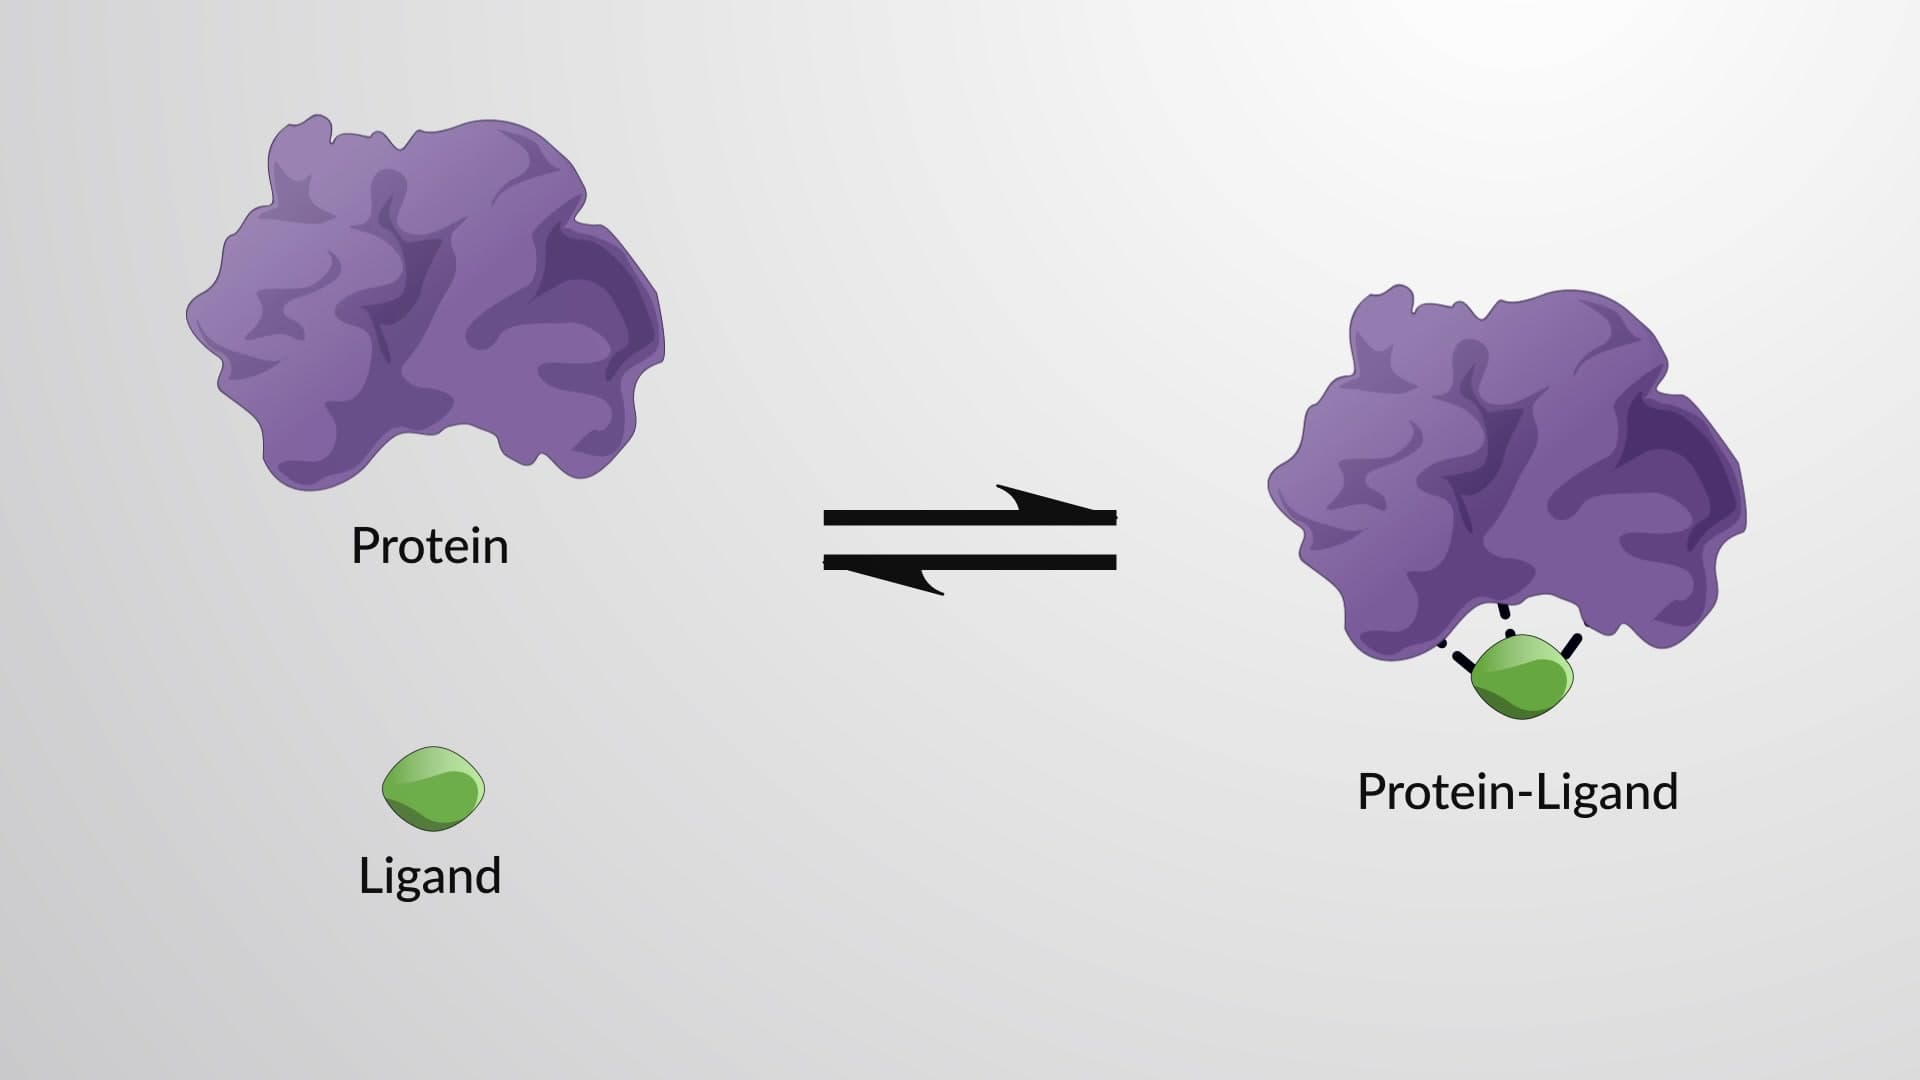
The Equilibrium Binding Constant and Binding Strength
 02:04
Protein-protein Interfaces
 01:22
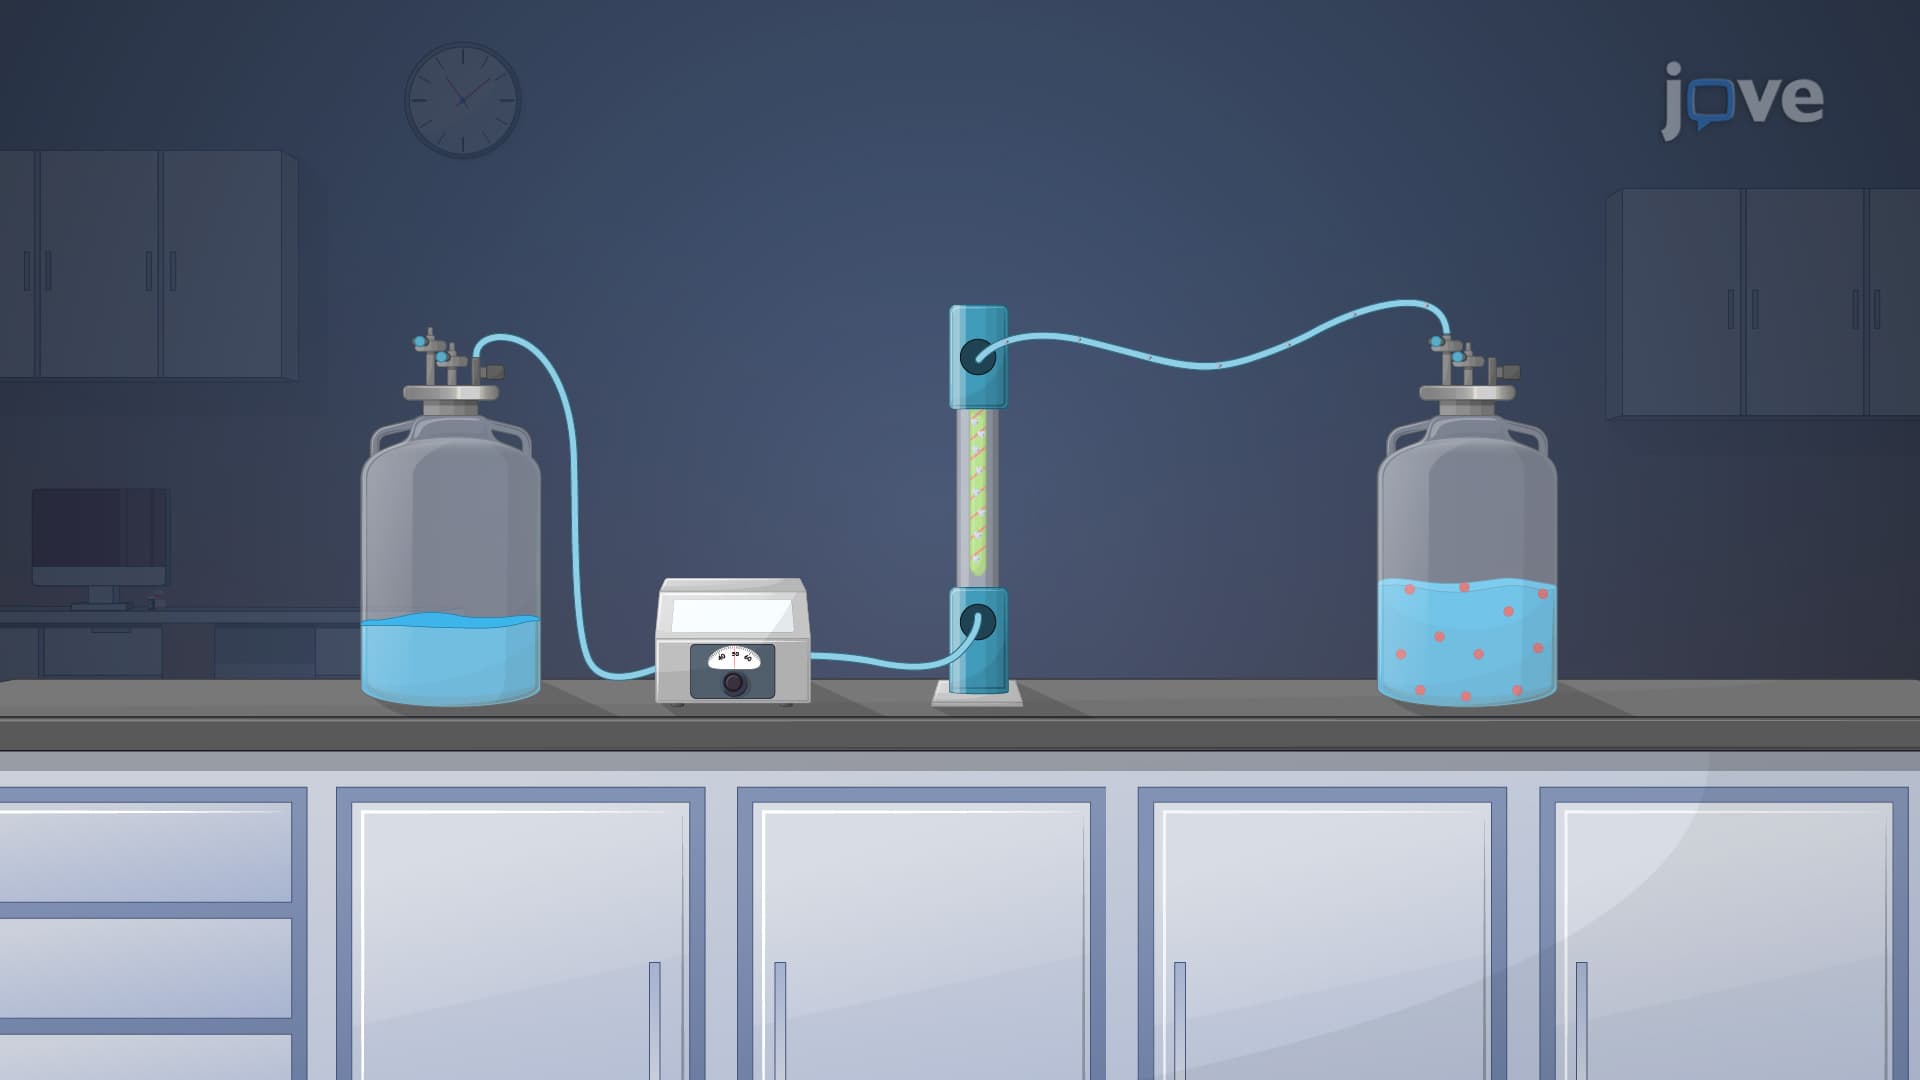
Protein-Drug Binding: Determination Methods
Indirect methods involve isolating the bound drug from its free form in biological samples such as blood, serum, or plasma. These techniques aim to measure the percentage of drugs bound to proteins. Equilibrium dialysis is a commonly used method where the free drug concentration at equilibrium is measured by separating the bound...
 02:29
Relative Strengths of Conjugate Acid-Base Pairs