Related Experiment Video
Updated: Mar 29, 2026

Line Shape Analysis of Dynamic NMR Spectra for Characterizing Coordination Sphere Rearrangements at a Chiral Rhenium Polyhydride Complex
Published on: July 27, 2022
Vibronic Quasi-Free Rotation Effects in Biphenyl-Like Molecules. TD-DFT Study of Bifluorene
Rémy Fortrie1, Henry Chermette1
1Université de Lyon, École Normale Supérieure de Lyon, Laboratoire de Chimie, UMR 5182 du CNRS, 46 Allée d'Italie, F-69364 Lyon Cedex 07, France, and Université Lyon 1 and CNRS UMR 5180 du CNRS Sciences Analytiques, Chimie Physique Théorique, Bat. Paul Dirac (210), 43 Boulevard du 11 Novembre 1918, F-69622 Villeurbanne Cedex, France.

Abstract:
In this paper, we investigate the vibronic shape of the lowest UV-visible absorption band of biphenyl-like systems, using the bifluorene molecule as a workhorse. The molecule is here regarded as a one-dimensional two-level system, whose ground and excited states are simulated with time-dependent density functional theory and semiempirical methods. The vibrational Schrödinger equation is then numerically solved along the torsional coordinate, and the vibronic shape of the absorption band is modeled. Comparisons with the harmonic approximation, with or without the Franck-Condon approximation, are performed. This study confirms that a vibronic effect is most likely responsible for the strong dissymmetry of the lowest UV-visible absorption band of biphenyl-like systems and that, for such systems, the experimental data should be extracted using the whole absorption band, instead of a Gaussian fit on the first part of the band, as it is often done when a superposition between several electronic transitions is suspected.
More Related Videos
 08:49
Author Spotlight: Unveiling the Potential of VSFG Microscopy in Studying Mesoscopically Heterogeneous Self-Assembled Structures
Published on: December 1, 2023
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
Related Concept Videos
 01:22
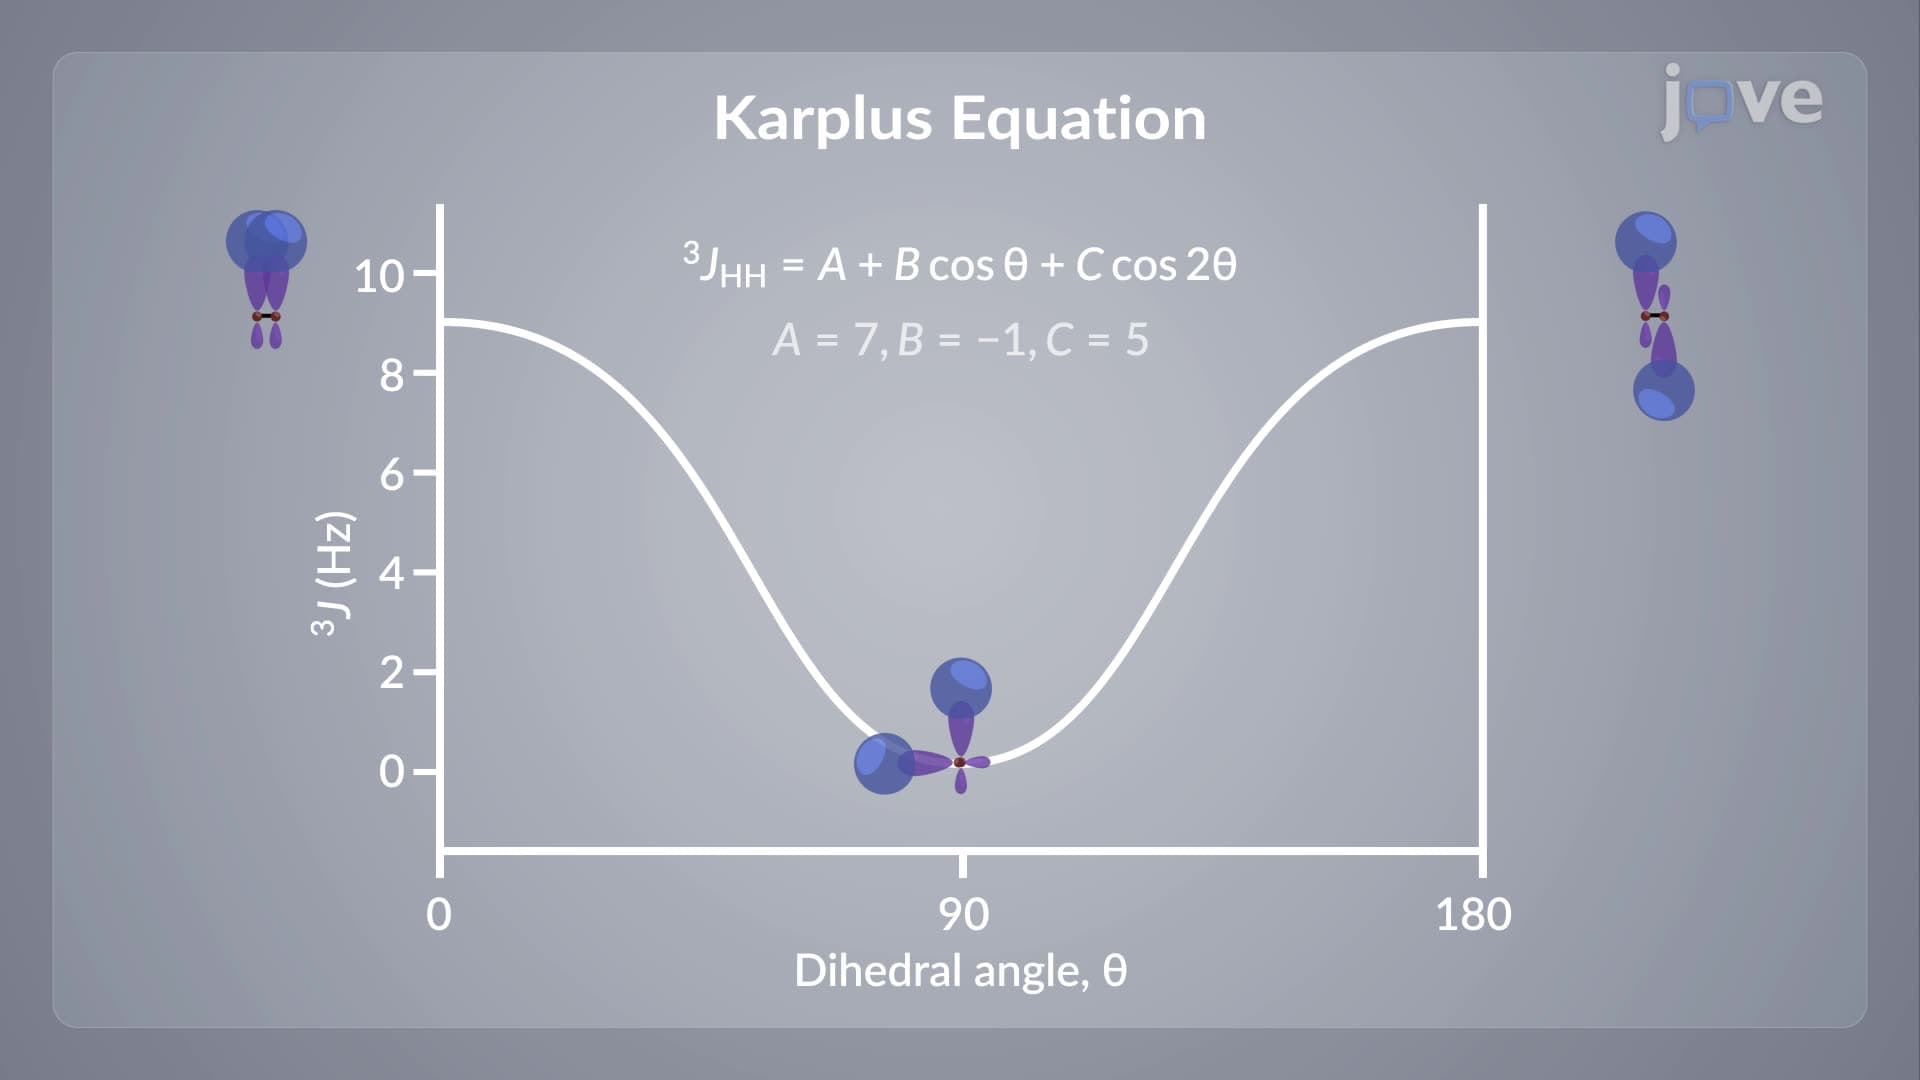
Spin–Spin Coupling: Three-Bond Coupling (Vicinal Coupling)
The extent of coupling depends on the C‑C bond length, the two H‑C‑C angles, any electron-withdrawing substituents, and the dihedral angle between the involved orbitals. The...
 01:14
π Electron Effects on Chemical Shift: Aromatic and Antiaromatic Compounds
 00:52
¹H NMR of Conformationally Flexible Molecules: Temporal Resolution
 01:18
Structure of Benzene: Molecular Orbital Model
 01:27
π Electron Effects on Chemical Shift: Overview
 01:08
Spin–Spin Coupling Constant: Overview
Qualitatively, any spin plus-half nucleus polarizes the spins of its electrons to the minus-half state. Consequently, the paired electron in the hydrogen–carbon bond must...