Related Experiment Video
Updated: Mar 29, 2026


Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Implementation and Optimization of DFT-D/COSab with Respect to Basis Set and Functional: Application to Polar
Roberto Peverati1, Kim K Baldridge1
1University of Zürich, Winterthurerstrasse 190, CH-8057 Zürich, Switzerland.

Abstract:
The implementation, optimization, and performance of DFT-D, including the effects of solvation, has been tested on applications of polar processes in solution, where dispersion and hydrogen bonding is known to be involved. Solvent effects are included using our ab initio continuum solvation strategy, COSab, a conductor-like continuum solvation model, modified for ab initio in the quantum chemistry program GAMESS. Structure and properties are investigated across various functionals to evaluate their ability to properly model dispersion and solvation effects. The commonly used S22 set with accurate interaction energies of organic complexes has been used for parametrization studies of dispersion parameters and relevant solvation parameters. Dunning's correlation consistent basis sets, cc-pVnZ (n = D, T), are used in the optimization, together with the Grimme B97-D exchange-correlation functional. Both water (ε = 78.4) and ether (ε = 4.33) environments are considered. Optimized semiempirical dispersion correction parameters and solvent extent radii are proposed for several functionals. We find that special parametrization of the semiempirical dispersion correction when used together in the DFT-D/COSab approach is not necessary. The global performance is quite acceptable in terms of chemical accuracy and suggests that this approach is a reliable as well as economical method for evaluation of solvent effects in systems with dispersive interactions. The resulting theory is applied to a group of push-pull pyrrole systems to illustrate the effects of donor/acceptor and solvation on their conformational and energetic properties.
More Related Videos
 10:52
Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
Related Concept Videos
 01:20
¹³C NMR: Distortionless Enhancement by Polarization Transfer (DEPT)
 01:18
Frost Circles for Different Conjugated Systems
 03:35
Hybridization of Atomic Orbitals II
 01:29
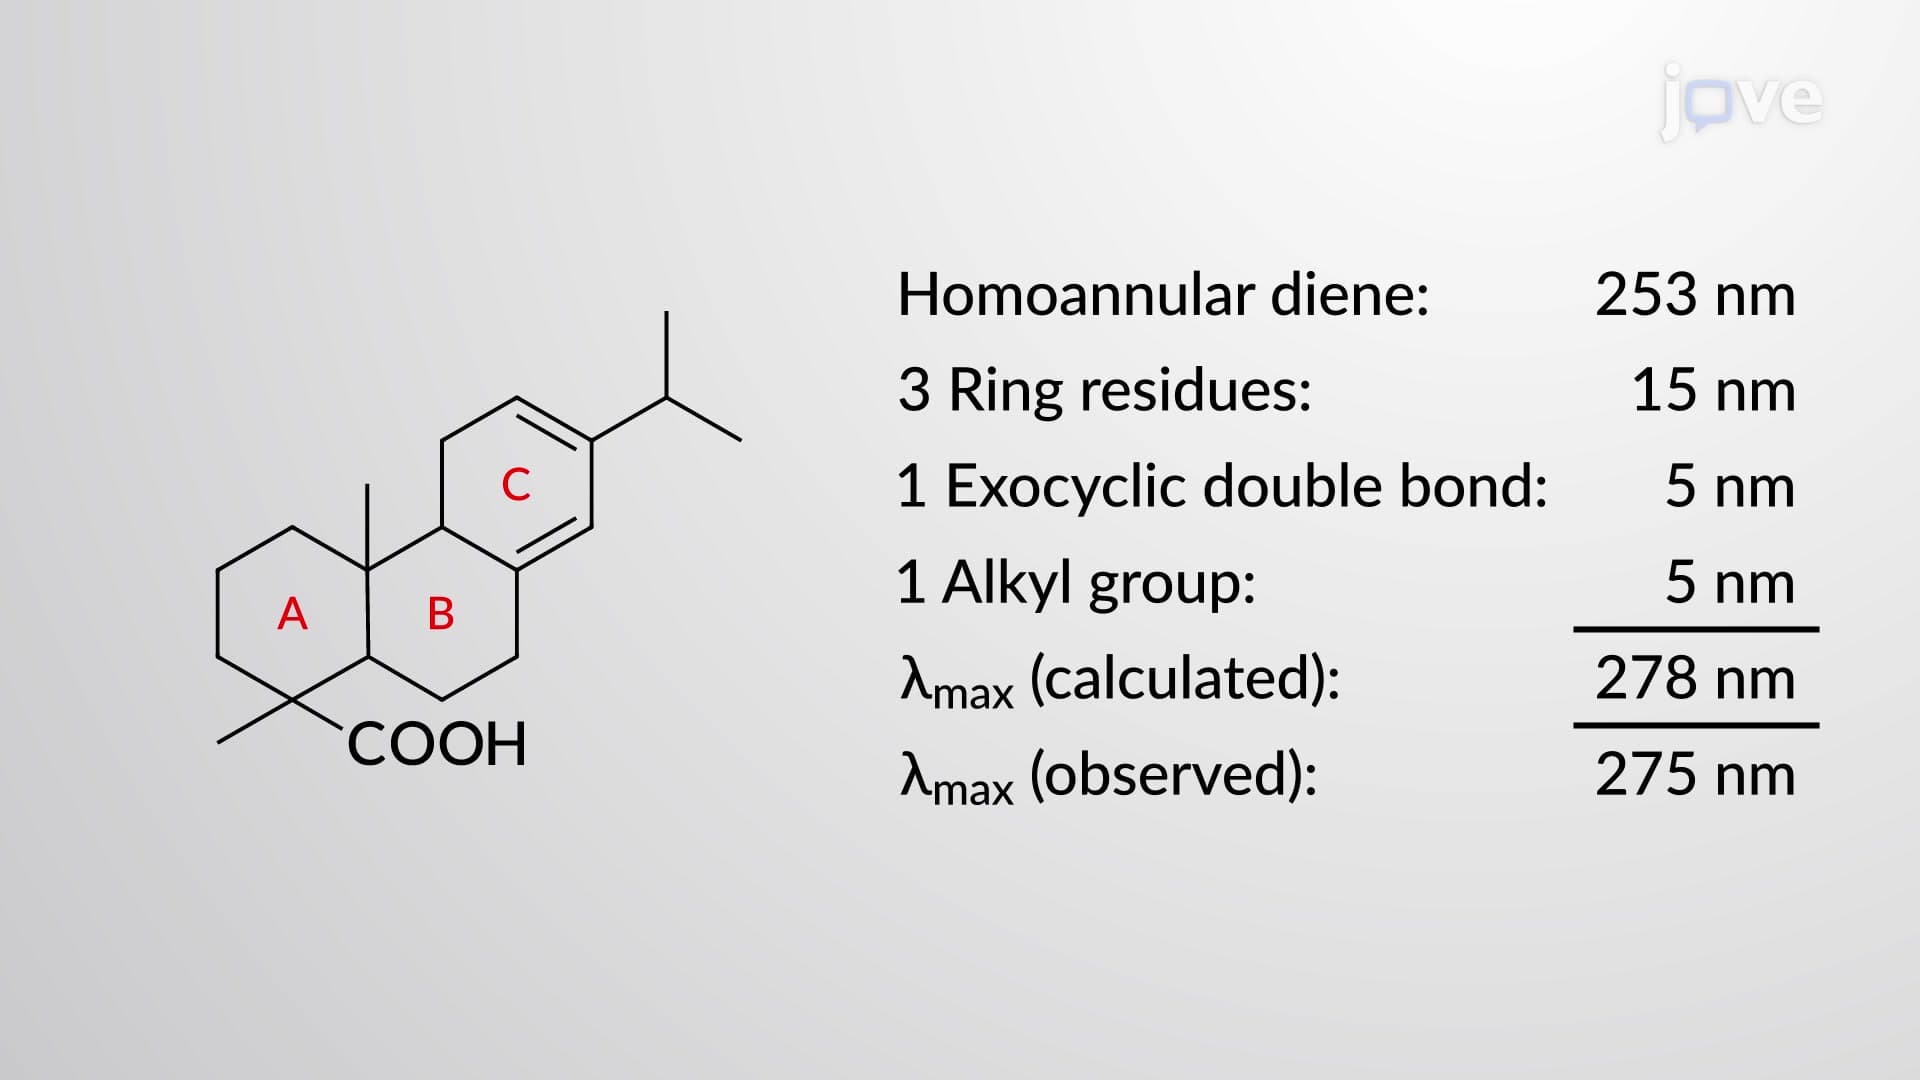
UV–Vis Spectroscopy: Woodward–Fieser Rules
 03:24
Hybridization of Atomic Orbitals I
 01:24
π Molecular Orbitals of 1,3-Butadiene
The simplest conjugated diene is 1,3-butadiene: a four-carbon system where each carbon is sp2-hybridized and has an unhybridized p orbital that contains an unpaired electron. According to molecular orbital theory, atomic orbitals combine to form molecular orbitals such that the number...