Effects of the Aluminum Content of a Zeolite Framework: A DFT/MM Hybrid Approach Based on Cluster Models Embedded in an Elastic Polarizable Environment
Related Experiment Video
Updated: Mar 29, 2026
Facile Preparation of Ultrafine Aluminum Hydroxide Particles with or without Mesoporous MCM-41 in Ambient Environments
Published on: May 11, 2017
Elena A Ivanova Shor1, Alexei M Shor1, Vladimir A Nasluzov1
1Institute of Chemistry and Chemical Technology, Russian Academy of Sciences, 660049 Krasnoyarsk, Russian Federation, Faculty of Chemistry, University of Sofia, 1126 Sofia, Bulgaria, and Department Chemie, Technische Universität München, 85747 Garching, Germany.

Abstract:
We report the first computational study with a sophisticated quantum mechanics/molecular mechanics (QM/MM) technique that addresses the effect of the aluminum content on the properties of acidic zeolites. To account for both electrostatic and mechanical interaction between the QM cluster and its MM environment, we used cluster models embedded in the covalent variant of the elastic polarizable environment (covEPE) [Nasluzov, V. A.; Ivanova, E. A.; Shor, A. M.; Vayssilov, G. N.; Birkenheuer, U.; Rösch, N. J. Phys. Chem. B 2003, 107, 2228]. For the practical application of the covEPE method, it was necessary to develop a new force field for Al containing zeolites. Two types of zeolite materials, FAU and MFI, were employed as examples. We modeled the variation of the Al content both in the MM environment and in the QM cluster, and we studied pertinent properties of bridging OH groups of the zeolite frameworks, OH vibrational frequencies, and deprotonation energies. The computational results suggest that the local structure and the location of the OH groups exert a stronger effect than the variation of the Al content of the framework.
Related Concept Videos
 03:24
Hybridization of Atomic Orbitals I
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 01:18
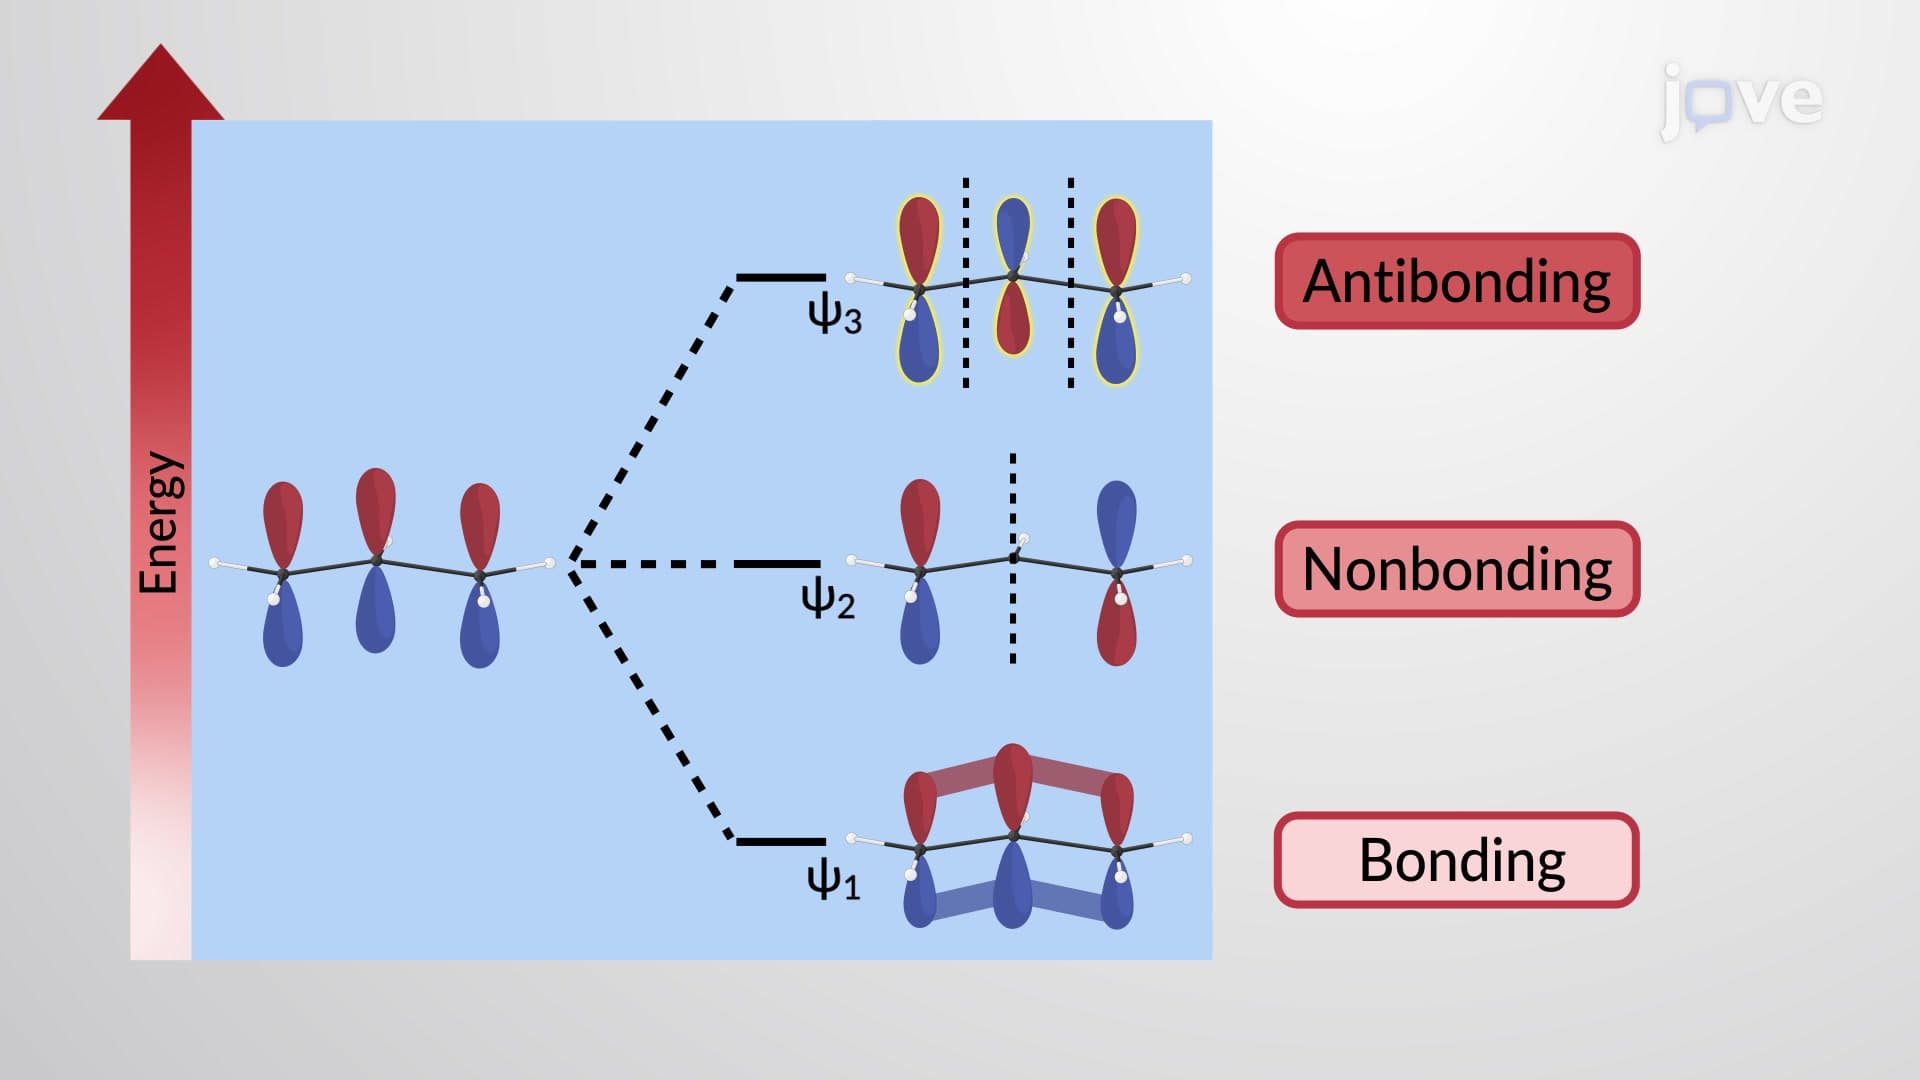
π Molecular Orbitals of the Allyl Cation and Anion
 02:47
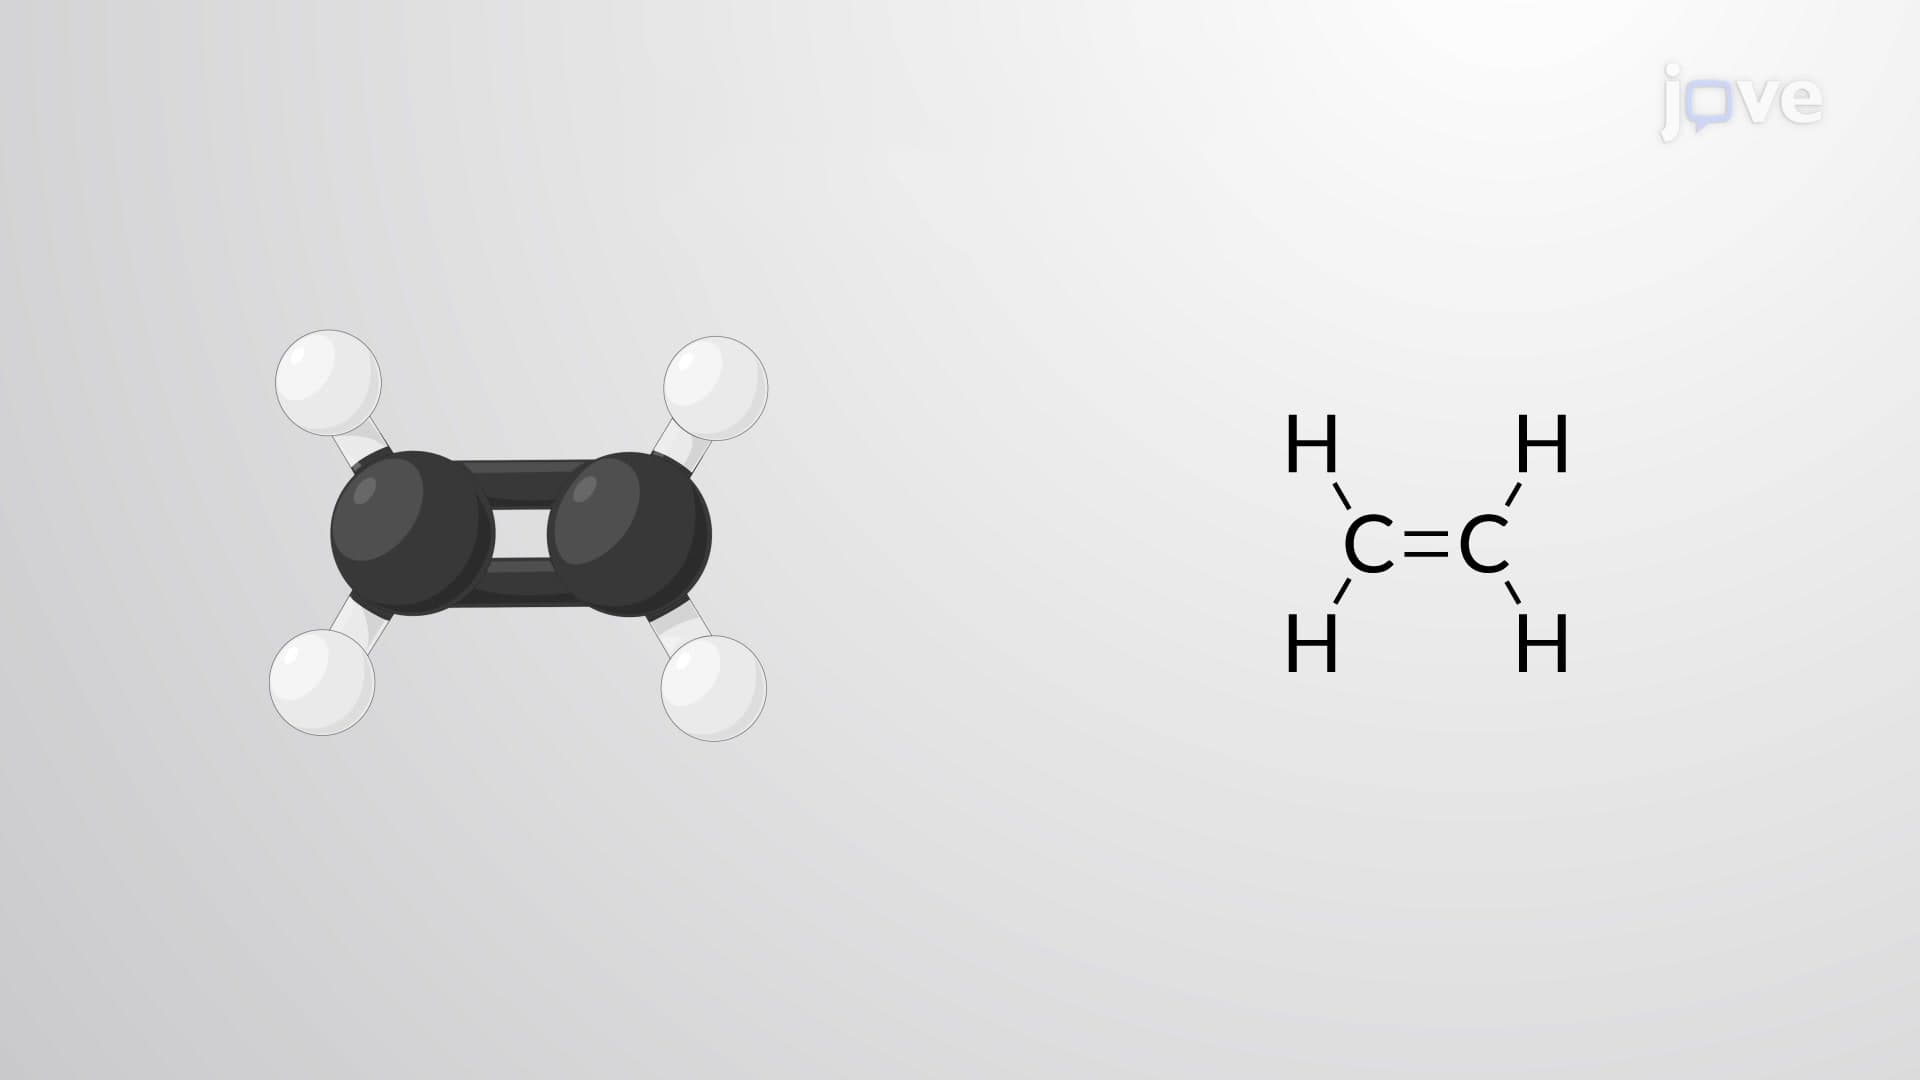
Structure and Bonding of Alkenes
Doubly bonded carbons are sp2 hybridized and have a trigonal planar geometry. The double bond is composed of a σ bond formed by the overlap of hybrid orbitals and a π bond produced by the lateral overlap of unhybridized 2p orbitals on both the carbons. Each carbon atom is...
 01:27
π Electron Effects on Chemical Shift: Overview
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...