Related Experiment Video
Updated: Mar 28, 2026
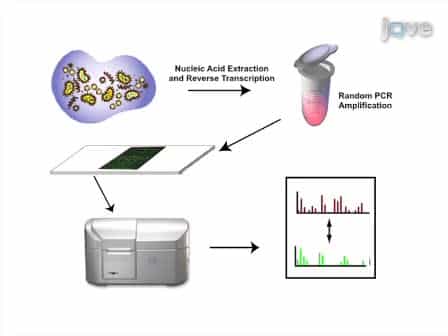
Using a Pan-Viral Microarray Assay Virochip to Screen Clinical Samples for Viral Pathogens
Published on: April 27, 2011
Comparison of microarray-predicted closest genomes to sequencing for poliovirus vaccine strain similarity and
Sebastian Maurer-Stroh1, Charlie W H Lee2, Champa Patel3
1Bioinformatics Institute (BII), A*STAR, 30 Biopolis St, #07-01 Matrix, 138671, Singapore; School of Biological Sciences, Nanyang Technological University (NTU), 60 Nanyang Drive, 637551, Singapore.

Abstract:
We evaluate sequence data from the PathChip high-density hybridization array for epidemiological interpretation of detected pathogens. For influenza A, we derive similar relative outbreak clustering in phylogenetic trees from PathChip-derived compared to classical Sanger-derived sequences. For a positive polio detection, recent infection could be excluded based on vaccine strain similarity.
Insights
PathChip sequencing data provides reliable epidemiological insights for pathogens like influenza A, enabling outbreak analysis comparable to traditional methods. It also aids in distinguishing between wild and vaccine strains for diseases such as polio.
Area of Science:
- Microbiology
- Epidemiology
- Genomics
Background:
- High-density hybridization arrays offer potential for pathogen detection and characterization.
- Epidemiological interpretation of sequence data is crucial for public health surveillance.
- Traditional sequencing methods like Sanger sequencing are well-established but can be resource-intensive.
Purpose of the Study:
- To evaluate the utility of sequence data generated by the PathChip high-density hybridization array for epidemiological interpretation.
- To compare the epidemiological clustering of influenza A outbreaks derived from PathChip data with that from Sanger-derived sequences.
- To assess the ability of PathChip data to differentiate between pathogen strains, using polio as a case study.
Main Methods:
- Utilized sequence data obtained from the PathChip high-density hybridization array.
- Performed phylogenetic analysis on influenza A sequences derived from PathChip and Sanger sequencing.
- Analyzed polio detection data to assess vaccine strain similarity and infer recent infection status.
Main Results:
- PathChip-derived sequences for influenza A demonstrated similar relative outbreak clustering in phylogenetic trees compared to Sanger-derived sequences.
- PathChip data allowed for the exclusion of recent infection in a positive polio detection case due to similarity with vaccine strains.
Conclusions:
- The PathChip high-density hybridization array is a viable tool for generating sequence data applicable to epidemiological interpretation.
- PathChip sequencing provides comparable results to traditional methods for outbreak clustering analysis.
- This array facilitates pathogen strain differentiation, aiding in public health investigations.
Related Concept Videos
 02:54
Evolutionary Relationships through Genome Comparisons
 01:28
Microbial Phylogeny
 00:36
Viral Mutations
 01:29
Modern Molecular Taxonomy